Working With Enthalpy Data in Practice
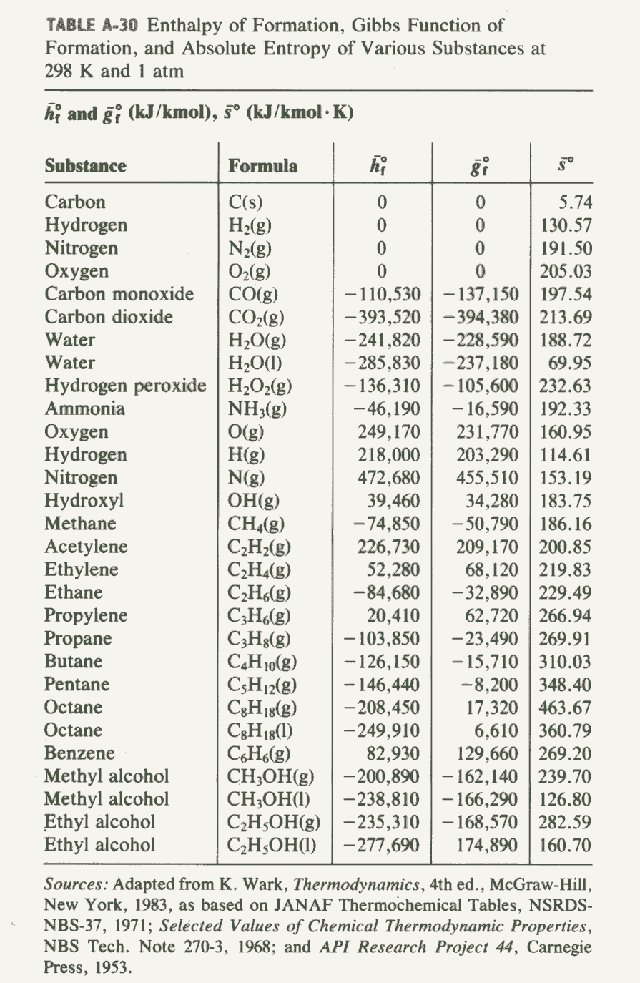
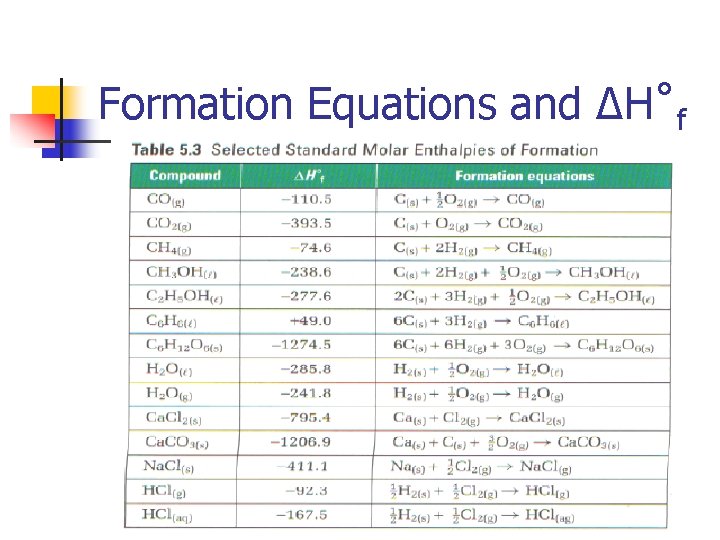
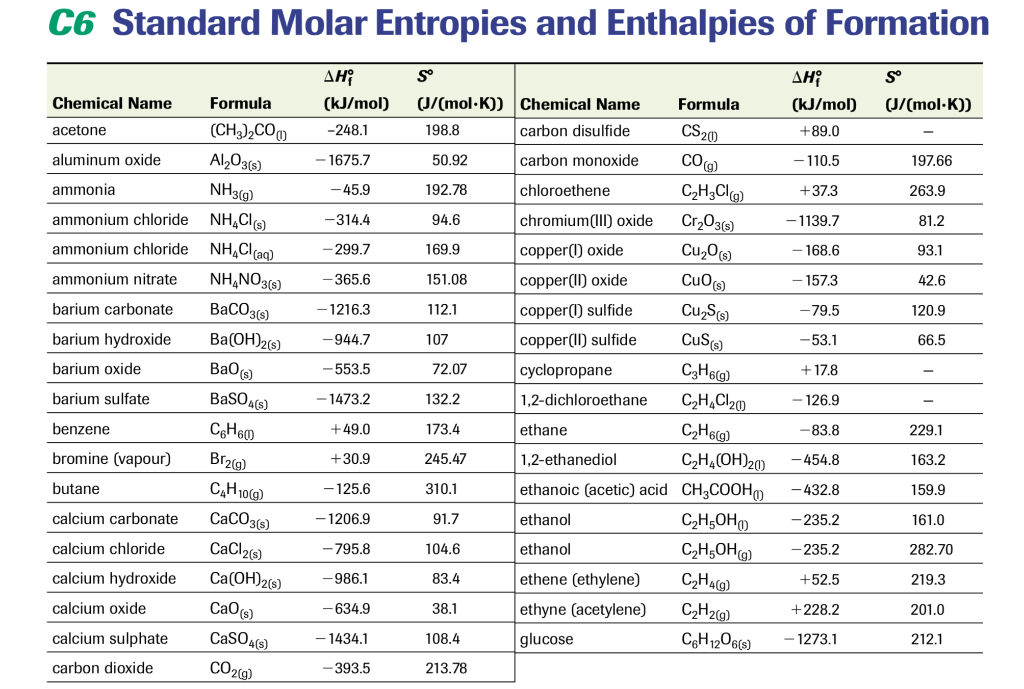
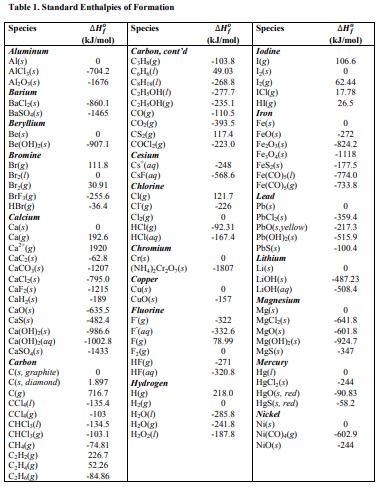
I spend most of my time doing reaction energy balances for industrial processes, and the data I rely on almost always traces back to tabulated Standard Enthalpies Of Formation. The concept itself is straightforward enough—define a reference point where every element in its most stable form at 298 K and 1 bar gets assigned a value of zero, then build everything else relative to that. The problem isn't the definition. It's what happens when you try to apply it to real systems, and the table doesn't cover the conditions you're actually working with. The calculation you'll use most looks like this: add up the formation enthalpies of everything on the product side, subtract the sum for everything on the reactant side, and multiply each value by its stoichiometric coefficient. Simple arithmetic on paper. Messy when you're dealing with streams that contain phases the table doesn't list cleanly, or when temperatures shift away from the standard reference point. That's where the actual work begins.
Standard Enthalpies Of Formation — What You Actually Need to Know
Every entry in a standard thermodynamic table carries a phase label. CO(g) is not the same number as CO(aq). Water vapor gives you 241.8 kJ/mol while liquid water at the same temperature sits at 285.8 kJ/mol. The difference is the enthalpy of vaporization, and skipping it is the single most common mistake I see in reports. Someone will pull a value from a handbook without checking the phase notation, run the balance, and wonder why their energy reconciliation is off by several megajoules per kilomole of fuel. Elements in their standard states carry a Hf° value of exactly zero by definition. O(g), N(g), C(graphite), Br(l), Hg(l). The rest don't. Iron solid, sulfur as S(s), phosphorus as P(s)—these are assigned non-zero values because the standard state of the element isn't the form you're working with. If you're modeling combustion with rhombic sulfur but your table lists an orthorhombic modification, you need the transition enthalpy added in. I ran into this last year on a biomass gasification project where the feedstock sulfur content was measured as elemental S but the process conditions stabilized it as monoclinic sulfur above 95°C. The tabulated difference between those two allotropes is about 0.3 kJ/mol, which looks negligible until you're balancing a reactor that handles tens of thousands of kilograms of sulfur per day. The accumulated error was showing up as a 2.1% discrepancy in the heat recovery calculation, and it took two days of tracing before I caught that the thermodynamic table I was using didn't account for the phase transition enthalpy. Another thing nobody emphasizes enough: Hf° values are strictly defined at 298.15 K and 1 bar. If your reaction runs at 800 K, you can't just plug those numbers into a Hess's law calculation and call it done. You need to bring everything to the operating temperature using heat capacity integrals, or use a thermodynamic table that already provides temperature-dependent entries. Some handbooks include Cp° polynomial coefficients alongside the formation values so you can do the integration yourself. Without those coefficients, you're stuck interpolation or looking up values at discrete temperatures, which introduces its own errors if the Cp curve isn't relatively flat across your range.
The Practical Workflow
Here's how I actually go about it when a new process comes across my desk. First, I write the balanced equation. Not the simplified version with whole numbers that looks nice in a textbook. The actual stoichiometry with every species labeled by phase, including the nitrogen that comes in with the air, the argon, the trace CO in the combustion air, and the water vapor that may or may not condense depending on whether I'm calculating higher or lower heating value. Then I pull the Hf° values from a reliable source. NIST Chemistry WebBook is the default. The DECHEMA tables are worth keeping around for organics that NIST hasn't well. The JANAF thermochemical tables are the gold standard when you need confidence intervals on the data. I keep a spreadsheet with columns for species, phase, stoichiometric coefficient, Hf° at 298 K, and the source reference. The source reference matters because different compilations can disagree by a few kilojoules per mole on borderline compounds, and that disagreement propagates directly into your final answer. I note which table I used so someone else can audit the work later. Once the table is populated, the calculation is mechanical. Product sum minus reactant sum. The sign convention trips people up occasionally—if your result is negative, the reaction is exothermic and heat is released. If positive, heat must be supplied. That's basic, but I've seen it reversed in process simulations because someone defined the system boundary backward. Double-check that the sign convention matches your energy balance equation before you send the numbers anywhere.
Get the Full Details
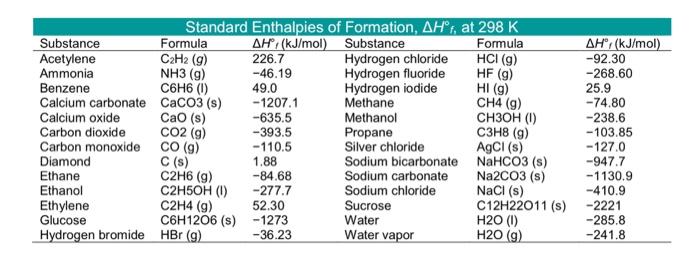
For reactions at temperatures other than 298 K, I adjust each species individually. The formula is H°(T) = Hf°(298) + [Cp°(T) dT from 298 to T]. The integral part is where the Cp polynomial comes in. Most modern handbooks give Cp° as a fourth-order polynomial in T, something like Cp° = a + bT + cT² + dT³. You integrate term by term, evaluate at your target temperature, and add the result to the standard formation enthalpy. Do this for every species, then run the product-minus-reactant calculation on the temperature-adjusted values. The total is your reaction enthalpy at the actual operating temperature.
Where the Method Breaks Down
Standard enthalpies of formation don't help you when the compound you're interested in doesn't have a tabulated value. This happens more often than you'd think with intermediate species in complex reaction mechanisms—radicals, transition states, unstable organic fragments. The NIST tables are comprehensive for common industrial chemicals, but they're not complete. When you hit a gap, you have three realistic options: estimate the value using group contribution methods like the ones in theJoback or Benson approaches, run a quantum chemistry calculation to get a computational estimate, or measure it experimentally. Each option has tradeoffs. Group contributions can be off by 10 to 20 kJ/mol for poorly characterized functional groups. Quantum calculations at the G4 level typically land within 5 kJ/mol for well-behaved organic molecules but can diverge significantly for transition metal complexes or species with significant multireference character. Calorimetry is the most accurate route but requires access to equipment and pure samples, which you often don't have in an early-stage process evaluation. Another hard limit: the standard state definition assumes ideal behavior at 1 bar. At high pressures, fugacity corrections become necessary, and the simple Hf° table loses relevance. Supercritical fluids, dense phase CO streams, high-pressure hydrogen systems—these require equation-of-state-based property packages, not a lookup table. I've watched junior engineers try to force a standard enthalpy approach onto a 50-bar ammonia synthesis loop and end up with energy balances that were off by 15% because they ignored the pressure dependence of enthalpy entirely. There's also the matter of solutions. Aqueous standard enthalpies of formation exist for many ions, but they're referenced to H(aq) = 0 by convention, which means you're working with a relative scale that has no absolute zero. Mixing two electrolyte solutions at non-dilute concentrations introduces activity coefficient effects that the standard tabulated values simply don't capture. If your process involves concentrated brines or molten salts, you need Pitzer models or similar frameworks, not a formation enthalpy table.
A Quick Worked Example
Let's run through methane combustion, because it's the simplest case and it exposes the phase issue I mentioned earlier. The balanced equation with standard states is CH(g) + 2O(g) CO(g) + 2HO(l). Using NIST values: Hf°(CH) = 74.8 kJ/mol, Hf°(CO) = 393.5 kJ/mol, Hf°(HO, liquid) = 285.8 kJ/mol, and Hf°(O) = 0 by definition. The calculation is [393.5 + 2(285.8)] [74.8 + 0], which gives 890.3 kJ per mole of methane. That's the higher heating value because the water is liquid. If the water stays as vapor, you use Hf°(HO, gas) = 241.8 instead, and the result is 802.3 kJ/mol. The difference, 87.6 kJ/mol, is exactly the latent heat of vaporization for two moles of water. Every time you write a combustion balance, you have to decide which heating value applies to your system. Condensing the water downstream recovers that extra energy, but only if your heat exchanger operates below the dew point. If it doesn't, you're leaving money on the table or misreporting your efficiency. That's the practical reality of working with Standard Enthalpies Of Formation. The tables give you a solid starting point, but the work is in the details—phase labels, temperature corrections, source verification, and knowing when the method stops being useful and you need a different approach.