Why Your Solutions Aren't Trustworthy Without This Step
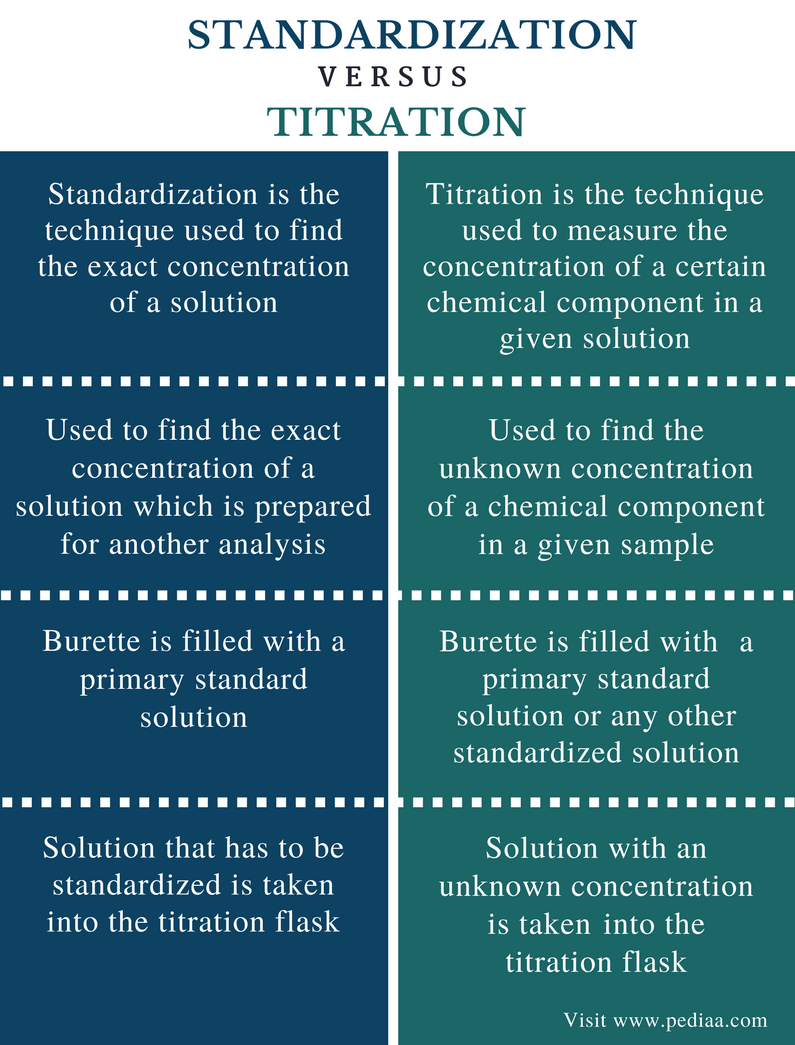
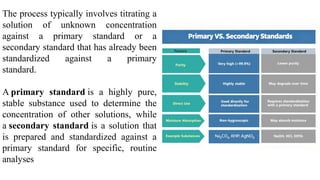
You've probably been told in lab that you can just weigh something out and dissolve it to get a known concentration. That's true for primary standards. But most reagents you grab off the shelf? They're unreliable. Sodium hydroxide absorbs water from the air. Hydrochloric acid is a gas dissolved in water, and the concentration drifts with temperature. So before you use any of these for actual work, you need to standardize them. An imprecise mass of a reagent that cannot serve as its own primary standard. You make up an approximate concentration first, then determine the exact value by titration against a known primary standard. The procedure is straightforward on paper, but the devil is in the execution. Here's how I do it. I start by making the approximate solution. Say I need 0.1 M NaOH. I calculate the mass needed for about 500 mL, weigh it on a balance that reads to 0.01 g, and dissolve it in freshly boiled and cooled distilled water. The boiling drives out dissolved CO2, which otherwise reacts with NaOH to form carbonate and throws off your results over time. I store it in a plastic bottle with a soda lime trap. Glass is fine for short-term work, but CO2 diffuses through glass over days.
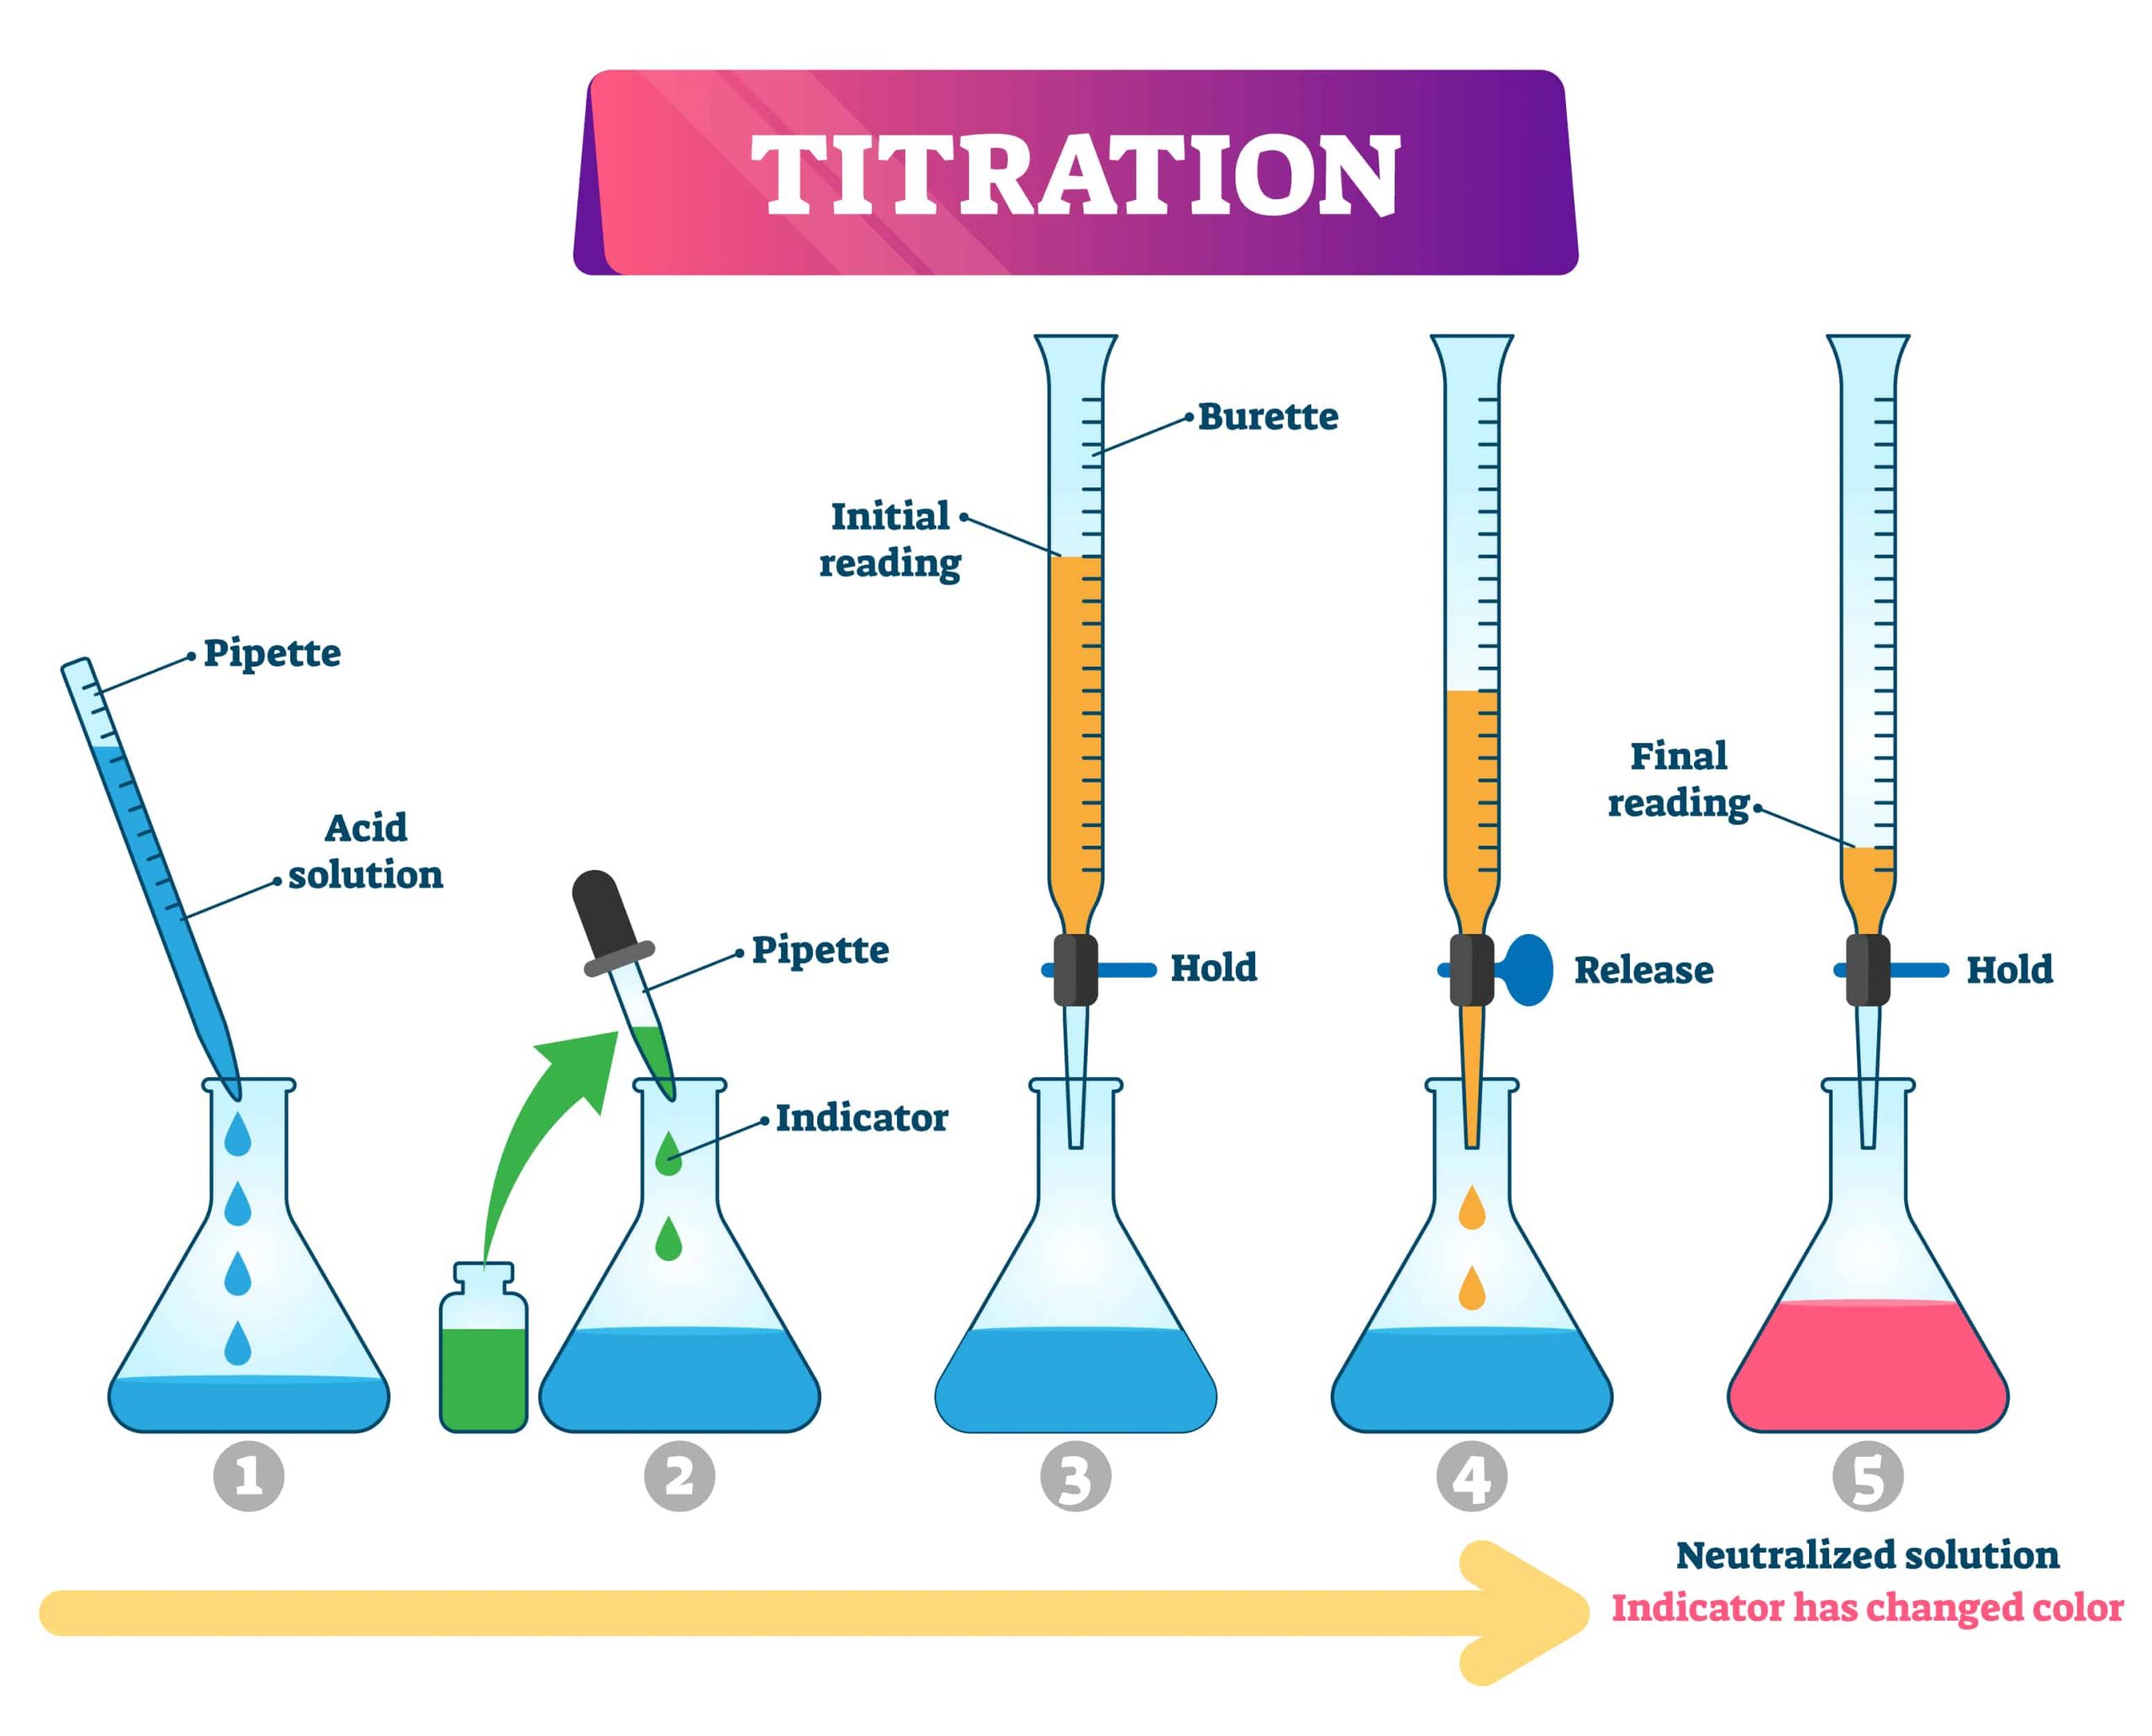
Next comes the titration part. I pick a primary standard appropriate for the solution I'm standardizing. For bases like NaOH, potassium hydrogen phthalate (KHP) is the go-to. It's stable, non-hygrosopic, and has a decent molar mass so weighing errors stay small. For acids like HCl, I might use borax or anhydrous sodium carbonate. The choice matters more than people admit. Sodium carbonate needs to be ignited at 270°C for two hours before use, or the moisture content varies between batches. I keep a bottle of pre-ignited carbonate in a desiccator and pull from it fresh each week. Old stock absorbs moisture fast once you open it. The actual titration requires a burette that's properly conditioned. Rinse it with the titrant solution at least three times before filling. Air bubbles in the tip are a common source of error that beginners miss. Tap the burette gently while you fill it, and run some solution through quickly to clear the tip. If you skip this, your first few readings will be off by a few tenths of a milliliter, which ruins precision at the 0.1 M level. I typically weigh out about 0.4 to 0.5 g of KHP into each of three Erlenmeyer flasks, dissolve in about 50 mL of distilled water with a couple drops of phenolphthalein, and titrate. The endpoint should be a faint pink that persists for thirty seconds. Not a deep magenta. If you overshoot, you're starting over. I always run a quick rough titration first just to get the approximate volume, then do the precise ones knowing roughly where the endpoint sits.
The math is simple. Divide moles of primary standard by liters of titrant used. Average the three trials and report to four significant figures if your balance and burette support it. In practice, you're usually looking at three reliable figures for 0.1 M solutions unless you're working in a controlled environment. One thing that catches people out: temperature. Titrant volumes expand and contract. The calibration of your volumetric glassware assumes 20°C. If your lab runs at 25°C, your actual volume is about 0.1% higher than what the meniscus reads. For high-precision work, apply a correction factor. For routine analysis, it usually doesn't matter unless you're chasing tight tolerances. Here's a specific problem I ran into that took me a day to figure out. I was standardizing a dilute perchloric acid solution in acetic anhydride for non-aqueous titration. The endpoints were inconsistent, drifting by several milliliters between trials even though I was using the same amount of sample. Turns out the glacial acetic acid I was using as solvent contained enough water to make the whole system unreliable. The perchloric acid was reacting with the water instead of doing what I wanted. I switched to freshly distilled acetic anhydride-grade acetic acid and the variability dropped to under 0.3%. It cost extra and required distillation, but it was the only fix that worked.
Get the Full Details

Another counter-intuitive point: not all primary standards are created equal, even within the same category. Some lots of KHP have been recrystallized from water and may retain trace moisture if not dried properly. Others are synthesized differently and contain different impurity profiles. Always check the certificate of analysis. If the vendor says dry at 110°C for two hours, do it. Don't assume the bottle sitting on the shelf has been treated correctly. There are cases where standardization is simply not practical. If you need a solution that's unstable on the timescale of your work—like iodine solutions, which slowly oxidize water and decompose under light—you standardize immediately before use and discard the rest. Iodine is often standardized against arsenic trioxide, which is toxic and requires careful handling, or against sodium thiosulfate that itself was standardized against potassium dichromate. It's a chain of dependencies, and each link adds uncertainty. For most teaching labs and routine quality control work, the approach I described covers it. Make the solution approximately, standardize against an appropriate primary standard using proper technique, and record everything. If your results show more than 0.5% relative standard deviation across your triplicate trials, something is wrong with your technique or your materials. Recheck your glassware, your drying procedure, and your endpoint detection before recalculating.
The bottom line is that standardization is not optional for anything that isn't a certified primary standard. The process exists because real-world reagents are messy, and your analytical results are only as good as the concentration you assign to your titrant.