What You Need to Know Before You Start Mixing These Up
I spent three hours once trying to figure out why my NMR spectrum didn't match the literature value for a compound I'd synthesized. The structure was right. The connectivity was right. But it was the wrong stereoisomer. That kind of mistake costs you more than time—it costs you credibility when you're publishing or pitching a molecule to someone who knows what they're looking at. So let's get this straight once and for all. The difference between a constitutional isomer and a stereoisomer isn't just academic. It changes how you isolate, characterize, and handle a compound. Getting it wrong early saves you from doing the same work twice later.
Stereoisomer Vs Constitutional Isomer
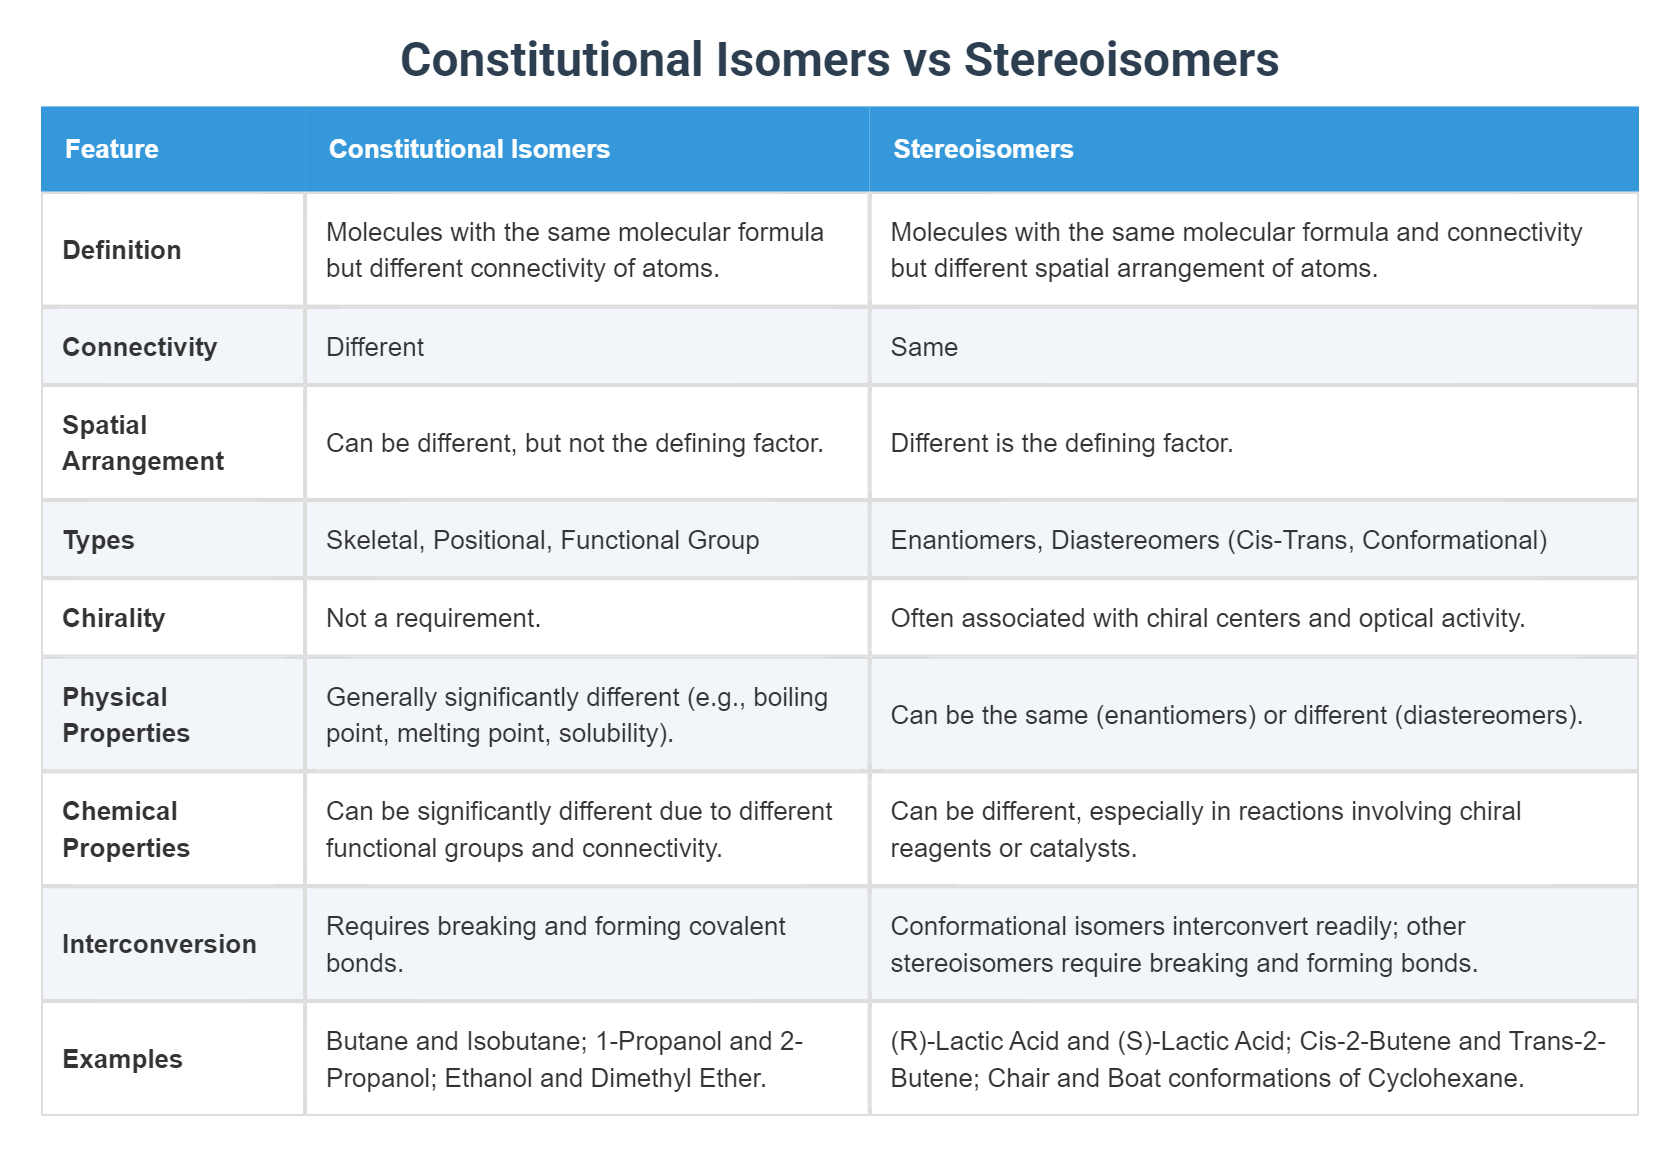
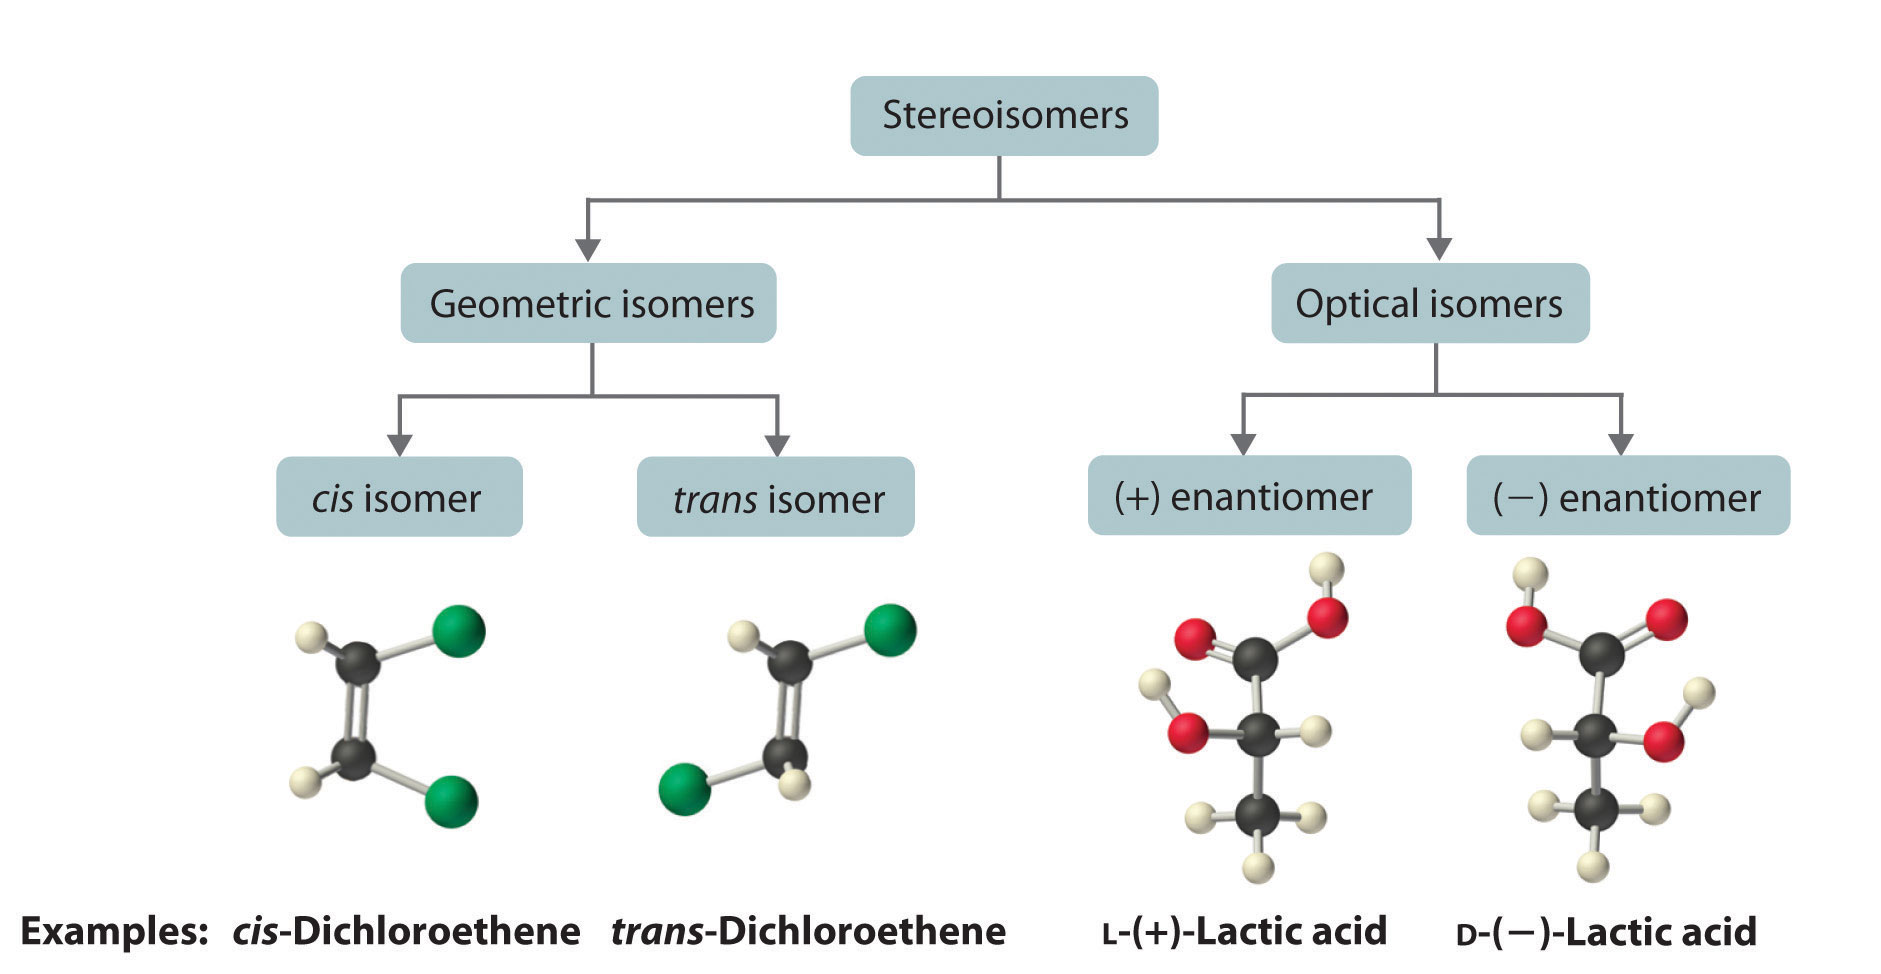
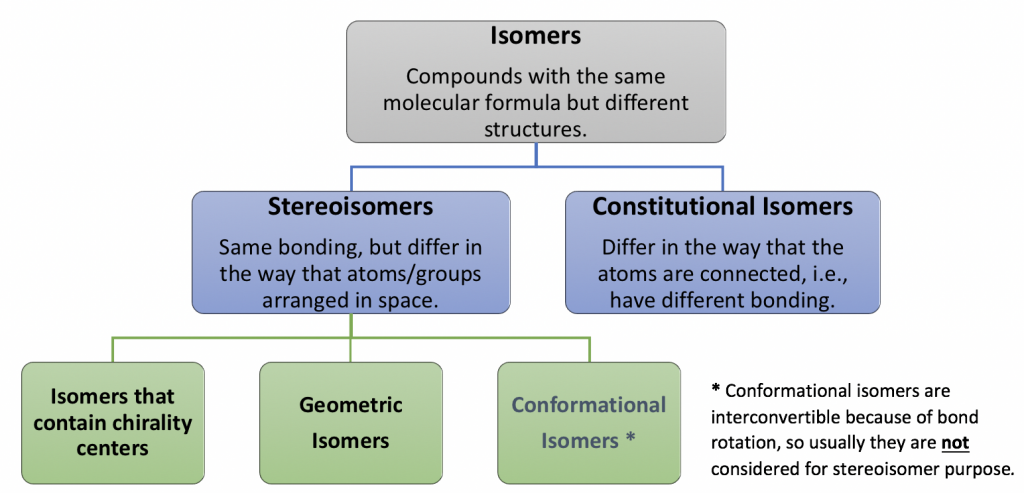
A constitutional isomer—sometimes called a structural isomer—means the atoms are connected in a different order. Same molecular formula, different bonding sequence. Take C4H10O. You could have butanol with the OH on the end carbon, or you could have sec-butanol with the OH on the second carbon. Or diethyl ether, where the oxygen sits between two ethyl groups instead of capping a chain. These are completely different molecules with different boiling points, different reactivity, different everything. You can usually separate them by standard distillation or column chromatography because their physical properties diverge significantly. A stereoisomer means the atoms are connected in the exact same order but arranged differently in three-dimensional space. Same bond connectivity. Different spatial orientation. The two most common types are enantiomers and diastereomers. Enantiomers are mirror images that are non-superimposable—like your left and right hands. Diastereomers are stereoisomers that aren't mirror images of each other, which includes cis-trans isomers and compounds with multiple chiral centers where some but not all centers differ. Here's where people mess up: you can't separate enantiomers by regular column chromatography on silica. They have identical physical properties in an achiral environment—same boiling point, same Rf value, same solubility in non-chiral solvents. You need a chiral stationary phase or you need to derivatize them into diastereomers first using a chiral resolving agent. I learned this the hard way when I tried to purify a racemic mixture through a standard flash column and got what I thought was a pure product, only to find later by chiral HPLC that it was still a 50-50 mix. Wasted a week of downstream work on a compound I thought I'd isolated cleanly.
Diastereomers, on the other hand, you can often separate by normal methods. They have different physical properties because they're not mirror images. Cis and trans alkenes are a classic example. The cis isomer might boil at 115°C while the trans boils at 121°C. That's enough for a good fractional distillation or a straightforward silica column to resolve them. When you're working with a new compound, the first thing you should always determine is whether you're dealing with constitutional isomers or stereoisomers. Run your HRMS to confirm the molecular formula. Then look at your NMR. For constitutional isomers, you'll see distinctly different chemical shifts and coupling patterns because the electronic environment around each proton is fundamentally different. For stereoisomers, the NMR differences are subtler. Enantiomers will give you identical 1H and 13C NMR spectra in an achiral solvent. Diastereomers will show slight but measurable differences in chemical shift, often in the 0.01 to 0.1 ppm range. X-ray crystallography is the gold standard for determining stereochemistry when you can get a good crystal. It tells you absolutely where every atom is in three dimensions. But not every compound crystallizes well, and preparing a sample for X-ray can take days or weeks depending on your molecule. I've had compounds that refused to form crystals no matter what solvent system I tried—methanol, ethanol, ethyl acetate, hexanes, combinations of all of them. In those cases, I fall back to NOE experiments in NMR or circular dichroism spectroscopy, though those methods come with their own interpretation challenges.
Get the Full Details

One thing most beginners miss is that constitutional isomerism and stereoisomerism aren't mutually exclusive categories in the way textbooks sometimes imply. A single molecular formula can give you both types simultaneously. C5H10, for example. You could have pent-1-ene and cyclopentane as constitutional isomers—completely different connectivities. But you could also have cis-pent-2-ene and trans-pent-2-ene as stereoisomers within the pent-2-ene constitutional isomer. So when you're analyzing an unknown, you need to establish the connectivity first before you start worrying about stereochemistry. Jumping straight to chirality without confirming the bond framework is a common error that leads to misidentification. Another thing worth noting: the number of possible stereoisomers grows exponentially with the number of chiral centers. One center gives you two enantiomers. Two centers give you up to four stereoisomers—two pairs of enantiomers, plus diastereomeric relationships between the pairs. Three centers and you're looking at up to eight. This matters practically because each additional chiral center roughly doubles the complexity of your purification problem if you're running a non-selective reaction. I've seen people synthesize molecules with three or four stereocenters and then wonder why their crude product was an unresolvable mess on the column. The yield of any single stereoisomer drops to 12.5% or lower, and separating them becomes a nightmare unless you're using chiral HPLC or crystallization-based resolution. If you need a practical workflow, start here. Run elemental analysis or high-resolution mass spectrometry to nail down the molecular formula. Then do a full NMR assignment—1H, 13C, and if you have a modern spectrometer, HSQC and HMBC to confirm connectivity. If the connectivity matches your target structure, you're dealing with either stereoisomers or you've made a constitutional isomer. Check your coupling constants. Large J values around 15 Hz for vinyl protons usually indicate trans geometry. Values around 10 Hz suggest cis. For cyclohexane systems, axial-axial couplings tend to be 8 to 12 Hz while axial-equatorial and equatorial-equatorial are smaller, around 2 to 5 Hz. These numbers won't tell you everything, but they'll narrow things down significantly before you invest time in more expensive characterization.
For stereochemical determination specifically, optical rotation gives you a quick screen. If your compound is optically active and you know the literature value for the expected enantiomer, a matching rotation is strong evidence you have the right one. But optical rotation alone is never conclusive—concentration, solvent, and temperature all affect the reading, and two different compounds can have similar specific rotations by coincidence. Pair it with polarimetry and chiral HPLC whenever possible, and you'll have a much more reliable answer. The bottom line is that constitutional isomers are fundamentally different molecules that you can usually separate and characterize with standard techniques. Stereoisomers share connectivity but differ in 3D arrangement, and separating them requires either chiral methods or clever use of diastereomer formation. Both types show up constantly in synthesis labs, and neither one cares how confident you feel about your reaction conditions. Your job is to prove which one you actually made before you move forward.