Let's talk about how Technology Transfer In Pharmaceutical Industry actually works when you strip away the regulatory brochures.
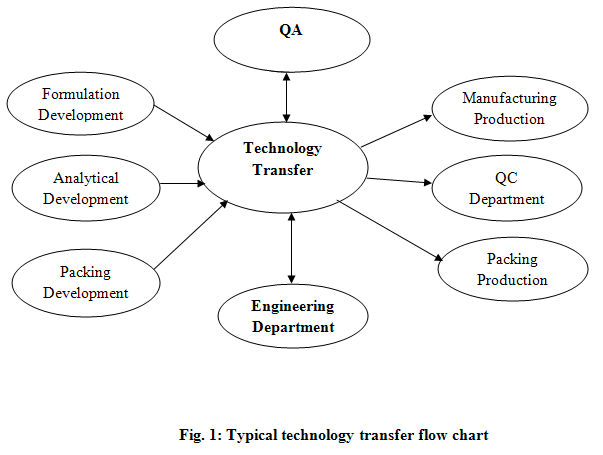
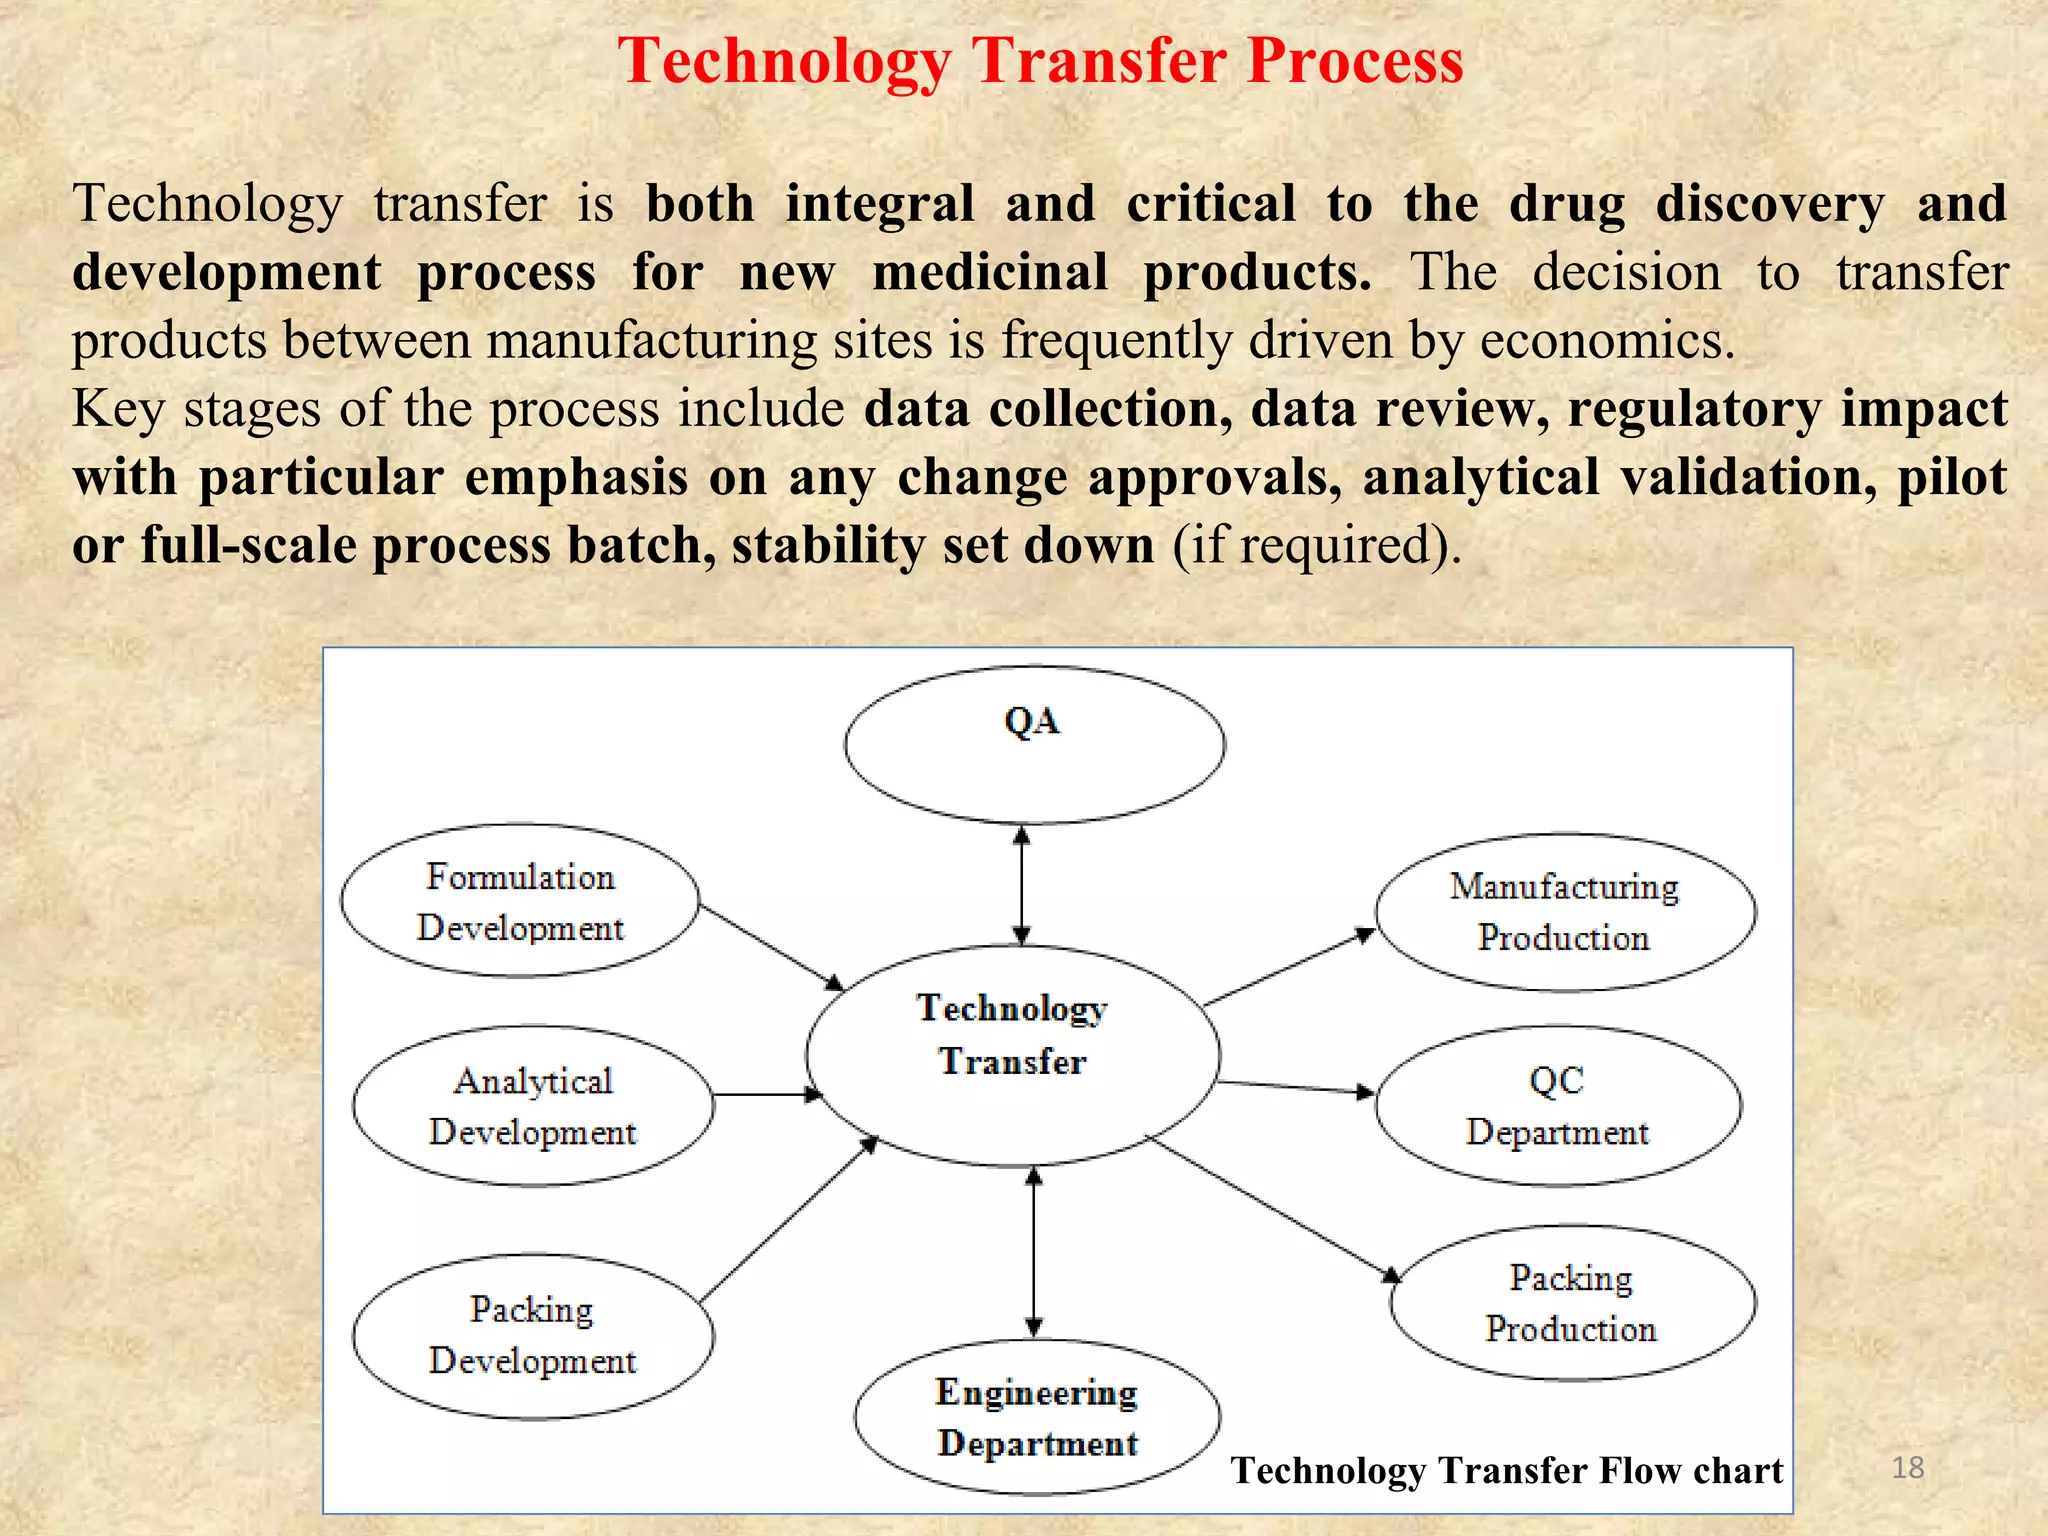
Most people treat tech transfer like a checklist exercise. They queue up protocol documents, fill in signatures, and move on. That approach works fine for simple formulation transfers between same-site lines. It falls apart quickly when you're dealing with complex biologics or novel delivery systems. I have learned this through repeated failures rather than any single dramatic incident. At its core, you are moving product knowledge from one controlled environment to another. That knowledge exists in multiple forms. Some of it is documented in standard operating procedures and batch records. Most of it lives in the heads of the people who built the process. The gap between those two repositories is where things go wrong. You need to capture tacit knowledge before people move on to other projects. A typical transfer involves three structural components. The source site possesses a validated or development-stage process. The receiving site has its own equipment, materials, and personnel. Your job is to demonstrate that the receiving site can reproduce the product to the same quality standards using its own resources.
How the process actually unfolds
Start with a transfer readiness assessment. This is not a perfunctory formality. You need to evaluate whether the receiving site has the appropriate equipment capabilities, analytical instruments, and raw material supply chains to handle the product. I once skipped this step on a semi-scaled bioprocess transfer and spent six weeks chasing a contamination issue that traced back to a HVAC system designed for a different product class. The assessment phase normally takes two to four weeks for straightforward chemical processes and up to twelve weeks for complex sterile biologics. After readiness comes the knowledge transfer phase. This is where most programs stall. You need documented protocols, but documentation alone does not transfer expertise. You need hands-on training sessions, video recordings of critical operations, and structured knowledge exchange meetings. The receiving team should operate the process under supervision before they ever attempt an independent run. Budget at least two weeks of on-site training for complex processes. The execution phase involves conducting pilot batches at the receiving site. You compare analytical results, yield data, and quality attributes against predetermined acceptance criteria. These criteria must be established before any batch is manufactured. Setting criteria after seeing the data is a regulatory red flag and a common pitfall that triggers inspection observations.
Documentation and closure round out the process. You compile a transfer report summarizing all activities, deviations, and outcomes. Regulatory filings may require updates depending on the scale change and jurisdiction.
Get the Full Details

A specific problem I dealt with
During a Technology Transfer In Pharmaceutical Industry project for an oral solid dose formulation, the receiving site reported inconsistent dissolution profiles across three consecutive pilot batches. The source site could not reproduce the issue because their mixing equipment and granulation parameters were essentially different. The problem was not the formulation itself. It was the shear intensity during the high-shear granulation step. The receiving site's mixer had a different impeller geometry that generated higher localized shear rates at equivalent power settings. My workaround was to characterize the mixing energy input directly. I used torque measurements and power consumption data from the receiving site's mixer alongside computational fluid dynamics modeling to map the shear distribution. We then adjusted the binder addition rate and granulation time rather than trying to match the source site's exact parameters. The revised approach brought dissolution profiles within specification on the fourth batch. This kind of root cause analysis typically adds four to eight weeks to a transfer timeline but prevents costly re-validation cycles later.
Counter-intuitive insights beginners miss
People assume that more data always leads to a smoother transfer. In practice, gathering excessive baseline data from the source site without a clear purpose creates analysis paralysis. You need targeted data that addresses specific risk areas identified during your risk assessment. A focused data collection plan covering critical quality attributes and process parameters is more useful than a comprehensive dataset that nobody knows how to interpret. Another misconception is that transfer success means the receiving site produces identical results. It does not. Acceptable transfer means producing product within specification using the receiving site's own process. Minor parameter variations are expected and often necessary. The goal is quality equivalence, not numerical identity. Regulatory agencies understand this distinction when it is properly documented and justified.
Common pitfalls and why they happen
The biggest failure mode is treating the transfer as a documentation exercise rather than a process adaptation exercise. When teams focus on completing forms instead of understanding process behavior, they miss subtle differences between sites that affect product quality. This pitfall is especially common when the source site has limited personnel available for training due to competing priorities. A second pitfall involves inadequate acceptance criteria. Criteria that are too broad allow a transfer to pass on paper while hiding quality issues that emerge during commercial production. Criteria that are too narrow create unnecessary reworks and delays. The sweet spot requires statistical justification based on historical source site performance and a clear understanding of the process design space. Here is a blunt observation: this process does not work well when the source and receiving sites are in different regulatory jurisdictions with divergent expectations. The additional burden of dual regulatory submissions and potential additional clinical studies can extend timelines by six to eighteen months depending on the complexity. In these cases, consider whether a license agreement with the original sponsor might be more efficient than a full transfer.

Tools and frameworks that actually help
The Quality by Design framework from ICH Q8 provides a structured approach to understanding your process. It forces you to identify critical material attributes and critical process parameters before you begin the transfer. Skipping this step and relying on trial-and-error optimization during the transfer phase is a reliable way to waste time and budget. Process analytical technology tools like near-infrared spectroscopy and inline particle size analysis can reduce the need for offline testing and provide real-time process understanding. These tools require upfront investment and method validation but typically pay for themselves within two or three successful transfers by reducing batch failure rates and shortening release timelines. Standard reference materials and certified reference standards are essential for ensuring comparability across sites. Without them, analytical method differences between laboratories can masquerade as process performance issues. Budget for SRM procurement early in the planning phase because lead times can reach twelve weeks for specialized standards.
What to do when transfer fails
Sometimes the process simply cannot be replicated at the target site with acceptable quality. This outcome is rare for well-understood conventional processes but more common with novel modalities like gene therapies or complex generics. When this happens, you have two realistic options. You can modify the process to fit the receiving site's capabilities, which may require regulatory filing changes. Or you can abandon the transfer and seek alternative manufacturing arrangements. Neither option is ideal, but proceeding with a compromised process without proper justification is worse for patient safety and regulatory compliance. The entire Technology Transfer In Pharmaceutical Industry process typically runs from six months to two years depending on product complexity, site readiness, and regulatory requirements. The variance is large enough that any timeline prediction should include explicit contingency buffers for the unknown unknowns that inevitably appear during execution.