Understanding Thermo When You Actually Need It
Thermodynamics in chemistry deals with energy transfer during physical and chemical changes. It is the framework that tells you whether a reaction will actually proceed, how much heat you need to apply, and where your system will settle before anything explodes. People tend to memorize equations for it. That usually does not help when you are standing at a fume hood trying to figure out why your yield dropped from 82% to 34% after a simple scale-up. I spent most of my early career doing process optimization, and thermodynamics was always the invisible bottleneck. You could balance every equation perfectly and still end up with a reactor that stalled halfway through because you ignored the temperature dependence of Gibbs free energy. The branch of chemistry that studies changes is called thermodynamics, but it only becomes useful when you treat it as a prediction tool rather than a worksheet exercise.
The Branch Of Chemistry That Studies Changes Is Called Thermodynamics
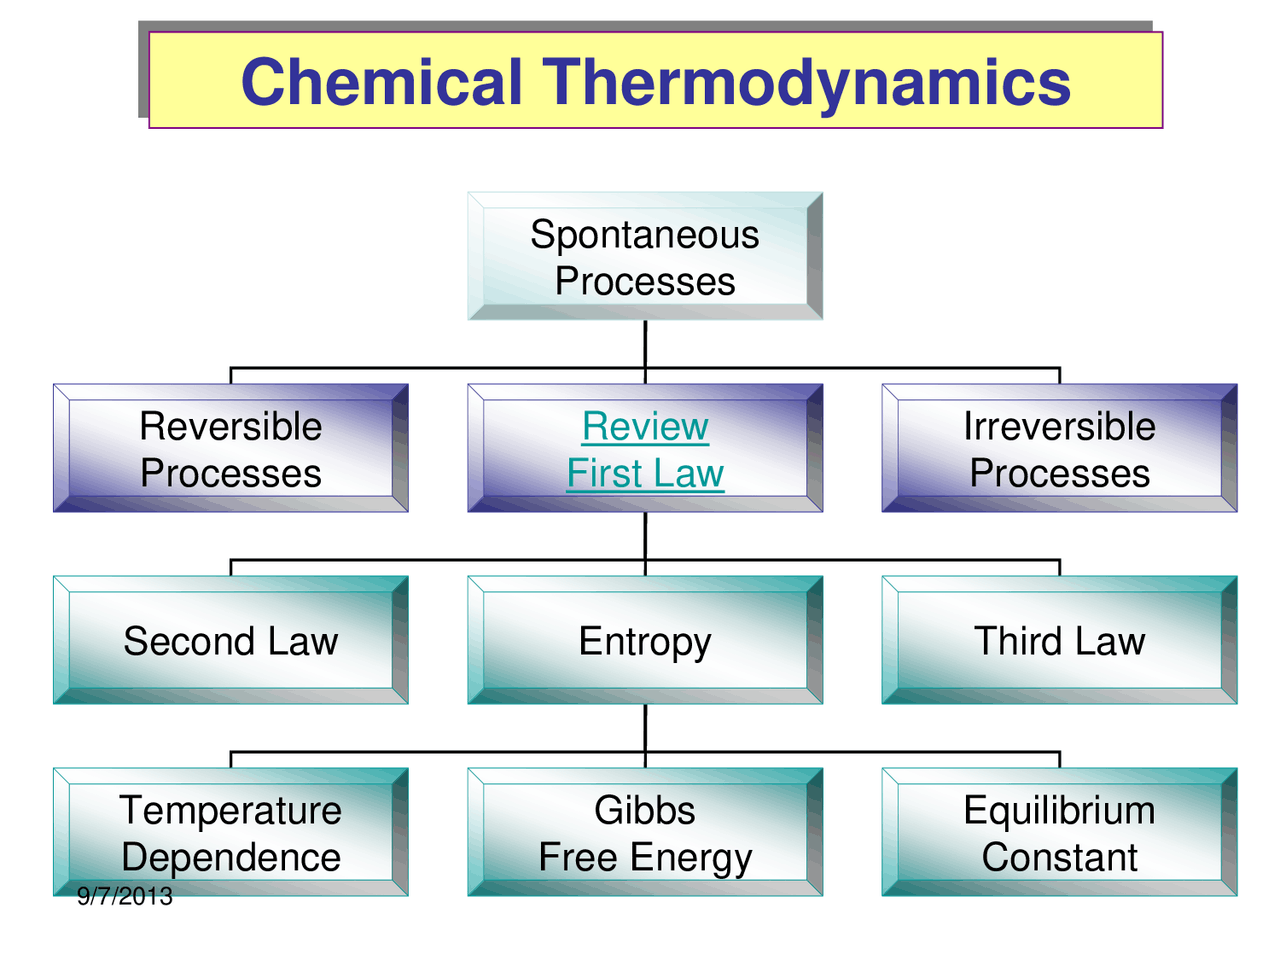
At the core, you are tracking three state functions: enthalpy, entropy, and Gibbs free energy. Enthalpy tells you the heat content change. Entropy measures the dispersal of energy and matter. Gibbs free energy combines the two and accounts for temperature. If Gibbs free energy is negative, the process is spontaneous under those conditions. If it is positive, you either need to change the conditions or accept that the reaction will not happen on its own. The standard equations are straightforward: G = H - TS
That equation alone explains a lot of the failures I saw in the lab. A reaction with a positive H and negative S will never be spontaneous at any temperature. A reaction with negative H and positive S is spontaneous at all temperatures. The interesting cases sit in between, where temperature decides the outcome. Most students stop there. They calculate a value, check a sign, and move on. The real work happens when you realize that standard values assume 298 K and 1 bar. Real reactors do not operate at standard conditions. When you run a reaction at 150°C or under high pressure, those standard tables become approximations at best. You need to correct for temperature using heat capacity data or rely on experimental measurements. I have seen multiple process failures traced back to someone plugging standard G values into an equation for a reaction running at 400 K without any adjustment. Another thing beginners miss is that thermodynamics tells you nothing about rate. A reaction can be wildly spontaneous and still take years to show any measurable progress. Kinetics and thermodynamics are separate conversations. I once had a colleague insist we switch catalysts because the Gibbs free energy was not favorable enough. The G was perfectly fine. The activation energy was the problem. Switching catalysts had nothing to do with the thermodynamic barrier.
Get the Full Details

How to Actually Apply This in Practice
Start by identifying what you are trying to predict. Are you checking if a reaction is feasible? Determining the equilibrium composition? Calculating the heat load for a reactor design? Your approach changes depending on the question. For feasibility checks, use standard Gibbs free energy of formation values from a reliable database. NIST Chemistry WebBook is one option. The critical part is verifying the phase of each compound. Using a gas-phase value for water when your reaction produces liquid water at the operating temperature will throw your calculation off by about 24 kJ/mol. That difference can flip a borderline result. When temperature matters, integrate the heat capacity over the range. The Kirchhoff equation gives you a quick approximation if Cp is roughly constant:
H(T) = H(T) + Cp(T - T) For Gibbs free energy at non-standard temperatures, use the Gibbs-Helmholtz relationship or tabulated values if available. Some software packages like Aspen Plus or Chemically can handle this automatically, but you should know what they are doing under the hood. Relying on a black box without understanding the assumptions is how you get wrong answers that look right. Equilibrium calculations require a different approach. Write the equilibrium expression, express concentrations or partial pressures in terms of a single variable, and solve. For gas-phase reactions, remember that Kp relates to Kc through the factor (RT)^n. Getting that conversion wrong is one of the most common errors I encounter. I corrected a student's calculation once where they had used Kp directly with concentration-based units. The result was off by several orders of magnitude. The reaction appeared non-spontaneous when it was actually heavily favorable.
For solution chemistry, activity coefficients matter. The ideal dilute solution assumption breaks down quickly as concentration increases. I worked on a precipitation process where the calculated solubility product predicted complete precipitation, but the actual yield was only 60%. The missing factor was the ionic strength effect. Once I applied the Debye-Hückel correction, the prediction aligned with the experimental data within 5%.
A Specific Problem That Taught Me Something
On a scale-up project involving an exothermic esterification, the thermodynamics indicated a manageable heat release. The lab-scale runs performed well. When we moved to the pilot reactor, the temperature ran away and the product degraded. The error was not in the thermodynamic calculation itself. It was in assuming the heat capacity of the reaction mixture stayed constant. As the reaction progressed, the composition shifted significantly, and the actual heat capacity dropped by about 18%. This meant the adiabatic temperature rise was much higher than predicted. We had used a fixed Cp value from the pure reagents, not the changing mixture. The fix was to measure Cp across the full composition range and feed that data into the energy balance. That single adjustment changed our cooling system specification by 40%. Thermodynamics has real constraints. It assumes equilibrium, which many real systems never reach. It ignores transport limitations. It does not account for metastable states, which are common in crystallization and phase separation processes. If you are working with electrochemical cells, overpotentials mean your measured voltage will always be lower than the thermodynamic prediction. For combustion systems, dissociation at high temperatures shifts the equilibrium composition in ways that simple stoichiometry misses entirely. Another limitation is data quality. Enthalpy and entropy values are measured experimentally, and the uncertainties add up. When you combine multiple substances in a reaction, the propagated error can be significant. For reactions involving organic molecules with similar structures, the tabulated values sometimes come from group contribution methods rather than direct measurement. Those estimates can deviate by 5 to 10 kJ/mol from experimental results. If your feasibility decision hinges on a G near zero, that uncertainty matters a lot.
If thermodynamic data is unavailable or unreliable for your system, the practical alternative is calorimetry. Titration calorimetry or differential scanning calorimetry can give you direct measurements of heat effects under your actual conditions. It is more expensive and slower than looking up a table, but it is often the only way to get trustworthy numbers for novel or complex reactions. Bottom line: thermodynamics gives you boundaries. It tells you what is possible and what is not. It does not tell you how fast, how clean, or what side products will form. Use it to filter out impossible routes early, then validate with experiment. The people who skip that second step tend to waste more time and money than the ones who spend extra time on the calculations upfront.