What "Concerted" Actually Means in Organic Chemistry
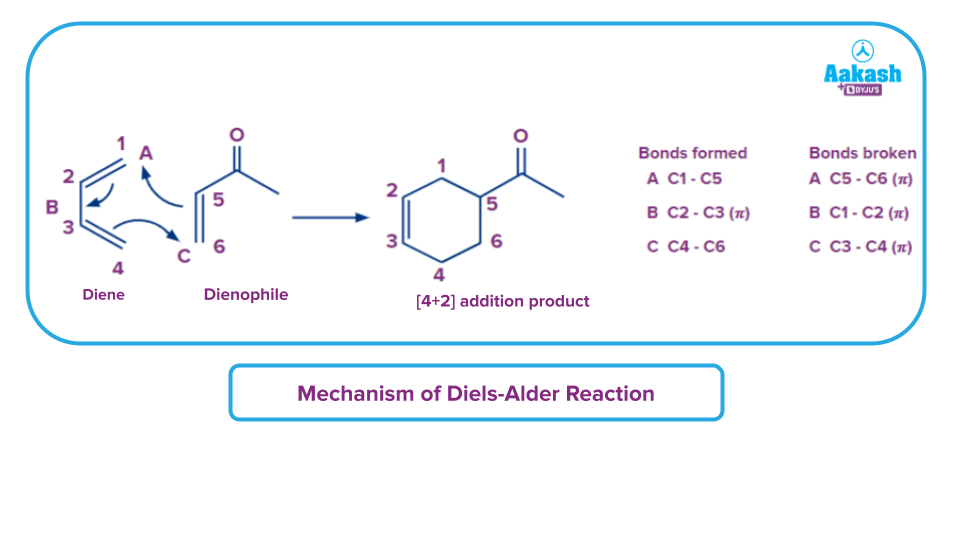
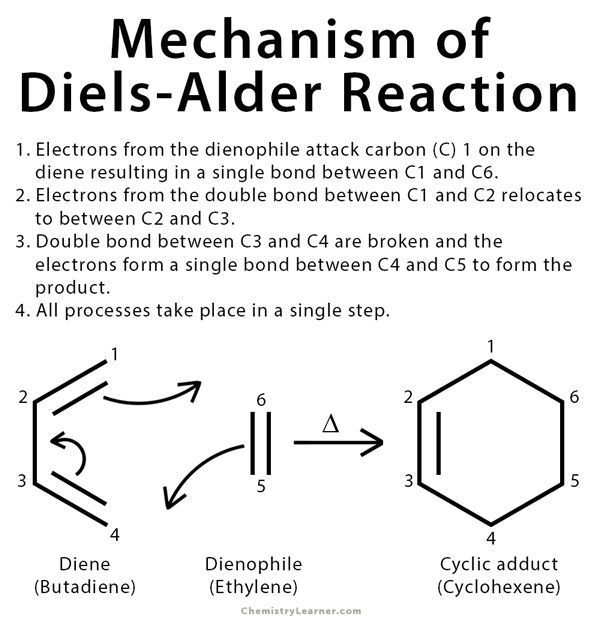
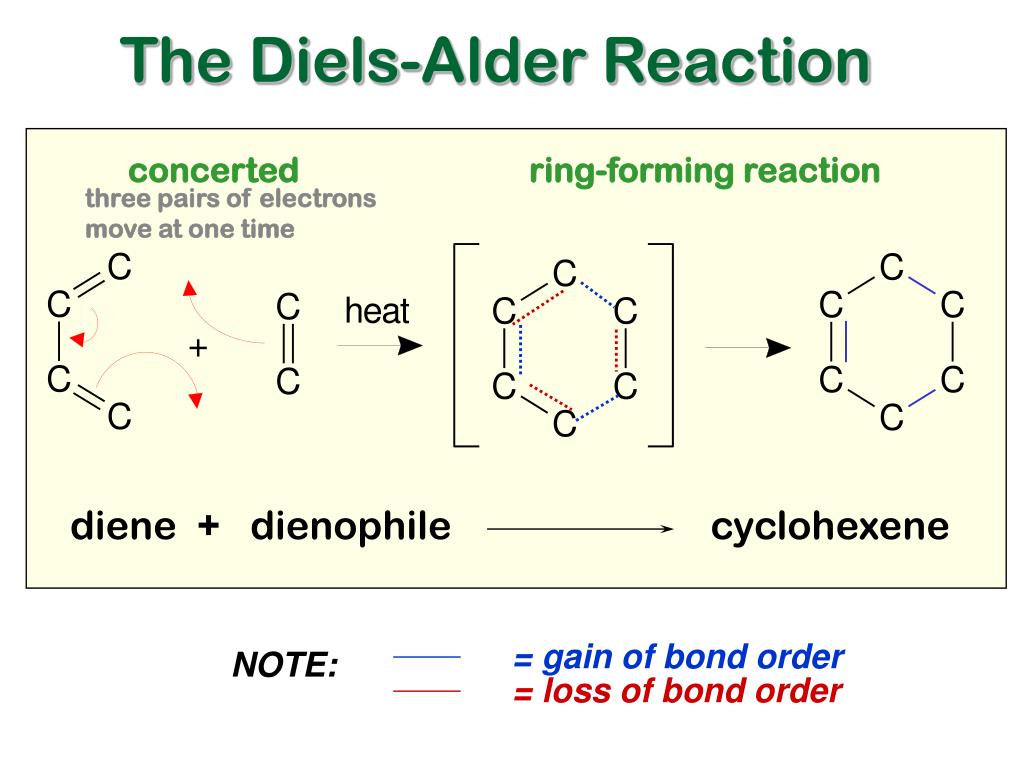
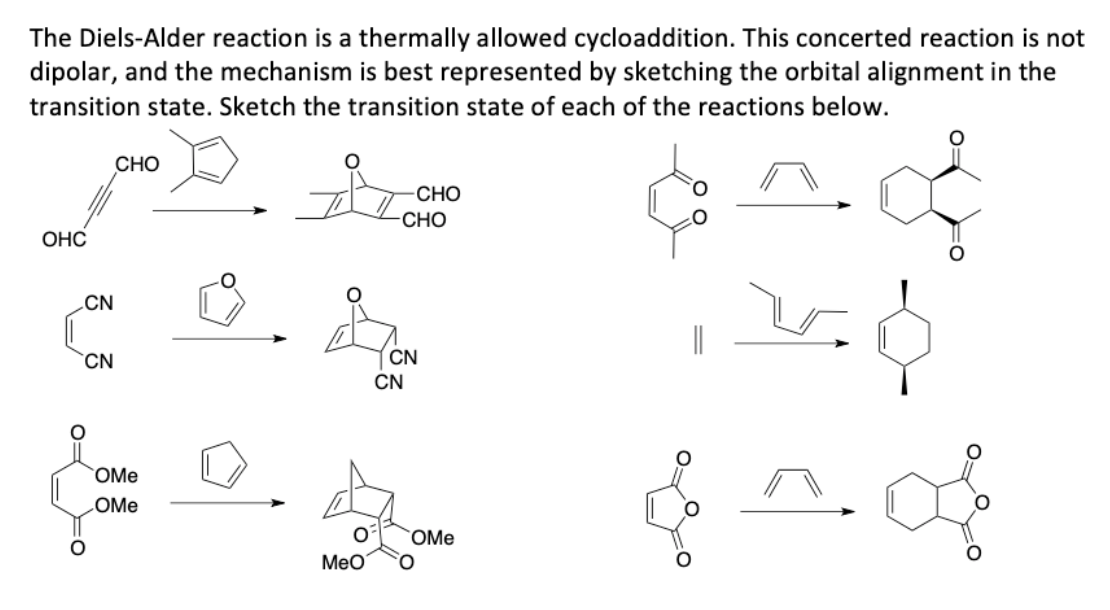
The word concerted shows up constantly when people talk about the Diels-Alder reaction, and it gets misused more often than you would think. A concerted reaction is one where every bond-making and bond-breaking event happens at the same time, through a single transition state, with no intermediate structures sitting in between. That is the textbook definition. What the definition does not tell you is why that matters in practice, or what happens when things do not go perfectly concerted. In the Diels-Alder, a conjugated diene and a dienophile come together to form a six-membered ring. Six pi electrons move around in a single cyclic reorganization. Two new sigma bonds form at opposite ends of the system while three pi bonds break and one new pi bond forms inside the ring. Nothing pauses in the middle. No carbocation. No diradical. Just one transition state connecting reactants to product. The pericyclic label comes from that same idea—the electrons circulate around a ring-shaped transition state, and that closure is what makes the whole thing concerted. I have seen students draw stepwise mechanisms for Diels-Alder because they are trying to make sense of regiochemistry, and that is a real problem. When you draw a zwitterionic intermediate to explain why electron-rich dienes react faster with electron-poor dienophiles, you are no longer describing a concerted process. You are describing something that looks like it could be ionic. The distinction matters because the kinetic isotope effects, the stereochemical retention, and the activation parameters all point to one transition state. If you invoke an intermediate, you have to explain why the reaction still shows complete stereospecificity. That explanation usually collapses under scrutiny.
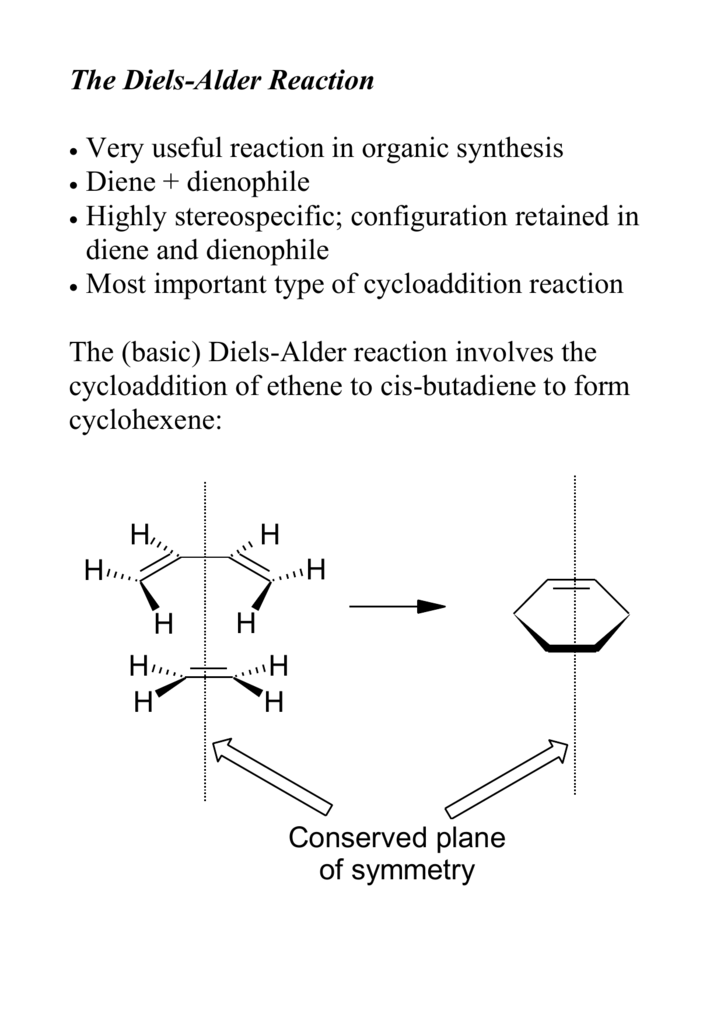
The practical consequence of concertedness is stereochemical memory. If your dienophile is cis, the product is cis. If it is trans, the product is trans. If you run a deuterium-labeled substrate, the labels end up exactly where the concerted mechanism predicts, not scrambled. I once spent two weeks trying to figure out why a particular brominated cyclohexene product had lost its stereochemical integrity. The answer turned out to be that the starting material was not pure—it had a tiny amount of isomerized dienophile contamination that I could not see on TLC. Once I ran GC-MS and corrected the starting material, the stereochemistry came back exactly as expected. Concerted reactions are strict about what you put in. They do not forgive sloppy materials. There are edge cases where the reaction is not perfectly concerted, and you need to know about them. Certain substrates with very poor orbital overlap can slip into stepwise pathways, especially if the resulting intermediate is stabilized by resonance or neighboring group participation. I encountered this with a highly electron-deficient fulvene derivative reacting with an unactivated alkene. The expected adduct formed slowly alongside a rearranged product that could only come from a zwitterionic intermediate. The workup was straightforward—column chromatography separated the two—but the mechanistic lesson stuck with me. You cannot assume concertedness just because you are running a Diels-Alder. You have to check the data: kinetic studies, isotope effects, and sometimes computational modeling to confirm that the transition state is indeed a single structure. The orbital symmetry explanation for why this works comes from the Woodward-Hoffmann rules. A suprafacial-suprafacial [4+2] cycloaddition is thermally allowed because the HOMO of the diene and the LUMO of the dienophile overlap in a bonding fashion across the entire system simultaneously. The frontier molecular orbital picture makes this concrete. The diene HOMO has two terminal lobes with the same phase, and the dienophile LUMO has two lobes with matching phases at the reactive carbons. When they approach, all four interactions are bonding at once. That simultaneous bonding is the quantum mechanical reason behind the word concerted. It is not just a label. It is a prediction about how the electron density moves.
One counter-intuitive point that beginners miss is that concertedness does not mean fast. Some Diels-Alder reactions require heat and pressure, sometimes several hours at elevated temperature. The activation barrier depends on the orbital energy gap between the diene HOMO and the dienophile LUMO, not on whether the mechanism is stepwise. An electron-poor dienophile like maleic anhydride reacts with cyclopentadiene at room temperature in minutes. A less activated pair might need 150 degrees Celsius in a sealed tube. Both are concerted. Both go through one transition state. The difference is the height of the barrier, not the nature of the mechanism. Another thing people overlook is that the concerted mechanism predicts endo selectivity through secondary orbital interactions. The endo product forms faster because the pi system of the dienophile's substituent overlaps with the back lobes of the diene's central double bond in the transition state. This is a kinetic effect, not a thermodynamic one. The exo product is often more stable, but it forms more slowly. At low temperature, you get the endo product. At high temperature with long reaction times, the product distribution can shift toward exo if the reaction becomes reversible. I learned this the hard way when a graduate student ran a Diels-Alder at 180 degrees for twelve hours and wondered why the endo ratio dropped from 95:5 to about 60:40. The reaction was retro-Diels-Alder happening in parallel with the forward reaction. The endo adduct decomposed faster because it was the kinetic product, and the exo adduct accumulated as the thermodynamic product. The limitations of relying on concertedness as a predictive tool are real. Concerted mechanisms assume that the reaction proceeds through a single transition state, but solvent effects, steric strain, and substrate flexibility can distort that picture. In polar solvents, some Diels-Alder reactions show rate enhancements that are difficult to explain without invoking some ionic character in the transition state. The reaction is still concerted in the sense that no intermediate forms, but the transition state may be asynchronous—one bond is more formed than the other. I have seen computational studies show bond lengths in the transition state that differ by 0.2 angstroms or more. That is still concerted, but it is not symmetric concerted. Beginners often treat concerted and synchronous as the same thing. They are not. Synchronous means both bonds form at exactly the same rate. Concerted means no intermediate forms. You can have a concerted but asynchronous reaction.
Get the Full Details
If you need to determine whether your reaction is truly concerted, the most reliable approaches are kinetic isotope effect measurements and computational transition state optimization. A primary kinetic isotope effect at the reacting carbons indicates that bond formation is advanced in the transition state. DFT calculations at the B3LYP/6-31G* level or higher will show you the transition state geometry directly. If the computed structure has one C-C bond at 2.2 angstroms and the other at 1.8 angstroms, the reaction is concerted but asynchronous. If the computation shows a stable intermediate between the two transition states, then you have a stepwise mechanism, and the concerted label does not apply. I used this approach to resolve a dispute in my group about whether a particular intramolecular Diels-Alder was stepwise. The kinetic data suggested concerted, but the product distribution looked stepwise. The DFT calculation showed a highly asynchronous concerted transition state that explained both the kinetics and the products without invoking an intermediate. The controversy resolved in about a week once we had the computation. The broader implication of understanding concertedness goes beyond the Diels-Alder. Once you internalize what concerted means, you start seeing it everywhere—in electrocyclic reactions, sigmatropic rearrangements, and 1,3-dipolar cycloadditions. The common thread is the pericyclic transition state: a cyclic array of overlapping p orbitals where electron reorganization happens in a single coordinated step. The mnemonic is simple—count the electrons involved. Four pi electrons give you a different symmetry rule than six pi electrons. Eight pi electrons follow the four pi electron rules because 4n and 4n+2 are different categories. This is not arbitrary. It comes from the phase relationships in the molecular orbitals, and it is the same orbital symmetry logic that makes the Diels-Alder concerted in the first place. For anyone actually running these reactions in the lab, the takeaway is practical. If your Diels-Alder is concerted, you can predict stereochemistry with confidence. If your starting materials are pure and your geometry is right, the product stereochemistry is locked in. You do not need to worry about racemization or scrambling at the reactive centers. But if your yields are low or your stereochemistry is wrong, do not immediately blame the mechanism. Check your materials, check your temperature, check for competing retro-reactions. Concerted mechanisms are elegant, but they do not make bad chemistry good.