Why You Are Probably Doing This Wrong
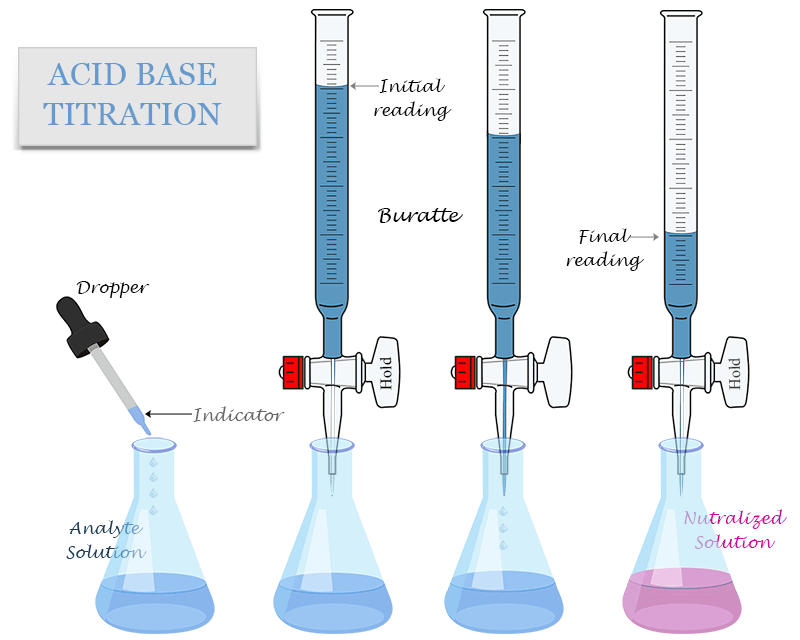
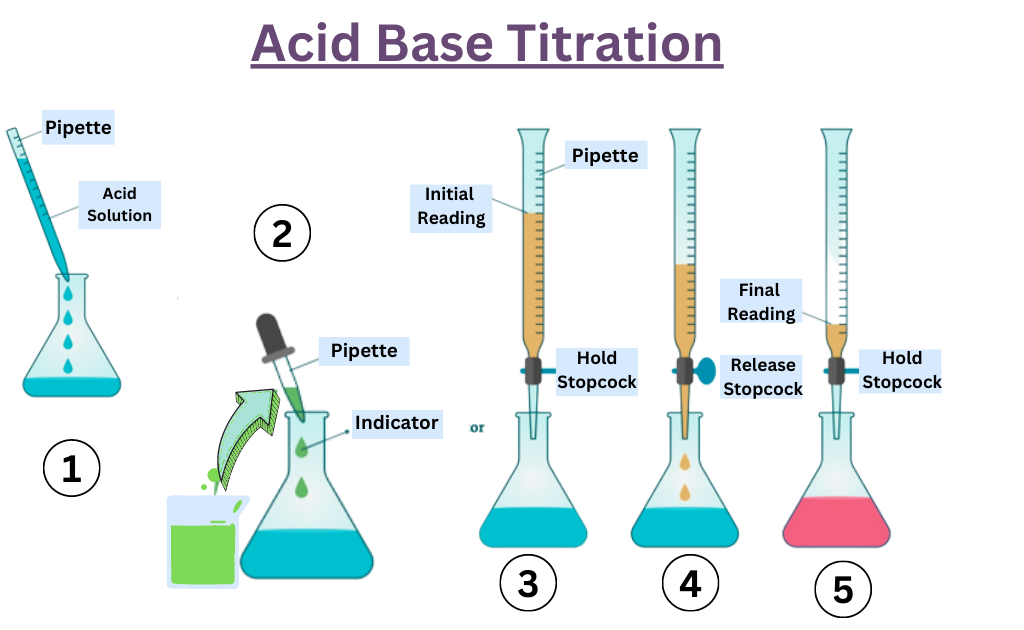
I spent about three years in an analytical chemistry lab before I ever cared about this stuff, and honestly, the titration of an acid and a base seems dead simple until you are sitting there at 2 AM with a burette that refuses to stop dripping and your endpoint keeps drifting by half a milliliter every time. The core concept is straightforward enough — you are figuring out how concentrated an unknown solution is by gradually neutralizing it with a solution of known concentration. The problem is that doing it well requires patience, steady hands, and knowing where most people screw up before they even start. The basic setup involves a burette clamped vertically above an Erlenmeyer flask. You fill the burette with your titrant, which is the solution of known concentration, usually a strong base like sodium hydroxyde if you are standardizing an acid. You then deliver that titrant drop by drop into the analyte — the solution you are trying to analyze — while constantly swirling the flask. An indicator, most commonly phenolphthalein for strong acid strong base work, changes color when the reaction reaches its equivalence point. You record the volume dispensed and calculate the unknown concentration from there. The math part is trivial. At the equivalence point, the moles of acid equal the moles of base. M_a times V_a equals M_b times V_b for a monoprotic acid and a monobasic compound. For diprotic acids like sulfuric acid, you need to account for the stoichiometry — two moles of base are required per mole of acid, so the calculation becomes M_a times V_a times 2 equals M_b times V_b. Get this wrong and your result is completely off, and you will not know it until you compare against a known standard and everything looks wrong.
Here is a practical problem I ran into repeatedly that nobody mentions in textbooks. When you are titrating a weak acid with a strong base, the equivalence point is not at pH 7. It is somewhere between 8 and 10 depending on the acid strength. If you use methyl orange as your indicator for a weak acid titration, you are going to get a premature endpoint and your results will be consistently too low. Phenolphthalein, which transitions around pH 8.2 to 10, is the correct choice here. I once spent two days troubleshooting what I thought was contaminated reagent before I realized I had been using the wrong indicator the entire time. The sodium hydroxide was fine. I was just reading the color change at the wrong pH range. Another thing that catches people out is carbon dioxide absorption. Sodium hydroxide solutions absorb CO2 from the air over time, forming sodium carbonate. This affects the titer of your standard base solution. If you prepare your NaOH and leave it sitting open for a week, the concentration shifts. The workaround is to standardize your base against primary standard potassium hydrogen phthalate immediately before use, and to keep the storage bottle capped with a soda lime trap if you need the solution to stay stable for more than a day or two. It is a small detail but it introduces systematic error that compounds with every titration you run after the solution degrades. The burette itself demands attention. Reading the meniscus correctly is non-negotiable. Your eye level needs to be exactly at the bottom of the meniscus, not above or below it. Parallax error alone can easily add 0.05 to 0.1 mL of uncertainty to each reading, which at the volumes you are typically working with translates to a significant percent error. Also, always rinse the burette with a small amount of the titrant before filling it. If you fill it with distilled water residue still inside, you are diluting your standard solution right at the start. Same rule applies to the pipette you use to measure your analyte — rinse it with the solution you are about to pipette, not just water.
Waiting for the color change to stabilize is another area where beginners rush and lose accuracy. After you add a drop of titrant and the pink color appears and then disappears with swirling, do not celebrate prematurely. Keep adding dropwise, and after each drop, swirl thoroughly and wait a full three seconds before deciding if the endpoint is truly reached. The color can fade as the local excess of base mixes into the bulk solution, giving you a false impression that you have overshot. I have seen people stop titration while the solution is still slightly acidic in the center of the flask because they could not see the mixing process clearly. Temperature is another factor that gets ignored. Reaction kinetics and indicator pKa values both shift slightly with temperature. If you are doing work that requires high precision, note the laboratory temperature and try to keep your standard solutions and analyte at roughly the same temperature. A five degree difference between your standard solution stored in a drawer and the analyte sitting on the bench is usually negligible for routine work, but it matters if you need results within 0.1 percent relative error. For very dilute solutions below 0.01 M, titration becomes unreliable. The volume readings require a microburette or at least a 10 mL burette instead of the standard 50 mL, and the indicator color change becomes vague rather than sharp. In those cases, potentiometric titration using a pH meter is the better approach. You plot pH versus volume added and identify the equivalence point as the inflection point on the curve. It takes more equipment and more time per run, but it gives you a clear, unambiguous endpoint regardless of solution color or transparency.
Get the Full Details

If you are looking for something to help you track your titration data and calculate results automatically, there are spreadsheets and small software tools you can find online, but honestly, the calculation itself is so simple that a basic sheet does more harm than good if you do not understand the chemistry behind the numbers. I would recommend learning to do the calculation by hand first, then automating it later. Most of the errors in undergraduate labs come from blind trust in a formula without understanding what each variable represents. The biggest single piece of advice I can give is this: run a rough titration first to find the approximate endpoint volume, then run at least two precise titrations within one drop of that rough estimate. A single careful titration is worth more than five rushed ones, and replicates tell you whether your technique is consistent or whether you are just getting lucky with one good read.