The Nucleus Is Where The Whole Thing Starts
If you are studying molecular biology, you already know transcription occurs in the nucleus of eukaryotic cells. That is the textbook answer. In prokaryotes it happens in the cytoplasm since they lack a nucleus. The process itself is straightforward until you actually try to work with it in a lab setting, which is where things get complicated. I spent a few years doing in vitro transcription assays and dealing with RNA isolation workflows. It is not glamorous work. RNA degrades fast. RNases are everywhere. I once spent two weeks tracking down why my transcripts kept coming back shorter than expected. The issue wasn't the polymerase or the template. It was contamination from a single used pipette tip that someone had left sitting out on the bench. RNases don't care about your experimental design. They just degrade whatever RNA they touch.
Transcription Occurs In The Nucleus (and Other Places Too)
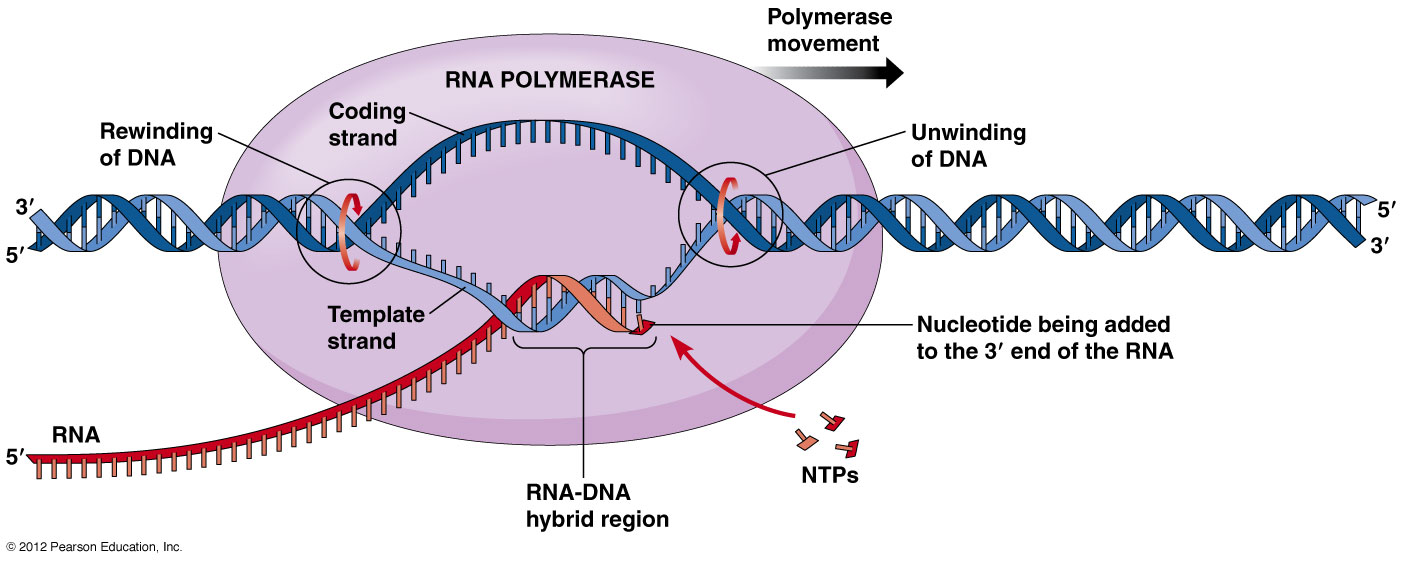
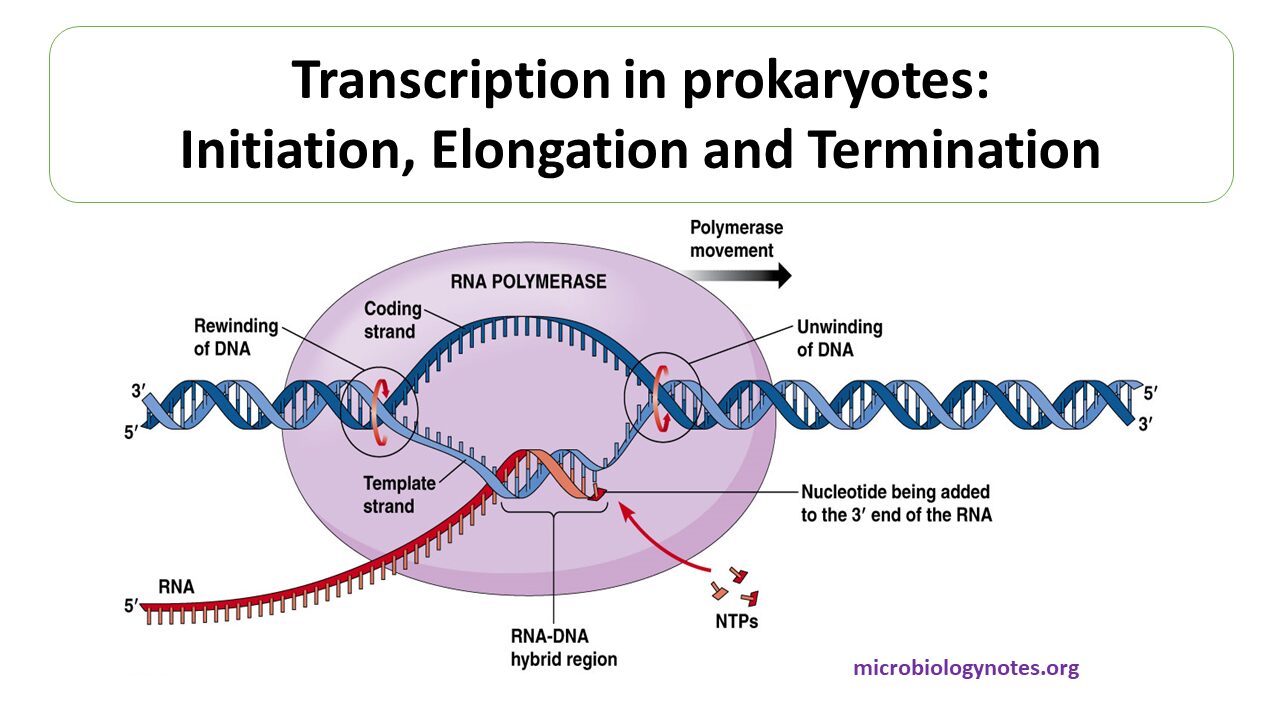
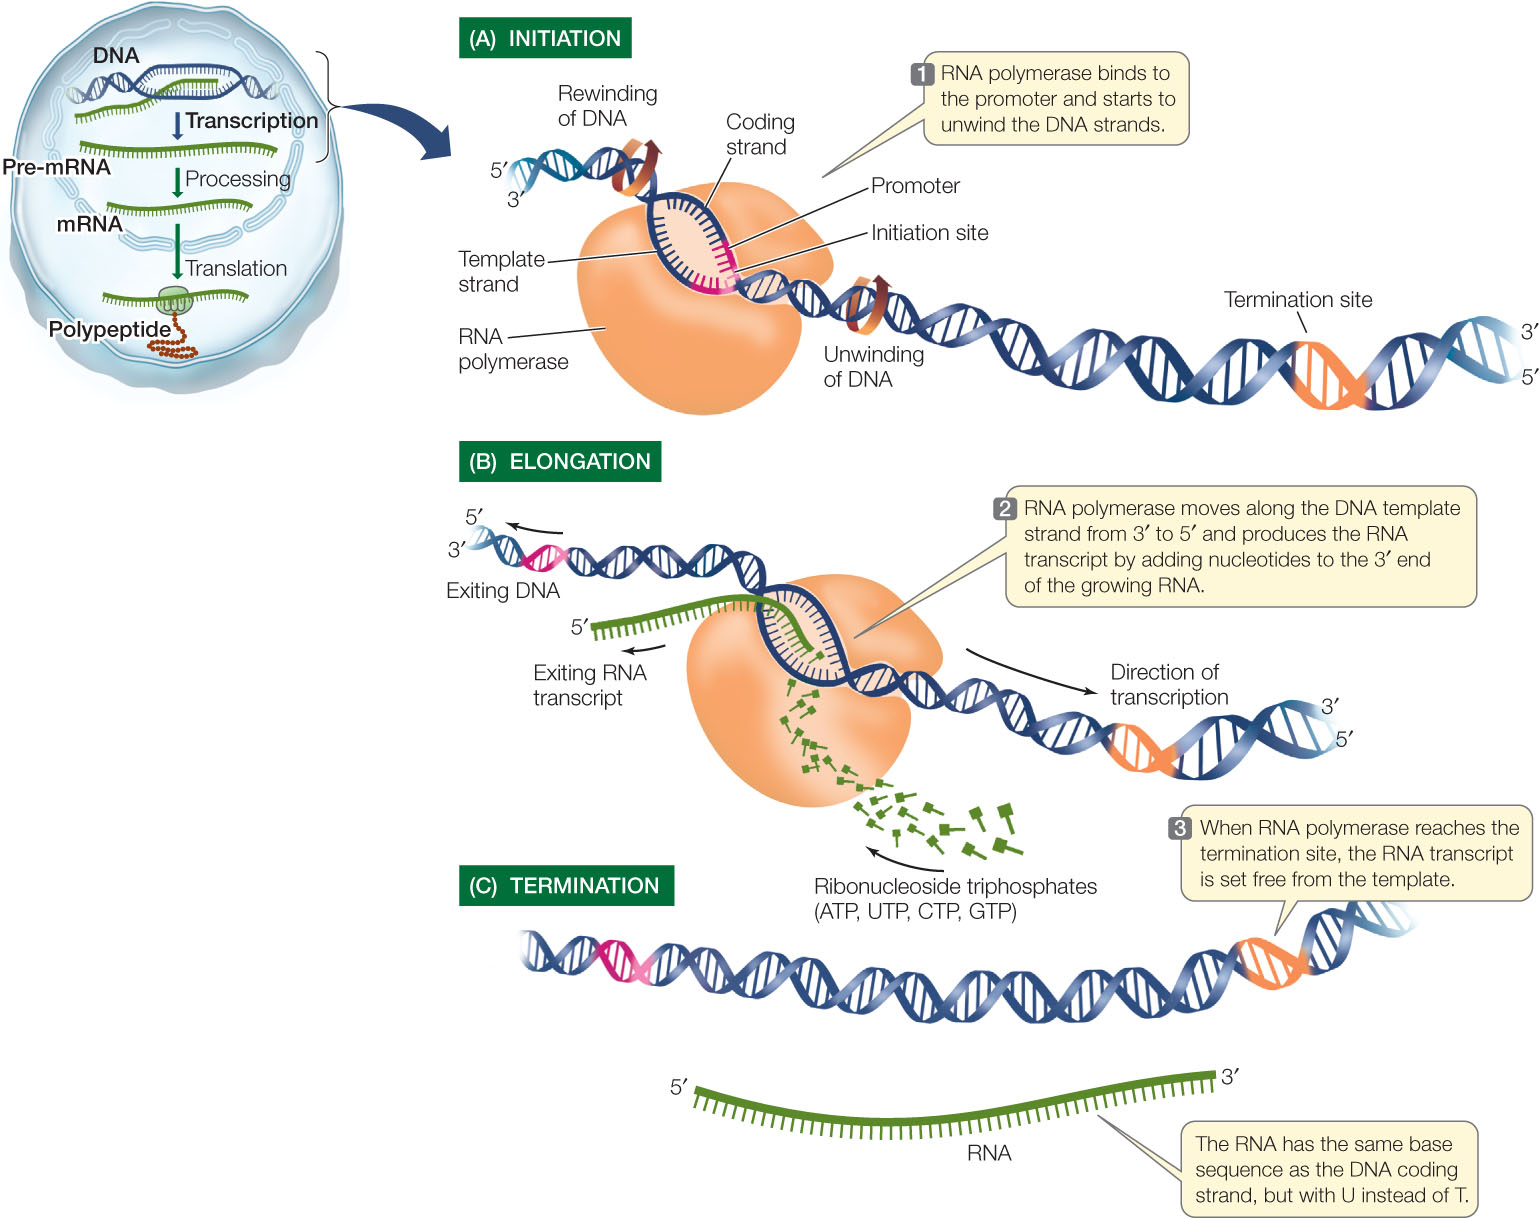
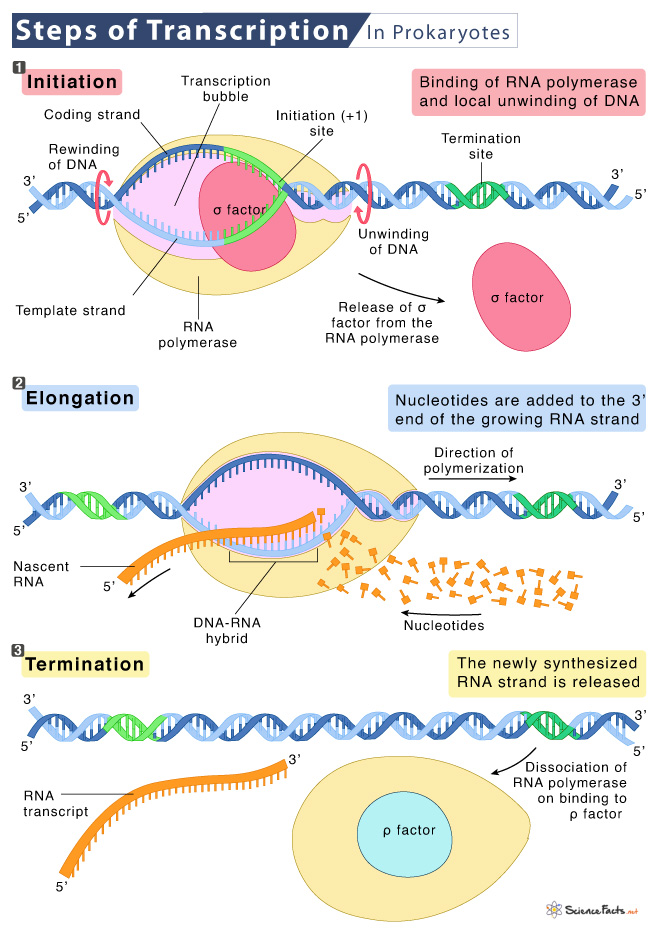
Eukaryotic transcription involves three main RNA polymerases. RNA polymerase II handles messenger RNA. RNA polymerase I takes care of ribosomal RNA. RNA polymerase III does transfer RNA and a few other small RNAs. Each one has its own set of general transcription factors and regulatory proteins. TFIIID, TFIIB, TFIIF, TFIIE, TFIIH — these are the ones you will encounter most often with Pol II. They assemble at the promoter in a specific order to form the pre-initiation complex before transcription begins. Prokaryotic transcription is simpler. One RNA polymerase handles everything. The sigma factor directs the core enzyme to the promoter. Once initiation happens, the sigma factor falls off and elongation proceeds. That is about it. The whole process from start to finish can take a bacterial cell only a few seconds to transcribe an operon. Here is something people often miss. Transcription and RNA processing are coupled in eukaryotes. The 5' cap is added to the nascent RNA almost immediately after the first twenty nucleotides emerge from the polymerase. Splicing begins before transcription is even finished. The poly-A tail gets added as well. These aren't separate steps you can neatly pull apart. They happen simultaneously on the same RNA molecule while it is still being synthesized. Trying to study them in isolation means you are looking at artifacts, not reality.
The Practical Side of Working With Transcription
If you are setting up an in vitro transcription reaction, you need a DNA template with a promoter sequence recognized by your chosen polymerase. T7, T3, and SP6 promoters are the standard choices for bacteriophage polymerase systems. These give you higher yields than using eukaryotic Pol II directly, which is why most commercial kits rely on them. A typical 20 microliter reaction might look like this: ten nanograms of linearized DNA template, one unit of T7 RNA polymerase, four millimolar NTPs, forty millimolar DTT, and the appropriate buffer. Incubate at thirty-seven degrees Celsius for one to two hours. The yield depends heavily on your template quality. Supercoiled plasmid DNA doesn't work well for in vitro transcription. You have to linearize it first, or the polymerase will run off the end of the circle and re-transcribe the same region repeatedly, producing unproductively long transcripts that are difficult to work with. I learned that the hard way during my first month in the lab. I pulled transcripts from a plasmid I hadn't checked for supercoiling integrity. The resulting RNA smear on a denaturing gel looked nothing like the clean band I was expecting. Linearizing the template solved it immediately. Now I check every template on an agarose gel before I ever set up a transcription reaction.
Get the Full Details
After transcription, you need to remove the DNA template and the enzymes. DNase I treatment is standard practice. A common pitfall here is adding too much DNase I or leaving it in the reaction too long. Some of your RNA transcript will get dragged along with the degraded DNA in subsequent purification steps if you aren't careful. I use a phenol-chloroform extraction followed by ethanol precipitation, or a silica column kit if I need something faster. The column kits lose about ten to fifteen percent of your yield compared to precipitation, but they save you at least an hour of hands-on time. One thing worth noting is that transcription efficiency drops significantly with longer templates. If you are trying to transcribe something over three thousand nucleotides, the polymerase will often fall off before finishing. This isn't a minor inconvenience. It means you get a population of full-length transcripts mixed with shorter abortive products. For long transcripts, it helps to add spermidine to the reaction at a final concentration of about zero point five millimolar. It stabilizes the polymerase-template interaction and improves full-length yield by roughly twenty to thirty percent.
Common Mistakes and What Actually Fails
People tend to overestimate how pure their RNA needs to be for downstream applications. If you are doing Northern blotting or RT-PCR, a little bit of leftover buffer salts or trace ethanol from precipitation won't kill the experiment. But if you plan to do anything with ribosome profiling or single-molecule sequencing, even small amounts of contaminating DNA or truncated transcripts will ruin your data. Always run a small aliquot on a denaturing gel before you commit the rest to an expensive downstream assay. Another frequent problem is ignoring the temperature sensitivity of your polymerase. T7 RNA polymerase works well at thirty-seven degrees, but extending the incubation beyond two hours at that temperature starts degrading your product through autolytic cleavage. Lower the temperature to thirty degrees for longer incubations if you need maximum yield. It usually takes about four hours at thirty degrees to get the same result you would in two hours at thirty-seven. There is also the issue of template degradation during repeated freeze-thaw cycles. I keep all my DNA templates at negative twenty degrees in TE buffer rather than water. The EDTA in TE chelates divalent cations and slows down any contaminating nucleases. Templates stored in water alone tend to degrade noticeably after three or four freeze-thaw cycles, which introduces variability between reactions that is hard to account for statistically.
The bottom line is that transcription is conceptually simple but practically finicky. The theory doesn't break. The reagents do. RNases, degraded templates, incorrect polymerase choices, and improper purification are the usual suspects when something goes wrong. Pinpointing which one it was takes patience and systematic elimination. It is a skill that only comes from doing it repeatedly over a long period of time.