So You're Working With DNA Transcription. Here's What Actually Happens.
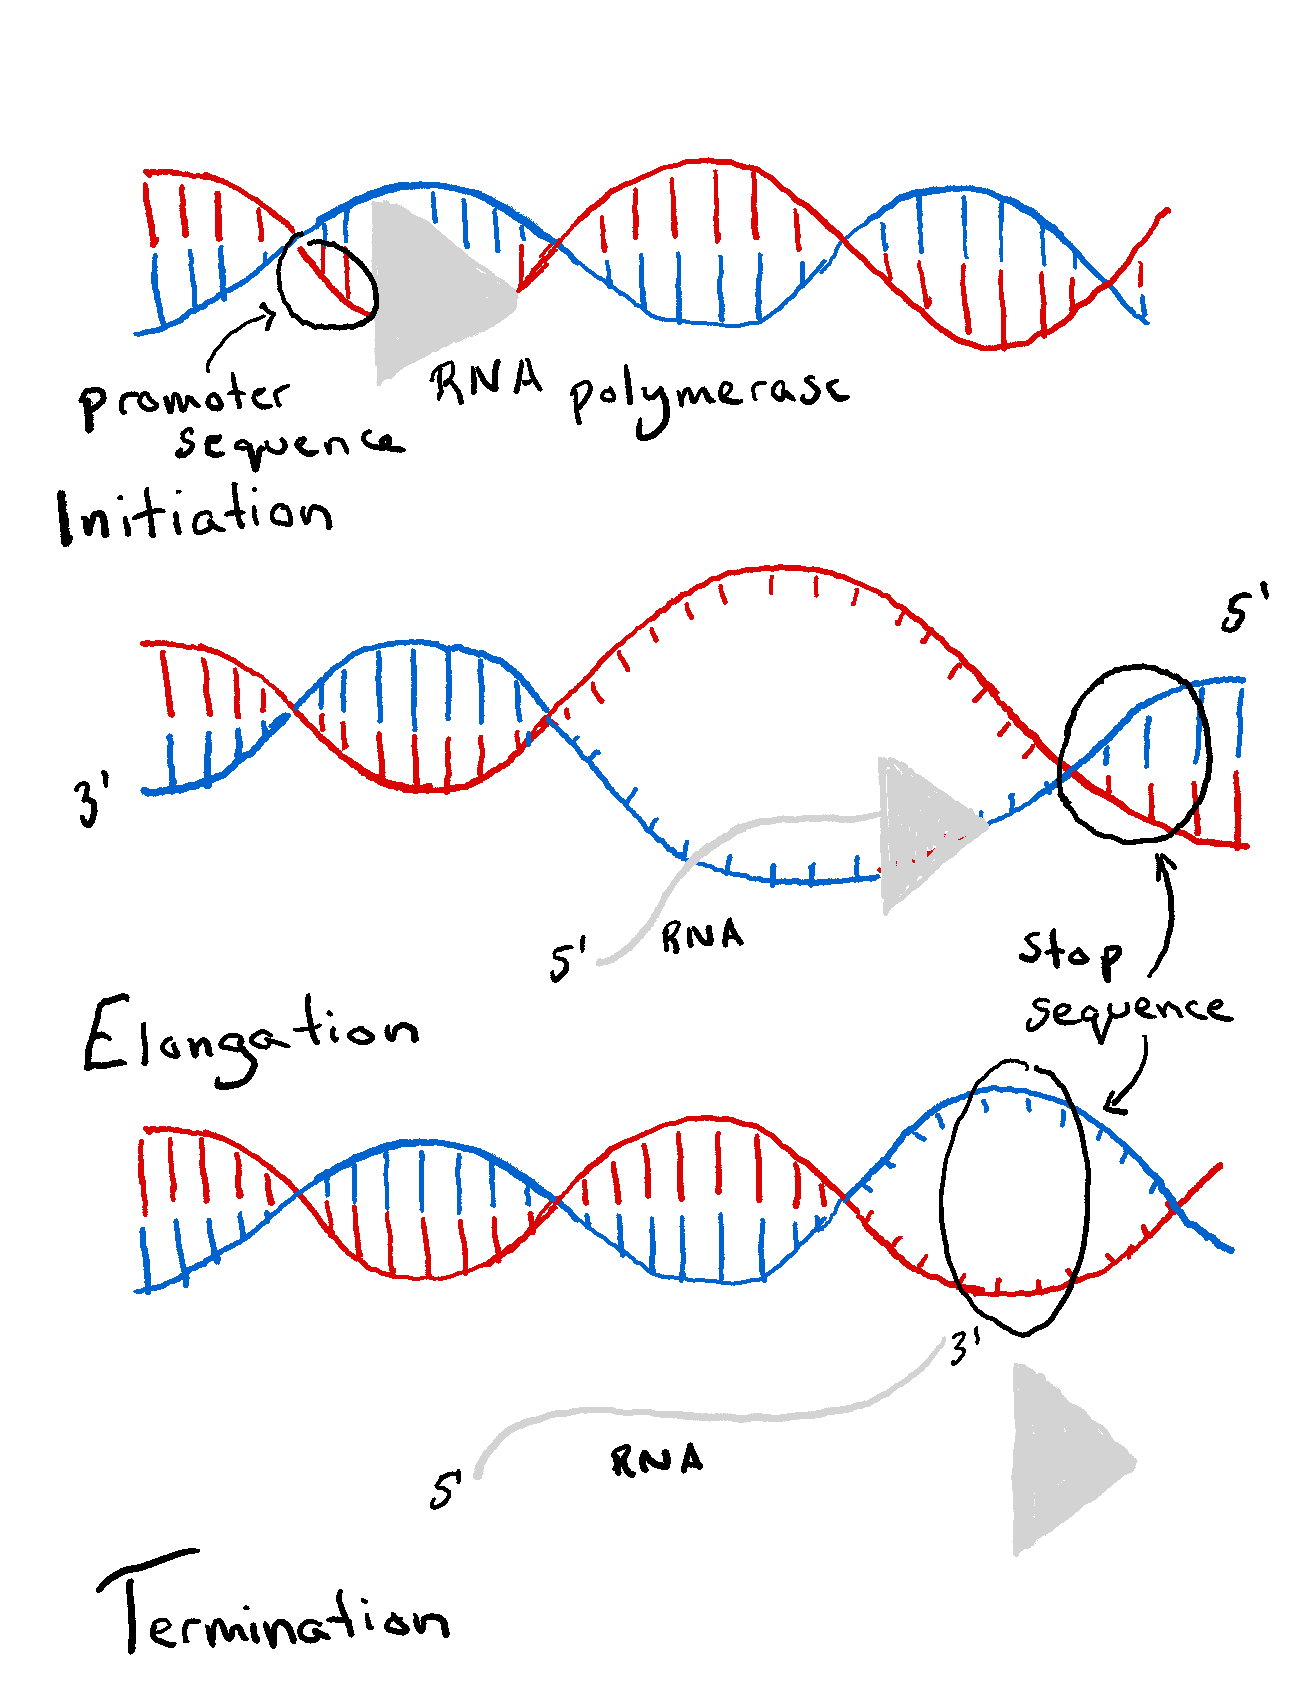
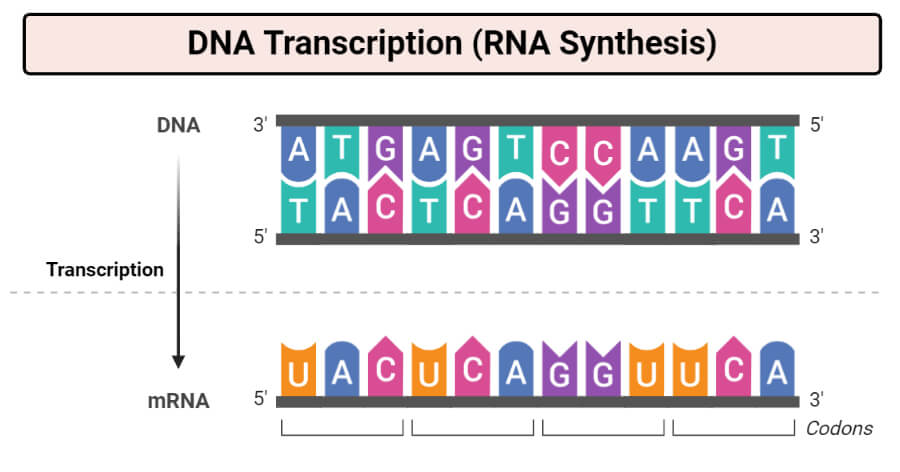
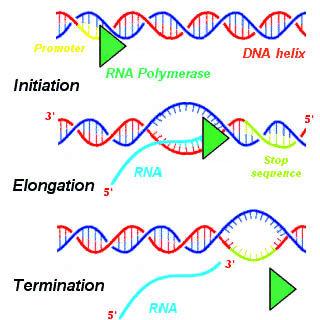
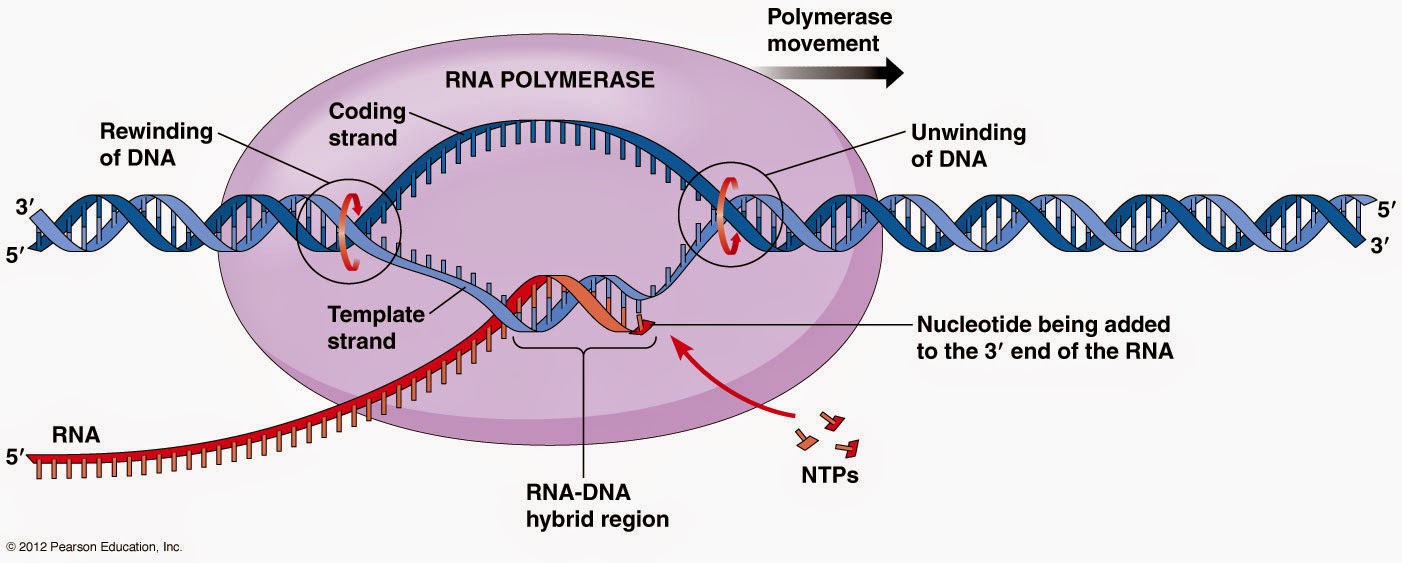
It starts with RNA polymerase binding to a promoter region, unwinding the double helix, and reading the template strand to build a complementary RNA copy. That's the textbook version. The real version involves a lot of failed experiments, misread base pairs, and polymers that just refuse to stay on track. Transcription of the dna is fundamentally about copying genetic information from DNA into RNA. Three main stages: initiation, elongation, and termination. In eukaryotes you also have splicing, capping, and polyadenylation layered on top, which is where things get messy fast. RNA polymerase II is the enzyme responsible for messenger RNA synthesis. It doesn't work alone. It needs transcription factors—TFIID, TFIIH, the whole complicated mess—to recognize the TATA box and start the whole process. Without those, nothing happens. Not even close.
One thing people get wrong is thinking transcription produces a perfect copy of the gene. It doesn't. The RNA transcript includes introns that need to be removed. You end up with a pre-mRNA that gets edited, capped at the 5' end with a modified guanine nucleotide, and polyadenylated at the 3' end. Skip any of those steps and your RNA is basically useless inside the cell.
What I Learned After Wasting Two Weeks On a Bad Protocol
I was running in vitro transcription assays with a plasmid template, trying to produce clean mRNA for an expression study. The polymerase kept falling off before it reached the full length of the insert. I thought it was a buffer issue. Switched suppliers. Tried different MgCl2 concentrations. Nothing. The problem turned out to be secondary structure in the DNA template itself. The region I was trying to transcribe had a strong hairpin-forming sequence that caused the RNA polymerase to stall and dissociate prematurely. I couldn't see it on the linearized plasmid map because I wasn't looking at the RNA product's folding potential. Running a quick RNAfold prediction on the expected transcript showed a stable stem-loop right around the drop-off point. I swapped the sequence for a codon-optimized version with less propensity to fold, and the full-length yield went from roughly 15% to about 85%. This kind of thing doesn't show up in any standard protocol sheet. You just hit it and spend a week troubleshooting before you figure out what's actually going wrong.
Get the Full Details
Things Nobody Warns You About
Promoter strength matters more than you'd think for in vitro systems. A T7 promoter isn't just a switch. Some plasmids have weak downstream elements or terminators that cause read-through into vector sequences, giving you a heterogeneous RNA population. Always verify your transcript ends where you expect them to. Run a denaturing gel. If you see smearing below your expected band, your termination isn't clean and you've got a cleanup step coming. DNase treatment after transcription is standard, but the inactivation step can be a trap. If you're using EDTA to chelate the magnesium for DNase inactivation, you'll also chelate the magnesium needed for any downstream translation or enzymatic reaction. I've wasted samples by forgetting this. Do a column purification instead if you're working toward a sensitive application. It removes the DNase, the salts, and the dead-end transcripts in one go. Another counter-intuitive point: higher temperature doesn't always mean better transcription. T7 RNA polymerase runs at 37 degrees Celsius, but that's not optimal for every template. Some templates with GC-rich regions produce significantly better full-length yields at 30 degrees. The polymerase moves slower, giving it more time to resolve local structural barriers. It trades speed for accuracy and completeness. Depends on what you need.
When Transcription Falls Apart Completely
There are scenarios where in vitro transcription just won't give you what you need. Long transcripts over 5 kilobases are rough. Yield drops exponentially with length. The polymerase stalls, the RNA degrades, and you're left with a mostly fragmented product. If you need long RNAs, consider running the transcription at reduced temperature with added BSA and longer incubation times, but don't expect full-length material above 6 to 7 kb. It's a hard limit with standard T7 systems. Heavy secondary structure in the template DNA itself can be another wall. If your gene has extensive palindromes or repetitive elements, the transcription machinery will choke. I've seen this with certain viral sequences and synthetic constructs. The workaround is usually — building the template in overlapping fragments, transcribing each piece separately, and then ligating the RNA products together. It's tedious and reduces overall yield, but it's often the only route. For cap analogs, think clearly about what you're using. Cap 0 analogs work fine for basic studies. If you're doing anything involving immune response or translational efficiency in mammalian cells, you need diethyl pyrocarbonate-treated cap analogs or structured cap analogs like ARCA. Standard cap analogs lead to significant fractions of uncapped transcripts, and those get degraded rapidly in cell lysates. I learned this the hard way after my protein expression numbers were a fraction of what the same construct gave with properly capped RNA.
Practical Checklist Before You Start
Verify your template is linearized downstream of your insert. Supercoiled plasmid gives you heterogeneous run-off products. Digest it, clean it up with a column, and measure concentration accurately — ethanol carryover inhibits polymerase activity noticeably. Include a proper negative control without the DNA template. You'll catch contaminating RNA in your reagents before it wastes a run. Use fresh DNase. Old aliquots lose activity and leave genomic DNA or plasmid backbone in your prep, which skews your yield calculations and interferes with downstream applications.
If you're producing mRNA for translation, treat the final product with a DNA removal column and then ethanol precipitate. It removes residual nucleotides, enzymes, and abortive transcripts. The pellet typically resuspends cleanly in RNase-free water or buffer, and you avoid the salt interference that ruins most downstream steps. Keep everything cold once the transcription is done. RNases are everywhere and they don't care about your timeline. Work on ice, use barrier tips, and don't reuse tubes. The cost of contaminated reagents is way lower than the cost of starting over. Yield expectations vary wildly based on template quality and transcript length. A clean 1 kb transcript from a well-verified template might give you 50 to 100 micrograms per milliliter of reaction. A 4 kb transcript with moderate structure might give you 10 to 20. Plan accordingly. Don't scale up a reaction blindly expecting linear yield increases — larger volumes introduce mixing and temperature gradient problems that offset the gain.