Getting Translation For Protein Synthesis Right in the Lab
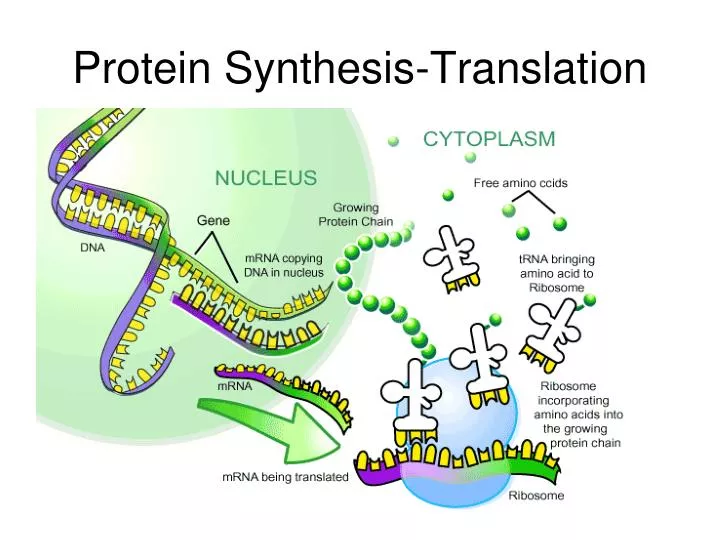
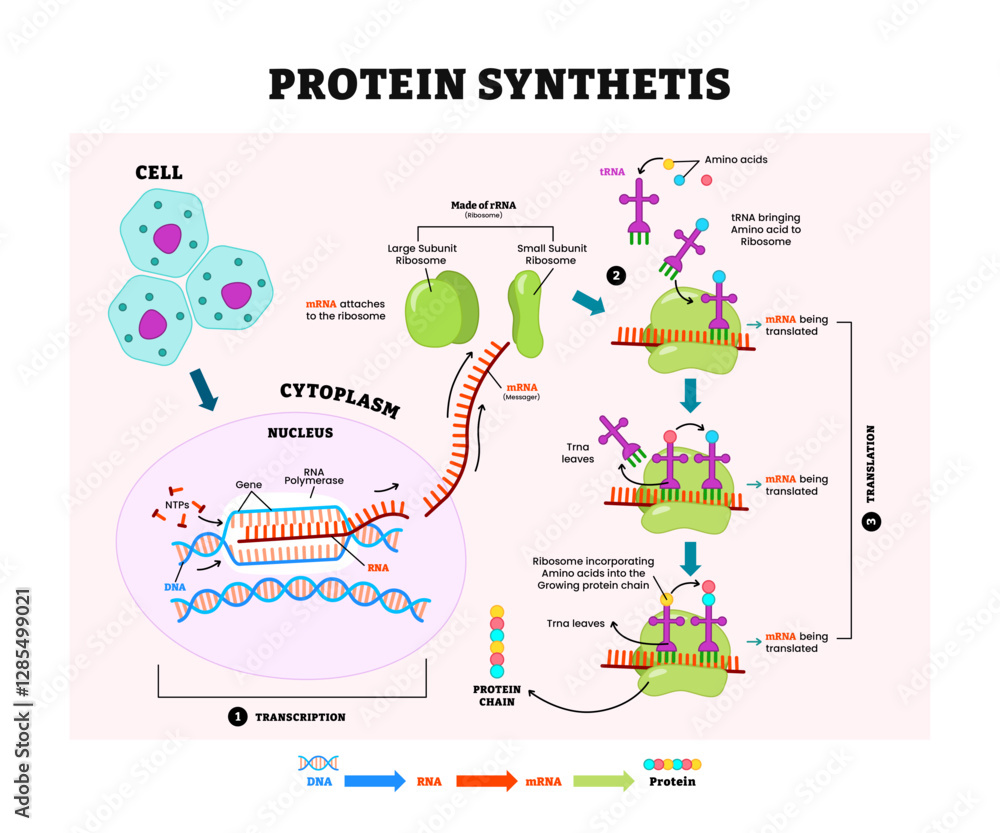
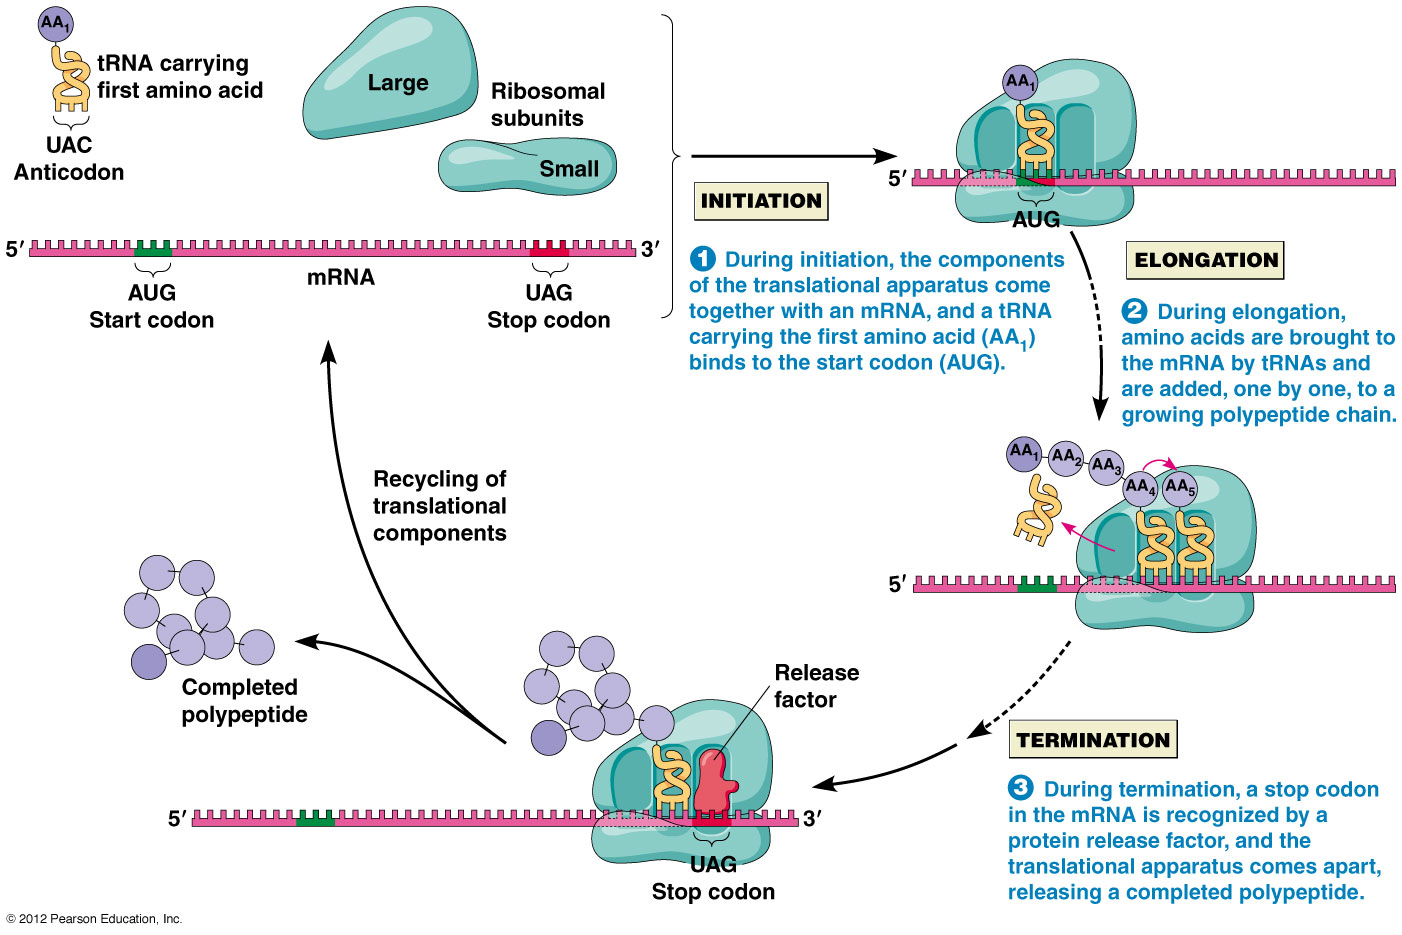
Protein synthesis through transcription and translation is one of those topics everyone learns in undergrad biology but almost nobody actually does by hand afterward. I've spent years running cell-free expression systems and in vitro translation assays, and the gap between textbook diagrams and what actually happens in a tube is significant enough that you'll waste reagents and weeks of your life if you don't understand the mechanics. Let's start with what's actually happening. DNA gets transcribed into mRNA, then ribosomes read that mRNA and assemble amino acids into a polypeptide chain. That's the textbook version. In practice, if you're doing this recombinantly, you're working with an in vitro transcription/translation kit like the PURE system or a commercial coupled transcription-translation mix, and the real work is in the design phase. The codon matters more than most people account for. E. coli-based expression systems prefer certain codons over others. If your gene has a stretch of rare codons — especially AGA, AGG, AUA, CUA, or CCC — the ribosome stalls, the protein misfolds, or you get truncated products. I spent three months troubleshooting a protein that wouldn't express past 15 kilodaltons before I realized the gene had six arginine codons in a row that the standard DH5 strain doesn't carry tRNAs for in sufficient quantity. Switching to a Rosetta (DE3) strain, which supplies supplementary tRNAs for those rare codons, solved it immediately. No optimization of temperature, no different tags, just the right strain.
Designing Your Construct Before You Touch the Bench
This is where most people fail. They order a gene, plug it into a vector, and hope. The success rate on that approach is roughly 30 percent if you're lucky. You need to think about the sequence first. Codon optimization is standard now, but don't blindly run your sequence through an online optimizer and accept the default output. Over-optimizing can create secondary structures in the mRNA that slow or block ribosome progression. I've seen optimized sequences with GC content over 70 percent in the 5' untranslated region that essentially formed hairpins strong enough to prevent ribosome binding. The fix was reducing the GC content in the first 50 codons while keeping the rest optimized. Expression went from undetectable to high yield in a single round of redesign. Also, the ribosome binding site needs to be appropriate for your system. The Shine-Dalgarno sequence should be positioned about 5-10 nucleotides upstream of the start codon in bacterial systems. Too close and the ribosome can't properly engage. Too far and translation initiation drops off significantly. I once saw a cloned construct where the RBS was shifted by 8 nucleotides due to a restriction site artifact, and the expression level was essentially zero. Re-cloning with the correct spacing fixed it.
The Actual Reaction Setup
Whether you're using a commercial kit or the PURE (Protein synthesis Using Recombinant Elements) system, the reaction is deceptively simple. You mix template, ribosomes, amino acids, energy regeneration components, and the appropriate buffer, then incubate. The complexity is in the variables you need to control. The mRNA template quality is critical. If you're doing in vitro transcription, the DNA template needs a proper T7 promoter upstream of your gene. Run your transcription product on a denaturing gel to check for full-length mRNA. Degraded templates produce truncated proteins or nothing at all. I typically get 50-100 nanograms per microliter of clean, full-length mRNA from a 20-micro liter transcription reaction, which is plenty for multiple translation reactions. For the translation step itself, the standard incubation is 37 degrees Celsius for 1-2 hours for E. coli-based systems. Lower temperatures like 25-30 degrees can help with solubility for difficult proteins, especially membrane proteins or ones prone to aggregation. If your protein isn't soluble after a standard 37-degree reaction, try a temperature gradient across multiple reactions. It's a cheap way to identify the right condition before committing to purification.
Get the Full Details

Common Pitfalls That Waste Time
Coupled transcription-translation systems are convenient but they have a fundamental limitation: the transcript and the protein are both being made in the same tube, and the mRNA is only present during active transcription. This means your protein yield is capped by the stability of the mRNA and the amount of template you can reasonably add. For high-yield applications, a two-step process — in vitro transcription first, then purification of the mRNA, followed by a separate translation reaction — will give you 3-5 times more protein. It takes longer but the yield difference is substantial. Another issue that catches people off guard: protease activity. Even in cell-free systems, residual proteases from the extract can degrade your protein over time. If you're expressing a protein that's known to be protease-sensitive, adding a protease inhibitor cocktail to the translation reaction or switching to a pure system like PURE, which has no endogenous proteases, will make a noticeable difference. PURE systems are more expensive per reaction but they're reproducibly cleaner, and you can customize exactly what's in the reaction, which matters if you're trying to incorporate non-canonical amino acids or do site-specific labeling. I also want to flag something that isn't obvious from the protocols: the concentration of magnesium. Magnesium is a cofactor for ribosome function, but the optimal concentration is a narrow window. Too low and ribosomes fall apart. Too high and you get non-specific translation and aggregation. Most kits recommend 5-10 millimolar MgCl2, but I've found that titrating down to 3-4 millimolar improved solubility for several difficult proteins without sacrificing yield significantly. It's worth running a small matrix with different magnesium concentrations if your protein tends to precipitate.
Checking Your Results
After the reaction, you need to verify what you actually made. Run a small aliquot on an SDS-PAGE gel alongside a molecular weight marker. If you have a tag, you can also do a quick Western blot. ThePURE system gives you a relatively clean background, but E. coli extracts will show you a smear of endogenous proteins alongside your product, so distinguishing your band might require a tag-based detection method. If your protein doesn't appear on the gel, don't assume it didn't express. Run a liquid scintillation count if you have access to it — adding a tiny amount of radiolabeled methionine to the reaction and spotting it on a filter tells you definitively whether translation occurred, even if the protein is degraded immediately after synthesis. I've used this trick more than once to catch cases where the protein was being made but was rapidly degraded by contaminating proteases, which a gel alone would have missed because there'd be no visible band.
When Cell-Free Isn't the Answer
There are honest limitations to in vitro translation. If you need large amounts of protein — grams rather than milligrams — or if your protein requires complex post-translational modifications that a bacterial system can't provide, you're better off moving to a cultured cell system. Mammalian expression in HEK293 or CHO cells handles glycosylation and other modifications that bacterial cell-free systems simply can't replicate. The tradeoff is time and cost. A mammalian expression project takes weeks to set up and optimize compared to hours for a cell-free reaction. But for quick tests, toxic proteins, membrane proteins, and early-stage construct validation, cell-free translation is still the fastest route from sequence to protein. The initial learning curve is steep, and the reagents aren't cheap, but once you have a working protocol for a particular protein, the turnaround is unmatched. You can go from a gene sequence to a microgram of protein in a single day, and that speed is why I keep coming back to it despite the frustrations.