Working With Trigonal Bipyramidal Geometry in Practice
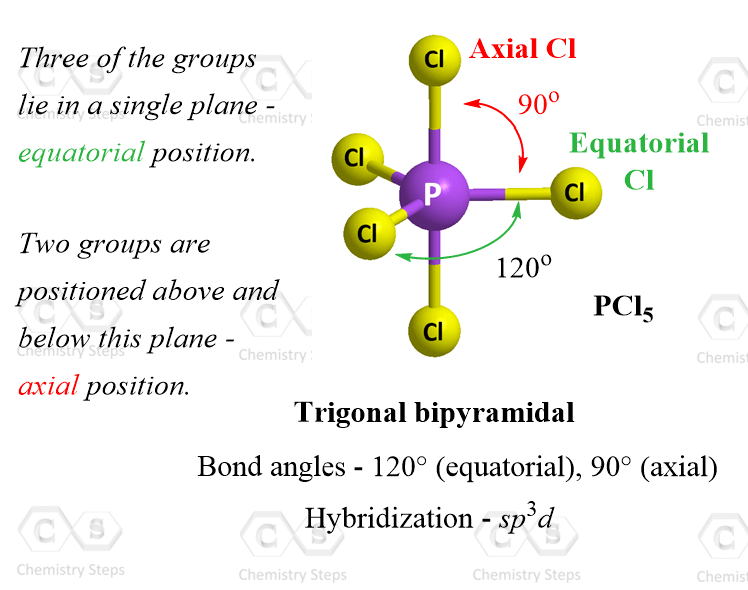
The Trigonal Bipyramidal Bond Angle comes in two forms: 90 degrees between axial and equatorial positions, and 120 degrees between equatorial positions. There is also a 180-degree angle between the two axial atoms. That is the basic textbook answer. The real problem starts when you try to apply this to actual molecules and their distortions. VSEPR theory predicts these angles for five-coordinate species, but the actual angles shift depending on electronegativity, lone pairs, and steric bulk. Take PF5 as the clean textbook example. The angles sit right at 90 and 120. Now look at something like SF4, which has one lone pair. The lone pair goes equatorial because it needs more room, and suddenly your axial bonds bend away from perfect 90-degree angles. The actual F(axial)-S-F(equatorial) angle drops to around 86-87 degrees. This is not a small correction. It matters if you are modeling this for anything beyond an undergrad homework problem. I spent a semester troubleshooting crystallographic data for a niobium complex with trigonal bipyramidal geometry, and the reported angles from the refinement were sitting at 84 degrees and 126 degrees instead of the expected 90 and 120. Turned out the crystal had significant thermal motion along the axial direction, and the displacement ellipsoids were stretching enough to make the equatorial ligands buckle. The fix was to restrain the refinement with geometric constraints rather than letting the angles float freely, which brought the model back into chemical reality without fabricating data.
Why Beginners Get This Wrong Every Time
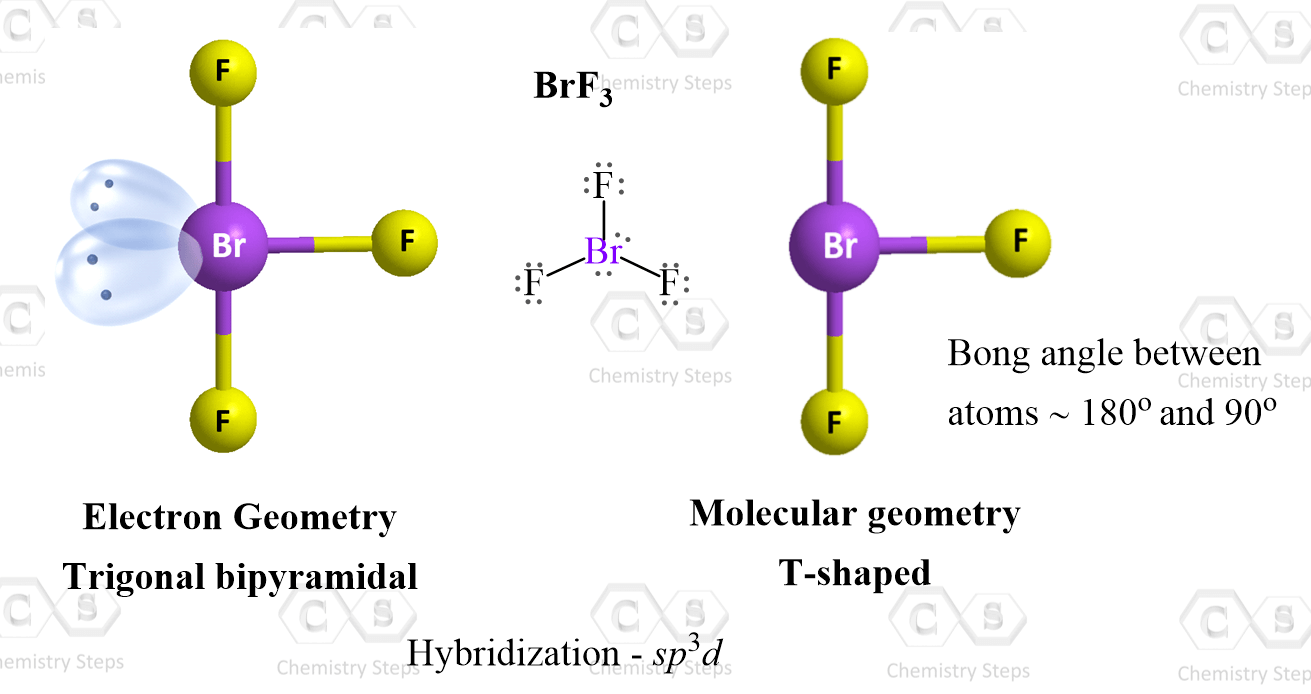
The biggest mistake people make is treating the Trigonal Bipyramidal Bond Angle as fixed across all five-coordinate molecules. It is not. Lone pairs on the central atom compress adjacent angles. Bulky substituents in equatorial positions push axial ligands closer together, sometimes dropping the axial-central-axial angle below 180 degrees. The Berry pseudorotation mechanism, where axial and equatorial ligands exchange positions through a square pyramidal intermediate, also means that in solution the angles are time-averaged for many fluxional molecules. If you are doing NMR interpretation, assuming static 90-degree and 120-degree angles will give you the wrong coupling patterns. Another thing nobody warns you about: computational chemistry software defaults often assume idealized geometries when generating starting coordinates. If you run an optimization on a molecule like ClF3 or a transition metal complex with d8 or d10 configuration and skip checking the initial guess, you might waste 20 minutes watching the optimizer fight against itself before converging to a structure that looks correct but actually started from a bad local minimum. I learned this the hard way with a bromine pentafluoride model. The initial guess had the axial bonds slightly bent, and the optimizer just propagated that error instead of correcting it. A quick manual adjustment of the starting angles to exactly 90 and 120 before launching the calculation cut the convergence time from over an hour down to under six minutes on a standard desktop machine. The main limitation with VSEPR predictions for Trigonal Bipyramidal Bond Angle is that it breaks down for heavier main group elements and most transition metal complexes. The theory assumes lone pairs occupy equatorial positions to minimize repulsion, but for metals with significant d-orbital participation, crystal field effects and ligand field stabilization energies dominate the geometry. In those cases you need DFT calculations or experimental data, not a worksheet prediction. Even then, basis set choice matters a lot. Using a minimal basis set on a five-coordinate zinc complex gave me bond angles that were off by four degrees compared to experiment, which sounds small until you are trying to match spectroscopic data.
For quick reference, here are typical angle ranges you should expect depending on the system: perfect five-coordinate with no lone pairs gives 90 and 120 degrees. One equatorial lone pair, like in SF4, shifts axial-equatorial angles down to roughly 86-88 degrees. Two lone pairs in equatorial positions, as in ClF3, compress things further with angles in the 85 to 105 degree range depending on which positions you are measuring. Four-coordinate square pyramidal intermediates during pseudorotation temporarily distort everything toward 90 degrees across the board.
Get the Full Details