What actually happens when you force a lone pair into a trigonal planar scaffold
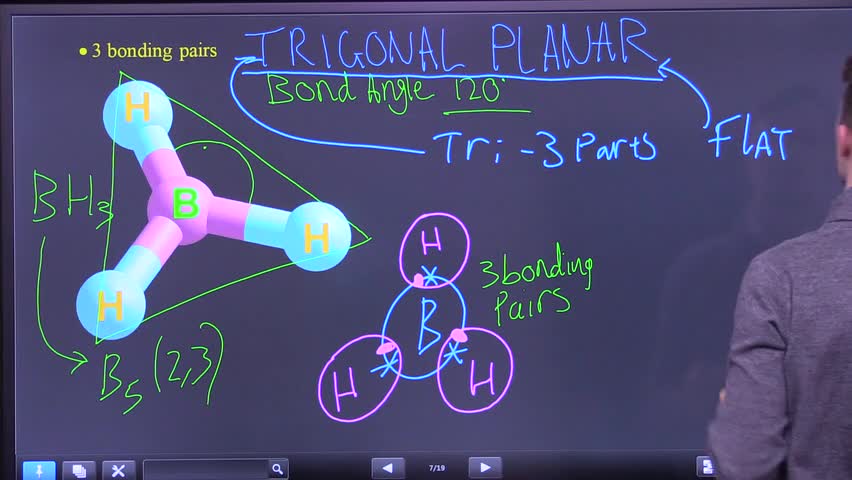
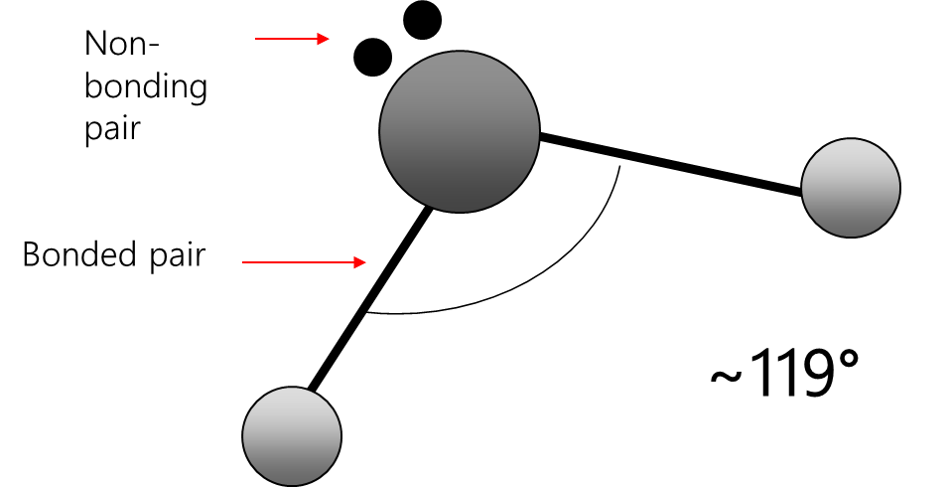
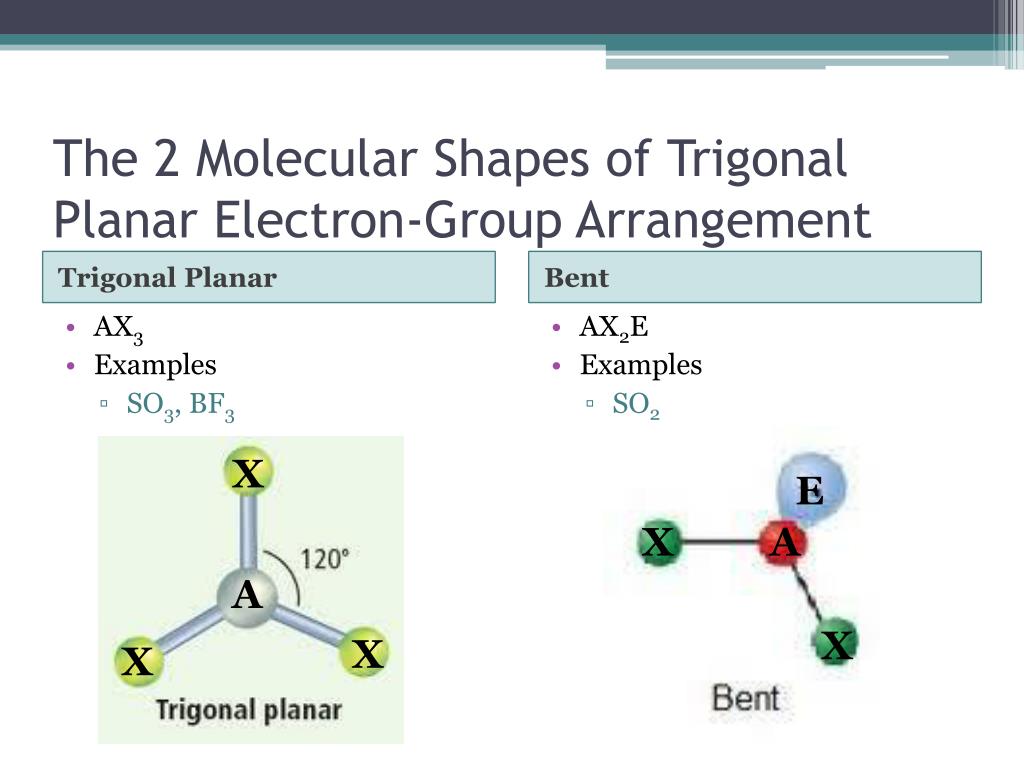
The basic textbook answer is straightforward. A trigonal planar center with three bonding domains gives you 120 degrees between each bond. Add a lone pair and one domain disappears from the bonding picture, the geometry bends, and the remaining bond angle shrinks somewhere below 120. That is VSEPR in its simplest form. But the real number depends on which molecule you are looking at, what atoms are attached, and how the lone pair is distributed across the electronic structure. The Trigonal Planar Bent Bond Angle is not a fixed constant. It is a range that shifts based on electronegativity, hybridization character, and the specific lone pair localization. I spend most of my time running geometry optimizations in Gaussian and ORCA, so I will describe the workflow from that angle. You build the initial structure with the atom in question roughly at 120 degrees, place the lone pair visually on the opposite side of the missing bond, and then run a full optimization at a reasonable level of theory. For most main-group molecules, B3LYP/def2-SVP gets you into the right neighborhood in about ten minutes on a standard workstation. If you need quantitative accuracy, switch to wB97X-D/def2-TZVP and expect the run to take closer to forty-five minutes for a small molecule. The output gives you the optimized Cartesian coordinates. From there, you calculate the angle between the two bonding vectors originating from the central atom. Most people use the built-in analysis tools or just pull the coordinates into a free program like Avogadro. The angle it reports is your answer. For sulfur dioxide, this comes out around 119 degrees. For the nitrite ion, it is closer to 115 degrees. These numbers are not arbitrary. They reflect the actual repulsion between the lone pair and the bonding electron domains after the geometry has relaxed.
Here is where things get messy in a way the textbooks do not warn you about. I was modeling a chlorinated sulfite intermediate last year, something with a central sulfur bonded to two oxygens and one chlorine, with a lone pair sitting on the sulfur. The initial guess with 120-degree angles converged to a bizarre structure where the O-S-O angle collapsed to roughly 103 degrees and the molecule essentially folded into a nearly perpendicular arrangement. The optimizer was pulling the heavy chlorine atom into the lone pair region because the initial geometry had placed it directly opposite the lone pair. The fix was simple once I understood what was happening. I reoriented the initial structure so the chlorine was positioned roughly in the plane of the two oxygens rather than opposite the lone pair, and I constrained the O-S-O angle to 118 degrees for the first few optimization steps before releasing the constraints. This took the calculation from failing to converge to finishing in under twenty minutes. The final angle settled at 116.4 degrees, which matched the expected range for a bent trigonal planar center with a moderately electronegative substituent. The general principle here is that your starting geometry matters more than people admit. If you seed the optimization with a perfectly symmetric 120-degree arrangement and the molecule has a strong asymmetry in its substituents, the optimizer may follow an unintended path. Constraining the angle during the early stages or manually adjusting the initial coordinates to reflect where the lone pair actually points is usually faster than rerunning the job three times and hoping for a sensible result.
Counter-intuitive points that come up repeatedly
The first thing beginners miss is that the lone pair does not always behave like a visible domain in the way VSEPR draws it. In molecules with significant pi-bonding character, the lone pair can be delocalized into nearby orbitals, which reduces its effective repulsive volume. This means the bond angle can end up larger than you would predict from a simple steric argument. Sulfur dioxide is a classic example. The simple VSEPR picture says one lone pair should compress the angle to somewhere around 117 degrees. The experimental value is 119.5 degrees because the lone pair participates in the pi system and is not purely localized on the sulfur. If you ignore this and model the molecule with a localized lone pair in a simple force field, your predicted angle will be wrong by several degrees. The second point is about how different computational methods handle this. Hybrid functionals like B3LYP tend to over-stabilize certain lone pair configurations, which can artificially inflate or deflate the bond angle depending on the system. Range-separated functionals like wB97X-D or M06-2X generally give more consistent results for bent trigonal planar geometries, but they are more expensive. The trade-off is real. For a quick screening of fifty similar molecules, B3LYP/def2-SVP is fine. For a single structure where you need the angle to within one degree, the extra computational cost of a range-separated functional is worth it. There is also a practical limitation that nobody emphasizes enough. When you have a very bulky substituent on the central atom, steric effects can override electronic ones. The bond angle may open up beyond what the lone pair repulsion alone would predict because the substituents are physically pushing against each other. I ran into this with a tin-based compound where the expected angle based on VSEPR was around 112 degrees. The actual optimized angle was 121 degrees because the two organic ligands on the tin were large enough that they could not fit at the narrower angle. Electronic effects and steric effects are not independent, and treating them as separate terms in your head is a shortcut that breaks down in real calculations.
Get the Full Details

If you are working with systems where the central atom is in a higher period, like phosphorus or sulfur, you should also be aware that d-orbital contributions are sometimes invoked in older literature to explain bond angle deviations. Modern computational chemistry largely treats these as artifacts of basis set choice rather than real physical effects. Using a well-balanced basis set like def2-TZVP or cc-pVTZ without adding diffuse functions unless your system actually has anions or Rydberg states will give you cleaner results than older protocols that relied on d-functions to compensate for poor basis sets. The takeaway is not that the concept is unreliable. It is that the Trigonal Planar Bent Bond Angle is a property that emerges from the full electronic structure, not a geometric rule you can memorize and apply without checking. The VSEPR model is useful for getting the right general direction. Computational chemistry or diffraction data is necessary for the actual number. And your initial guess for the geometry determines whether the calculation finishes in twenty minutes or you spend an hour debugging a convergence failure.