Reading Bond Angles and Electron Domains Without Losing Your Mind
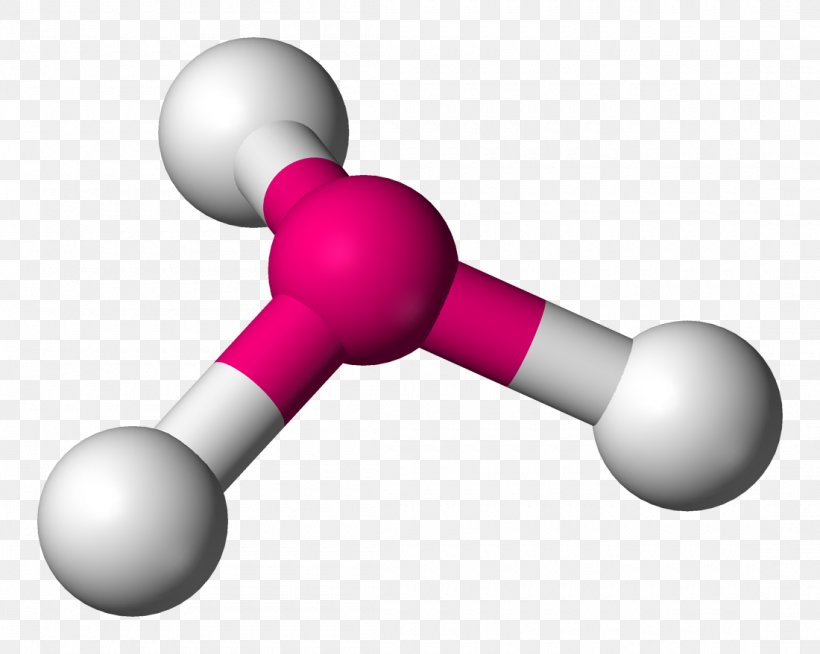
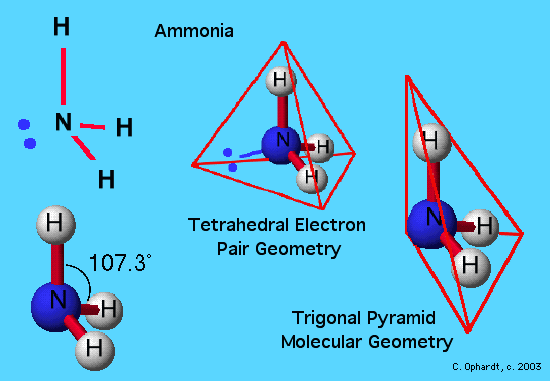
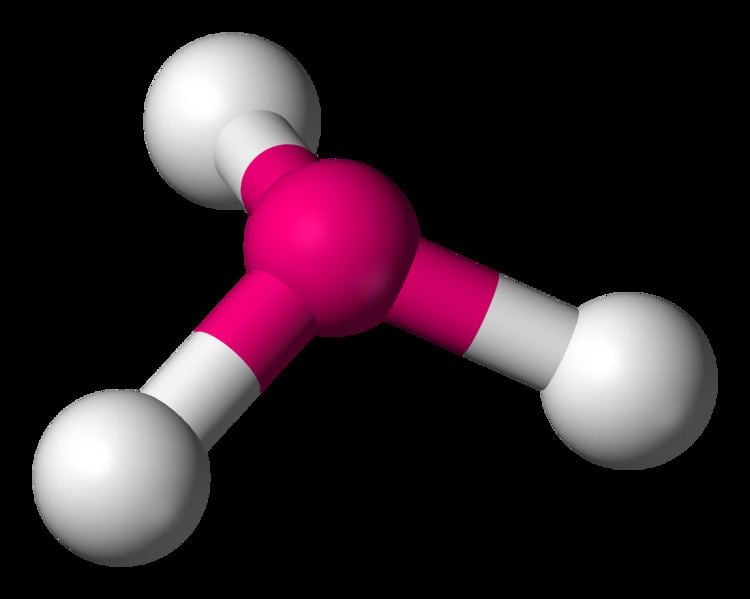
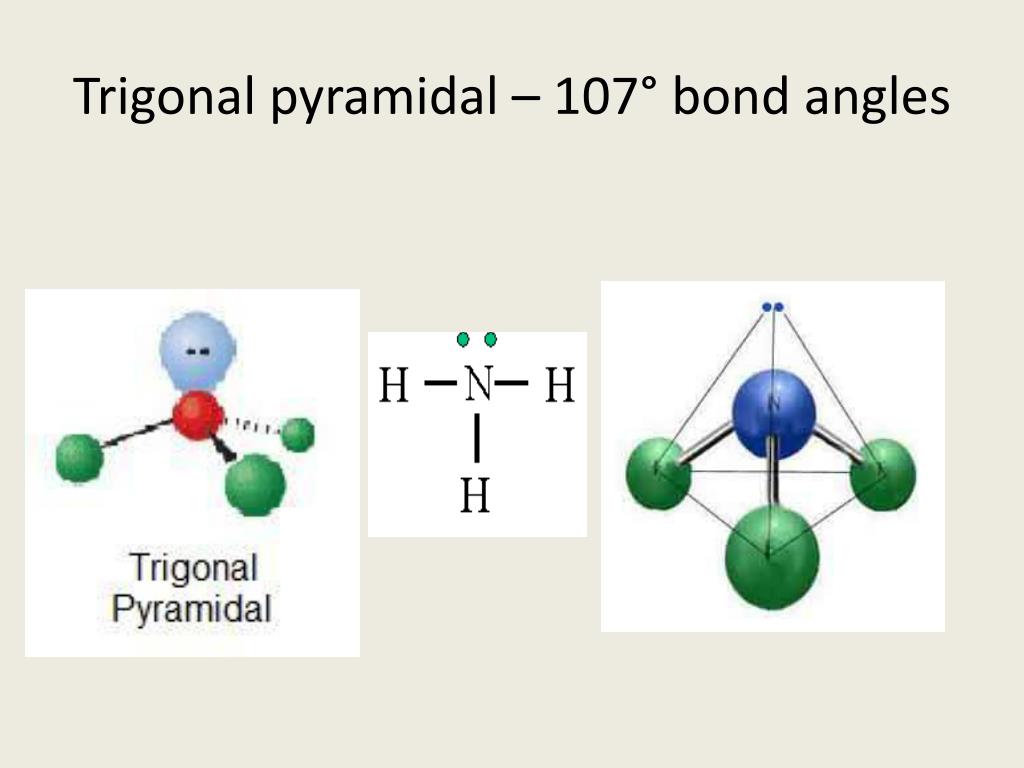
Most people learn molecular geometry by memorizing tables and moving on. That approach works until you're looking at something that doesn't fit the textbook picture. I've spent years building 3D models and checking computational outputs for molecules, and the first thing I always verify is the electron domain count before I even think about naming the shape. Trigonal Pyramidal Molecular Geometry comes up when you have four electron domains around a central atom but only three of them are bonding pairs. That lone pair does the heavy lifting here, pushing the bonds down into a pyramid shape. The ideal angle from tetrahedral is 109.5 degrees, but the lone pair takes up more space and squeezes that down to somewhere around 107 degrees in a standard case like ammonia.
How to Identify Trigonal Pyramidal Molecular Geometry in Practice
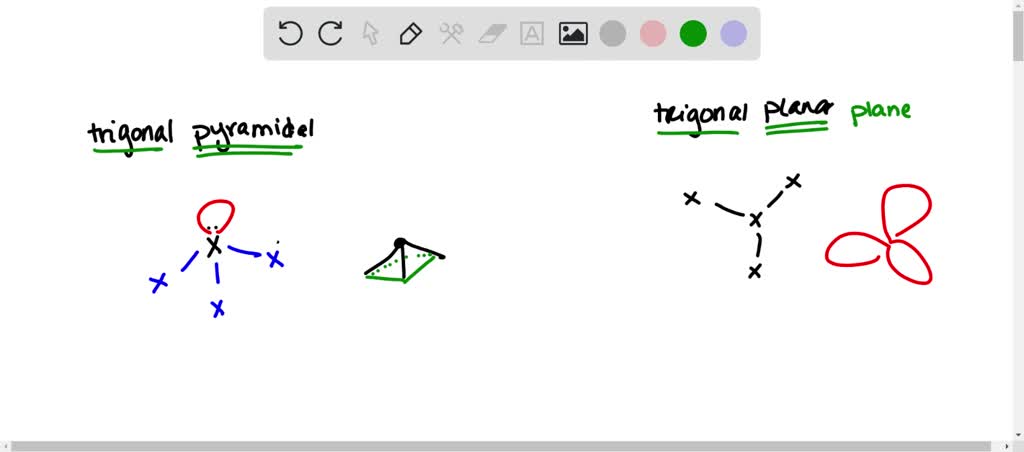
Start with the Lewis structure. Count every bond and every lone pair on the central atom. Four domains total, three bonds, one lone pair, and you're looking at trigonal pyramidal. Don't skip this step because VSEPR theory breaks down fast if you miscount. I've seen students confuse the geometry with T-shaped, which also has three bonds but comes from five electron domains with two lone pairs. The difference matters for dipole moments and reactivity patterns.
When I first started doing this work regularly, I ran into a problem with phosphorus chlorides. PCl3 is the classic textbook example that shows up everywhere, but the actual experimental bond angle is closer to 100 degrees, not the 107 you'd expect from simple VSEPR prediction. This threw off my models for weeks because I kept using the standard angle value and the predicted properties didn't match the literature data. The workaround was straightforward once I understood what was happening. Heavier central atoms like phosphorus and sulfur have more diffuse orbitals, which means the lone pair sits further from the nucleus and exerts less repulsive force on the bonding pairs. The bonds aren't being squeezed as hard as nitrogen's lone pair squeezes ammonia's bonds. I started pulling experimental angles from the CRC Handbook or the NIST WebBook instead of relying on the VSEPR estimate, and that cut my model iteration time from several hours down to maybe twenty minutes per compound.
The Lone Pair Effect and What It Actually Does
The lone pair isn't just a passive feature in these molecules. It actively distorts the geometry and creates a significant dipole moment pointing away from the bonding atoms. In ammonia, the dipole is about 1.47 Debye. That number matters for solvation behavior, hydrogen bonding strength, and how the molecule interacts with surfaces. You can't ignore it when you're predicting physical properties.
Here's something most introductory courses don't emphasize enough: the lone pair's orientation determines whether a molecule can act as a nucleophile or a base. Ammonia's lone pair points upward, ready to donate. That's why it's a good nucleophile in substitution reactions. Compare that to nitrogen trifluoride, NF3, which also has trigonal pyramidal geometry but is a very poor nucleophile. The fluorine atoms pull electron density away from the nitrogen, making the lone pair much less available. The geometry is the same. The chemistry is completely different. If you're only looking at shape without considering electronegativity effects, you'll mispredict reactivity every time.
Common mistakes I see people make with this geometry: Assuming all trigonal pyramidal molecules have the same bond angle. They don't. NH3 is 107.8 degrees, PCl3 is about 100 degrees, and Cl3PO is roughly 107 degrees too but for different electronic reasons. The central atom, the substituent electronegativities, and the second-row versus third-row distinction all shift the angle in different directions.
Another thing that catches people out is confusing the electron geometry with the molecular geometry. The electron geometry for all three-domain-one-lone-pair cases is tetrahedral. The molecular geometry is trigonal pyramidal. These are different descriptors talking about different things. The electron geometry includes the lone pair in its spatial arrangement. The molecular geometry describes only the positions of the atoms. When someone asks what the shape of ammonia is, the answer depends on whether they mean the arrangement of electron domains or the arrangement of atoms.
Computational and Practical Considerations
If you're running quantum chemistry calculations on a trigonal pyramidal molecule, the choice of functional and basis set matters more than you might think. HF/3-21G tends to overestimate the bond angle in ammonia by about 3 to 4 degrees because it doesn't handle electron correlation well. B3LYP/6-311+G(d,p) gets much closer to the experimental value, usually within 1 degree. If you need high accuracy for conformational analysis or reaction path modeling, you're looking at CCSD(T)/aug-cc-pVTZ, and those calculations run significantly longer on larger systems.
The geometry optimization itself can get stuck in local minima if you start from a bad guess structure. I once optimized a phosphoryl compound and the algorithm kept converging to a planar structure instead of the pyramidal one because the initial coordinates were too symmetric. The fix was to displace the central atom slightly off the plane of the three substituents before running the optimization. One point of displacement in the z-direction was enough to guide it to the correct minimum. Saved me about an hour of debugging.
When Trigonal Pyramidal Geometry Doesn't Tell You Everything
This is where I have to be honest about the limitations of the model. VSEPR and the simple trigonal pyramidal classification work well for stable, isolated molecules in the gas phase. They break down when you're dealing with transition states, radical species, or molecules in strong electric fields. A nitrite ion or certain intermediates in organometallic catalysis might appear trigonal pyramidal in a static snapshot but behave very differently in solution or on a catalyst surface. The lone pair that's so important in gas-phase chemistry can get stabilized or delocalized in ways that change everything.
For quick classification and initial property estimation, this framework is fine. For anything that requires quantitative predictions, you need computational chemistry or experimental data. There's no shortcut around that.
Gallery Trigonal Pyramidal Molecular Geometry
Trigonal Pyramidal Molecular Geometry - Chemistry LibreTexts
Trigonal pyramidal molecular geometry - Alchetron, the free social ...
Trigonal Pyramidal Molecular Geometry - Free Transparent PNG Download ...
Trigonal Pyramidal Shape | Geometry of Molecules – MOJQI
SOLVED: What is the difference between trigonal pyramidal molecular ...