Understanding Ring Strain Before You Waste a Week on a Reaction That Makes No Sense
I spent three days trying to figure out why a Suzuki coupling on a cyclopropyl boronate was giving me 12% yield instead of the 80% I was expecting. Turns out the answer was sitting right in front of me the whole time: I hadn't properly accounted for the combined angle and torsional strain destabilizing the sp2-hybridized carbon in the transition state. The aryl group was trying to adopt a geometry that the ring literally wouldn't allow. This is exactly why you need to actually understand what is going on with Types Of Strain Organic Chemistry before you start drawing mechanisms on a whiteboard. Most students memorize definitions and then move on without really internalizing how these strains compound in real molecules. Let me walk through what matters practically.
Types Of Strain Organic Chemistry
Angle Strain: The One That Kills Cyclopropane Derivatives
Angle strain happens when bond angles are forced away from their ideal hybridization geometry. For sp3 carbons the ideal is 109.5 degrees. In cyclopropane the internal angles are 60 degrees. That is a deviation of roughly 48.5 degrees per angle, and with three angles in the ring you are looking at around 146 degrees of total angular distortion. The Baeyer strain model from 1885 predicted this accurately even though he assumed all rings were planar, which they are not. The practical consequence is that cyclopropane derivatives are dramatically more reactive than you might expect. The C-C bonds have significant pi character because the orbitals cannot align head-on. This is called bent bonding or banana bonding. The bonds are weaker and more accessible to electrophilic attack. I have seen people use cyclopropyl ketones as masked vinyl anions in conjugate additions, and it works precisely because the ring strain energy (about 27.5 kcal/mol) provides the thermodynamic driving force for ring opening. One thing beginners miss: cyclobutane is not as strained as cyclopropane but it is not dramatically less strained either. The angle strain is only about 5 kcal/mol lower. Cyclobutane adopts a puckered conformation to reduce torsional strain, but this means the bond angles open to about 90 degrees, still far from ideal. The total strain energy is around 26.3 kcal/mol. The takeaway is that going from three to four members does not give you the relief you would naively expect.
Torsional Strain: Why Your NMR Looks Wrong
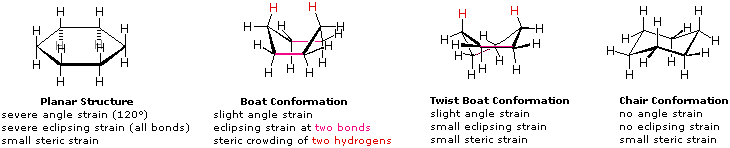
Torsional strain is the resistance to eclipsing interactions between substituents on adjacent atoms. In ethane the staggered conformation is about 2.8 kcal/mol more stable than the eclipsed form. This might seem small until you realize that cyclobutane has four eclipsing interactions locked in by the ring, and cyclopropane has six. Here is where it gets interesting. Cyclohexane avoids both angle and torsional strain almost entirely by adopting the chair conformation. All bond angles are near 109.5 degrees and all hydrogens are staggered. The strain energy is essentially zero. But put a methyl group in the axial position and you immediately introduce 1,3-diaxial interactions. Each axial methyl costs about 1.74 kcal/mol due to gauche butane-like interactions with the hydrogens on C3 and C5. A tert-butyl group in the axial position costs roughly 5 kcal/mol, which is why tert-butylcyclohexane exists almost exclusively with the bulky group equatorial. I once had a student try to predict the product distribution of an elimination reaction on a substituted cyclohexane and got it completely wrong because they drew the ring flat on paper. The actual conformer that reacts has the leaving group axial and the beta-hydrogen anti-periplanar to it. In a chair, only certain conformations satisfy the stereoelectronic requirement for E2 elimination. Drawing flat hexagons hides this entirely.
Get the Full Details

Steric Strain: The One People Ignore Until It Ruins Their Synthesis
Steric strain, sometimes called van der Waals strain, arises when atoms are forced closer than their combined van der Waals radii allow. This is distinct from torsional strain because it involves non-bonded atoms rather than eclipsing bonds. In practice the two often occur together and it is hard to separate them energetically. Neo-pentane is the classic example. The central carbon is bonded to four methyl groups that want to occupy the same space. The molecule relieves this by slightly bending the C-C bonds outward, which reintroduces angle strain as well. The total strain is about 7 kcal/mol. In larger molecules this shows up as A(1,3) strain, or allylic strain, where a substituent on an sp2 carbon clashes with a group two atoms away along the chain. This is relevant for controlling E versus Z selectivity in eliminations and for explaining why some Grignard reagents fail to form. A real problem I ran into: trying to run a Dauben-Emmons olefination on a substrate with a quaternary center adjacent to the reaction site. The yield was terrible because the bulky group created so much steric strain in the oxaphosphetane intermediate that it preferentially decomposed back to starting material instead of collapsing to the alkene. Switching to a Wittig variant with a stabilised ylide and running it at higher dilution (0.02 M instead of 0.1 M) improved the yield from 15% to 62%. The higher dilution reduced intermolecular side reactions that were competing with the desired pathway.
Transannular Strain: The Sneaky One in Medium Rings
Transannular strain appears in rings of about eight to eleven members where atoms on opposite sides of the ring are forced into close proximity. This was predicted by Hausinger and Pitzer and later confirmed experimentally. Cyclooctane has a strain energy of about 9.7 kcal/mol, which is unexpectedly high given that cyclohexane is nearly strain-free and cyclodecane is not much worse. The boat-chair conformation of cyclooctane brings hydrogens across the ring into van der Waals contact. This is not the same as simple steric strain because the atoms are not directly bonded and the interaction is through space across the ring cavity. Cyclooctatetraene adopts a tub conformation partly to relieve this kind of strain, and doing so also avoids the anti-aromaticity that a planar geometry would impose.
How to Estimate Strain Without Running a Computation
You do not always need DFT calculations to get a reasonable picture. Hydrogenation data gives you heats of hydrogenation per CH2 group. Cyclopropane releases 166 kJ/mol, cyclobutane 106 kJ/mol, cyclopentane 53 kJ/mol, and cyclohexane 33 kJ/mol. The reference value for an unstrained cycloalkane is approximately 33 kJ/mol per CH2, which is what cyclohexane gives. Anything above that baseline is ring strain. Benson group additivity can also give you estimates for more complex systems. You assign strain contributions based on the number and type of interactions present: gauche butane interactions for 1,3-diaxial, eclipsing interactions for torsional strain, and van der Waals repulsion terms for steric overlap. It is not precise but it is fast and usually within a few kcal/mol of experimental values. One counter-intuitive point: smaller is not always more strained. Cyclohexane is less strained than cyclopentane despite having more atoms. Cyclopentane has 5 CH2 groups contributing roughly 5 x 33 = 165 kJ/mol of reference strain but the actual heat of combustion corresponds to about 26 kJ/mol of ring strain. Cyclohexane has essentially zero. The odd-numbered rings cannot adopt the same perfect staggered geometry that even-numbered rings can.

When Your Strain Analysis Fails
The big limitation of classical strain theory is that it treats strain as additive and static. Real molecules are dynamic. At room temperature cyclohexane is rapidly interconverting between chair forms. The strain energy you measure is a Boltzmann-weighted average over all accessible conformers, not a single fixed value. In flexible molecules this matters less. In constrained systems like bridged bicyclics or fused ring systems the picture gets complicated fast. Bredt's rule is a good example. You cannot place a double bond at the bridgehead of a small bicyclic system because the required planar geometry would introduce impossible strain. But this is not a hard rule. In norbornene the bridgehead double bond is forbidden, but in larger bicyclic systems like decalin derivatives it becomes accessible. The rule breaks down because the strain calculation depends on ring size in a way that simple hybridization arguments do not capture. Another failure mode: using strain energy to predict reactivity. A highly strained molecule is not automatically reactive. The reaction must provide a pathway to release that strain and the activation barrier depends on the transition state geometry, not just the ground state. Spiro[2.2]pentane has enormous strain energy but it is kinetically stable at room temperature because there is no low-barrier pathway to release it. It decomposes above 100 C. Strain energy tells you about thermodynamics. It does not tell you about kinetics.
A Quick Reference for Common Values
Cyclopropane ring strain: 27.5 kcal/mol. Mainly angle strain with a torsional component from three eclipsed interactions. Cyclobutane: 26.3 kcal/mol. Puckering reduces torsional strain but angle strain remains significant. Cyclopentane: 6.5 kcal/mol. The envelope conformation mostly relieves torsional strain, leaving a small angle strain penalty. Cyclohexane: 0 kcal/mol. Chair conformation eliminates both major strain types. Cycloheptane: 6.5 kcal/mol. Twist-chair conformation keeps things manageable but not perfect. Cyclooctane: 9.7 kcal/mol. Transannular interactions become noticeable. Cyclodecane: 12.5 kcal/mol. Medium ring strain is dominated by transannular effects and conformational rigidity. Cyclododecane: about 4 kcal/mol. Large enough rings can adopt strain-free conformations again. The numbers shift slightly depending on the source and measurement method, but the trends are consistent. The minimum is at six members. Three and four are bad. Five and seven are moderate. Eight through eleven get worse again. Twelve and above stabilize out.
What to Do When You Are Stuck
If your reaction yield is inexplicable, draw the actual 3D conformer, not the flat structure. Check whether the reacting centers can achieve the geometry required by the mechanism. For SN2 reactions the nucleophile needs a clear backside attack trajectory. For E2 it needs anti-periplanar alignment. For pericyclic reactions it needs the proper orbital overlap in three dimensions. A flat drawing on paper lies to you every time you look at a cyclic or constrained system. When the molecule is complex enough that mental rotation is unreliable, use molecular modeling software. Even a simple MM2 calculation will give you a reasonable geometry and a strain energy estimate in about two minutes. Chem3D, Avogadro, or even the free web-based Molden will do. You do not need a quantum chemistry package for most undergraduate-level strain analysis. For actual research-grade predictions where the strain is subtle or the system is novel, DFT at the B3LYP/6-31G(d) level with geometry optimization and frequency calculation is the standard workhorse. It gives you the correct conformer, the strain energy, and the vibrational frequencies to confirm it is a minimum and not a transition state. The whole process takes about thirty minutes on a modern laptop if you know what you are doing.


