What Variation Actually Means When You're Reading Papers
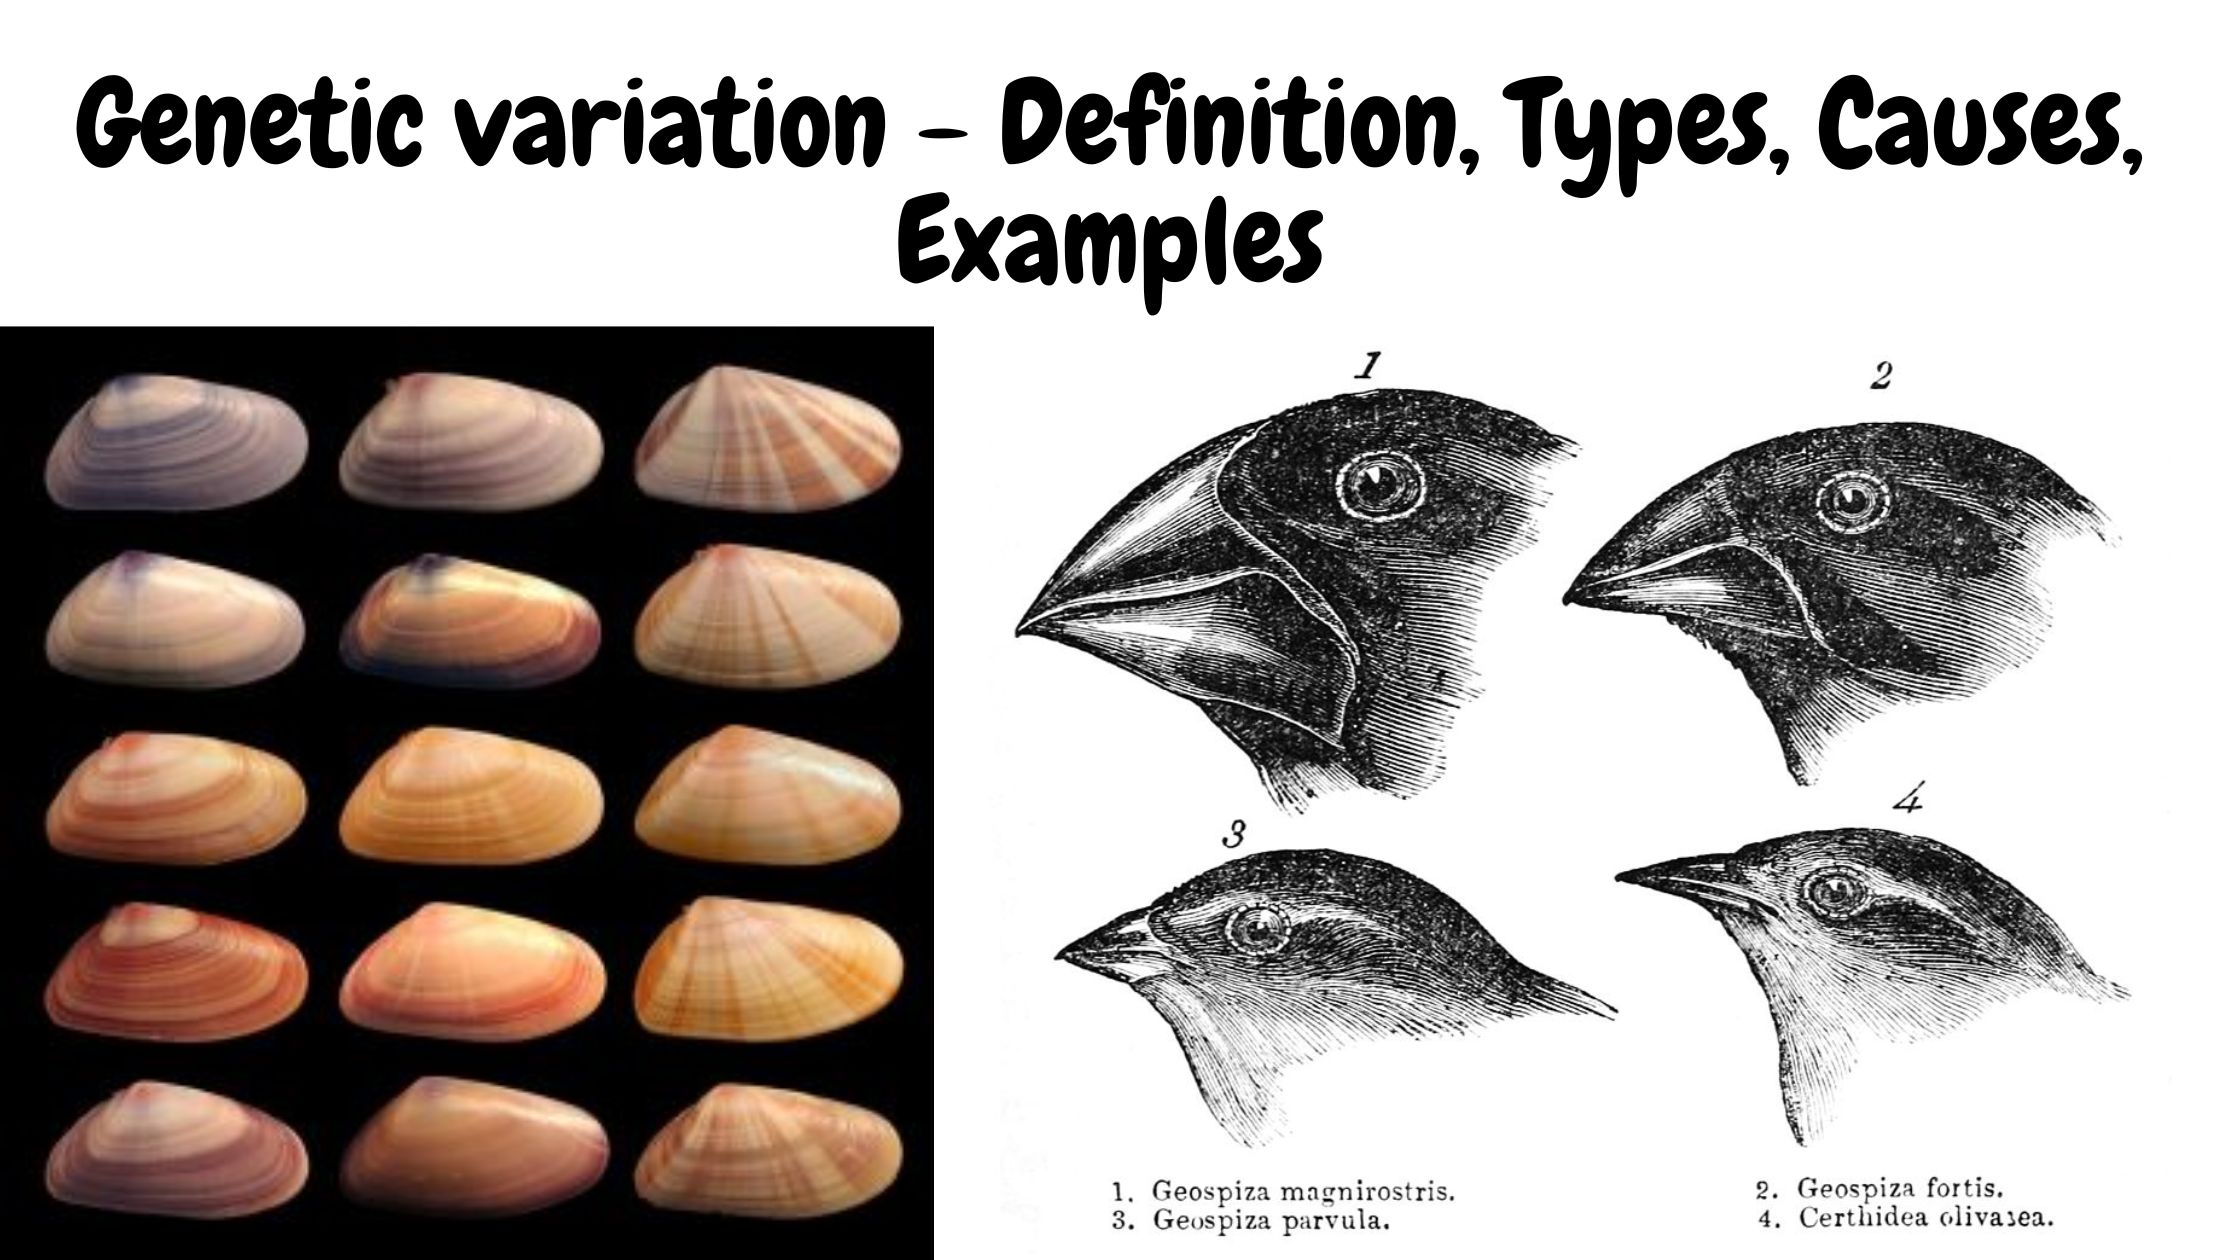
Variation Definition In Biology refers to the differences in traits among individuals within a population or species. That is the textbook version. The real thing is messier. In practice, variation is not just a list of differences — it is the raw material that selection acts on, and how you measure it changes your entire analysis. I spent years working with phenotypic data from wild bird populations, and the first time I really understood variation was when my lab tried to track beak depth across three generations during a drought. We thought we had a clean signal. We did not. The variation we measured was heavily confounded by environmental plasticity, and we nearly published a conclusion that was wrong because we did not separate genetic variation from the environment before running any statistics.
The Variation Definition In Biology You Actually Need
There are two major categories that matter most: phenotypic variation and genetic variation. Phenotypic variation covers observable traits — size, color, behavior, physiological rates. Genetic variation is the underlying differences in DNA sequences among individuals. Both feed into each other, but they are not the same thing, and confusing them is the most common mistake I see in early-career researchers. Beyond that, variation breaks down further. Additive genetic variation is the portion of genetic variance that responds predictably to selection. Dominance variation comes from interactions between alleles at the same locus. Epistatic variation comes from interactions between alleles at different loci. Environmental variation covers everything that is not genetic — temperature, nutrition, social context, developmental noise. The total phenotypic variance is usually expressed as Vp = Vg + Ve + Vgxe, where Vgxe is the genotype-by-environment interaction term. I learned the hard way that ignoring Vgxe is how you get burned. In my fieldwork, we had two populations of the same songbird species living in adjacent valleys with slightly different food availability. The phenotypic variation in body mass looked enormous between them. When we ran a common garden experiment and raised nestlings from both populations in identical conditions, the between-population difference dropped by roughly sixty percent. The remaining variation was largely additive genetic. That experiment took fourteen months and cost about twelve thousand dollars in labor and housing, but it was the only way to know what we were actually looking at.
Heritability is the metric most people reach for when they want to quantify variation, and it is also the metric most people misuse. Broad-sense heritability is H² = Vg/Vp. Narrow-sense heritability is h² = Va/Vp. Narrow-sense is usually more useful for predicting response to selection because it isolates the additive component. The response to selection equation R = h²S is not clever — it is just basic quantitative genetics. S is the selection differential, R is the change in the mean trait value next generation. It works when the assumptions hold, and the assumptions rarely hold cleanly in natural populations. One counter-intuitive point that beginners miss: high phenotypic variation does not automatically mean high evolutionary potential. If most of that variation is environmental or non-additive, selection has almost nothing to work with. I have seen graduate students treat a large standard deviation in a field dataset as proof that a trait could evolve rapidly. It was not. The trait was highly plastic, heritability was close to zero, and the population was not going to adapt to the new condition — it was just going to acclimate until something else killed it. Another pitfall: people conflate variation with diversity. Genetic diversity often refers to measures like heterozygosity or nucleotide diversity (), which are population-level summaries. Variation can be discussed at the individual, population, or species level. The definition shifts slightly depending on context, and in conservation biology, losing variation within a small population is not the same as losing a distinct adaptive lineage elsewhere. Both matter. They are not interchangeable.
Get the Full Details

If you are working with molecular data, variant calling introduces its own variation-related problems. Sequencing errors inflate apparent variation. Alignment artifacts in repetitive regions create false SNPs. I once ran a population genomics project where about eight percent of the reported variants turned out to be mapping errors in a segmentally duplicated region. We caught it only because the allele frequency spectrum looked biologically impossible — too many rare variants at a locus that should have been under purifying selection. The workaround was to mask the region entirely and re-run the diversity statistics, which cut the apparent nucleotide diversity at that locus by roughly half and brought the Tajima's D value back into a plausible range. For practical measurement of phenotypic variation, the standard approach is to collect trait data across a representative sample, calculate variance or standard deviation, and then partition it using ANOVA or mixed models if you have a pedigree or known relatedness. In wild populations without pedigrees, you can still estimate narrow-sense heritability using the animal model, which is a quantitative genetics framework implemented in software like MCMCglmm in R. It accounts for relatedness through a Gaussian graphical model and gives you posterior distributions for variance components rather than point estimates. It is computationally heavier — a typical run with five thousand individuals and ten thousand Markov chains takes roughly six to ten hours on a standard workstation — but it is the most reliable method I have used for non-model organisms. When you do not have controlled breeding data, which is the case for most ecological studies, you should be explicit about what kind of variation you are actually measuring and what you cannot claim. Saying a trait is "highly variable" without stating whether that variability is genetic, environmental, or both is not helpful. It is also not wrong — it is just incomplete, and incomplete claims cost you credibility in peer review faster than anything else.
The biggest bottleneck in variation studies is sample size relative to the number of variance components you want to estimate. If you are trying to partition Vg, Va, Vd (dominance), and Vgxe simultaneously, you need a lot of data and a well-designed breeding or family structure. Most published studies on wild populations estimate only additive genetic variance and environmental variance. That is fine. It is also honest to say that the non-additive components remain unknown. I recommend starting with a clear question: are you trying to quantify how much variation exists, or are you trying to understand its source? The answer determines your sampling design, your statistical model, and the conclusions you are allowed to draw. Mixing those two goals in a single study without separating them analytically is where most problems begin.