A Practical Look at How Molecules Stick Together
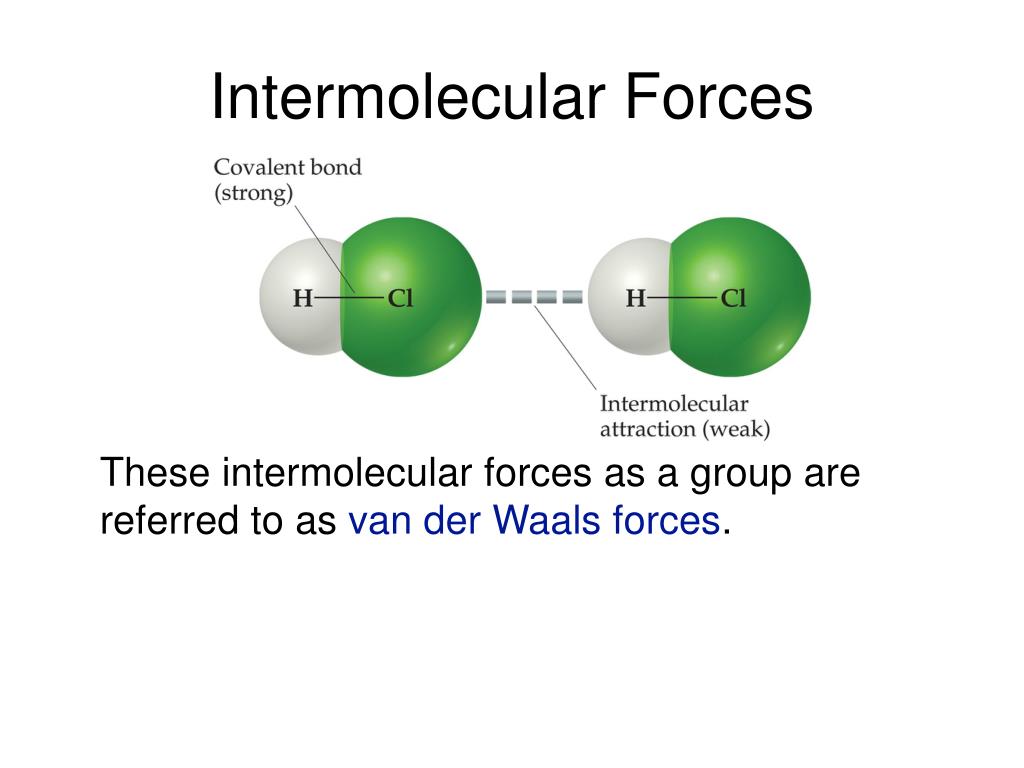
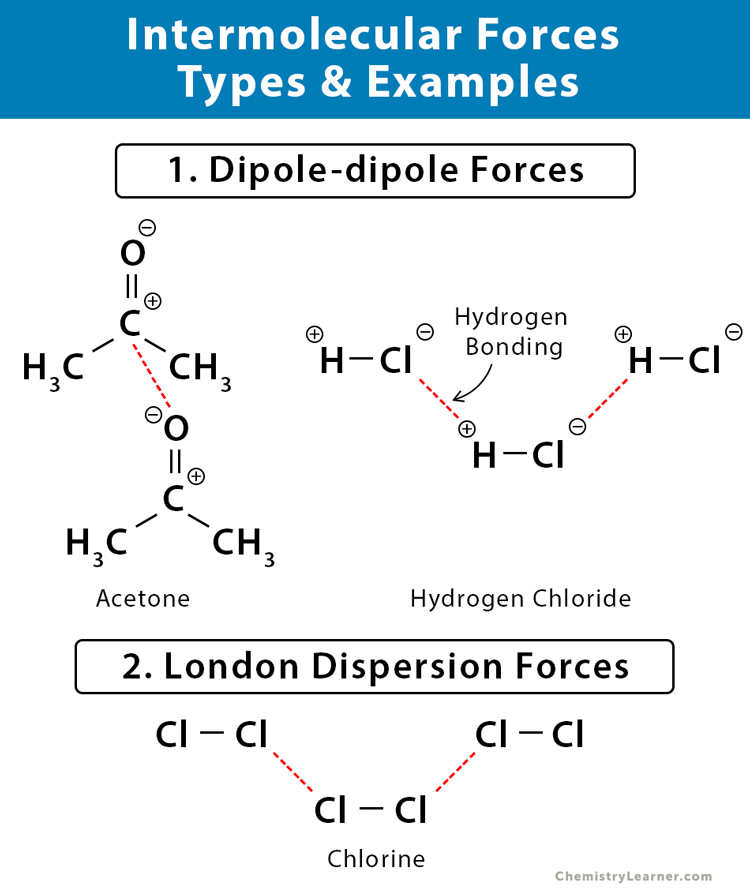
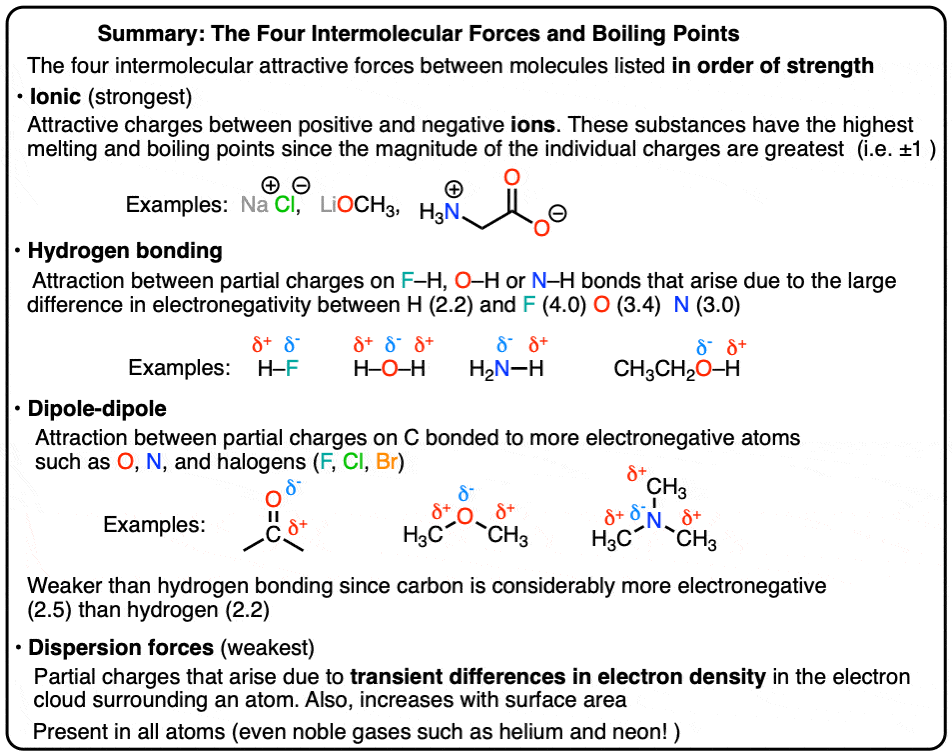
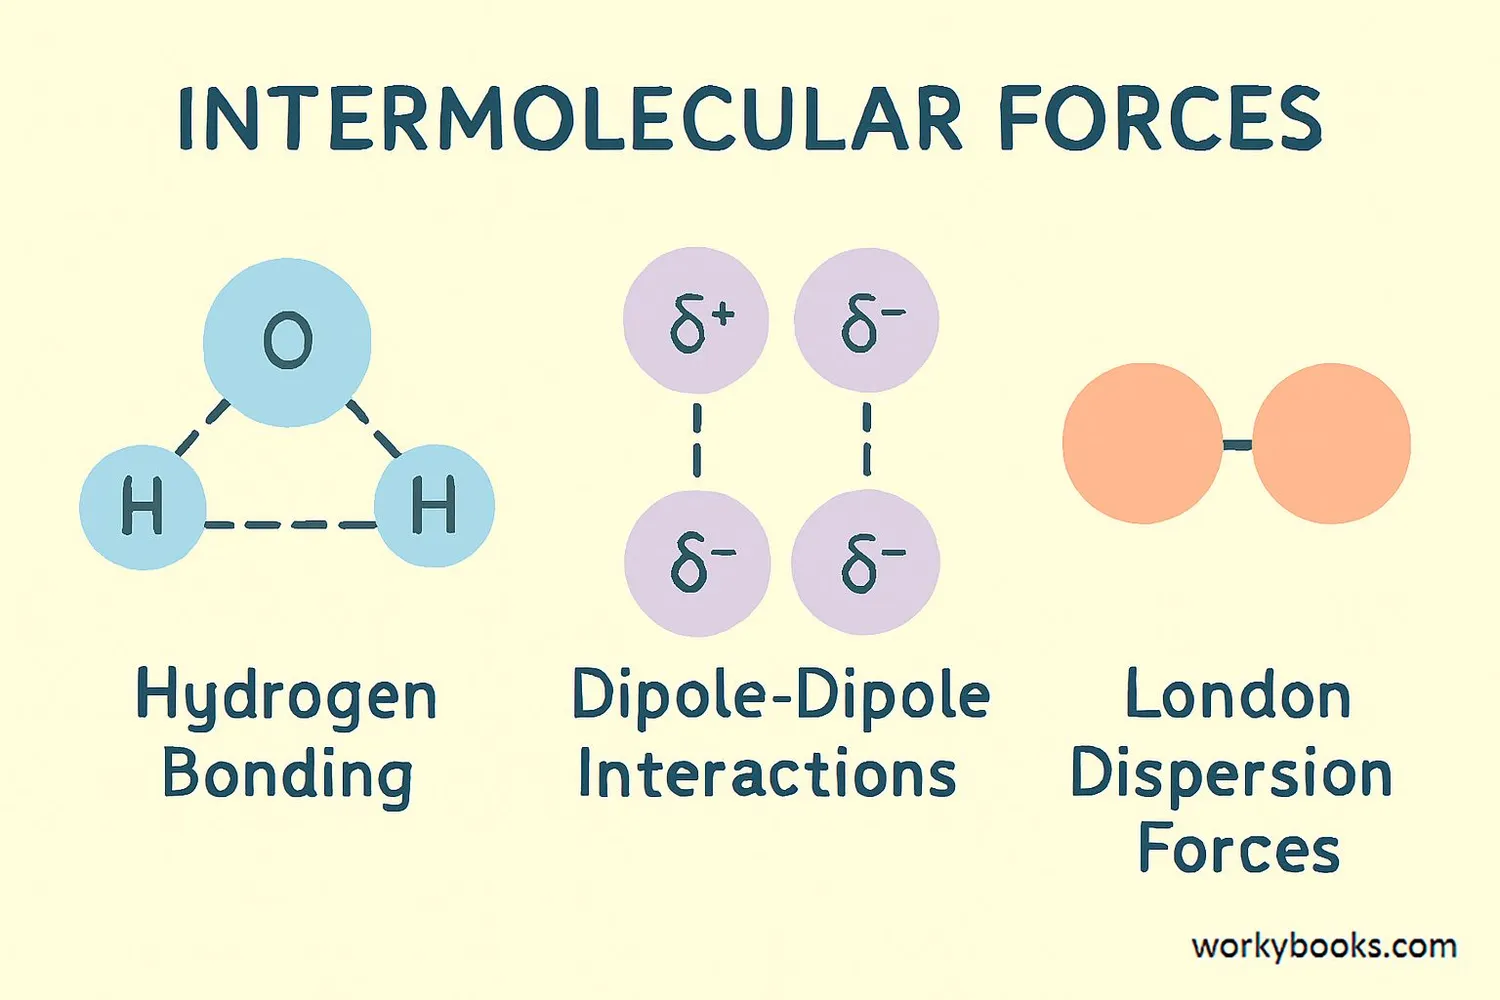
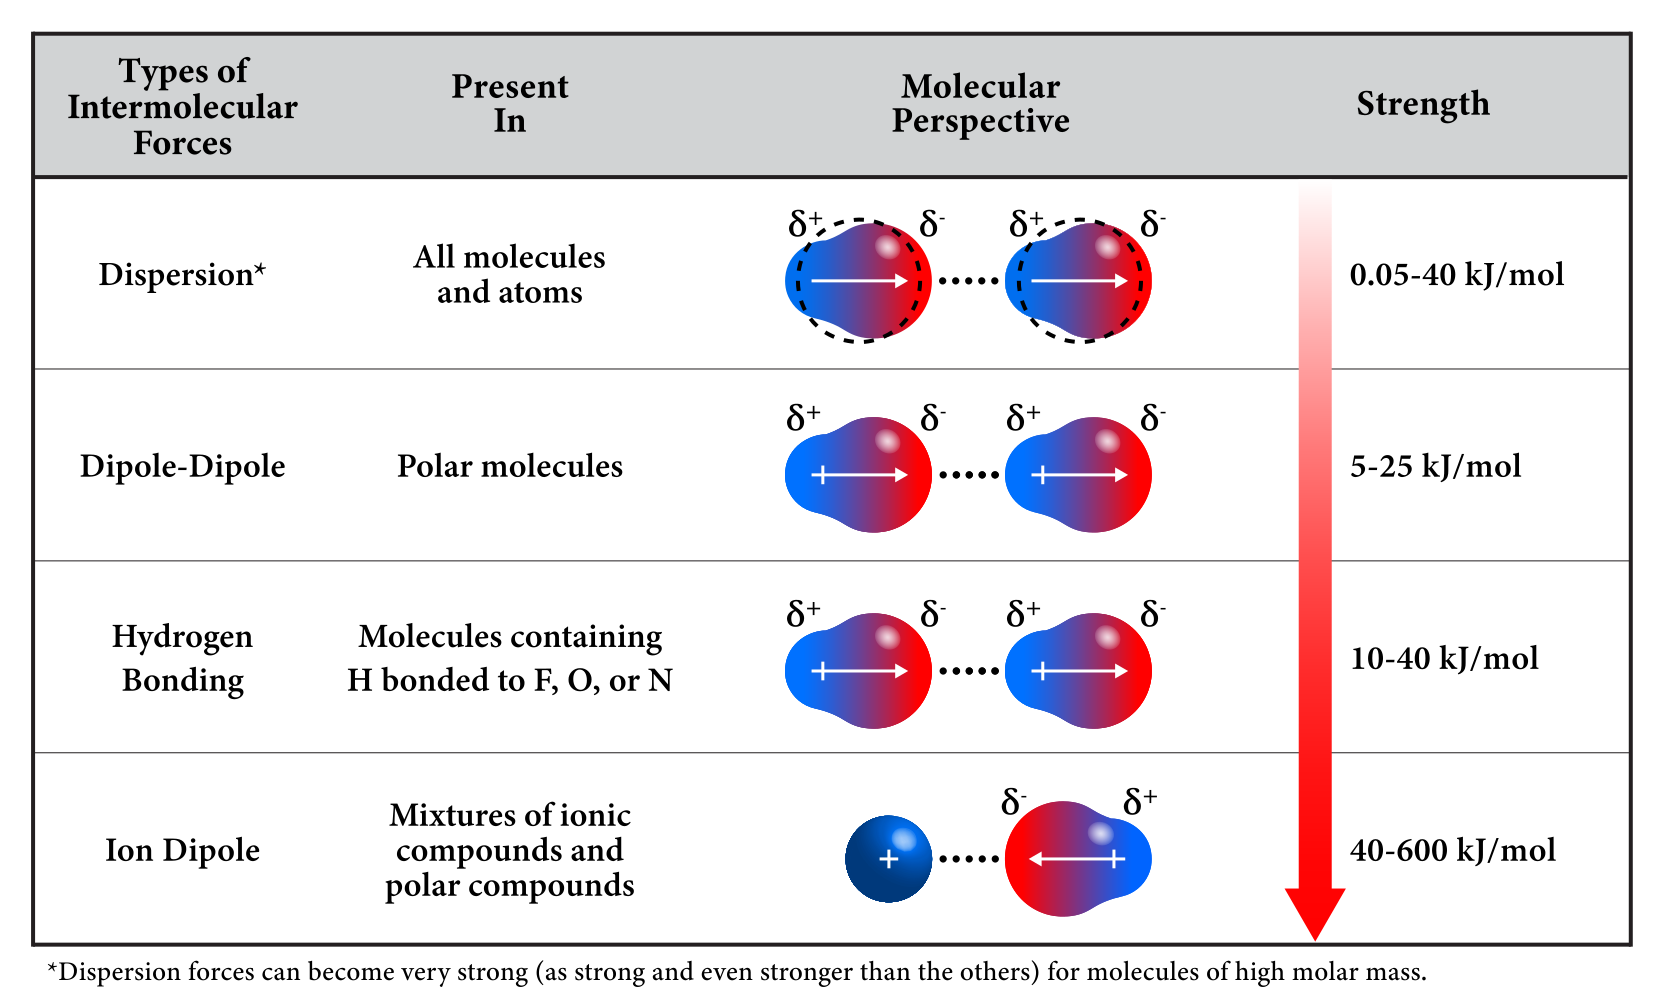
Intermolecular forces are the attractions between separate molecules, and they govern almost everything you deal with in practical chemistry — boiling points, solubility, separation techniques, formulation stability. If you have ever spent hours watching a compound refuse to move on a silica column, these forces are the reason. Let me walk through what they are and how they actually play out when you are working with real samples. There are three main categories you need to keep straight. London dispersion forces exist between every molecule, regardless of polarity. They come from temporary fluctuations in electron density that create momentary dipoles. The strength depends on polarizability, which scales with molecular size and surface area. A large hydrocarbon has much stronger dispersion forces than a small one, even though neither has a permanent dipole. Dipole-dipole interactions occur between molecules that have permanent dipoles. A chlorine atom pulls electron density away from hydrogen in HCl, creating a partial positive end and a partial negative end. Those ends align across molecules and create attraction. These forces are stronger than dispersion but weaker than hydrogen bonds.
Hydrogen bonding is a special case of dipole-dipole interaction that shows up when hydrogen is bonded directly to nitrogen, oxygen, or fluorine. The small size of hydrogen and the high electronegativity of those three atoms create an unusually strong local dipole. Water is the classic example, and its hydrogen bonding network is why it has a boiling point so much higher than hydrogen sulfide, which is heavier but cannot form hydrogen bonds. Here is the part most people miss. These forces do not operate in isolation. Any real molecule experiences all three simultaneously, and the total intermolecular attraction is the sum of everything happening at once. That is why simple rules of thumb often fail when you move beyond textbook examples. I learned this the hard way while developing a purification protocol for a moderately polar natural product. The compound had a molecular weight around 350 daltons, a few hydroxyl groups, and a long aliphatic chain. I expected it to elute reasonably fast on silica using a hexane-ethyl acetate gradient. Instead, the compound stuck to the column like it was glued. The molecular weight and the nonpolar chain meant strong dispersion interactions with the silica surface, and the hydroxyl groups added hydrogen bonding on top of that. Running pure ethyl acetate worked, but it co-eluted with a bunch of impurities I needed to separate. I ended up switching to a different stationary phase and using a gradient that started with a small amount of methanol to compete with the hydrogen bonding sites. That took the separation time from hours down to about twenty minutes per column.
The Real Pitfalls You Will Hit
Predicting intermolecular forces from structure alone sounds straightforward, but it is not. Two molecules can have the same functional groups and wildly different boiling points because of how their shapes affect surface contact. Linear chains pack better and have stronger dispersion forces than branched isomers. Neopentane boils at 9.5°C while n-pentane boils at 36.1°C, and the only difference is branching. That is a dispersion effect that polarity charts will not show you. Another issue is solvent competition. When you add a solvent, you are not just dissolving your compound. The solvent molecules are forming their own intermolecular interactions with each other and with your solute. Acetone disrupts hydrogen bonding networks much more effectively than dichloromethane because acetone is both a hydrogen bond acceptor and has significant dipole character. If you are trying to dissolve a polymer that relies on internal hydrogen bonding, picking the wrong solvent means you will waste days watching it sit there intact. There is also the matter of concentration. Intermolecular forces become much more complicated in concentrated solutions or in pure liquids. Activity coefficients deviate significantly from ideal behavior, and simple polarity-based predictions break down. If you are working with formulations or concentrated reaction mixtures, you should expect deviations from rules that assume dilute conditions.
Get the Full Details
One more thing that trips people up is temperature dependence. The strength of intermolecular attractions decreases as temperature increases, but not in a linear way. Entropy effects become more important at higher temperatures, which is why some compounds that seem insoluble at room temperature dissolve readily when heated. This is practical information when you are deciding between reflux conditions and ambient temperature procedures.
When the Theory Stops Working
Intermolecular force concepts assume you are dealing with discrete neutral molecules. That assumption falls apart for ionic liquids, molten salts, and highly concentrated electrolyte solutions. In those systems, electrostatic interactions dominate everything, and calling them intermolecular forces is misleading. The terminology breaks down because the interactions are fundamentally different — you are dealing with ion-ion and ion-dipole forces on a scale that dwarfs dispersion or hydrogen bonding. Supercritical fluids present another problem. Carbon dioxide above its critical point has solvent properties that change dramatically with small adjustments in pressure and temperature. The solvation power depends on density, which controls how closely molecules can approach each other and how strong the dispersion interactions become. You cannot predict extraction behavior from structure alone in supercritical CO. You need empirical data or at minimum a thermodynamic model calibrated to your system. Polymeric systems are also problematic. Intermolecular forces within a polymer chain interact with forces between chains, and the macromolecular architecture adds entanglement effects that have nothing to do with simple pairwise attractions. Glass transition temperatures, crystallinity, and mechanical properties emerge from all of this simultaneously. If you need to predict polymer behavior from intermolecular forces alone, you will be wrong.
What Actually Works in Practice
For everyday laboratory work, start with boiling point and solubility trends. Compounds with similar intermolecular force profiles tend to be miscible. "Like dissolves like" is an oversimplification, but it is directionally correct. Nonpolar solutes dissolve in nonpolar solvents because dispersion forces are comparable on both sides. Polar solutes dissolve in polar solvents because dipole-dipole and hydrogen bonding interactions are favorably exchanged. When you need more precision, use group contribution methods or activity coefficient models. UNIQUAC, NRTL, and UNIFAC can estimate non-ideal behavior in mixtures when you do not have experimental data. These are not perfect, but they are far better than guessing. For phase equilibrium calculations in process work, equations of state like Peng-Robinson give you reasonable results for hydrocarbon systems and moderate pressure ranges. If you are working with unusual systems — ionic liquids, deep eutectic solvents, supercritical fluids, or highly concentrated mixtures — the only reliable approach is experimental characterization. Differential scanning calorimetry, vapor pressure measurements, and solubility determination will save you more time than any theoretical model. I have seen people spend weeks trying to predict separation behavior with computational methods only to find that a single experimental run at the right temperature gave them the answer in an hour.
The bottom line is that intermolecular forces are real and they matter, but they are not a crystal ball. They give you a framework for understanding why things behave the way they do. They do not replace observation. The best practitioners use the theory to form hypotheses and then test those hypotheses against actual data instead of trusting the model to tell the whole story.