Measuring What It Takes to Pull an Electron Away
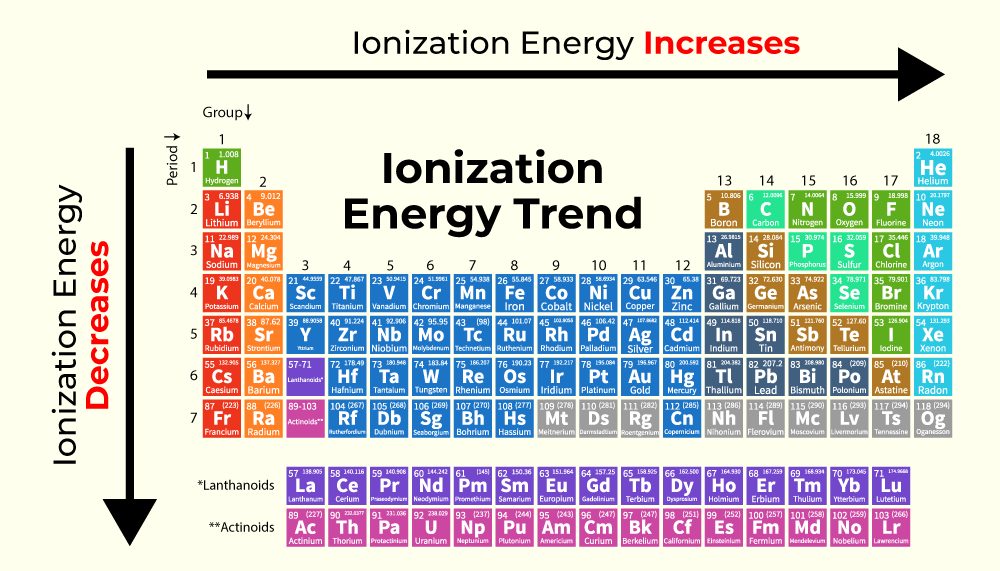
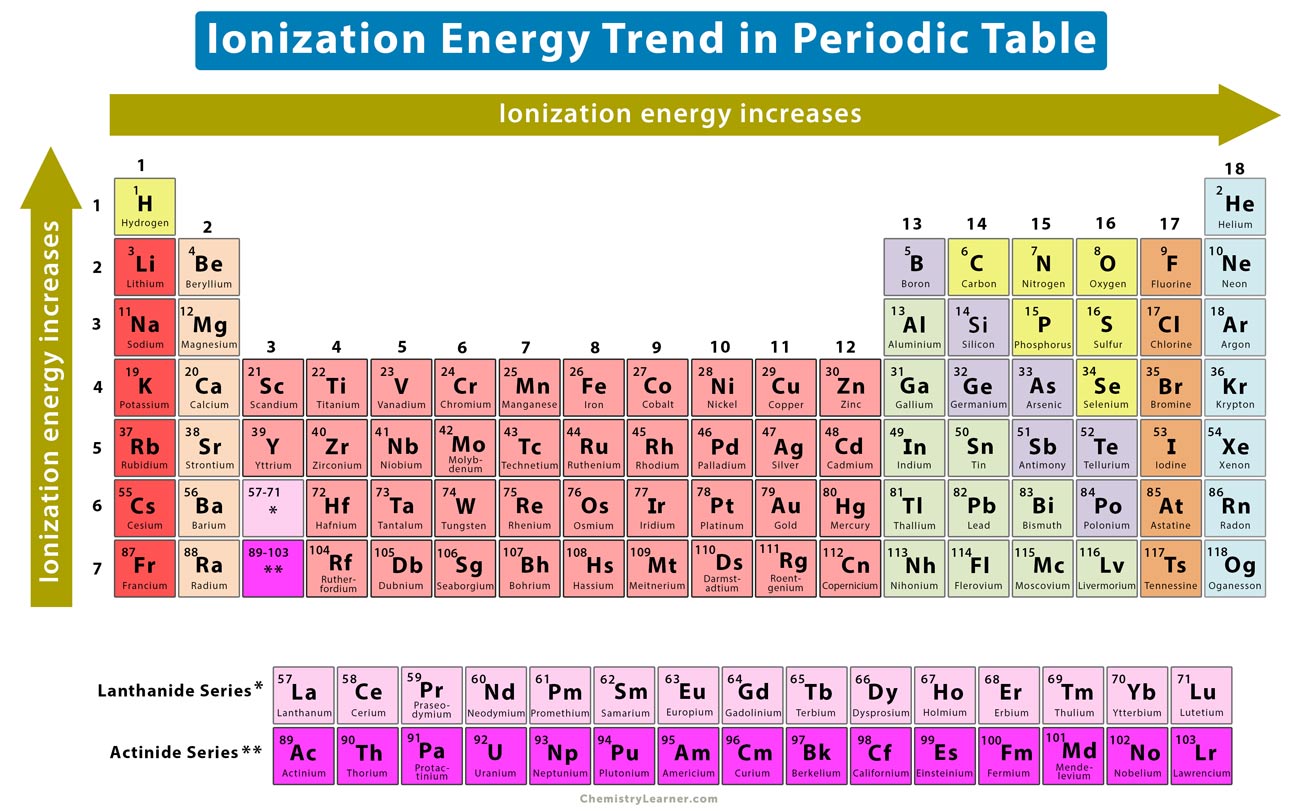
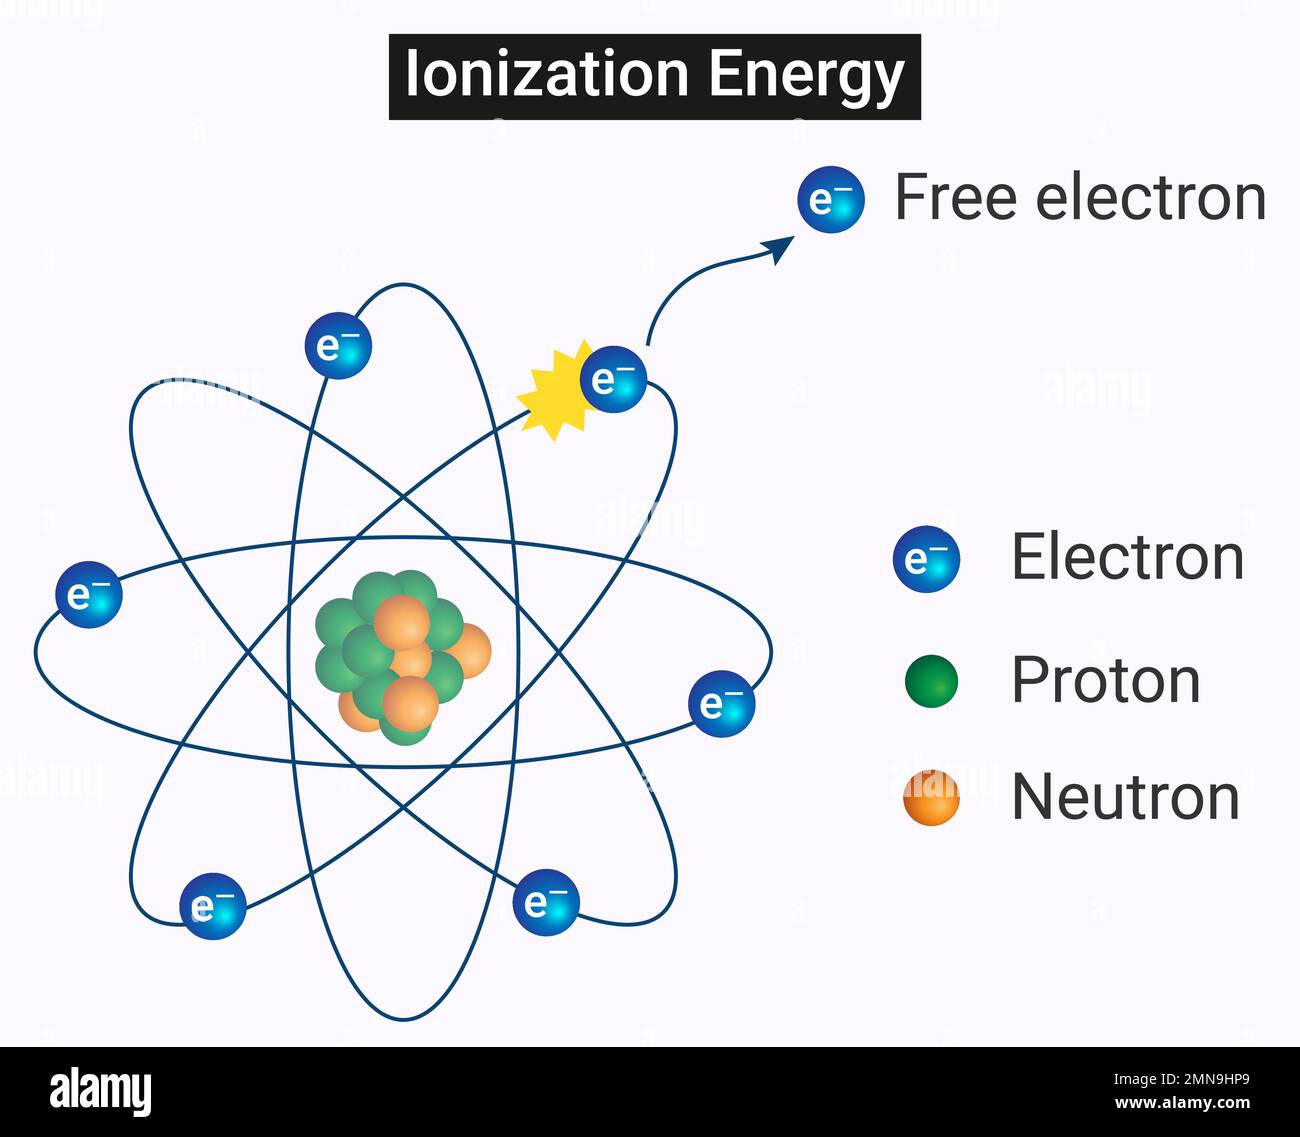
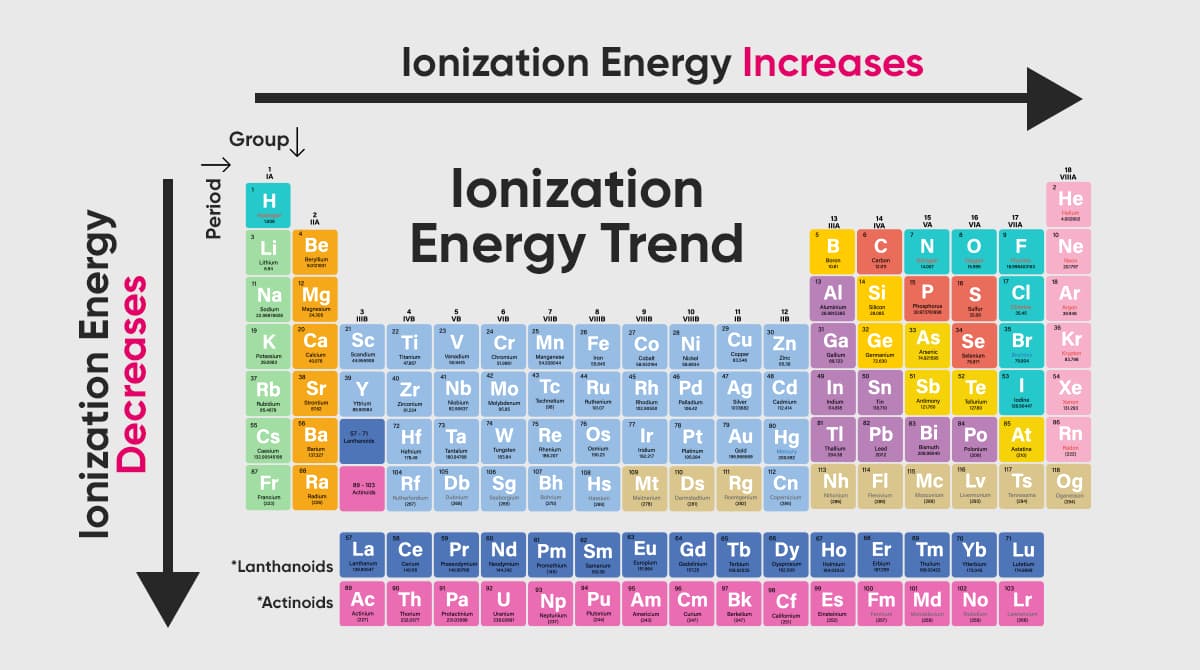
The first ionization energy is the minimum energy required to remove the most loosely held electron from one mole of neutral gaseous atoms. The second ionization energy removes a second electron, and so on. These values aren't abstract—they're measured in kilojoules per mole or electron volts, and they sit at the center of everything from predicting how an element bonds to designing plasma etching recipes for semiconductor fabs. I spent two weeks troubleshooting why a sputtering deposition tool kept yielding inconsistent film stoichiometry on a titanium target. The problem wasn't the power supply or the chamber pressure readings. It came down to the second and third ionization energies of titanium—717.7 kJ/mol and 1364 kJ/mol—interacting with the argon plasma in a way that favored selective ejection of certain species. Once I recalibrated the target conditioning cycle to account for that energy gap, the stoichiometry stabilized within two hours of running. That's the kind of thing ionization energy data actually solves. You look at a periodic table and see trends. Sodium drops to 495.8 kJ/mol, fluorine climbs to 1681.0 kJ/mol. You memorize that it increases across a period and decreases down a group. That's textbook level and it's correct, but it doesn't tell you what happens when you're looking at transition metals with partially filled d-orbitals or lanthanides where the 4f electrons create subtle shielding effects that shift everything by a few dozen kilojoules.
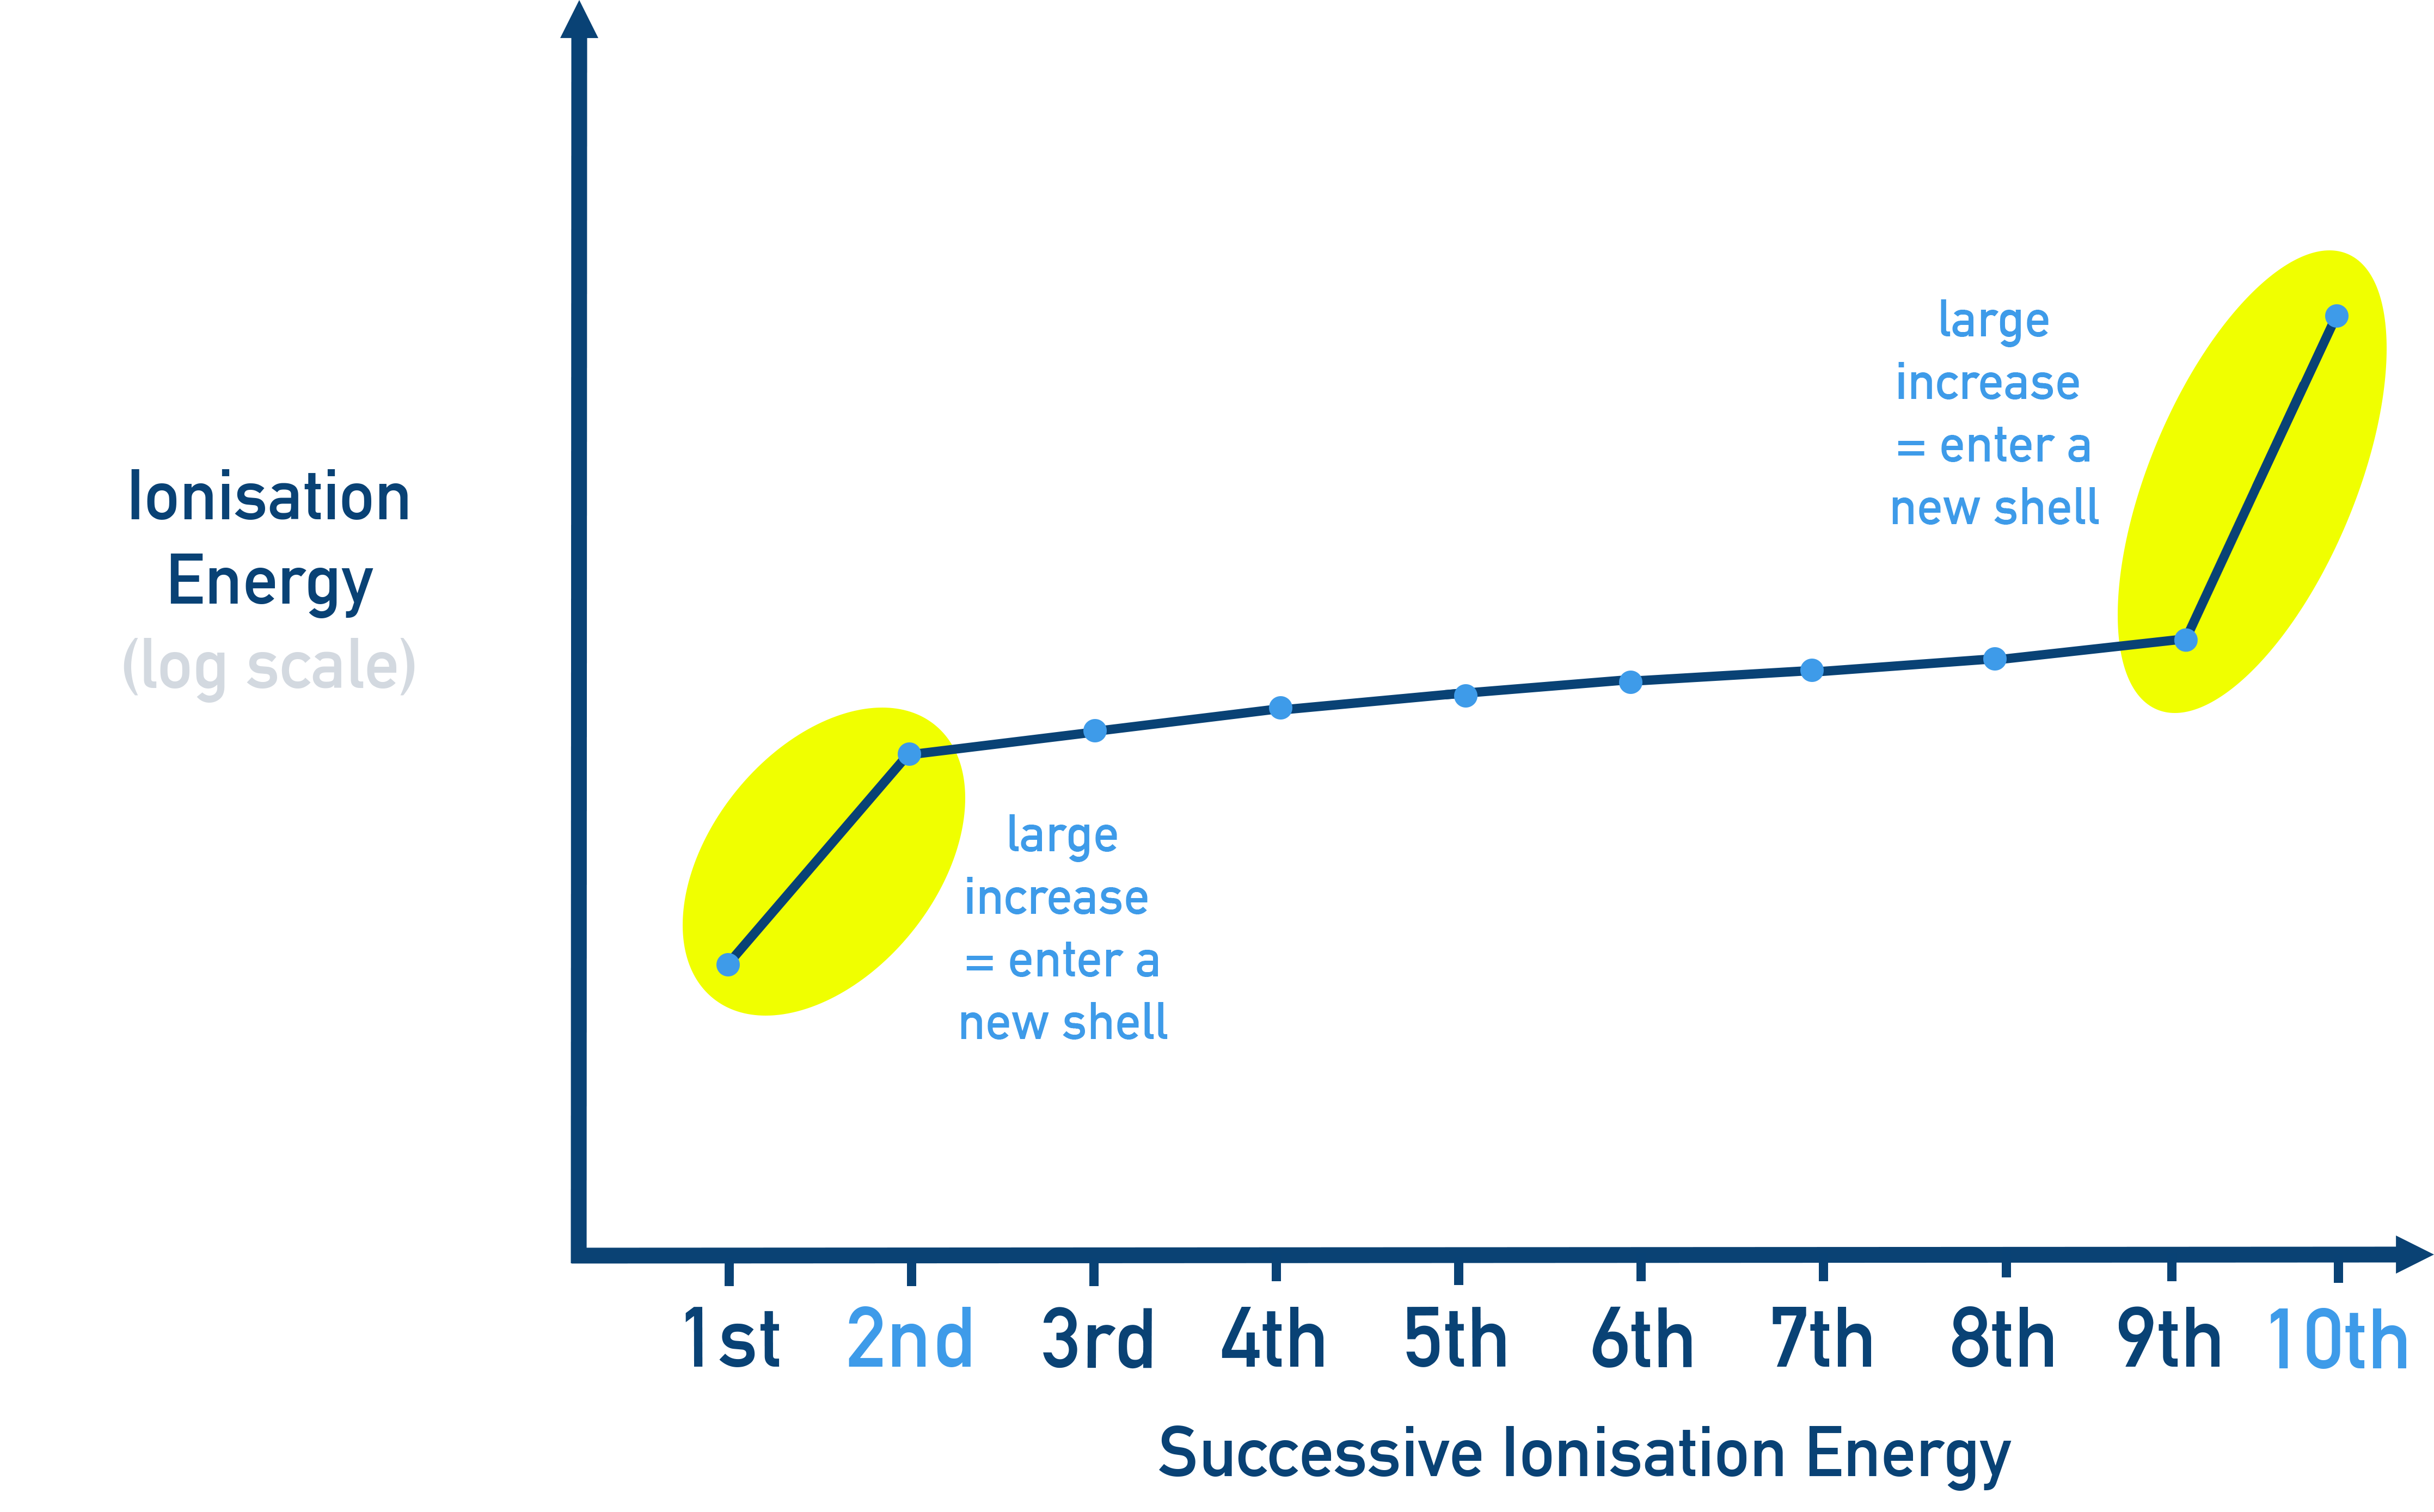
Here's something most introductory courses gloss over: the jump between successive ionization energies isn't smooth. It spikes sharply whenever you break into a core shell. Look at magnesium. The first ionization energy is 737.7 kJ/mol. The second is 1450.7 kJ/mol. The third is 7732.7 kJ/mol—that's a five-hundred-percent increase because you're now pulling an electron out of the neon-like 1s²2s²2p core. This jump is actually the most reliable experimental evidence we have for electron shell structure. It's not theoretical. It's data you can plot on graph paper and read directly. Another thing beginners miss: ionization energy and electronegativity are related but they're not the same thing, and conflating them causes real mistakes. Electronegativity is a bonded-atom property measured on arbitrary scales like Pauling or Mulliken. Ionization energy is a gas-phase atomic property measured in absolute energy units. The Mulliken scale actually combines ionization energy and electron affinity, but that's a derived quantity, not the ionization energy itself. If you're calculating bond character using only ionization energy values and ignoring electron affinity, your dipole moment predictions will be off by meaningful margins, especially for halogens.
How to Read and Use Ionization Energy Tables Correctly
When you're pulling values from NIST's Atomic Spectra Database or the CRC Handbook, pay attention to what state the atom is in. Most tabulated first ionization energies assume the ground electronic state of the neutral atom. But for transition metals, the ground state configuration can involve multiple terms close in energy, and the ionization can originate from different subshells depending on which term you start from. Chromium is a good example. Its ground state is [Ar] 3d 4s¹, not [Ar] 3d 4s² like you might expect from a naive Aufbau reading. That means the first electron removed comes from the 4s orbital, and the ionization energy is 652.9 kJ/mol. If you'd assumed the wrong configuration, you'd still get the right number for chromium, but you'd be explaining it wrong, and that wrong explanation compounds when you try to predict properties of Cr or Cr². The lanthanide contraction is another area where tables can mislead if you don't understand what's happening. You'd expect ionization energies to decrease steadily across the lanthanide series as atomic number increases and shielding gets better. They don't. Cerium is 534.4 kJ/mol, lutetium is 523.5 kJ/mol, but europium sits at 547.1 kJ/mol and ytterbium at 603.4 kJ/mol. Those bumps come from half-filled and fully filled f-subshell stability. When you're modeling a lanthanide-containing material and you ignore these local maxima, your thermodynamic calculations drift, sometimes by enough to flip a predicted reaction direction. There's also the issue of relativistic effects in the heavy elements. Gold has a first ionization energy of 890.1 kJ/mol, which is higher than silver's 731.0 kJ/mol despite silver being above gold in the group. The 6s electrons in gold are contracted by relativistic effects—they move fast enough near the nucleus that their effective mass increases and their orbital shrinks. That makes them harder to remove. If you're working with any element past mercury and you use non-relativistic trends to estimate ionization energy, your error margins can exceed ten percent. For routine lab work that might not matter. For computational chemistry workflows where you're feeding ionization potentials into DFT codes, it absolutely matters.
Get the Full Details
Where the Data Gets Messy and How to Handle It
Not every element has well-determined ionization energies for all twelve or more successive removals. For some heavy or exotic isotopes, only the first two or three are measured experimentally. The rest are calculated using Hartree-Fock or coupled-cluster methods, and those calculations carry uncertainty estimates that vary widely depending on the software package and the correlation treatment. When I've had to work with elements like francium or meitnerium, the literature values are often derived from one or two theoretical studies with error bars of several tens of kJ/mol. That's not trivial when you're trying to model something precise. A practical workaround for sparse data is to use the Slater-Condon rules combined with established screening constants to estimate missing values, then cross-reference against isoelectronic sequences. If you know the ionization energy of Fe², you can infer things about Co³ and Ni because they share the same electron configuration. This isoelectronic extrapolation usually gets you within five to eight percent for consecutive elements in a sequence, which is far better than nothing when you're doing preliminary screening. Another common problem: published values sometimes disagree between sources. The NIST value for nitrogen's first ionization energy is 1402.3 kJ/mol, but you'll find older textbooks listing 1400 or 1405 depending on when and how the measurement was remade. These discrepancies come from differences in the experimental technique—photoelectron spectroscopy versus threshold ionization—and the treatment of vibrational and rotational contributions in the neutral versus ionized state. Always check the source and the year. For most applications, the spread across major references is under one percent, but if you're calibrating instrumentation or validating a computational method, that one percent is your limiting factor.
Using Ionization Energy to Predict Bonding Behavior
The most direct application is predicting whether an element tends to form cations or anions. Low ionization energy elements like the alkali metals readily lose electrons and form ionic bonds. High ionization energy elements like the noble gases and halogens hold onto their electrons tightly. But the real utility shows up in the intermediate cases—the metalloids and transition metals where ionization energy alone doesn't give you a clean answer. You need to combine it with electron affinity, atomic radius, and lattice energy or bond enthalpy data. For example, aluminum has a first ionization energy of 577.5 kJ/mol, which is moderate. You might expect it to form Al³ ions easily. It does, in compounds like AlF, but in AlCl the third ionization energy—2744.8 kJ/mol—is so high that the lattice energy or covalent bond energy available doesn't compensate for removing that third electron. The result is that aluminum chloride exists as a dimer with significant covalent character rather than as a simple ionic lattice. This is a case where looking at only the first ionization energy would lead you astray. You have to consider the cumulative cost of removing multiple electrons and whether the bonding environment can pay that cost. Photoelectron spectroscopy is the experimental technique that directly measures ionization energies by firing photons at atoms and measuring the kinetic energy of the ejected electrons. The binding energy is simply the photon energy minus the kinetic energy. Modern instruments can resolve features down to a few meV, which means you can see fine structure splitting in the ionization peaks—spin-orbit coupling effects that separate p/ and p/ electrons, for instance. If you're interpreting XPs data for a transition metal compound, you'll see multiple peaks corresponding to different d-orbital ionization channels, and the relative intensities and positions of those peaks encode information about oxidation state, ligand field strength, and even surface contamination. That's why ionization energy measurements aren't just tables in a handbook—they're a diagnostic tool.
What Are Ionization Energies Beyond the Periodic Table Trends
They're fingerprints. Each element has a unique sequence of ionization energies that reflects its electron configuration, nuclear charge, and the quantum mechanical rules governing orbital energies. That sequence is reproducible to high precision and it doesn't change based on sample preparation or environmental conditions, which is why ionization energy data remains one of the most trusted references in analytical chemistry and materials science. When someone tells you the ionization energy of an unknown sample matches iron within 0.1 percent, that's a confident identification, not a guess.