Enzyme kinetics basics for people who just need to get it done
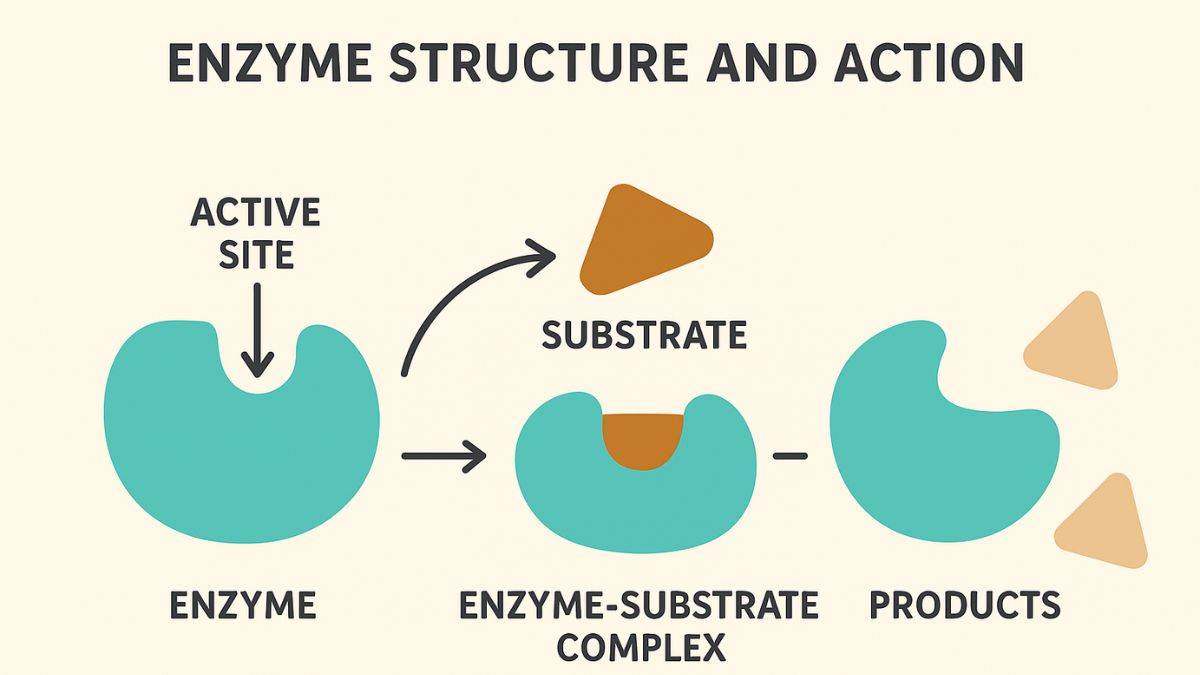
A substrate is whatever molecule an enzyme acts on. That's the textbook answer. The enzyme binds the substrate at its active site, stabilizes the transition state, and converts it into product. The whole system gets measured by how fast that happens and how much substrate is sitting around when you start. Here's what most people miss when they first work with enzymes. A substrate isn't just a label you throw on a molecule. The same molecule can be a substrate for one enzyme and an inhibitor or product for another. Urea is the classic example. It's the substrate for urease, but in other systems it acts as a competitive inhibitor at certain concentrations. The identity of the substrate is tied to the enzyme, not the molecule itself. I learned this the hard way in my first year running kinetic assays. I was measuring urease activity with a standard colorimetric kit, and the Michaelis-Menten curve looked completely wrong at substrate concentrations above 50 mM. I spent three days troubleshooting a broken spectrophotometer, recalibrating the pH meter, and checking reagent expiry before I realized the high urea concentration was actually inhibiting a contaminating phosphatase in my enzyme prep that was eating the detection reagent. The substrate wasn't behaving badly. The assay wasn't built to handle that concentration range of it. I switched to a coupled enzyme assay with glutamate dehydrogenase instead and got clean data in an hour. Not a single thing was broken.
The mechanics of how it actually works
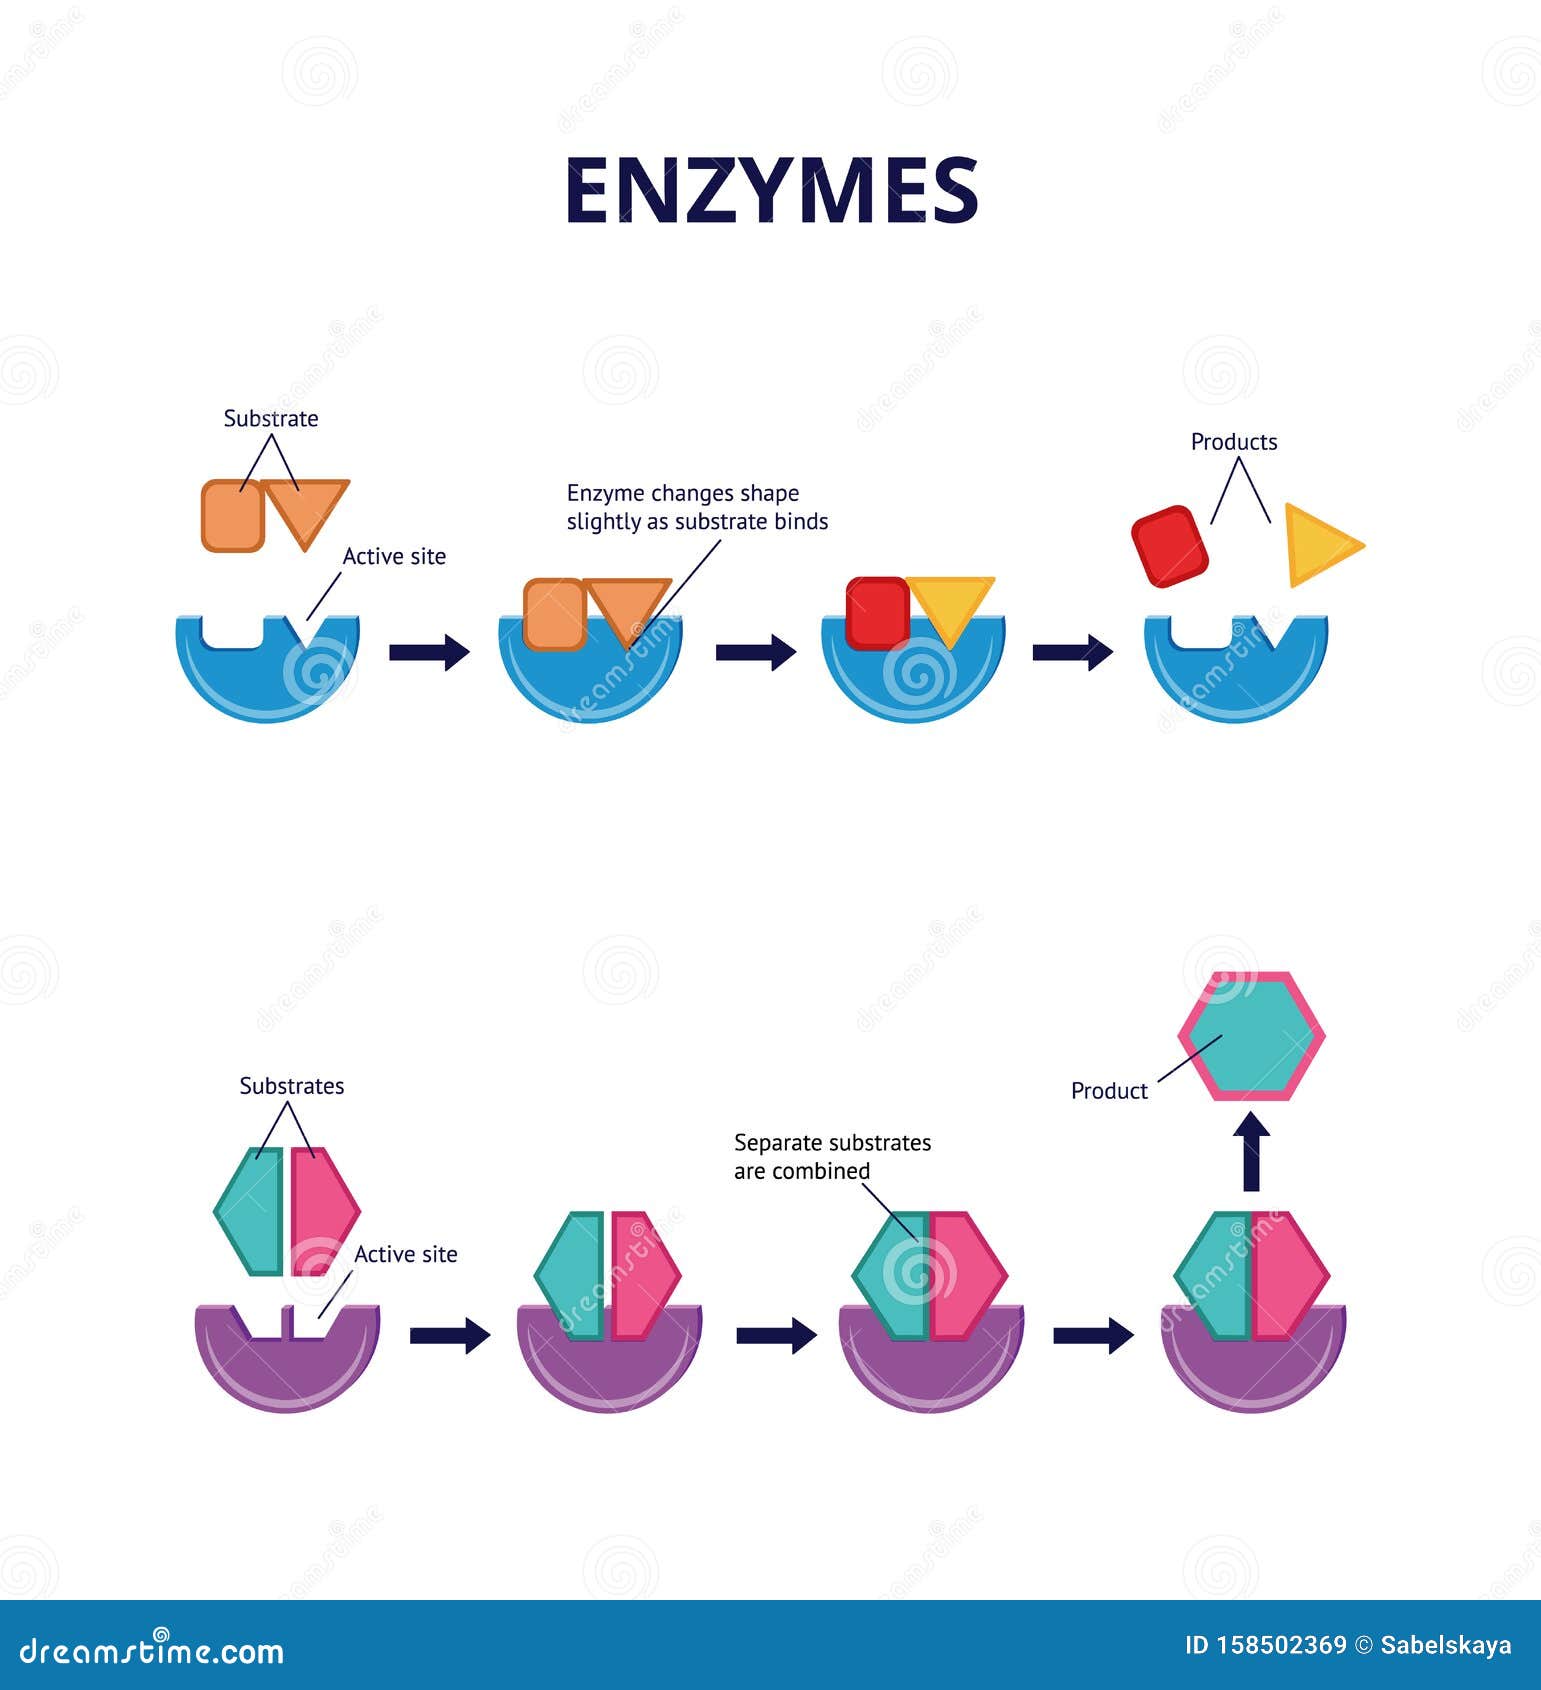
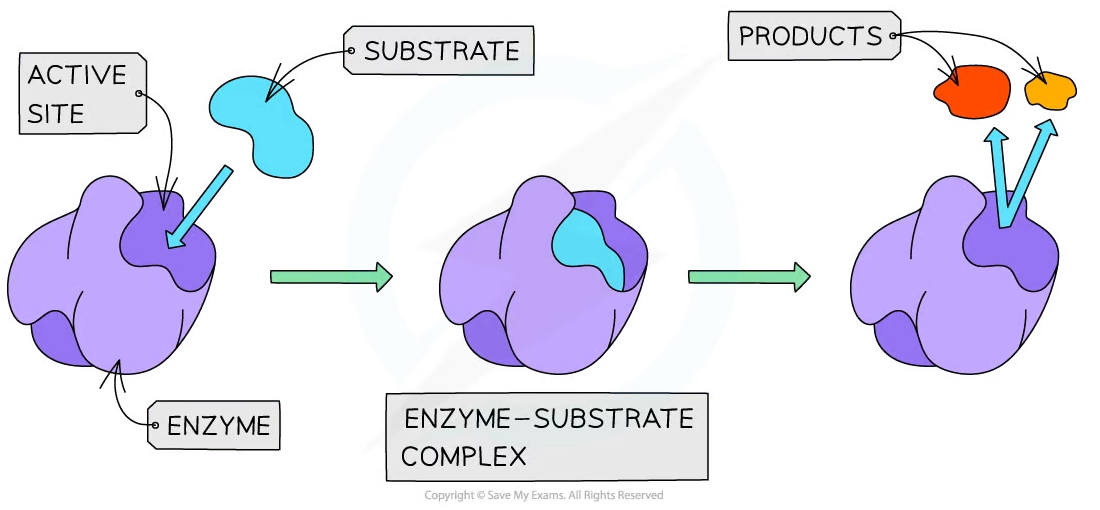
When you add substrate to an enzyme solution, you're setting up a competition. The enzyme either binds the substrate and goes through the catalytic cycle, or it stays empty waiting for something else. The turnover number tells you how many substrate molecules each active site can process per second. The Michaelis constant tells you how much substrate you need to reach half of maximum velocity. These numbers aren't fixed properties. They shift with temperature, pH, ionic strength, and the presence of any other molecules in solution. I've seen Km values for hexokinase change by a factor of four when the potassium concentration dropped below 50 mM. That matters if you're comparing literature values to your own data, because the buffer composition people report in methods sections is often incomplete or simplified. The induced fit model is where most beginners get stuck conceptually. The enzyme doesn't just have a rigid lock waiting for a matching key. Binding of the substrate causes conformational changes that reshape the active site around the substrate. This means substrate analogs that look right on paper but don't trigger the same conformational shift can bind tightly without getting converted to product. You end up with a dead-end complex that looks like catalysis in structural studies but produces zero turnover. If you're doing inhibitor screening, this is exactly why Ki values from binding assays don't always predict IC50 values from activity assays.
I ran into this with a purine nucleoside phosphorylase project. We had a substrate analog that bound in the crystal structure at the active site with sub-micromolar affinity, but in solution it showed no inhibition at concentrations up to millimolar range. The crystal structure didn't capture the Mg2+ ion that was required for the conformational change. Without the metal, the enzyme couldn't close around the analog properly, and the analog just sat on the surface. We added a stoichiometric amount of MgCl2 and suddenly the inhibition constants made sense. Structural data alone would have sent us down the wrong path for weeks.
Get the Full Details

What to watch out for
Substrate depletion is the most common practical problem. If you start with too much substrate relative to enzyme, the initial velocity assumption breaks down within seconds. You're measuring steady-state conditions instead of initial rates, and your kinetic parameters become unreliable. For a typical enzyme with a kcat in the 10 to 100 per second range, you want the substrate concentration to drop less than five percent during the time you're collecting data. That usually means using enzyme concentrations in the low nanomolar range for purified preparations. Substrate solubility is the second problem. I've lost more samples to precipitation than I care to admit. Dimethyl sulfoxide stocks are convenient until the aqueous buffer dilutes them and the substrate crashes out mid-assay. Always make fresh dilutions from concentrated stocks, never from a stock that's been in the buffer for more than a few hours. Check the solution under a microscope if you're unsure. Cloudy isn't cloudy enough to skip. Multiple substrates complicate everything. Sequential mechanisms require all substrates to bind before any product leaves. Ping-pong mechanisms release one or more products before the second substrate even binds. Running a double-reciprocal plot with one substrate held at multiple fixed concentrations of the second substrate will tell you which one you're dealing with, but only if you actually run the full matrix of concentrations. Most people run one or two and extrapolate. That's how you get the wrong mechanism assigned and the wrong kinetic model fitted to your data.
There's also the problem of substrate inhibition, where high substrate concentrations actually slow the reaction. This happens when a second substrate molecule binds at a secondary site and locks the enzyme in an inactive conformation. It's easy to miss if you only measure at a narrow concentration range and mistake it for experimental noise. I saw this with lactate dehydrogenase in a routine assay. The activity curve peaked at around 10 mM pyruvate and then dropped sharply. We thought the enzyme was denaturing. It wasn't. It was substrate inhibition, and the IC50 was roughly 25 mM. If you'd only measured between one and five millimolar, you'd have reported a normal Michaelis constant and missed the whole phenomenon.
The edge case that ruins everything
Product inhibition is the silent killer in substrate assays. The product of your reaction often binds back to the active site with affinity comparable to the substrate. As product accumulates, it competes with substrate for binding and slows the reaction rate independently of substrate depletion. This is particularly problematic when you're measuring initial velocity over anything longer than thirty seconds. The product inhibition makes your velocity curve bend downward before you've even started collecting meaningful data. The workaround is to use a coupled assay where the product is immediately consumed by a second enzyme. For dehydrogenases, this means coupling NADH production to something that oxidizes it back to NAD+. The coupling enzyme should have a much higher Vmax than your test enzyme so it doesn't become rate-limiting. I've seen this approach reduce the effective product inhibition from minutes to sub-second timescales, which is what you actually need for accurate kinetic measurements. Another issue that comes up more often than it should is that the substrate itself can interfere with detection. Fluorescent and absorbance assays both suffer from this. If your substrate or product absorbs light at the same wavelength as your reporter, you're not measuring enzyme activity. You're measuring the concentration of a compound that may or may not be related to the reaction rate. This sounds obvious until you're looking at a data set that makes perfect sense and then realize your "signal" is entirely from substrate autofluorescence that decreases over time due to photobleaching. The curve looked exactly like product formation. It wasn't.
There's no perfect way to handle all of these problems at once. You pick the assay format that minimizes the ones relevant to your system and live with the rest. The alternative is spending months trying to eliminate every source of error and realizing you still can't publish because the data has error bars that span an order of magnitude. I stopped chasing perfection around 2008 and started focusing on identifying which assumptions were actually violating my results. The difference in productivity was enormous. Below is a basic reference table for common substrate kinetic parameters. These are literature values for purified enzymes under standard conditions. Your actual values will differ based on buffer, temperature, and purity. Treat them as starting points, not targets.
Common Enzyme-Substrate Pairs and Reference Parameters
Enzyme: Carbonic anhydrase. Substrate: CO2. Km: approximately 12 mM. kcat: roughly 1.4 million per second. Enzyme: Catalase. Substrate: H2O2. Km: approximately 25 to 40 mM. kcat: around 40 million per second. Enzyme: Chymotrypsin. Substrate: N-acetylanthranilamide. Km: about 0.75 mM. kcat: approximately 150 per second.
Enzyme: DNA polymerase. Substrate: dNTP. Km: varies widely by enzyme and template, typically in the low micromolar range. kcat: roughly 50 to 100 per second for most replicative polymerases. Enzyme: Hexokinase. Substrate: glucose. Km: around 0.1 mM. kcat: approximately 100 per second. Enzyme: Lysozyme. Substrate: bacterial cell wall polysaccharides. Km: difficult to define precisely due to insoluble substrate, apparent Km in the milligram per milliliter range. kcat: roughly 0.5 per second.
Enzyme: Peroxidase. Substrate: H2O2 plus a reductant like ABTS. Km for H2O2: approximately 0.5 mM. kcat: around 50 per second. Enzyme: Pepsin. Substrate: denatured proteins. Km: broad range depending on substrate specificity, typically 0.1 to 1 mg/mL. kcat: roughly 10 to 20 per second. Enzyme: RNA polymerase. Substrate: NTPs. Km: varies, commonly 10 to 100 micromolar. kcat: around 50 per nucleotide added.
Enzyme: Trypsin. Substrate: BPTI or synthetic peptides. Km: depends on peptide sequence, generally low micromolar to low millimolar. kcat: around 100 to 500 per second for small peptide substrates. These numbers are for context. If your lab is measuring anything off by more than a factor of two from published values, check your temperature first, then your pH, then your enzyme activity. Those three variables account for most of the discrepancy in my experience.
