Covalent Bonds Are Nothing Fancy, They're Just Overlapping Orbitals
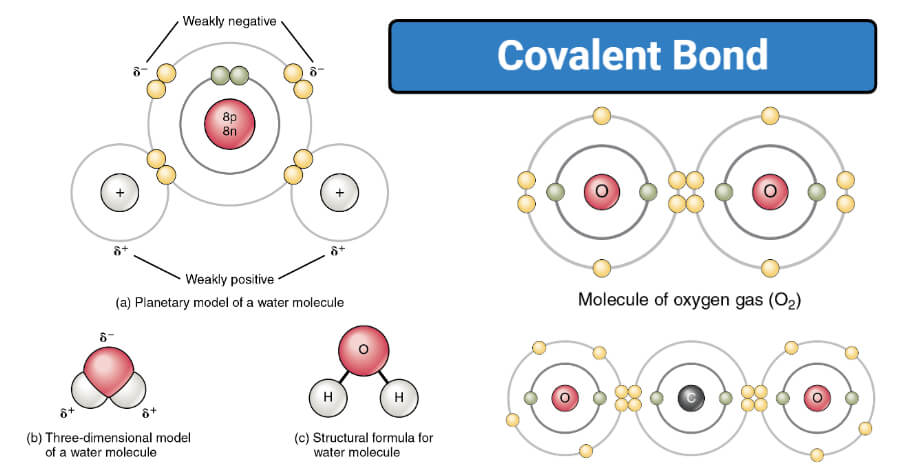
You pick up any intro chemistry textbook and it'll hand you a neat little list of properties. Share electrons, directional, saturable. Simple enough until you actually try to use this stuff in the lab or on a real problem set, where the details matter more than the definitions. Here they are, stripped of the gloss. First, directionality. Covalent bonds form along specific axes determined by the overlapping orbitals. This isn't just trivia. It's why water bends at 104.5 degrees instead of sitting flat, and why proteins fold the way they do. The bond angle comes from hybridization—sp3, sp2, sp—and if you ignore that, your molecular models will look right on paper and fall apart the moment you try to build something with real geometry. Second, saturation. An atom can only form so many covalent bonds before it runs out of valence electrons to share. Carbon makes four. Nitrogen three. Oxygen two. This is the octet rule doing its job, and it's what keeps organic chemistry from descending into chaos. But here's the thing most people miss: saturation is about available orbitals, not just electrons. Transition metals break this rule constantly because they've got d-orbitals sitting around that main-group elements don't bother with. If you're working with organometallics and expect simple saturation behavior, you're going to have a rough time.
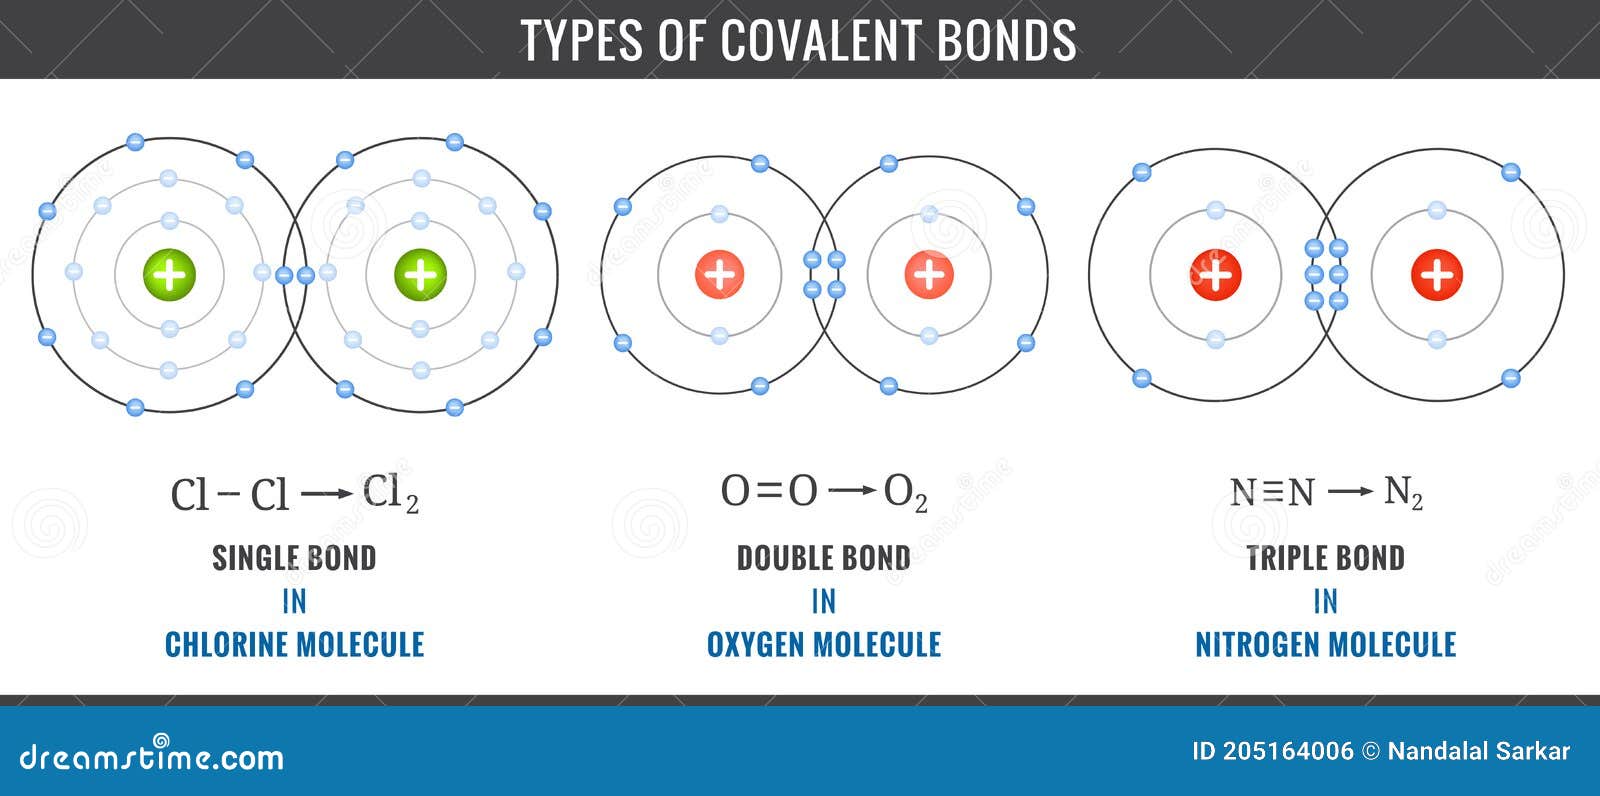
Third, bond strength, measured as bond dissociation energy. A C-C single bond sits around 347 kJ/mol. A C=C double bond is roughly 614 kJ/mol. Not double the single bond, by the way—the pi bond is weaker than the sigma bond. That matters when you're predicting whether a reaction will actually happen or just sit on your bench looking smug. I spent a few years debugging why certain elimination reactions gave unexpected product ratios, and the answer always came back to subtle differences in bond energies across similar molecular environments. A C-H bond next to an electronegative group isn't the same strength as one buried in the middle of a hydrocarbon chain. My workaround was stopping my reliance on tabulated average bond energies and switching to group-additivity methods for thermochemical estimates. It added about twenty minutes to my calculation time but cut my error rate significantly. You can look up the Benson method if you want to dig into that. There's a common pitfall worth mentioning. People treat covalent bonds as if they're rigid sticks connecting atoms. They're not. Bonds stretch, bend, and vibrate. The Morse potential describes this better than Hooke's law for anything beyond tiny displacements. If you're modeling anything where bond breaking actually happens—reaction dynamics, thermodynamics, spectroscopy—using a simple harmonic approximation will give you wrong answers faster than you'd expect.
Another thing beginners overlook: covalent bonds exist on a spectrum. The difference between a "pure" covalent bond and an ionic bond isn't a hard line. Pauling's electronegativity scale puts it roughly around 1.7 difference, but that's a guideline, not a law. Aluminum chloride is a classic case—it covalently dimerizes to Al2Cl6 in the gas phase but behaves ionically in solution. The bonding doesn't change categories, the environment just reveals different aspects of it. So three properties: directionality from orbital overlap, saturation from valence electron limits, and measurable bond strength that varies with molecular context. That's the framework. Everything else is just applying it to whatever problem you're actually facing.
Get the Full Details