Understanding Chemical Equilibrium Without the Textbook Fluff
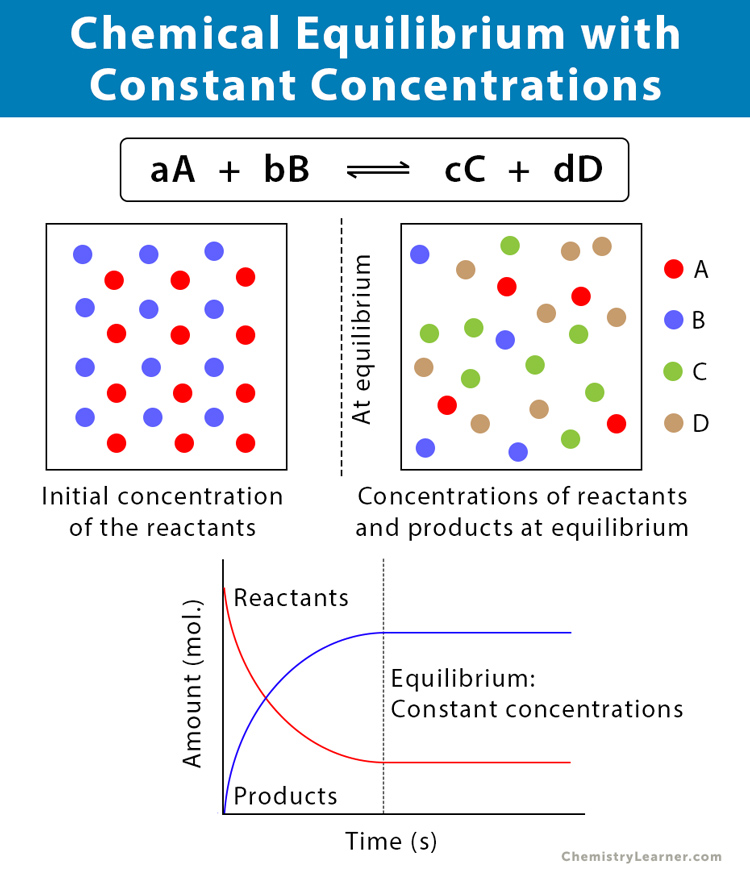
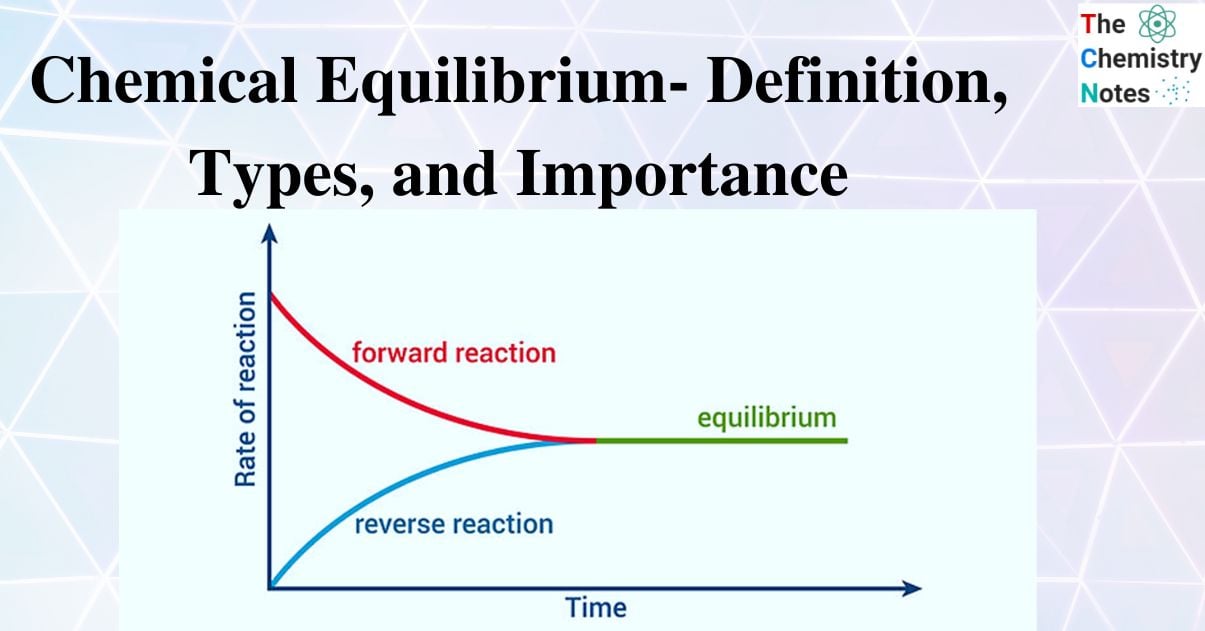
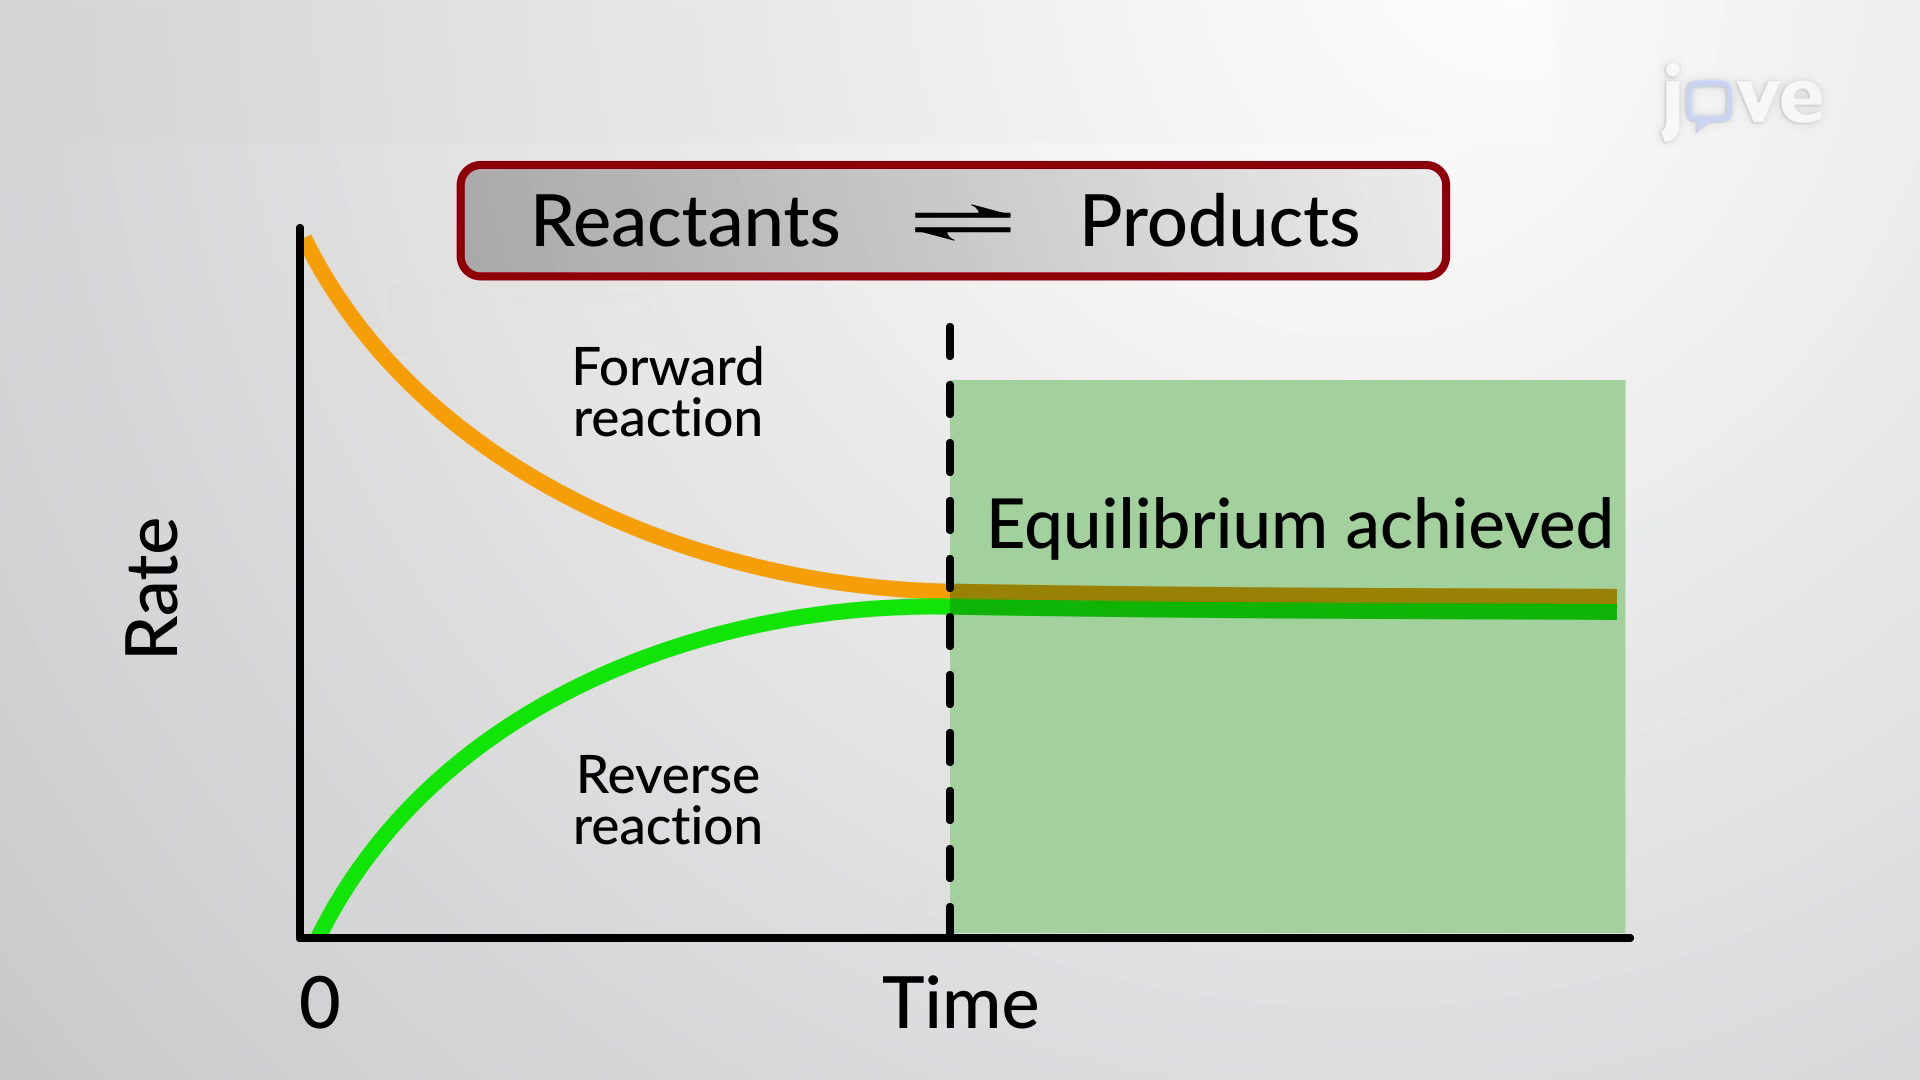
Chemical equilibrium is the point at which the forward and reverse reaction rates in a reversible process become equal, meaning the concentrations of reactants and products stop changing over time. That does not mean the reaction has stopped. Molecules are still reacting in both directions, but they are doing so at the same rate. The system is dynamic, not static. When I first encountered this concept in an undergraduate lab, I treated it like the answer key said: write the equilibrium expression, plug in numbers, get K. That approach works until you run into something that actually exists in the real world.
What Is A Chemical Equilibrium and How Do You Actually Calculate It
The standard method is setting up an ICE table—initial, change, equilibrium—and solving for the unknown. You write the balanced equation, assign initial concentrations, express changes in terms of x, substitute into the equilibrium constant expression, and solve. For simple cases with small K values, you can often skip the quadratic formula by assuming x is negligible compared to the initial concentration. That shortcut saves maybe five minutes per problem and introduces an error margin of roughly 5 percent or less when the assumption holds. It fails badly when K is large or when the initial concentrations are very small. I learned that the hard way during a process engineering internship. We were modeling an ammonia synthesis loop running at moderate pressure, and I defaulted to the small-x approximation on a system where the equilibrium constant was actually closer to 1. The calculated conversion was off by about 18 percent, which meant our reactor sizing was way too small. We had to rerun the full quadratic and then verify with a numerical solver. That cost us roughly a day of recalibration. The workaround I use now is straightforward: check the 5 percent rule before you ever assume anything away. Take your initial concentration, divide by K, and if the result is less than about 400, do not bother with the approximation. Just solve the full equation or use an iterative method. It takes longer on paper but saves you from rebuilding your entire model later.
Here is a practical example. Consider the decomposition of dinitrogen tetroxide: N2O4(g) 2NO2(g). The Kc at 25°C is approximately 4.6 × 10^-3. If you start with 0.10 M N2O4, setting up the ICE table gives you Kc = 4x² / (0.10 - x). Solving this properly yields x 0.0104 M, so the equilibrium concentration of NO2 is about 0.0208 M. If you had used the small-x approximation, you would have gotten x 0.0107 M, which looks close but pushes your error right to the edge of acceptability. In a well-controlled lab setting that difference matters. Another thing people miss is the distinction between K and Q. The equilibrium constant K is a fixed value at a given temperature. Q is the reaction quotient, calculated the same way but using whatever concentrations exist at any arbitrary moment. If Q is less than K, the reaction proceeds forward. If Q is greater than K, it proceeds in reverse. This matters more than you would think when you are troubleshooting a system that seems like it should be at equilibrium but is not. I once spent three hours chasing a contaminant in a spectrophotometry setup before realizing the cuvette solution had simply not been allowed to reach equilibrium. The absorbance readings were drifting slowly, and I had recorded them too early. Checking Q against K would have flagged the issue immediately. Le Chatelier's principle is usually taught as a qualitative tool, but it has real quantitative limits. Increasing pressure shifts the equilibrium toward the side with fewer gas moles, yes, but the magnitude of that shift depends entirely on the specific K value and the initial conditions. It is easy to overestimate how dramatic these shifts are. In practice, a pressure change might move the equilibrium position by a few percent, not the wholesale conversion people expect from textbook diagrams.
Get the Full Details
Temperature changes actually alter the value of K itself, unlike concentration or pressure changes. This is another area where intuition fails. For an exothermic reaction, raising the temperature decreases K. For endothermic, it increases K. The van 't Hoff equation relates the change in K to the change in temperature and the enthalpy of reaction, but most students never see it applied outside of a derivation exercise. Using it to estimate how much K shifts over a 50-degree temperature range is something I do regularly when scaling up reactions from bench to pilot scale. It usually changes K by 10 to 30 percent per 50°C, depending on the reaction enthalpy. The biggest limitation of equilibrium calculations is that they assume the system has actually reached equilibrium. In industrial reactors, flow rates, mixing efficiency, and mass transfer limitations can prevent that from happening. A reactor might be designed for a certain conversion based on equilibrium calculations, but if the residence time is too short or the catalyst is poisoned, you will never get there. I have seen entire production lines underperform because someone optimized for equilibrium yield without accounting for kinetic constraints. The fix was always the same: measure the actual conversion under operating conditions and work backward from there instead of trusting the textbook number. Heterogeneous equilibria add another layer of complexity. Solids and pure liquids do not appear in the equilibrium expression, which simplifies things on paper but creates confusion in practice. If you are working with a system involving a solid precipitate or a pure liquid solvent, you need to make sure you are not accidentally including their concentrations. A common mistake is writing K expressions that include water in aqueous solutions when water is the solvent. It cancels out, and including it throws off every calculation downstream.
Gas-phase equilibria expressed in terms of partial pressures require converting between Kp and Kc using the relationship Kp = Kc(RT)^n, where n is the change in moles of gas. This conversion is straightforward but easy to get wrong if you mix up the sign of n or use the wrong value of R. I keep a reference sheet with the common constants because relying on memory for those details has cost me points on exams and confidence in design calculations. If you want to practice these calculations, there are several free online calculators and worksheets available. Look for ones that show the ICE table setup rather than just giving you the answer. The process of building the table yourself is where the understanding actually happens. Apps and solvers are useful for verification, but they will not teach you to spot when a problem is set up incorrectly or when your answer is physically unreasonable. The bottom line is that chemical equilibrium is a framework, not a law of nature that applies perfectly to every real system. It works well when you respect its assumptions and know when those assumptions break down. Most mistakes come from applying the model beyond its valid range, not from misunderstanding the model itself.