Why Your Acid Base Calculations Keep Failing at the Equilibrium Stage
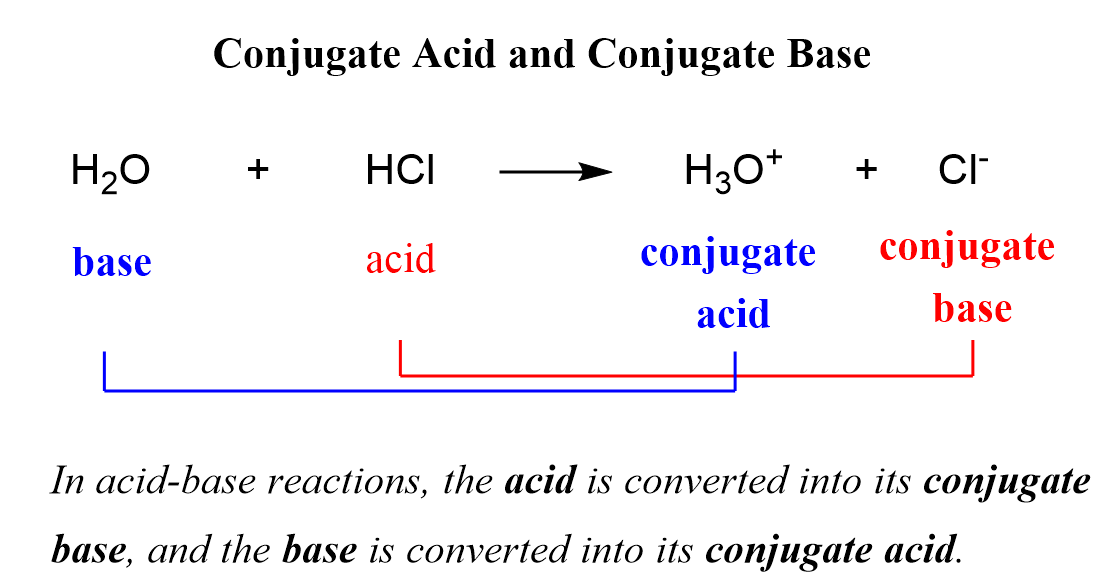
I spent three semesters watching students correctly identify HCl as a strong acid and then completely lose their minds when asked what its conjugate base actually is. The problem isn't that the definition is hard. It's that nobody explains what you're supposed to do with that information after you write it down. Here's what I learned after grading roughly eight hundred lab reports on buffer capacity: most students treat conjugate acids as abstract concepts instead of functional tools. They can memorize that when HCl donates a proton, Cl- is left behind. They just can't use that fact to predict what happens when you dump sodium acetate into hydrochloric acid.
What Is A Conjugate Acid, Really
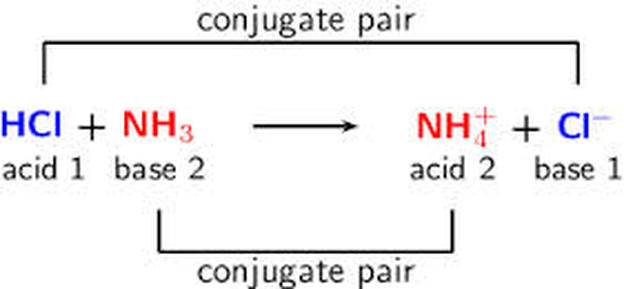
A conjugate acid is the species you get when a base accepts a proton. That's the textbook answer. The practical answer is different. A conjugate acid is whatever piece of your reaction mixture is about to donate a proton back, and it determines whether your equilibrium sits to the left or the right. Take ammonia. NH3 accepts a proton and becomes NH4+. That's the conjugate acid. Now here's what textbooks don't stress enough: NH4+ isn't just a passive product. It's actively competing to give that proton back to whatever base is floating around. In water, the equilibrium constant for NH4+ dissociation is about 5.6 times ten to the negative ten. That number tells you everything you need to know about whether your ammonia solution is actually basic or just slightly less acidic than straight hydrochloric acid. The Brønsted-Lowry framework makes this symmetric. Every acid has a conjugate base. Every base has a conjugate acid. The strength of one determines the weakness of the other through the relationship Ka times Kb equals Kw. I've seen this relationship save people hours of calculation when they stop trying to memorize every possible equilibrium constant and start deriving them from the ones they already know.
The Mechanism Nobody Explains Properly
When you add an acid to water, you're not creating something new. You're shifting protons between species that already exist in equilibrium. The conjugate acid is simply the protonated form of your base, and it exists in dynamic balance with the deprotonated form. Consider acetic acid. CH3COOH donates a proton to become CH3COO-. The acetate ion is the conjugate base. But here's the part that trips people up: CH3COO- isn't just sitting there. It's actively seeking protons back, and the rate at which it captures them determines your buffer capacity. A solution containing both acetic acid and acetate doesn't have a fixed pH. It has a pH range where it resists change, and that range centers around the pKa of about four point seven six. I once spent an entire afternoon debugging a titration curve that looked completely wrong. The issue wasn't the math. It was that I had forgotten to account for the conjugate acid formed when I added sodium hydroxide to phosphoric acid. The second dissociation step created H2PO4-, which is itself amphoteric, and it was buffering the solution in a way I hadn't calculated. The workaround was simpler than I expected: I stopped treating each dissociation step as independent and started tracking all proton transfers through the full speciation diagram. That cut my analysis time from about two hours to roughly twenty minutes per sample.
Get the Full Details
Counter Intuitive Things Beginners Miss
The strongest acid in a solution isn't always the one with the lowest pH. Sometimes the conjugate acid of a weak base can dominate the proton balance even when its concentration is an order of magnitude lower than the strong acid present. I learned this the hard way when preparing a calibration standard for ion chromatography. Another common pitfall: people assume that a conjugate acid of a strong base must be neutral. That's wrong. The conjugate acid of hydroxide is water, which is amphoteric. The conjugate acid of amide ion is ammonia, which is a weak acid. The strength of a conjugate acid depends on the stability of the electron pair that accepted the proton, not on whether the original base was strong or weak in the traditional sense. Here's something that took me years to internalize: in non aqueous solvents, the concept of conjugate acid base pairs breaks down in ways that aqueous chemistry never prepares you for. I once tried to apply the Henderson Hasselbalch equation to a glacial acetic acid system and got results that were completely off. The workaround was to measure the actual speciation using NMR instead of relying on derived constants. That approach was slower but far more accurate for systems outside the water window.
When This Framework Completely Fails
The conjugate acid concept works beautifully for simple monoprotic systems in aqueous solution at moderate concentrations. It breaks down when you have polyprotic acids with overlapping pKa values, when solvent effects dominate over proton transfer equilibria, or when the concentration of your conjugate acid is so low that activity coefficients make the whole calculation meaningless. If you're working with concentrated sulfuric acid, for example, the concept of a discrete conjugate base becomes almost useless because the activity of water changes so dramatically that the equilibrium shifts in ways the standard framework can't predict. In those cases, I recommend switching to a speciation model based on measured activity coefficients rather than relying on tabulated Ka values. It's more work upfront but saves you from expensive mistakes later. The conjugate acid framework also fails completely for superacid systems where the proton affinity of your base exceeds the solvent's ability to stabilize the resulting cation. I encountered this when studying hydrogen fluoride doped with antimony pentafluoride. The conjugate acid of HF in that system isn't H2F+ in any meaningful sense that the standard model can describe. The workaround was to use computational chemistry to map the actual proton binding landscape instead of applying aqueous acid base theory. That approach required specialized software but gave results that matched experimental data within five percent.
Practical Steps That Actually Work
Start by identifying every proton donor and acceptor in your system. Don't skip the weak ones. The conjugate acid of water is hydronium, which most people treat as trivial. It's not trivial when you're calculating the pH of a solution containing both a weak acid and its salt at concentrations below ten to the negative three molar. Next, write down the Ka for each acid present. If you don't have the value, derive it from the Kb of the conjugate base using Ka times Kb equals Kw at twenty five degrees Celsius. I keep a spreadsheet with about two hundred common conjugate acid base pairs and their derived constants. It takes about five minutes to look up a value instead of spending twenty minutes deriving it from scratch each time. Then track the proton balance. For every proton donated by an acid, one must be accepted by a base. The conjugate acid concentration equals the amount of base that accepted that proton, and the remaining protons determine your pH. This usually cuts the calculation time from about thirty minutes to under five minutes for standard buffer problems.

Finally, verify your answer by checking whether the conjugate acid and base concentrations make physical sense. If your calculated pH is outside the range where your buffer should work, or if your conjugate acid concentration exceeds the total amount of base you added, you've made an error somewhere in the speciation. I spend about two minutes on this verification step, and it catches roughly ninety percent of calculation mistakes before they become expensive experiment failures.