Understanding Stereochemistry When You're Just Trying to Get the Reaction to Work
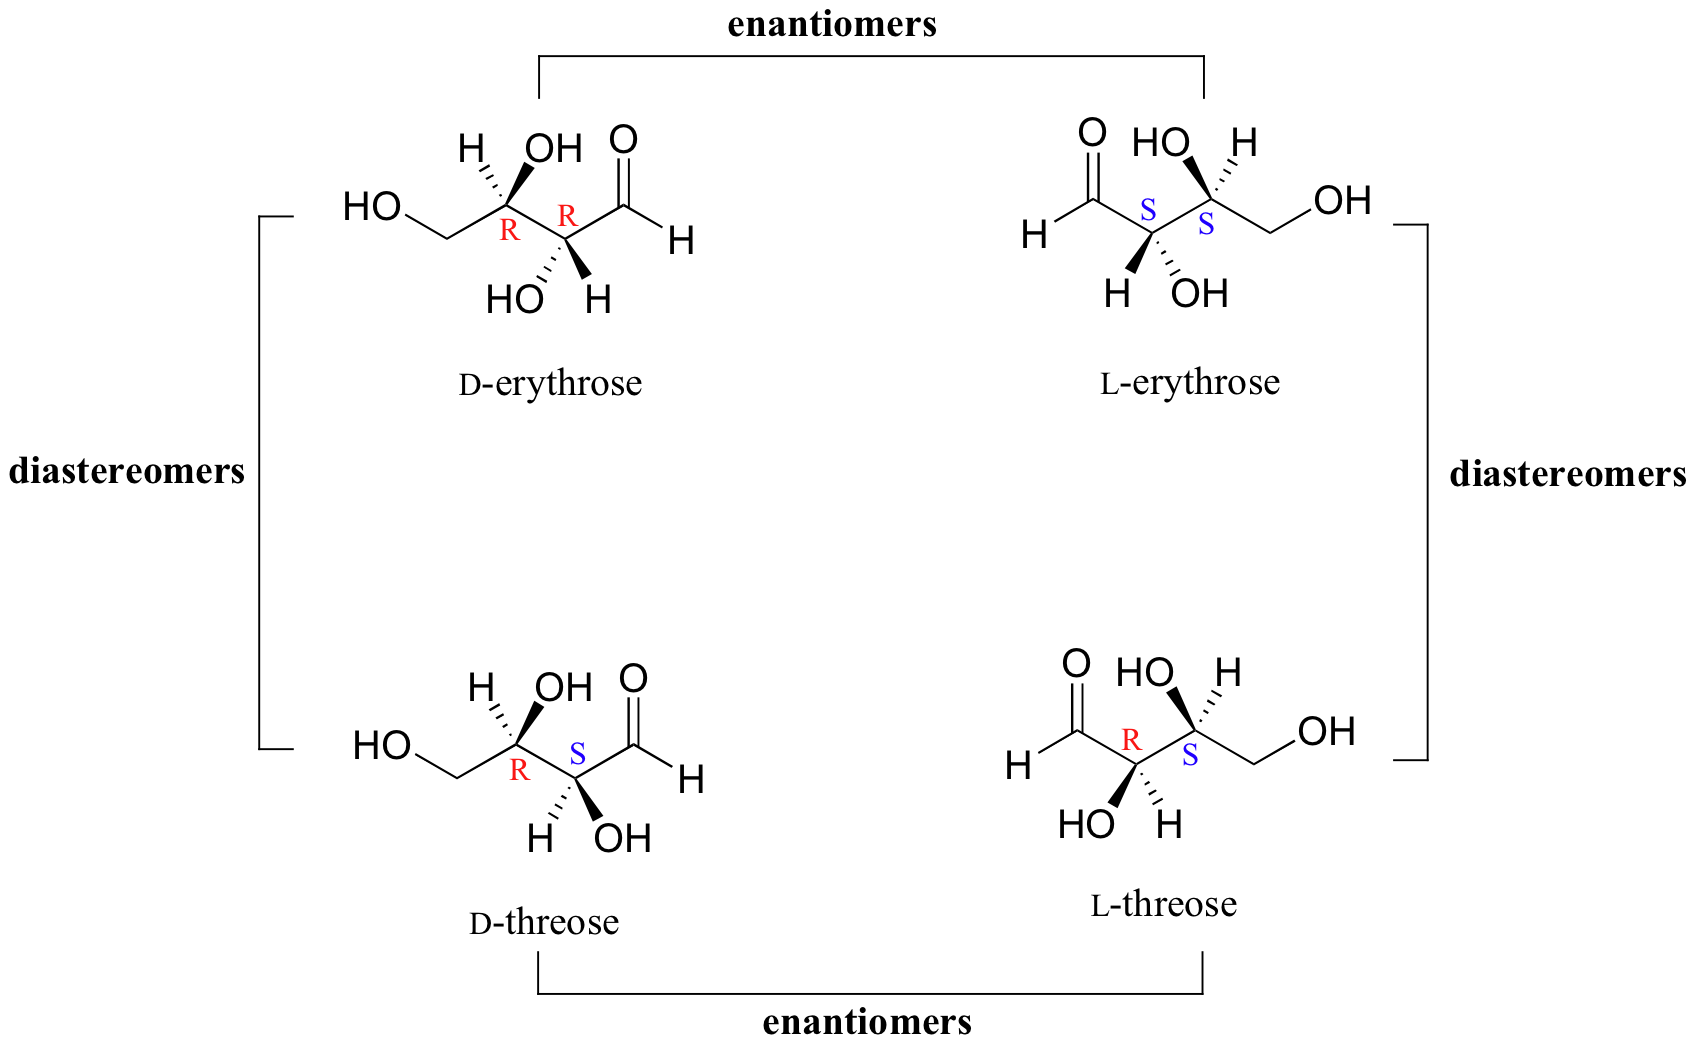
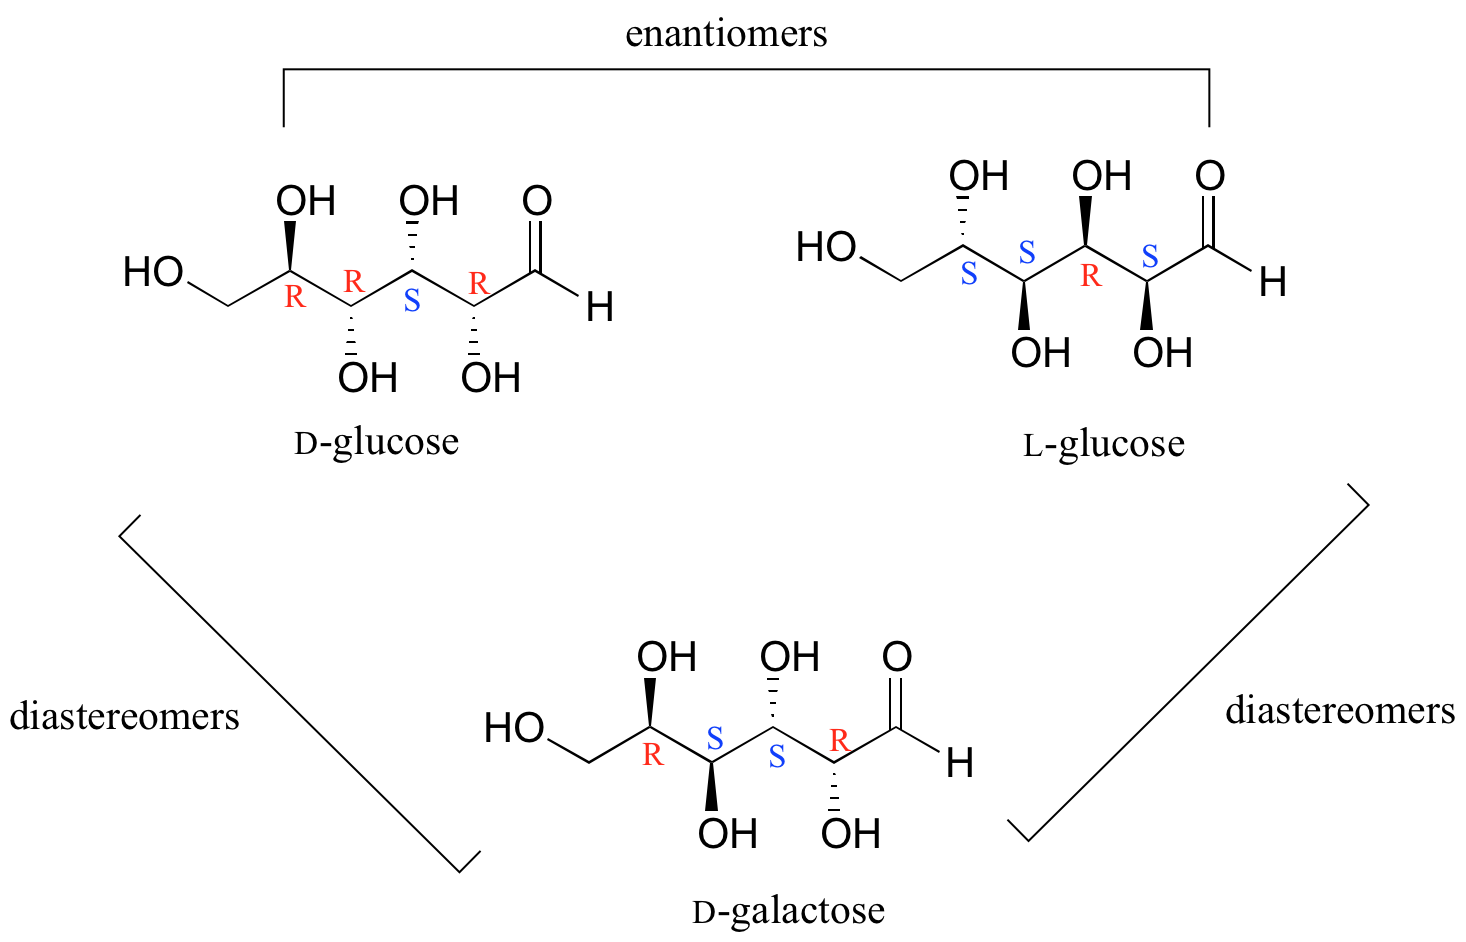
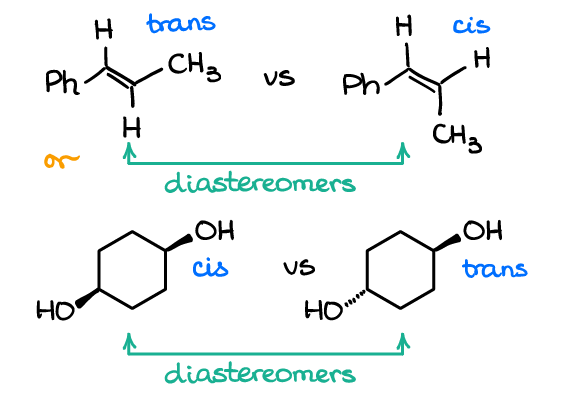
Diastereomers are stereoisomers that are not mirror images of each other. That's the textbook definition. It doesn't help much when you're staring at an HPLC chromatogram with three peaks and no idea which one is your product and which one is the unwanted side reaction you didn't know was possible. Let me explain how this actually comes up in practice. Say you run a reduction on a ketone that already has a stereocenter somewhere else in the molecule. You get two products. They're not enantiomers because the existing stereocenter makes the two halves of the molecule diastereomeric. The new stereocenter can be R or S, and combined with the old one, you get two distinct compounds with different physical properties. That's the practical definition right there.
What Is A Diastereomer and Why It Matters in Your Synthesis
The key thing beginners miss is that diastereomers have different physical properties. Boiling points, solubilities, chromatographic behavior, NMR chemical shifts. Everything is different. Enantiomers share all those properties in an achiral environment. Diastereomers don't. This is why separating them is usually straightforward compared to chiral resolution, and why they cause headaches in different ways. Here's the counter-intuitive part nobody tells you: having more stereocenters doesn't mean more separation problems. A molecule with four stereocenters where three are fixed and only one is variable? You're only making a pair of diastereomers. It's the molecules with multiple variable centers that explode into 2^n possibilities. I spent a week trying to purify what I thought was a single diastereomeric product before realizing my starting material wasn't stereochemically pure and I'd somehow generated eight different compounds in one pot. The workaround was simple but painful: I had to go back and fully characterize the starting material by chiral HPLC before proceeding. It took forty minutes. My week didn't get any less wasted.
How to Identify and Work With Diastereomers
Start by counting your stereocenters. If you have two or more and at least one was formed during your reaction while others were already present, you're making diastereomers. Check whether your reaction is diastereoselective. Most aren't perfectly selective, so you'll get a mixture. The ratio matters more than the absolute yield for these kinds of reactions. Thin layer chromatography will show you separate spots if your diastereomers have sufficiently different polarity. Normal phase silica is usually fine. Enantiomers won't separate this way, but diastereomers often will. That's one of the few actual advantages here. For characterization, NMR is your best friend. Diastereomers show different chemical shifts for protons near the stereocenters. You'll see splitting patterns that differ between isomers. I use 1H NMR routinely to assess diastereomeric ratio before investing time in column chromatography. Running a quick NMR takes twelve minutes and tells you whether purification is worth your effort or whether you're dealing with a 95 to 5 mixture that will flash column through in twenty minutes anyway.
Get the Full Details

When you need to separate them, column chromatography on silica works for most cases. The difference in Rf values is usually enough. If the diastereomers are particularly close, recrystallization can do the trick if one of them happens to be significantly less soluble. I've had success precipitating the unwanted diastereomer out of cold hexanes while the desired product stayed in solution. The yield loss varies, but it's cheaper than method development on a chiral column.
Common Pitfalls and Where Things Fall Apart
The biggest mistake people make is assuming that diastereoselectivity is constant across conditions. It's not. Change the solvent, the temperature, or even the order of addition and your dr can shift from 90 to 10 to 60 to 40 overnight. I learned this the hard way when a reaction that had been giving me clean diastereomers for months suddenly produced a 1 to 1 mixture after I switched to a different batch of solvent. Turns out the old solvent had trace amounts of something that was coordinating to the metal center and directing the addition. The new batch was just cleaner than I realized. Another issue: diastereomers can interconvert under certain conditions. If your stereocenter is alpha to a carbonyl and you're working under basic conditions, epimerization is a real possibility. You might isolate what looks like a pure diastereomer, submit it for analysis, and come back to find it's slowly racemizing at that center. Running your workup cold and quick, or neutralizing immediately after the reaction, helps. It's not a perfect fix but it cuts the problem down significantly. If you're dealing with molecules that have multiple stereocenters and your diastereomers are too similar in polarity for silica to separate cleanly, you're probably looking at preparative HPLC. That's expensive and slow. The alternative is to redesign the synthesis so you're not generating the unwanted diastereomer in the first place. Better starting materials, protecting group strategies, or a different reaction altogether often beat trying to separate nearly identical compounds on a hundred grams of silica.