
Understanding Atoms in Computational Chemistry
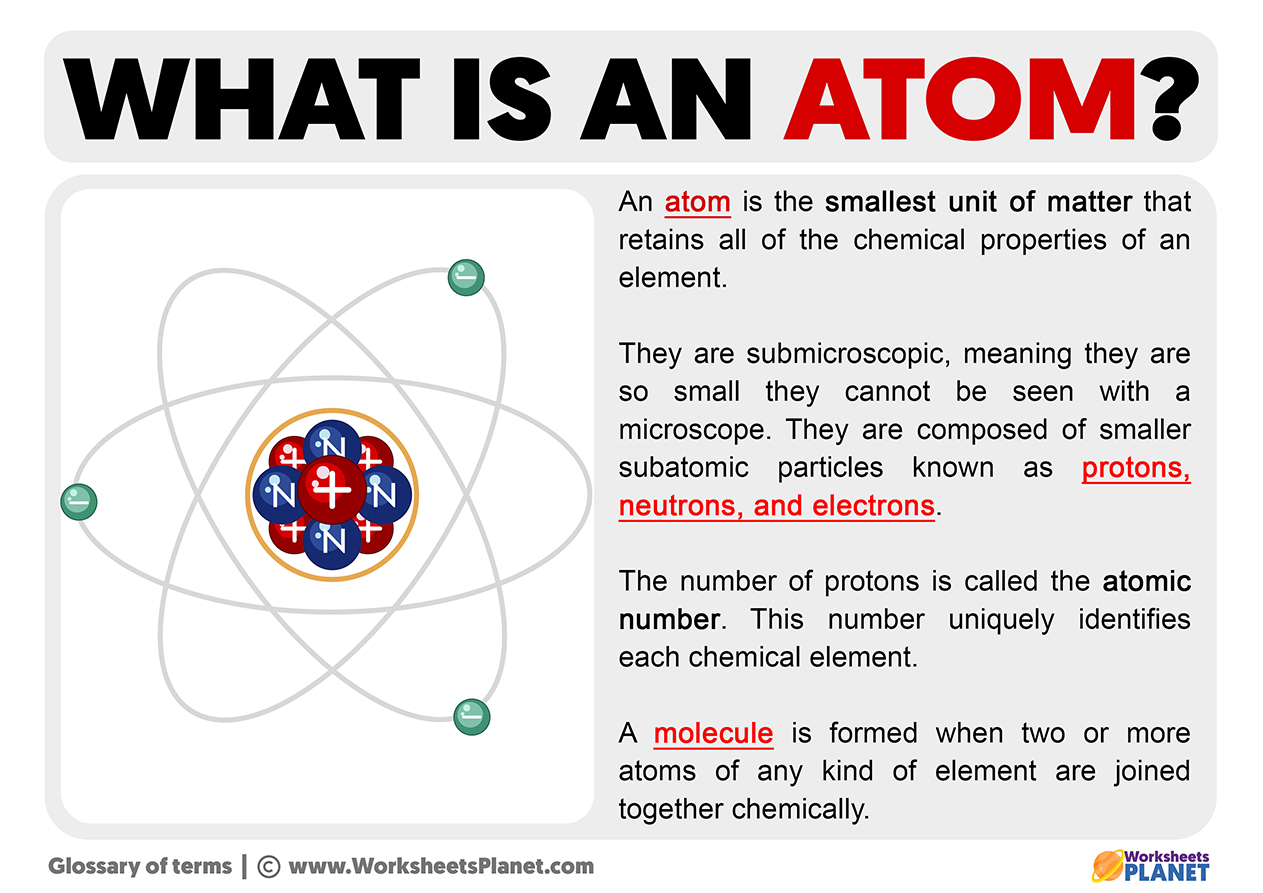
An atom is a neutral chemical species consisting of a nucleus containing protons and neutrons, surrounded by electrons occupying quantized orbitals. That is the textbook version. The version that matters when you are actually running calculations involves electron configuration, effective nuclear charge, and how basis sets approximate the wavefunction around that nucleus. Most people learning this subject hit a wall pretty quickly because they try to memorize definitions instead of understanding what happens when you feed atomic parameters into a program like Gaussian, ORCA, or Psi4. The theory is fine until it is not, and the problems show up in your output files, not in your textbook.
What Is An Atom and Why Does It Matter in Simulations
When you run a quantum chemistry calculation, you are essentially telling the computer where each nucleus sits and what kind of atom it is, then asking the program to solve the Schrödinger equation approximately. The input file for a simple water molecule looks almost nothing like the finished result. You define atoms by element symbol and Cartesian coordinates, and the program builds a basis set around each one. How well that basis set represents the actual electron density determines everything about your final numbers. I spent three weeks trying to debug why my benzene binding energy was off by about 4 kcal/mol compared to the literature value. Turned out the default basis set was missing diffuse functions on the oxygen atoms in my counterion model. Adding the ++ symbol to the basis set designation fixed it immediately. This is not something most tutorials mention upfront. The core concept is that an atom in a molecule is not the same as a free atom. Electrons redistribute, orbitals hybridize, and the effective potential changes depending on what neighbors are around. This is why atomic positions in your geometry matter as much as the element type itself. A carbon atom in a CH3 group behaves differently from one in a C=O group, and your calculation needs to capture that distinction.
Setting Up an Atomic Calculation From Scratch
Start with ORCA since it is free for academic use and handles most standard atomic calculations without a license server headache. Download it from the official server and make sure your system has OpenMPI installed. The binary runs on Linux, macOS, and Windows through WSL2. Create a input file with this structure for a single atom calculation: * xyz 0 2
Get the Full Details
C 0. 0. 0. * This tells ORCA you are calculating a carbon atom with a doublet spin state. Then add your method and basis set lines below it. A reasonable default for testing is BP86 with the def2-SVP basis set. For production work you would want def2-TZVP at minimum, ideally with RIJCOSX approximation enabled to cut runtime significantly.
The command to run it is straightforward: orca input.inp. The output will contain your final energy, optimized geometry if you asked for it, orbital energies, and a whole lot of numbers you probably do not need to read manually.
Common Pitfalls and How to Avoid Them
Convergence failure is the most frequent issue beginners encounter, and it usually has nothing to do with the atom itself. It is almost always a problem with your initial guess or your geometry. If you start with coordinates that put atoms too close together, the SCF cycle will oscillate and crash. Always check your interatomic distances before running anything. A C-C bond shorter than 1.0 Angstroms is a red flag. Another problem that trips people up repeatedly is spin contamination in unrestricted calculations. When you run an UHF or UDFT calculation on an open-shell atom like nitrogen, the expected spin state is a quartet, but the actual calculated S^2 value might come out to 2.1 or 2.4 instead of the ideal 1.875. This means your wavefunction is mixing in higher spin states and your energies are unreliable. The fix is usually to use a broken-symmetry approach or switch to a restricted open-shell method if your program supports it. ORCA handles this with the %uhf block and the noscfmix true option. Here is a specific edge case I ran into last year. I was calculating the ionization energy of several transition metal atoms, and the values were consistently 0.3 eV too low compared to NIST reference data. The issue turned out to be that the default grid size in ORCA was too coarse for the steep electron density near the transition metal cores. Switching the integration grid from C_Def2_Standard to C_Def2_Tight improved the agreement to within 0.05 eV. This is a settings detail that is never mentioned in the documentation unless you dig through the mailing list archives.
When Standard Approaches Break Down
DFT works reasonably well for main group atoms but has well-known failures with transition metals and lanthanides. Self-interaction error causes excessive electron delocalization, which makes your calculated atom sizes too large and your binding energies too weak. If you are working with systems where these errors matter, you need to either use a hybrid functional with some exact exchange mixed in, or go to a wavefunction-based method like CCSD(T). Both options cost significantly more computational time. A single-point energy calculation on a mid-size transition metal complex that takes 30 minutes with BP86 can take 8 hours with CCSD(T). Pure DFT also struggles with van der Waals interactions between atoms. If your system involves noble gas atoms or weakly bound complexes, standard functionals will give you essentially wrong answers. You need a dispersion-corrected functional like BP86-D3(BJ) or a range-separated hybrid. The difference in result can be 5 to 10 kcal/mol for a typical non-covalent interaction, which is enormous in chemical terms. There is no universal solution here. You pick the method that matches your system and accept the trade-offs. For most atoms and molecules in organic chemistry, a hybrid functional with a triple-zeta basis set and dispersion correction will give you results good enough for publication. For anything involving excited states or strong correlation, you are entering territory where the answers come with bigger error bars and longer run times regardless of what you do.
Reading Your Results Correctly
The most important number in your output is the final SCF energy, usually labeled as -E or total energy. Do not compare this directly to literature values from other programs or different basis sets. It means nothing in absolute terms. What matters is energy differences between similar calculations, like reaction energies or ionization potentials, where systematic errors cancel out. Orbital energies from DFT are sometimes interpreted as ionization potentials through Koopmans' theorem, but this is only an approximation and often off by several eV. For accurate vertical ionization energies, use Delta-SCF by calculating the neutral and ionized states separately and taking the difference. It takes twice the computational effort but gives you results that actually match experiment. If you need reference data to validate your calculations against, the NIST Computational Chemistry Comparison and Benchmark DataBase is the standard source. It lists experimental and high-level theoretical values for atom and molecule properties that you can use to check whether your method and basis set choice are reasonable for your specific application.


