So You Need to Put DNA Into Bacteria. Here's How It Actually Goes.
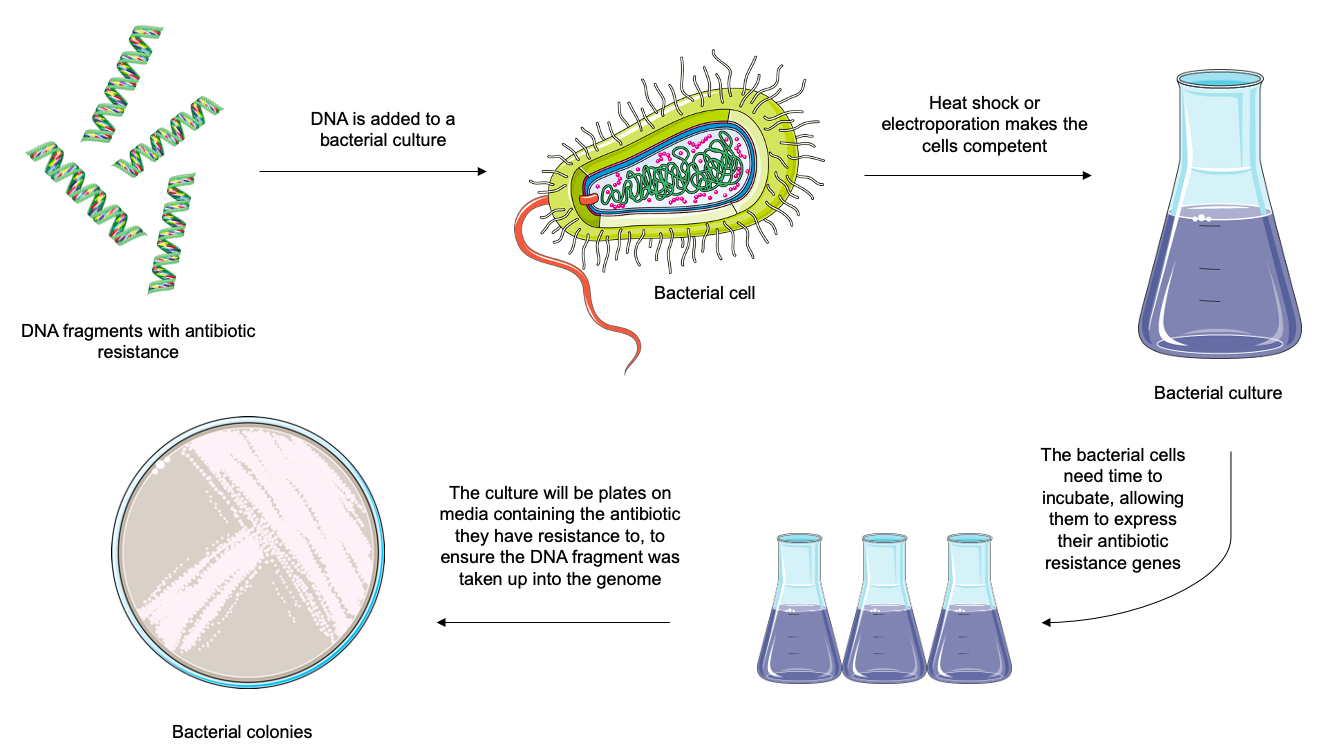
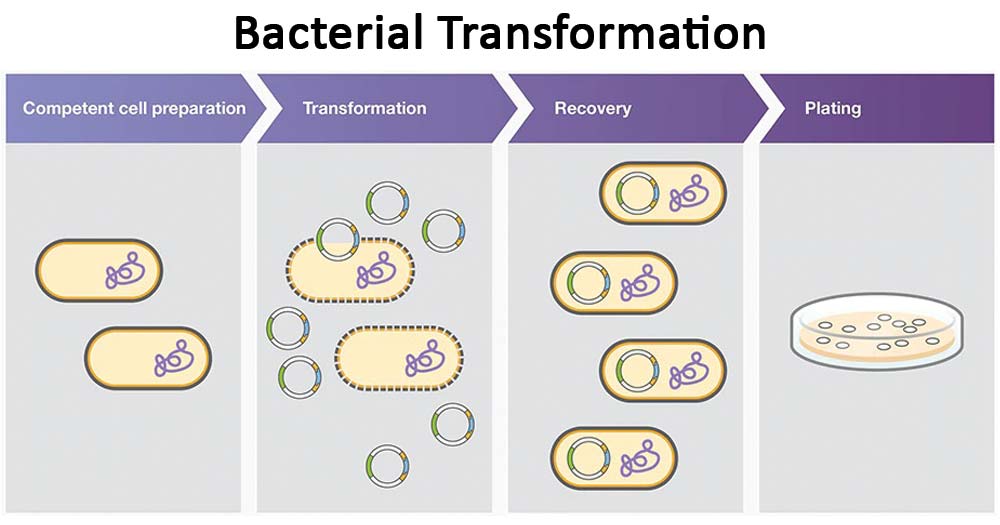
The first thing that goes wrong is always the competence. You buy a kit, you follow the protocol, and half the time your cells just won't take up any plasmid. Bacterial transformation is the process of introducing foreign DNA into a bacterial cell, usually E. coli, so that the cell expresses whatever gene you stuck on a plasmid. That's the textbook version. In practice it's a lot more fiddly than one sentence makes it sound. At its core, you're making cells permeable to DNA, letting a circular plasmid slip inside, and then giving the bacteria a chance to recover and express the new genetic material. The plasmid carries an antibiotic resistance gene, which is how you screen for success later. You plate the cells on agar with that antibiotic, and only the ones that actually picked up your plasmid will grow colonies. Everything else dies. Simple in theory. Not simple when you're waiting twelve hours for results and nothing is growing. Here's the standard heat-shock method, the way most people do it in a teaching lab or a basic research setup.
You grow your E. coli strain to mid-log phase, usually an OD600 between 0.4 and 0.6. Harvest the cells by centrifugation at around 4000 rpm for ten minutes. Resuspend them in cold calcium chloride, typically 50 to 100 millimolar, on ice for at least thirty minutes. This is what makes the cells chemically competent. The calcium ions neutralize the negative charges on both the cell membrane and the DNA phosphate backbone, which reduces electrostatic repulsion and lets the plasmid get close enough to the membrane to enter. Add your plasmid DNA, somewhere between 1 and 100 nanograms, and incubate on ice for another twenty to thirty minutes. Then heat shock at 42 degrees Celsius for exactly thirty to forty-five seconds. This thermal gradient creates a temporary imbalance across the membrane that drives the DNA inside. Immediately put the tube back on ice for two minutes. Then add about 500 microliters of rich media like SOC or LB and shake at 37 degrees for about an hour. This recovery period is non-negotiable. The bacteria need time to express the antibiotic resistance gene before you expose them to the selective agent. Skip this and you'll get zero colonies even if the transformation worked perfectly. Plate whatever volume makes sense on your selective agar and incubate overnight at 37 degrees. Usually 16 to 18 hours depending on how fast your strain grows and how much DNA you started with.
I once spent three days troubleshooting why a perfectly good plasmid prep yielded zero colonies. The protocol was correct, the cells were fresh, the heat shock was right on time. Turns out the ampicillin in my agar plates had degraded. Old stocks break down, especially if they've been sitting at room temperature or cycling through freeze-thaw. The colonies should have been there. I made fresh plates with a new ampicillin stock and got five hundred colonies on the first try. Worth mentioning because this is the kind of thing that will waste your week if you don't catch it. There are some things about transformation efficiency that people don't tell you until they've burned through enough reagents. Competent cell quality degrades faster than you think. Even stored at minus 80, aliquots lose efficiency after about three months. If you're doing something that requires high efficiency, like library construction or rare cloning events, buy fresh cells or make your own. Home-made calcium chloride competent cells will give you somewhere around 10 to the 6th to 10 to the 7th efficient transformants per microgram of DNA. Good commercial preps can hit 10 to the 8th or 9th. That difference matters when you're trying to clone something difficult.
Get the Full Details

The morphology of your plasmid matters too. Supercoiled plasmid transforms far more efficiently than linearized or nicked DNA. If your prep looks smearable on a gel instead of showing a clean supercoiled band, your transformation numbers are going to suck. I've seen people blame their technique when the real problem was a degraded plasmid prep. Run a gel before you waste a whole day on transformation. Another thing that bites people: the volume of your ligation product. If you're doing a direct ligation and transformation, don't add more than about 10 microliters of ligation mix to your competent cells. Too much buffer from the ligation reaction, especially if it contains PEG or high salt, can inhibit the transformation itself. Dilute it or clean it up if you need to add more DNA. Electroporation is the other major route and it's worth knowing when to use it instead. Chemical competence is fine for routine cloning, but if you're working with large plasmids above 10 kilobases, or trying to transform hard-to-transformation strains like certain BAC hosts or recA-minus strains for specialized applications, electroporation gives you orders of magnitude better efficiency. You need electroporation-competent cells prepared in pure water with no salt, a 2 kilovolt pulse in a 0.1 centimeter cuvette, and immediate recovery in warm media. The yield is better but the setup is more finicky and the cells are more fragile.
If you're working with Gram-positive bacteria or other non-E. coli hosts, none of this applies directly. Those organisms have completely different cell wall structures and usually require completely different protocols, often involving lysozyme treatment or protoplast formation. Don't try to force an E. coli protocol onto Bacillus or Streptomyces and wonder why it fails. The quick summary: grow cells to the right density, keep everything cold until the heat shock, recover properly, use fresh antibiotic, and verify your plasmid quality before you blame your technique. That covers the vast majority of problems people run into.