Working With Enthalpy Changes in Practice
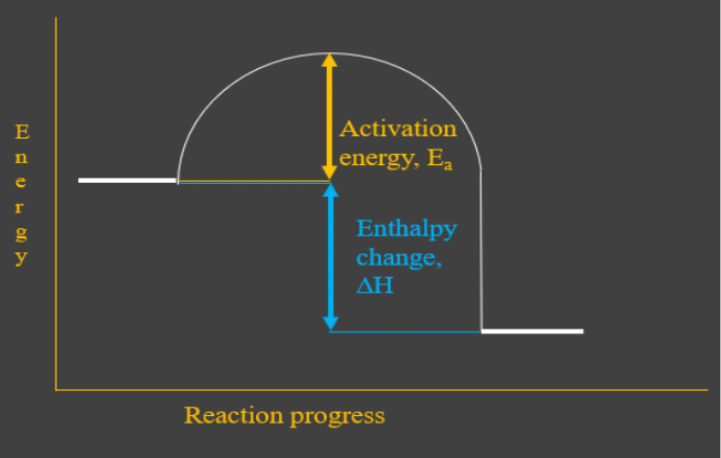
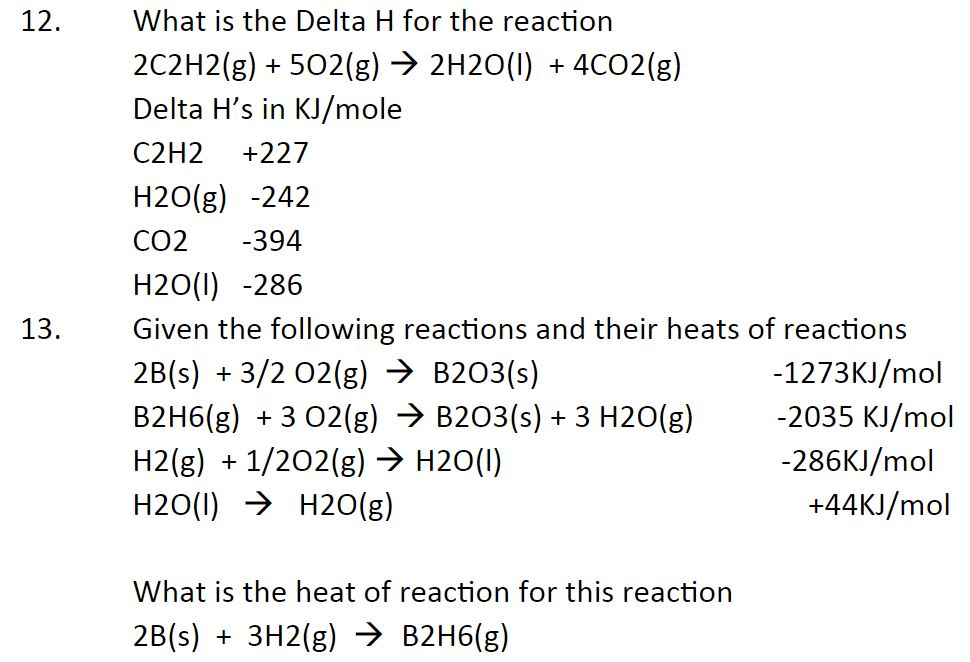
When you're running calorimetry experiments or pulling data from thermodynamic tables, understanding enthalpy change is less about memorizing a definition and more about knowing where the numbers lie and what they don't tell you. Most people first encounter this in a general chemistry class, which is where they also learn to treat it as some abstract symbol on a whiteboard. The gap between that and actually using it in a lab or process design is significant enough that it trips people up constantly.I remember working on a project calculating the heat release from a neutralization reaction, and the literature value said it should be around -57 kJ/mol for strong acid and strong base. The experimental result came out at -43 kJ/mol. Not a dramatic deviation on paper, but enough to completely mess up a downstream heat exchanger calculation. The problem turned out to be that the concentrated stock solutions weren't at standard conditions, and the heat capacity of the resulting salt solution was different than what the dilute-solution assumption predicts. You have to account for that. It's easy to overlook until you've been burned by it once. Delta H, written as H, represents the change in enthalpy of a system during a process at constant pressure. Enthalpy itself is a state function combining internal energy with the product of pressure and volume (H = U + PV). The delta tells you how much heat is absorbed or released. Positive values mean endothermic — the system takes in heat. Negative values mean exothermic — the system releases it. That's the textbook version. Here's the practical version: when you're designing a reactor or sizing a cooling system, getting the sign wrong means your equipment either can't handle the thermal load or it's massively oversized and wasting capital. One thing beginners consistently miss is that H is path-independent. The actual process your reaction takes — whether it happens in one violent step or slowly through intermediates — doesn't change the overall enthalpy change. What matters is the initial and final states. This is why Hess's Law works. You can add and subtract known reactions to find an unknown H without ever running that unknown reaction in the lab. I use this all the time when a direct measurement would be impractical or dangerous.
Another nuance that doesn't get enough attention: H depends on temperature. The value you look up in a table at 298 K isn't necessarily the right value for your process running at 450 K. To correct for that, you need the heat capacities of reactants and products as functions of temperature. The relationship is the Kirchhoff equation: H(T) = H(T) + Cp dT from T to T. If Cp is roughly constant over your temperature range, you can simplify it to H(T) H(T) + Cp(T - T). Most tables give you Cp values, so this is straightforward. The trap is assuming Cp is zero just because you don't have data for it. That assumption introduces error that compounds fast at higher temperatures.
How to Calculate It Without Making Expensive Mistakes
There are three main routes to getting a H value, and each has its own failure modes. Route one: standard enthalpies of formation. Look up Hf° for every reactant and product, then subtract: H°rxn = nHf°(products) - mHf°(reactants). Standard conditions here mean 1 bar pressure and usually 298.15 K. The NIST Chemistry WebBook is the most reliable source I've found. Be careful with phases — the Hf° for HO(l) is -285.8 kJ/mol while HO(g) is -241.8 kJ/mol. That 44 kJ/mol difference is the latent heat of vaporization, and using the wrong one is the single most common error I see in student lab reports and early-career engineering calculations alike. Route two: bond energies. This works for gas-phase reactions where you're breaking and forming bonds. Sum the bond energies of bonds broken (positive, energy in) and subtract the sum of bond energies of bonds formed (negative, energy out). It's approximate because bond energies are averages across many molecules. A C-H bond in methane isn't identical to a C-H bond in ethanol. If you need precision, don't use this method. But for quick estimates, it's useful and faster than looking up formation data for every compound.
Get the Full Details
Route three: calorimetry. This is the direct measurement approach. q = mcT for solution calorimetry, where m is mass, c is specific heat capacity, and T is the temperature change. For a coffee-cup calorimeter at constant pressure, q equals H. Bomb calorimeters run at constant volume, so you get U instead and need to convert: H = U + n_gRT. That n_g term is the change in moles of gas. People forget it exists. When I'm doing bomb calorimetry work, I always double-check that conversion because the difference can be several kilojoules per mole for reactions involving gases.
When This Approach Breaks Down
Constant pressure is baked into the definition of H, which means any process where pressure varies significantly — shock waves, rapid explosions, some high-pressure geochemical systems — H alone doesn't fully describe the energy dynamics. In those cases you need to go back to first principles with internal energy and work terms. Also, for non-ideal solutions, especially at high concentrations, the simple Hf° tables assume infinite dilution behavior. Real concentrated electrolyte solutions can deviate substantially. Activity coefficients matter there, and you'd need excess enthalpy data from specialized sources rather than relying on standard tables. Another hard limitation: phase transitions near critical points. As you approach the critical temperature and pressure of a substance, the distinction between liquid and gas vanishes, and the usual tabulated values for enthalpy of vaporization go to zero. Equations of state like Peng-Robinson or Soave-Redlich-Kwong become necessary instead of simple table lookup. I ran into this explicitly when modeling a supercritical CO extraction process — the enthalpy calculations near the critical point were throwing off mass and energy balances by 15 percent or more until I switched to a proper EOS-based property package.
Practical Workflow I Use
Start by identifying the reaction and the conditions. Check whether standard tables apply or whether you need temperature corrections. Pull Hf° values from NIST, verify the phases match your actual system, and compute the standard reaction enthalpy. If your operating temperature differs from 298 K, gather Cp data for each species — polynomial coefficients are typically available in the same databases — and apply Kirchhoff's equation. Validate against any available experimental data. When nothing matches, flag the uncertainty and document which assumptions drove the result. That documentation step is what separates a back-of-the-envelope guess from something you can stand behind when a design review comes along.