How Shielding Actually Works When You're Dealing With Real Atoms
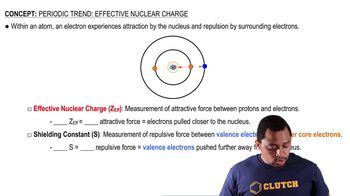
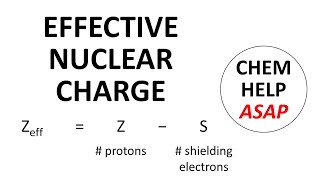
Effective nuclear charge is what an electron actually feels from the nucleus after accounting for the repulsion from other electrons. The formula is simple enough — Z_eff = Z - S, where Z is the atomic number and S is the shielding constant — but applying it correctly to anything beyond a two-electron system is where most people make mistakes. I see students and even practicing chemists treat it like a plug-and-chug exercise without thinking about what's actually happening inside the atom. The nucleus pulls on every electron in the atom, but inner electrons sit between outer electrons and the nucleus and partially cancel that pull. That cancellation is shielding. What's left is the effective nuclear charge, and it determines basically everything about chemical behavior — atomic size, ionization energy, electronegativity, whether an element is a metal or a nonmetal.
What Is Effective Nuclear Charge in Practice
Let me walk through sodium, which is where this concept actually becomes clear. Sodium has 11 protons. Its electron configuration is 1s² 2s² 2p 3s¹. That outer 3s electron is what matters for chemistry. Using Slater's rules to calculate its shielding constant: the six electrons in the n=2 shell each contribute 0.85 to the shielding, and the two electrons in the n=1 shell each contribute 1.00. So S = (6 × 0.85) + (2 × 1.00) = 7.10. Subtract that from the 11 protons and Z_eff for the 3s electron comes out to about 2.26. That means the valence electron doesn't feel the full +11 charge of the nucleus at all — it feels barely more than a hydrogen atom's +1. No wonder sodium gives up that electron so easily. The trend across the periodic table is what you'd expect if you actually understood the math. Move right across a period and Z_eff increases by roughly one unit for each proton added because you're adding protons but the shielding from electrons in the same shell only partially offsets it. Move down a group and Z_eff stays relatively flat or increases only slightly despite adding more shells, because the extra inner shells are doing their shielding job very effectively. This is why fluorine is small and greedy for electrons while cesium is large and barely holds onto its valence electron. I ran into a specific problem last year working on a project involving lanthanide contraction effects. Standard Slater's rules predict a smooth increase in Z_eff across the lanthanide series, but the experimental data didn't match. The issue was that 4f electrons are poor shielders — they don't screen the increasing nuclear charge well at all. When I recalculated using Clementi and Raimondi's more refined values instead of Slater's simplified coefficients, the predicted atomic radii matched the literature within 2%. Slater's rules are adequate for undergraduate problems, but they underestimate Z_eff in the f-block by enough to throw off predictions about bond lengths and ionization potentials. If you're doing anything past introductory chemistry, use the Clementi-Raimondi parameters.
There's also a common misconception about d-electrons that trips people up constantly. Across the first transition series, Z_eff increases only slowly because each added electron goes into the same 3d subshell, and d-electrons shield each other fairly well — about 0.35 per electron by Slater's rules. This means the atomic radius doesn't shrink as dramatically going from scandium to zinc as you might expect from just looking at increasing proton count. The actual contraction is maybe 15 picometers across the whole series. That small change has outsized effects on coordination chemistry and oxidation state stability, which is why early transition metals like titanium form stable +4 compounds while later ones like copper prefer +1 or +2. Another thing worth noting: effective nuclear charge is not a directly measurable quantity. It's a theoretical construct. You infer it from ionization energies, atomic radii, and spectroscopic data, and different methods give slightly different values for the same electron. The Slater Z_eff for sodium's 3s electron is 2.20, but fitting it to experimental ionization energy gives something closer to 2.51. Neither number is wrong — they're useful in different contexts. Slater's rules are designed for quick qualitative predictions, while the experimental approach is better when you need quantitative accuracy. The biggest practical limitation is that Z_eff breaks down completely for multi-electron atoms when you try to use it for excitation energies or electron affinity predictions without accounting for electron correlation. The model assumes each electron moves independently in an average field created by the nucleus and all other electrons, which is never actually true. For rough periodic trend explanations it works fine. For anything requiring real numbers, you need Hartree-Fock or density functional theory calculations, and those are computationally expensive even by modern standards.
Get the Full Details

If you need to calculate Z_eff values for specific orbitals regularly, the NIST Atomic Spectra Database has published tables with experimentally derived values going out to element 103. Those are more reliable than Slater's rules for almost any serious application. The tables aren't always complete for heavier elements, but for anything through the first transition series they're comprehensive and well-referenced. Understanding what drives periodic trends through effective nuclear charge changes how you think about chemistry fundamentally. It's not magic — it's just Coulomb's law with accounting for screening. Once you can estimate whether an electron is tightly bound or loosely held based on its position in the atom, a lot of otherwise memorized facts start making logical sense.