Understanding Enzyme Biology
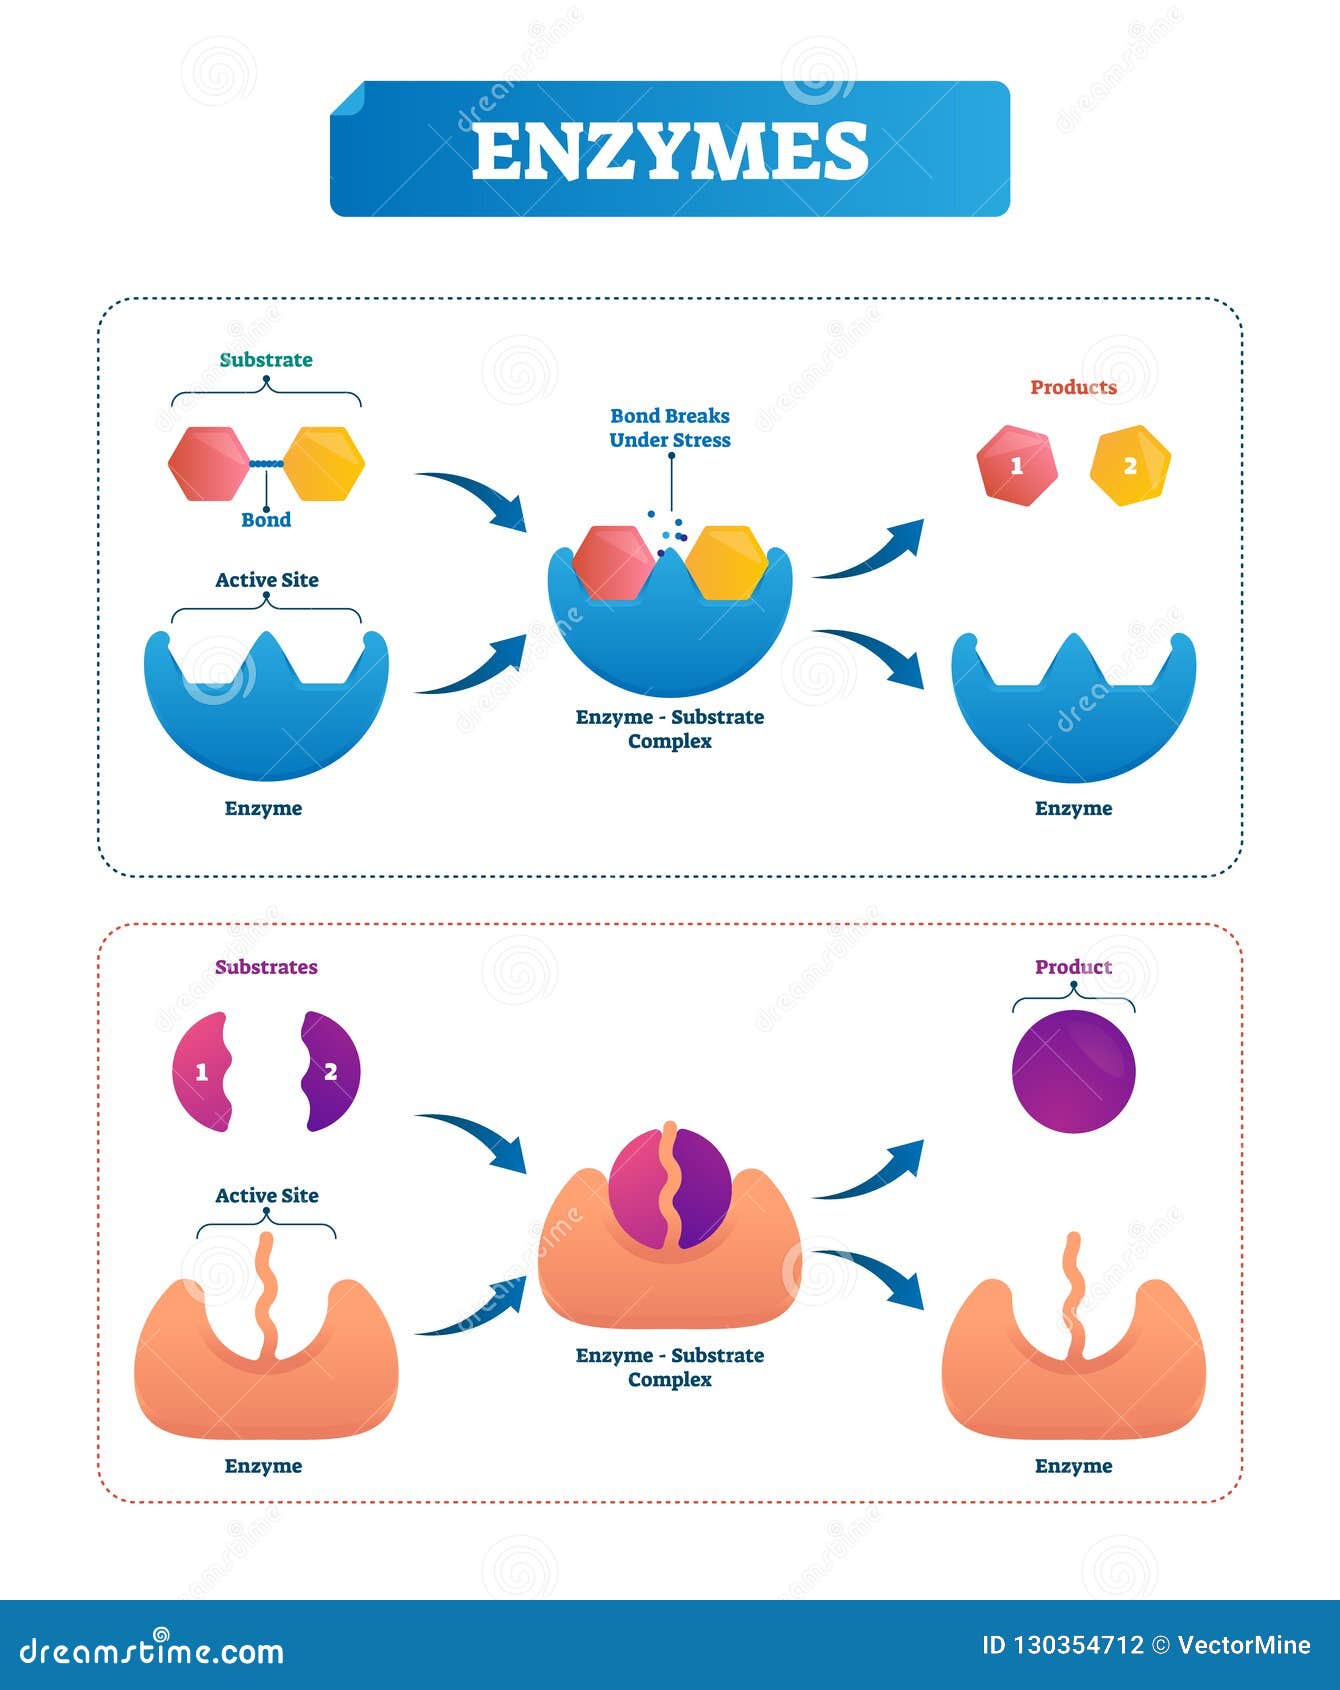
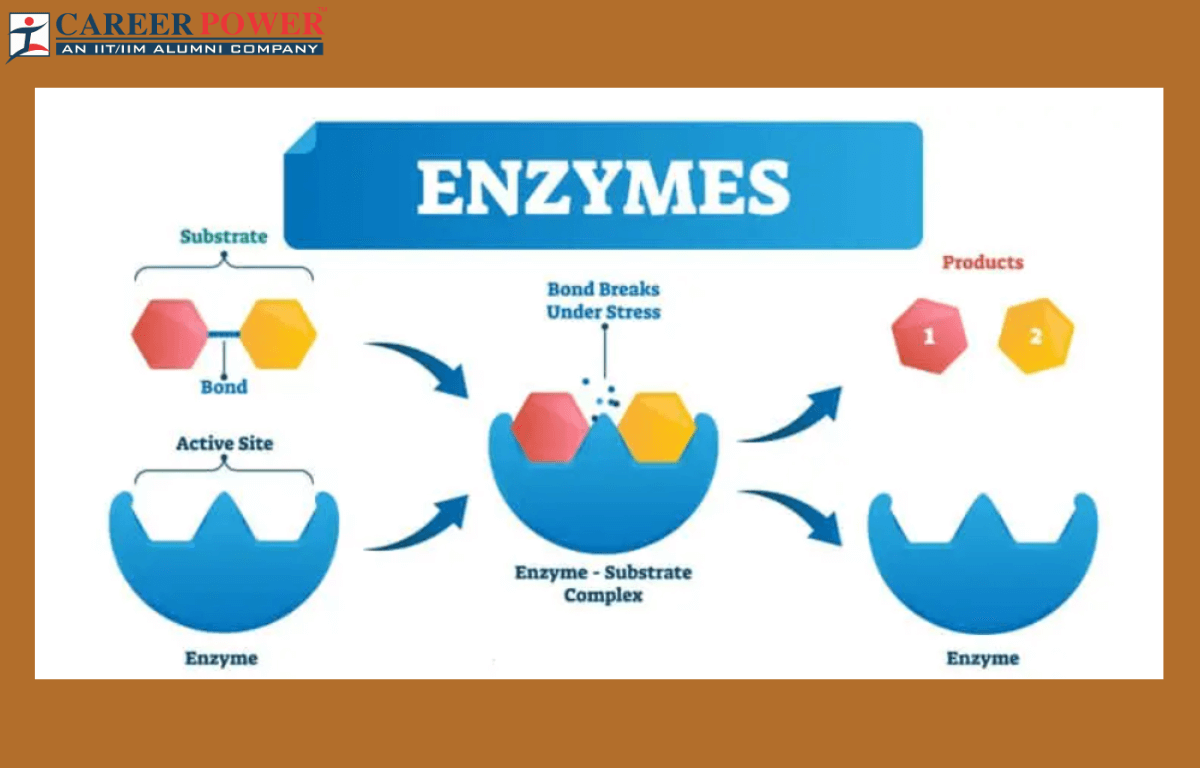
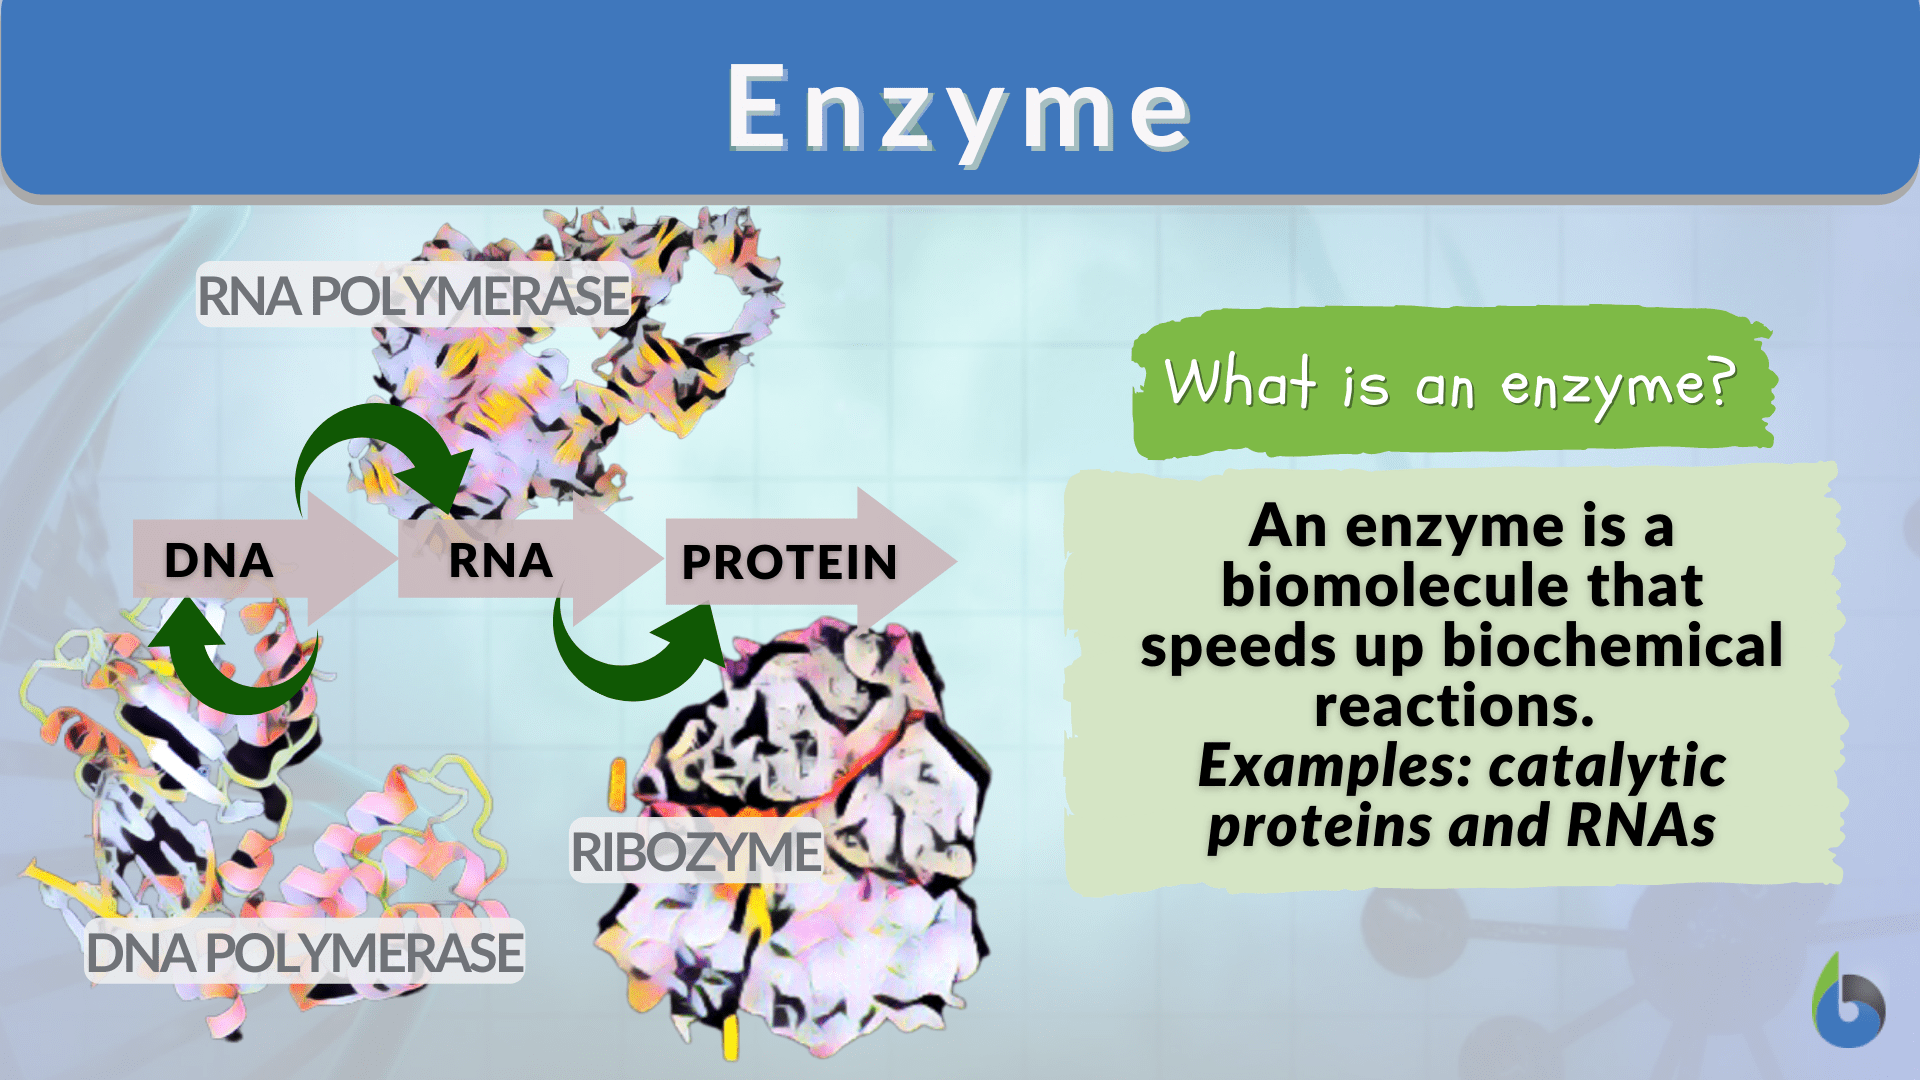
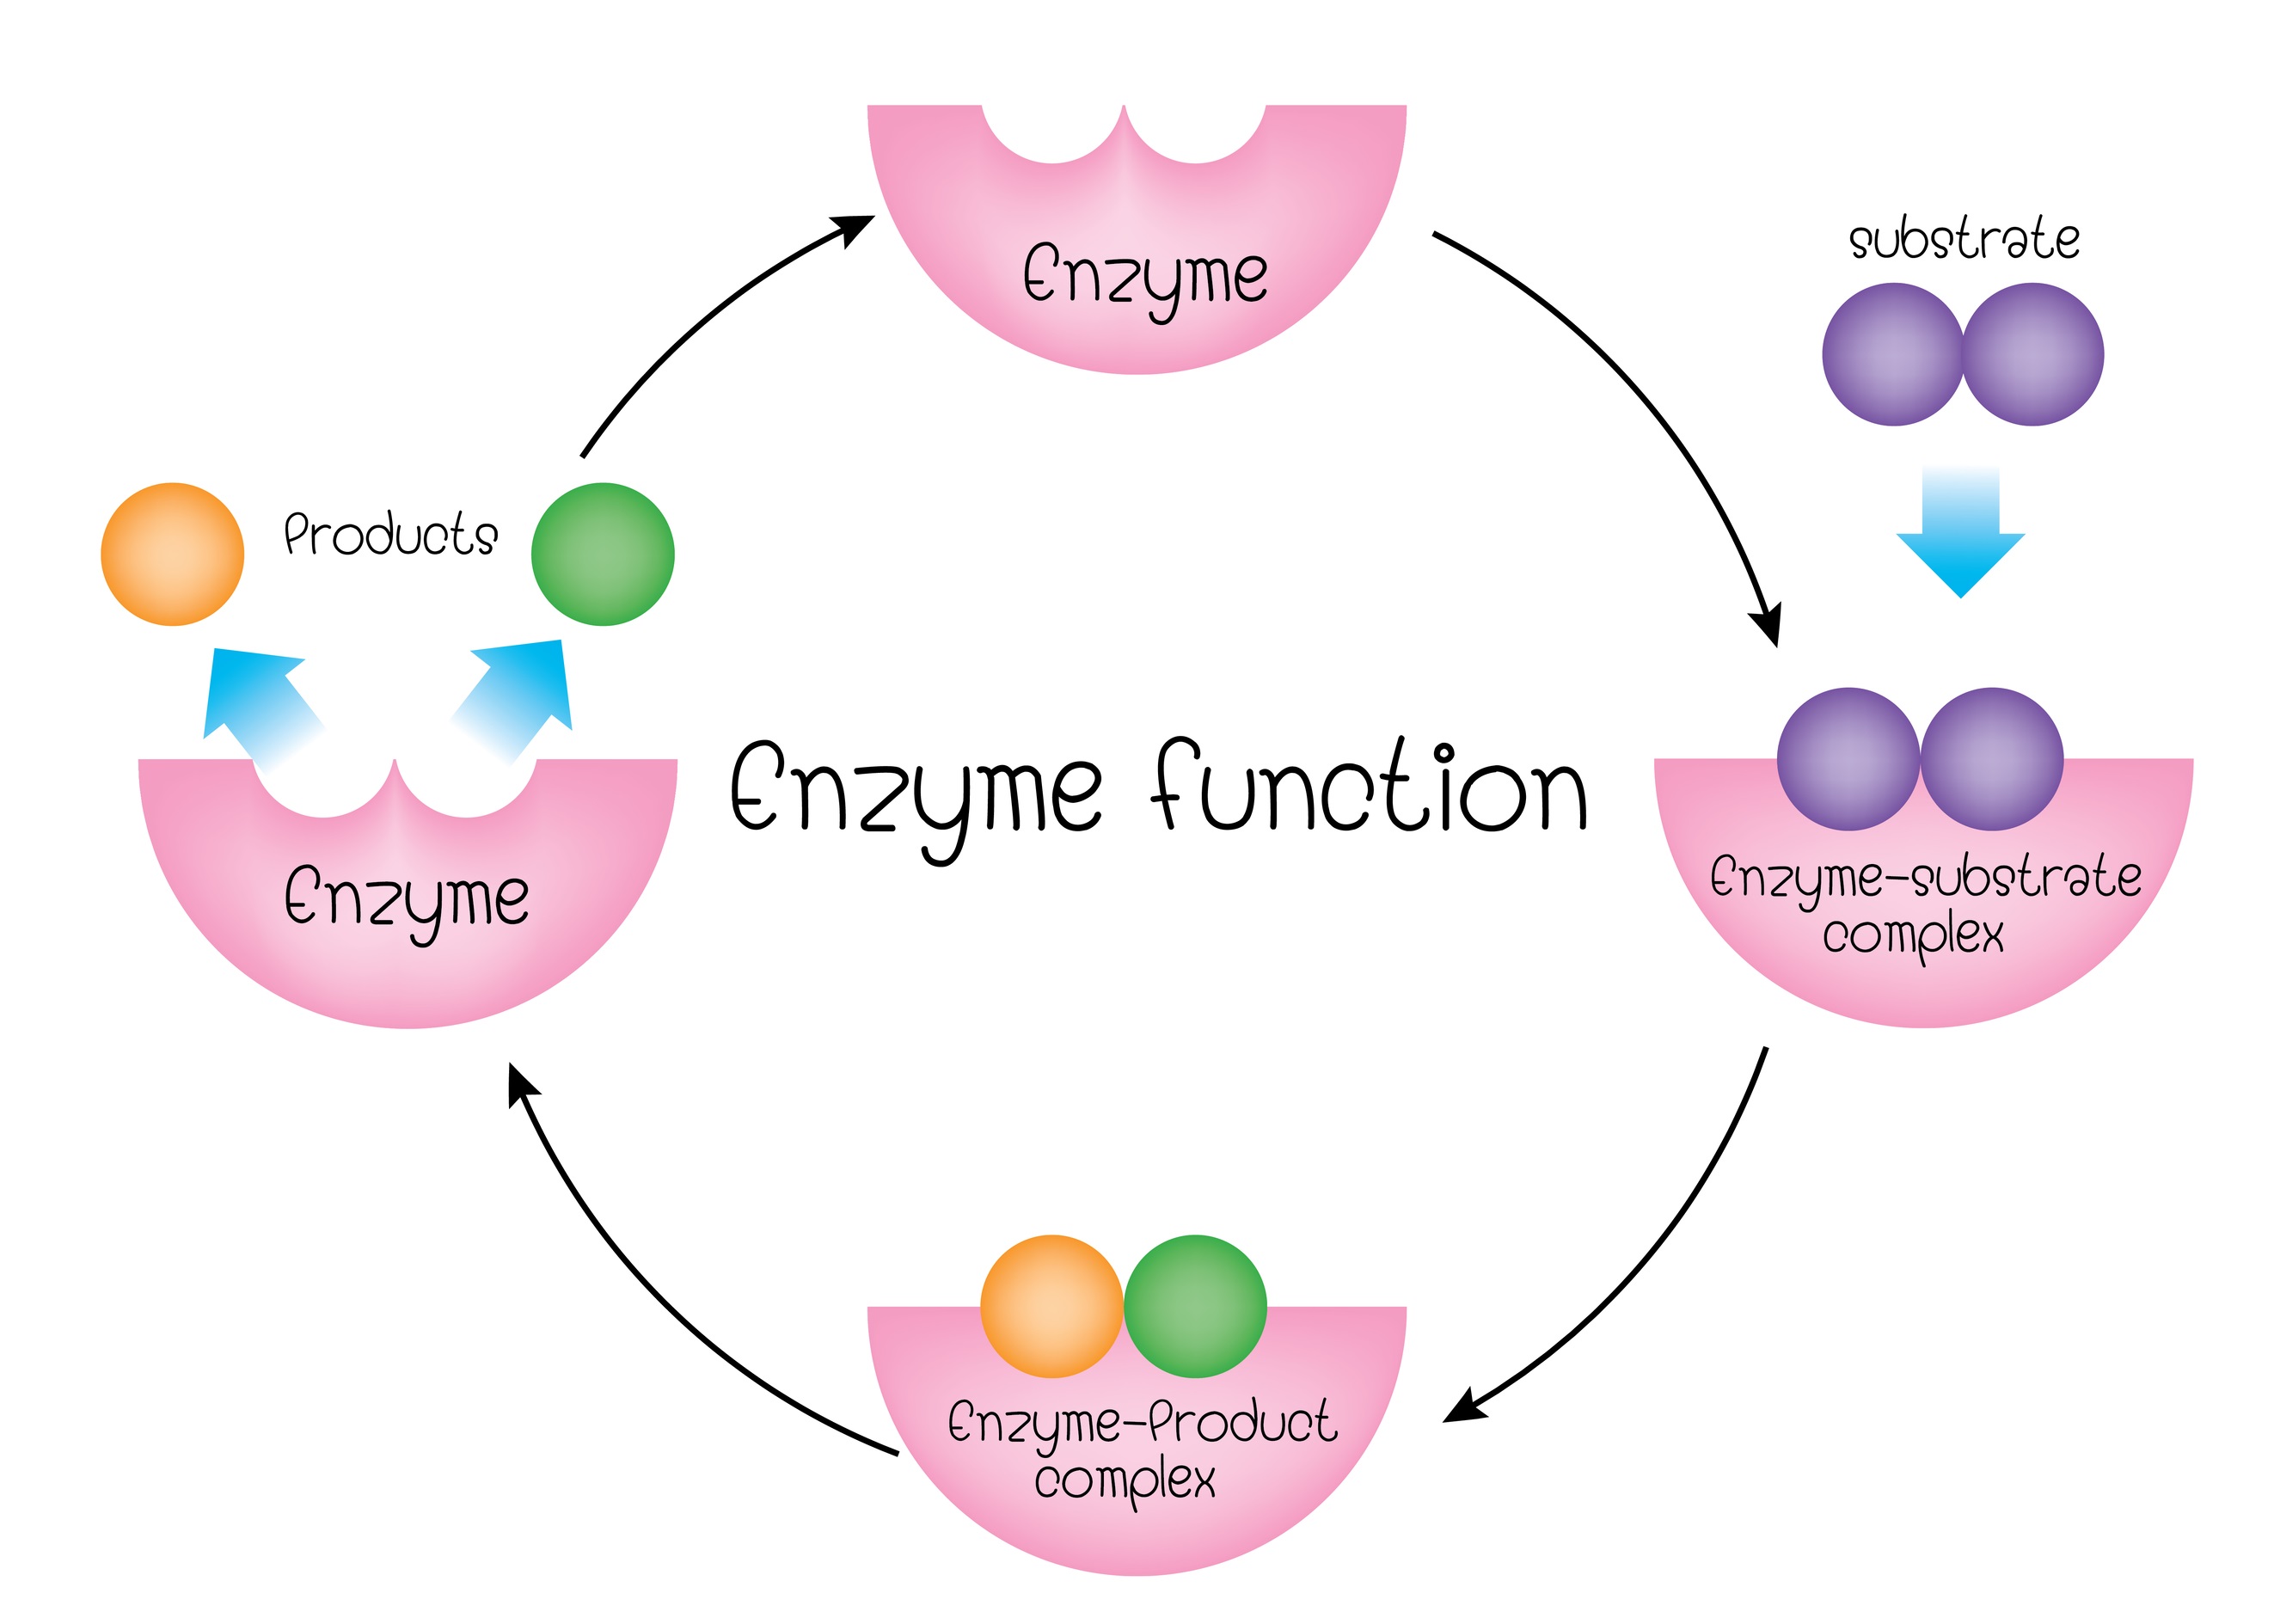
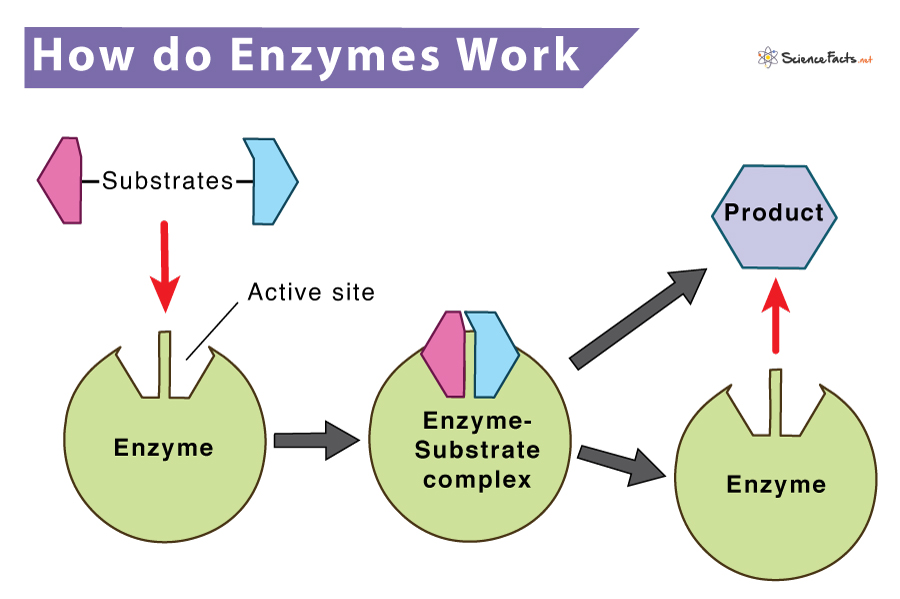
Enzymes are proteins that catalyze chemical reactions. That's the textbook version, but it leaves out most of what actually matters when you're working with them day to day. Enzyme biology is the study of how these molecules work, how they're regulated, and why they sometimes refuse to behave the way protocols say they should. I've spent years in labs running enzymatic assays, and the gap between theory and practice is where the real learning happens. At its core, enzyme biology covers the structure, mechanism, kinetics, and regulation of enzymes. Enzymes lower activation energy for reactions. They bind substrates at active sites. Products form. The enzyme is released unchanged and ready to catalyze again. Simple on paper. The details are where things get complicated. Kinetics is the first thing you need to understand properly. Michaelis-Menten kinetics describes how reaction velocity changes with substrate concentration. The Km value tells you the substrate concentration at which the reaction runs at half its maximum velocity. A low Km means high affinity. A high Km means the enzyme needs more substrate to get going. Vmax is the maximum rate when all enzyme active sites are saturated. These parameters aren't just numbers on a graph. They determine whether your assay will even work under given conditions.
One thing most introductory courses gloss over is that Km isn't a fixed constant. It changes with temperature, pH, ionic strength, and the presence of other molecules. I once spent three days troubleshooting an enzyme that appeared to lose activity across a board of reactions. Turned out the buffer I was using had a different pKa shift than expected at the incubation temperature, which meant the pH inside the tubes was off by almost a full unit from what I calculated. The enzyme wasn't degraded. It was just operating at a pH where it worked poorly. Always measure pH at the actual working temperature, not room temperature. This is something I learned the hard way, and it costs time and reagents if you get it wrong.
Practical realities of working with enzymes
Enzyme stability is a constant problem. Most enzymes lose activity over time, and the rate of degradation varies enormously between preparations. Lyophilized enzymes stored at minus eighty degrees Celsius can remain stable for months. Once you reconstitute them, the clock starts ticking faster. Aliquot them. Avoid repeated freeze-thaw cycles. If an enzyme is supposed to work at thirty-seven degrees Celsius, don't assume it'll stay active for more than an hour at that temperature without checking. Some enzymes, especially those from thermophilic organisms, are remarkably stable. Others fall apart within minutes under the same conditions. There's no universal rule. You test it. Inhibition is another area where textbook descriptions fall short. Competitive inhibitors bind the active site and can be overcome by adding more substrate. Noncompetitive inhibitors bind elsewhere and reduce Vmax regardless of substrate concentration. Uncompetitive inhibitors bind only to the enzyme-substrate complex. Mixed inhibition is a combination of competitive and noncompetitive effects. The real world has a fifth category: mechanism-based inhibition, also called suicide inhibition. The enzyme catalyzes a reaction that inactivates itself. This happens with drugs like penicillin binding to transpeptidase, and it's much harder to detect because standard kinetic assays don't flag it immediately. The activity drops irreversibly over time, and the effect looks superficially similar to enzyme degradation. If your enzyme seems to lose activity in a time-dependent manner that can't be explained by storage conditions or temperature, mechanism-based inhibition should be on your list of possibilities. I ran into this once with a phosphatase assay. The enzyme looked fine on initial testing. Activity declined linearly over the course of the reaction in a way that suggested poor stability. I ruled out temperature issues, buffer problems, and storage degradation. The breakthrough came when I added excess substrate before the reaction started. Activity held steady. With lower substrate concentrations, the decline was dramatic. The inhibitor was the product itself, binding irreversibly after the catalytic step. I had to redesign the assay to include a coupled reaction that continuously removed the product. This added complexity but made the measurements reliable. Standard protocols rarely mention this scenario because it doesn't fit the standard inhibition models taught in undergrad biochemistry.
Get the Full Details
Regulation and control in biological systems
Enzymes don't just operate at a fixed rate. They're regulated allosterically, through covalent modification, by proteolytic cleavage, and through interactions with regulatory proteins. Allosteric enzymes have sites separate from the active site where molecules bind and change the enzyme's conformation. This can increase or decrease activity. Hemoglobin isn't technically an enzyme, but it's the classic example of allosteric regulation, and the same principles apply to enzymes like aspartate transcarbamoylase and phosphofructokinase. Covalent modification is one of the fastest ways cells regulate enzyme activity. Phosphorylation adds a phosphate group, usually to serine, threonine, or tyrosine residues. Kinases add the phosphate. Phosphatases remove it. This can activate or inhibit the enzyme depending on the specific system. Glycogen phosphorylase is activated by phosphorylation. Glycogen synthase is inhibited by it. The same second messenger, cAMP, triggers both effects in different tissues through the same kinase cascade. This is efficient. It's also why signaling pathways can have broad and sometimes unintended effects. An inhibitor targeting one kinase will affect every substrate it phosphorylates, not just the one you're interested in. Zymogens are another important concept. Many enzymes are synthesized in inactive forms and activated only when needed. Digestive enzymes like trypsin and chymotrypsin are prime examples. Pepsinogen becomes pepsin in the acidic environment of the stomach. If these enzymes were active as soon as they were synthesized, they would digest the cells that produce them. The cleavage that activates them is usually irreversible. Once the bond is cut, the enzyme stays active until it's degraded. This is why protease inhibitors are essential in any extract preparation where you want to preserve native enzyme activity. A single protease contaminant can degrade your entire sample if left unchecked.
Assay design and common pitfalls
Designing an enzyme assay sounds straightforward. Mix enzyme with substrate. Measure product formation. Calculate activity. The reality involves far more decisions. You need to choose a detection method, determine the linear range of the reaction, establish the optimal conditions, and validate that you're measuring what you think you're measuring. Spectrophotometric assays are the most common. NADH absorbs at three hundred forty nanometers. NAD+ doesn't. Any reaction that produces or consumes NADH can be monitored this way. Coupled assays use a second enzyme to link an invisible reaction to this visible signal. The problem is that coupled assays introduce additional variables. The coupling enzyme must be in excess, or it becomes the rate-limiting step. If it isn't, your measured rate reflects the coupling enzyme's kinetics, not your target enzyme's. I've seen this mistake in published papers where the coupling enzyme concentration was never reported, making it impossible to verify the data. Fluorometric assays offer greater sensitivity but are more prone to interference. Fluorescent compounds in biological samples can quench signals or produce background fluorescence that overlaps with your readout. If you're working with cell lysates or tissue extracts, always run a no-enzyme control containing everything except the enzyme. This tells you whether your sample matrix is interfering with the signal. Don't skip this step. The background from a complex sample can be higher than you expect, and it changes between batches.
Temperature control is critical and often underestimated. Enzyme reaction rates approximately double with every ten-degree Celsius increase, at least until the enzyme starts denaturing. A difference of two degrees between reactions can produce a ten to fifteen percent difference in measured activity. Water baths fluctuate. Block heaters have hot spots. If you're comparing conditions, use the same instrument for all samples, and pre-equilibrate buffers to the reaction temperature before starting. I once compared two enzyme variants side by side and concluded one had significantly higher activity. The only difference was their position in the thermal cycler. One was closer to the heating element. The apparent difference vanished when I used a calibrated water bath and staggered the plate positions.
Why enzyme biology matters beyond the bench
Understanding enzymes isn't just useful for biochemists. Drug development depends on it. Most pharmaceuticals target enzymes. Statins inhibit HMG-CoA reductase. ACE inhibitors block angiotensin-converting enzyme. Tyrosine kinase inhibitors are a major class of cancer drugs. The effectiveness of these drugs depends on how well they bind to their target enzyme, how selective they are, and how they behave in the presence of natural substrates and competing molecules. Industrial biotechnology relies heavily on engineered enzymes. Amylases in detergents. Lipases in food processing. Polymerases in PCR. The enzymes used in these applications are often modified through directed evolution or rational design to improve stability, alter specificity, or increase activity under non-physiological conditions. Understanding the relationship between enzyme structure and function is what makes this possible. Diagnostic medicine uses enzyme assays constantly. Liver function tests measure ALT and AST. Pancreatic function can be assessed with amylase and lipase. Many immunoassays use enzyme labels like horseradish peroxidase or alkaline phosphatase to generate detectable signals. The reliability of these diagnostics depends entirely on the consistency and quality of the enzymes involved. Batch-to-batch variation in enzyme preparations can introduce errors that propagate through clinical interpretation.
The field continues to evolve. Cryo-EM has revealed enzyme structures at near-atomic resolution that were previously inaccessible. Computational methods like AlphaFold predict enzyme structures with remarkable accuracy, though predicting catalytic mechanism from structure alone remains difficult. Single-molecule enzymology has shown that enzyme activity can be heterogeneous even within a pure population, challenging the assumption that all enzyme molecules in a sample behave identically. These advances are changing how we think about what enzymes do and how they do it.