Le Chatelier's Principle Explained Like You're Trying To Run A Reactor Without Burning It Down
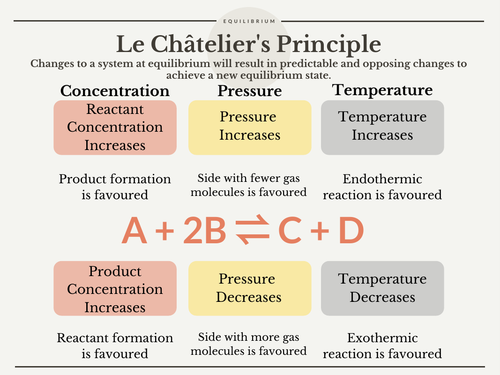
I keep seeing people treat this as some elegant law of nature, like the universe has moral opinions about chemical equilibria. It doesn't. What Is Le Chatelier S Principle really boils down to is this: when you push a system that's sitting at equilibrium, the system responds in a direction that partially opposes whatever you just did to it. That's it. End of story. The rest is just applying math to figure out which direction "partially opposes" actually points. The principle says that if you disturb a system at chemical equilibrium by changing concentration, pressure, volume, or temperature, the system will shift its equilibrium position to counteract, at least in part, that change. This isn't about stopping the change entirely. The system never fully negates your disturbance. It just moves in a direction that reduces the magnitude of whatever you introduced. Here's how it works in practice for each variable.
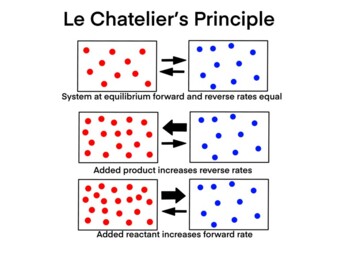
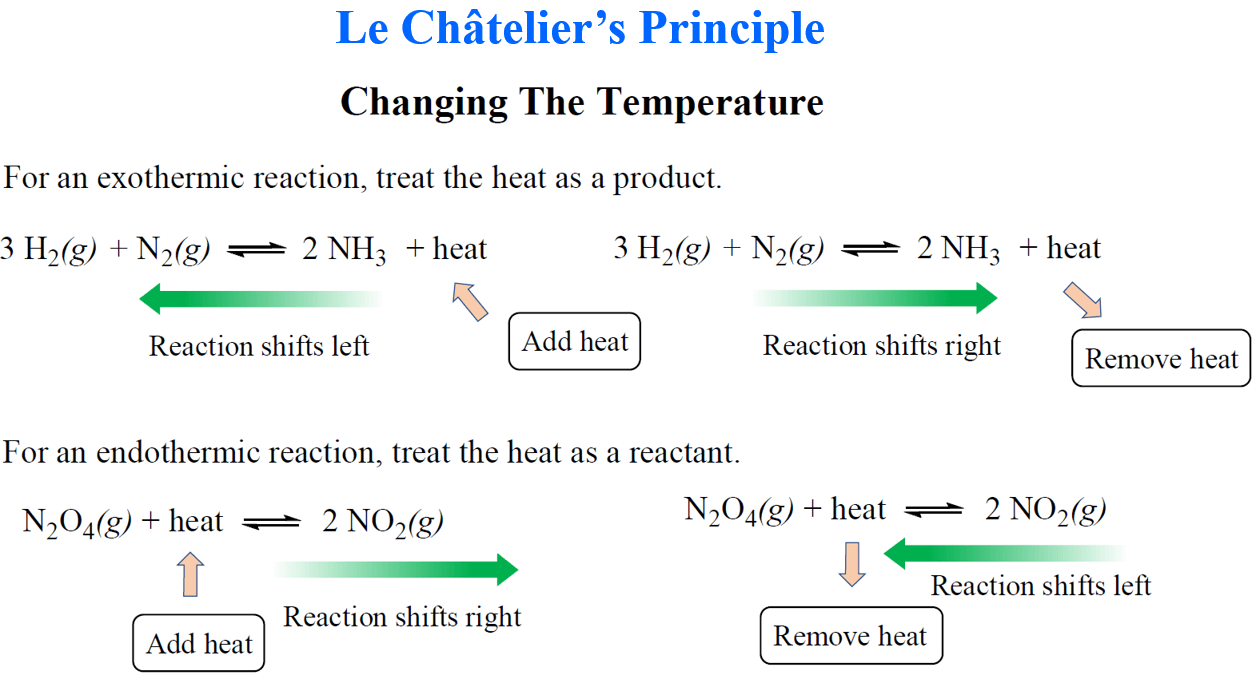
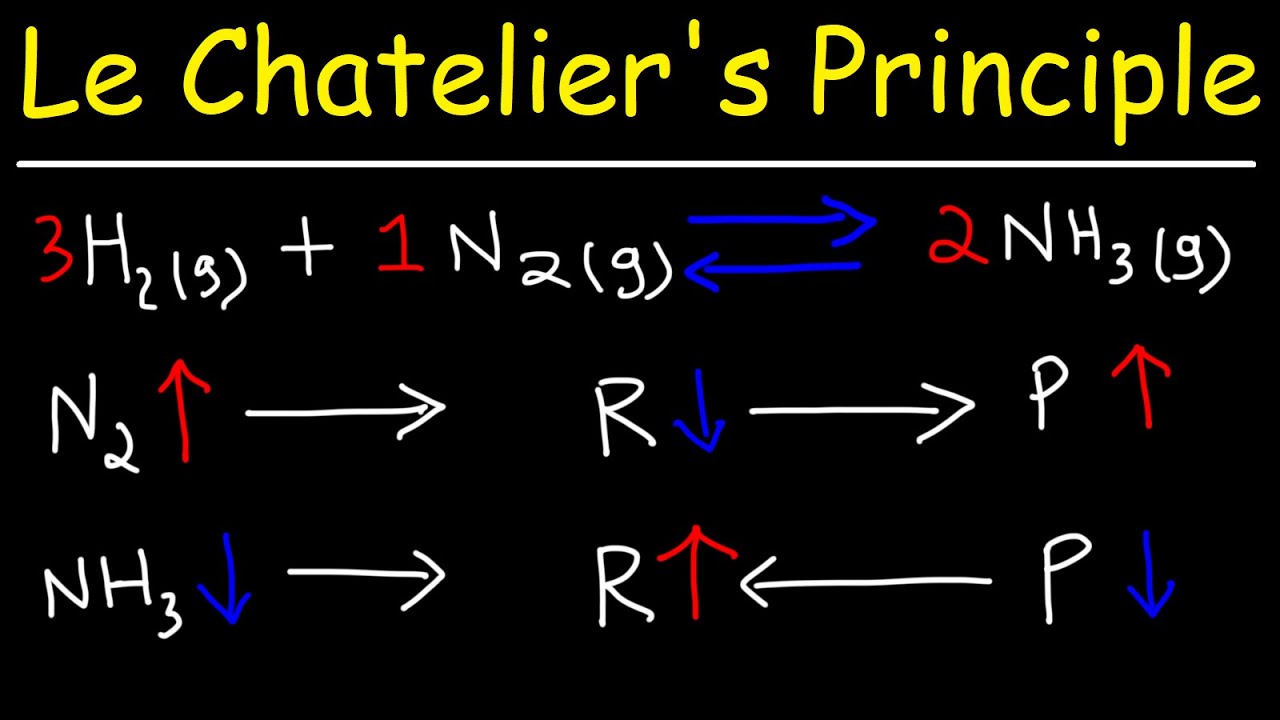
Concentration changes are the straightforward ones. If you add more reactant, Q drops below K, and the system shifts right to produce more product until Q equals K again. If you remove product, same thing. Remove reactant, shift left. These are the easiest to calculate because you're literally just comparing the new reaction quotient against the equilibrium constant. Pressure and volume changes only matter when gases are involved, and only when the number of moles of gas differs between reactants and products. If you increase pressure by decreasing volume, the equilibrium shifts toward the side with fewer gas molecules. Decrease the pressure, and it shifts toward the side with more gas molecules. If the moles of gas are equal on both sides, changing pressure does absolutely nothing to the equilibrium position. I've seen people waste hours troubleshooting a process where they'd cranked up the pressure hoping for a yield boost, only to discover the reaction had the same number of moles on each side. Pointless energy expenditure. Temperature is the variable that actually changes the equilibrium constant itself, which makes it qualitatively different from concentration and pressure changes. For an exothermic reaction, increasing temperature decreases K, so the equilibrium shifts toward the reactants. For an endothermic reaction, increasing temperature increases K, and the equilibrium shifts toward the products. You can remember this by treating heat as either a product (exothermic) or a reactant (endothermic) in the equation, but honestly it's simpler to just memorize which direction the shift goes and move on.
The Counter-Intuitive Stuff Nobody Teaches You
Most textbooks present Le Chatelier's principle as universally reliable. It's not. Here's where it gets messy. Inert gas additions are the classic trap. If you add an inert gas like argon or nitrogen to a system at constant volume, nothing happens to the equilibrium. The partial pressures of the reacting species don't change, so Q doesn't change, and there's no shift. Students always assume adding gas means adding pressure means shift happens. It doesn't. The inert gas contributes to total pressure but not to the partial pressures that matter for the equilibrium expression. If you add the inert gas at constant total pressure instead, then you're increasing the total volume, which dilutes all the reacting species. In that case, the equilibrium shifts toward the side with more gas molecules. Same disturbance, different boundary condition, opposite result. This distinction trips people up constantly in exams and in real plant operations.
Get the Full Details
Catalysts don't shift equilibria at all. They speed up both the forward and reverse reactions equally, so equilibrium is reached faster, but the position stays identical. I once watched a process engineer insist that adding a catalyst would improve yield on an equilibrium-limited reaction. It won't. It reduces time to equilibrium, which can matter enormously for throughput, but the final conversion number doesn't budge. He saved the company money on energy by reducing residence time, but he was wrong about why. Here's another one that causes real problems in industry. When you have multiple equilibria happening simultaneously, Le Chatelier's principle becomes unreliable as a predictive tool. Consider a system where A decomposes to B and C, and B further reacts to form D. Changing the concentration of D affects the second equilibrium, which changes B's concentration, which then feeds back into the first equilibrium. The net effect isn't obvious from simple qualitative reasoning. You have to write out the full set of equilibrium expressions and solve them. The principle gives you the right direction for each individual equilibrium in isolation, but the coupled behavior can produce unexpected results when you combine them.
Where I Got Burned By This
Early in my career, I was working on a gas-phase synthesis where we were trying to maximize product yield by manipulating pressure. The reaction involved three moles of gas on the reactant side and two on the product side, so Le Chatelier's principle clearly predicted that higher pressure would favor product formation. We designed the whole process around operating at 200 atmospheres. The problem was that our reactor had a significant pressure drop across its length due to flow resistance. The inlet was at 200 atm, but by the time the gas exited, pressure had dropped to about 140 atm. More importantly, the reaction was exothermic, and the temperature profile along the reactor wasn't uniform. The inlet zone was cooler, the middle ran hot from the reaction heat, and the outlet cooled again. Pressure was highest at the inlet and lowest at the outlet, while temperature followed the opposite pattern in the middle section. So locally, in different zones of the reactor, Le Chatelier's principle was giving contradictory guidance. The high pressure at the inlet favored products, but the lower temperature there meant the equilibrium constant was actually favorable for products too since it's exothermic. In the hot middle section, the equilibrium constant had dropped significantly, and even though pressure was still moderate, the unfavorable K dominated. The outlet had restored temperature but lost pressure. The net yield was nowhere near what we'd calculated using a single temperature and pressure assumption.
The workaround was to model the reactor as a series of small differential elements, each at its own temperature and pressure, and compute the local equilibrium composition at each point. Then I integrated along the length. This gave us a realistic prediction that was about 20 percent lower than our original design. We ended up running at 250 atm at the inlet instead, which brought us back to target yield. The principle itself was correct at every point. Our mistake was treating a spatially distributed system as if it were uniform.

What Actually Happens When You Apply This
Let me walk through a concrete example with numbers so you can see how the principle translates into calculation. Consider the reaction N(g) + 3H(g) 2NH(g), which is exothermic with H = 92 kJ/mol. At 500 K, Kp happens to be 1.7 × 10. Suppose the system is at equilibrium at a total pressure of 100 atm with the following partial pressures: P(N) = 18.2 atm, P(H) = 54.5 atm, P(NH) = 27.3 atm. You can verify that Q = (27.3)² / [(18.2)(54.5)³] 1.7 × 10, which matches Kp. Now you double the total pressure to 200 atm by compressing the system. The mole fractions stay the same initially, so the new partial pressures are double: P(N) = 36.4 atm, P(H) = 109 atm, P(NH) = 54.6 atm. Calculating the new Q gives you a value roughly one-quarter of the original, since the denominator has terms raised to higher powers than the numerator. Q is now much smaller than K, so the system shifts right, producing more NH and consuming N and H until Q equals K again. Le Chatelier's principle predicted exactly this: increase pressure, shift toward fewer gas moles (2 moles of product versus 4 moles of reactants).
If instead you raise the temperature to 600 K, Kp drops significantly because the reaction is exothermic. Let's say Kp falls to about 3.0 × 10³ at that temperature. The system is no longer at equilibrium, and Q is now larger than K. The reaction shifts left, consuming NH and producing N and H. Again, the principle predicted this correctly. But here's where reality diverges from the textbook problem. In an actual industrial reactor running the Haber process, you don't operate at 500 K because the reaction rate is impractically slow. You run closer to 700 K with an iron catalyst, where Kp is much smaller and the equilibrium yield is poor. You accept low per-pass conversion and recycle the unreacted gases. Le Chatelier's principle tells you the right direction, but it doesn't tell you that running at a temperature where the kinetics are viable forces you to operate far from the thermodynamic optimum. That's a tradeoff the principle doesn't address at all.
When Le Chatelier's Principle Fails Completely
The principle is strictly applicable only to systems at equilibrium. If you perturb a system that's not at equilibrium, or if the perturbation happens so fast that the system can't relax before you perturb it again, the principle has nothing to say about what occurs. This comes up in flow reactors where the residence time is shorter than the time needed to reach equilibrium. The gas enters, gets perturbed by temperature or pressure changes, and leaves before the composition has time to adjust. In those cases, the reactor is kinetically controlled, not equilibrium-controlled, and Le Chatelier's principle gives you the wrong answer. Another failure mode is when the system undergoes a phase change. If you compress a system and one component condenses out of the gas phase, the equilibrium expression changes fundamentally because that component is no longer in the gas phase. The principle doesn't account for this kind of boundary condition shift on its own. You need to recognize that the system has crossed a phase boundary and re-evaluate the equilibrium framework. Heterogeneous equilibria also require care. Solids and pure liquids don't appear in the equilibrium expression, so changing the amount of a solid reactant or product has no effect on the equilibrium position. Adding more solid doesn't shift anything. The principle works fine here, but students routinely apply it incorrectly by treating solids the same way they treat gases.
Practical Rules That Actually Hold Up
When you're working with this in the lab or on process equipment, keep these points in mind and you'll avoid most of the common mistakes. Always check whether the system is actually at equilibrium before applying the principle. If you're dealing with a flowing system, a transient process, or something that's been perturbed recently, verify that equilibrium has been established. A quick way to do this is to measure the relevant concentrations or pressures and calculate Q. If Q equals K within experimental uncertainty, you're at equilibrium and the principle applies. If not, you're dealing with kinetics, not equilibrium. For gas-phase systems, distinguish between total pressure changes caused by volume change versus total pressure changes caused by adding inert gas. Only volume-driven pressure changes affect the equilibrium position. Inert gas addition at constant volume is a non-event for equilibrium.
When temperature changes are involved, remember that you're changing K itself, not just Q. This means the direction of shift depends on whether the reaction is exothermic or endothermic, and you can't predict it solely from the stoichiometry. You need the sign of H. If you're working with coupled equilibria, write out all the equilibrium expressions simultaneously and solve them as a system. Don't try to reason through the interactions qualitatively unless you're very confident in your understanding of the coupling. The feedback between equilibria can produce results that contradict what you'd expect from applying the principle to each reaction independently. For industrial applications, remember that Le Chatelier's principle gives you thermodynamic limits, not practical yields. The maximum possible conversion at a given temperature and pressure is determined by equilibrium, but achieving that conversion requires sufficient reaction time, proper mixing, adequate catalyst activity, and heat management. In many processes, the bottleneck is kinetics or heat transfer, not equilibrium. Pushing pressure or adjusting temperature according to the principle might move the equilibrium in the right direction, but if your reactor is fundamentally limited by mass transfer or reaction rate, you'll see diminishing returns quickly. I've seen plants where engineers kept increasing pressure hoping for better yields, only to discover that the reaction was already mass-transfer limited and the extra pressure was just burning compressor power for no gain.
The principle remains useful as a first-order guide for understanding which direction changes push a system. But it's a guide, not a calculator, and it's definitely not a substitute for actually solving the equilibrium expressions or running the experiment. Treat it as a sanity check on your predictions, not as the prediction itself.