Getting Your Mass Spec Data Not to Look Like Garbage
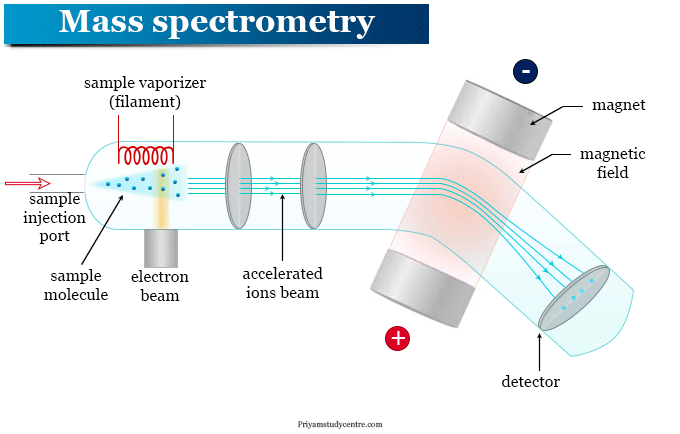
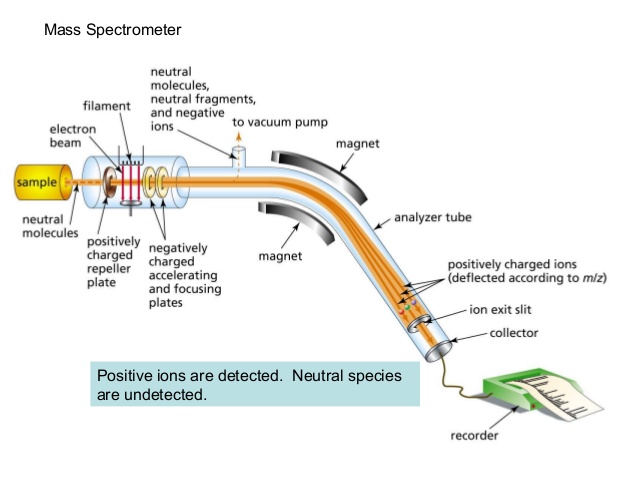
I spent six months fighting a baseline drift problem on an Orbitrap before I realized I'd been calibrating it wrong. That's pretty much the entire experience with mass spectrometry. You learn by losing samples and wasting instrument time until the machine behaves. Mass spectrometry measures the mass-to-charge ratio of ions. That's the textbook answer. The actual process is more like shearing molecules apart and timing how fast the pieces fly through a vacuum. You introduce a sample, ionize it, separate the ions by their m/z values, and detect them. The output is a spectrum that tells you what's in your sample and usually how much of it is there. The ionization method matters more than most beginners realize. Electrospray ionization (ESI) is soft and keeps molecules mostly intact, which is why it pairs with liquid chromatography for proteomics. Electron impact (EI) smashes molecules into fragments so thoroughly that you can match the pattern against libraries like NIST. Choose wrong and you're not just getting bad data, you're getting data that looks good but means nothing.
How the Instrument Actually Works
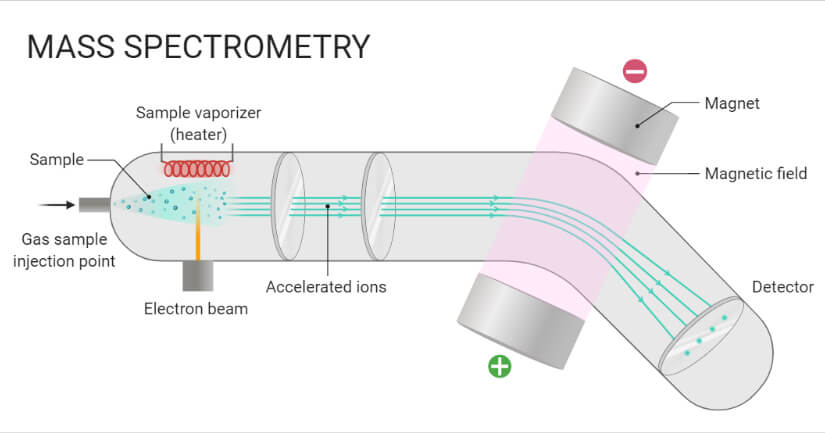
Here's what happens inside, stripped of the vendor brochures. Your sample enters the ion source. In ESI, a high voltage is applied to a liquid flowing through a tiny needle, creating a mist of charged droplets. Solvent evaporates, droplets shrink, ions pop off into the gas phase. Those ions then enter a vacuum system where electric fields guide them toward the mass analyzer. The analyzer sorts them. A quadrupole uses oscillating electric fields to let only certain m/z values through at a time. A time-of-flight tube measures how long ions take to travel a fixed distance. An orbitrap traps ions in an oscillating cloud and measures the frequency of their motion around a central electrode. Each approach has tradeoffs in resolution, speed, and cost that aren't always obvious until you're reading the manual at 11 PM. After separation, ions hit a detector. Most modern instruments use electron multipliers or microchannel plates. The detector generates a current proportional to the number of ions arriving, and the software builds a spectrum from those signals. Simple in theory. Painful in practice.
Where People Go Wrong
The biggest mistake I see is treating mass spectrometry like a black box you load samples into and expect truth to come out. It's not a scale. Contamination from the previous run, ion suppression from co-eluting compounds, and matrix effects can completely alter your results without any warning labels on the screen. I once had a client who was quantifying a drug metabolite in plasma and getting inconsistent recovery across batches. The standard calibration curve looked fine. We ran blanks and found that the plastic centrifuge tubes were leaching phthalate plasticizers at exactly the m/z of the internal standard. We switched to glass and the variance dropped from 18% to under 5%. Took three weeks to figure out. The instrument wasn't broken. The sample prep was. Another thing nobody warns you about: detector fatigue. When you run high-concentration samples back-to-back without enough wash cycles, the electron multiplier degrades. Sensitivity drifts downward slowly enough that you don't notice it day to day. You only see it when you run a quality control sample and it reads 30% lower than yesterday's value. Running a mid-level standard every ten injections and tracking it on a Levey-Jennings chart catches this before it ruins a full batch.
Get the Full Details

Practical Workflow That Actually Works
Start with clean samples. If you're doing LC-MS, filter through 0.2 micron PTFE or nylon filters. Particulate matter clogs columns and creates spike artifacts in your spectra. Don't skip this step because the sample looks clear. You can't see 0.5 micron particles with your eyes. Calibration should happen at the start of every session and after any maintenance event. Use the manufacturer's calibration solution, but also run an external lock mass compound to verify accuracy throughout the day. I use sodium formate clusters for positive mode and a phosphazide solution for negative mode as secondary checks. This catches calibration drift that the auto-calibration routine misses. For quantitative work, always use isotopically labeled internal standards when possible. They compensate for ion suppression and injection variability better than anything else. If you can't afford labeled standards, at minimum run a post-column infusion experiment to map out where suppression is happening in your chromatogram. It takes twenty minutes and will save you from publishing wrong numbers.
Database searching deserves more care than it gets. When you're identifying unknowns, a narrow mass tolerance of five ppm combined with proper fragment ion matching reduces false positives dramatically. Broad tolerances of twenty or fifty ppm will give you hits, but half of them will be wrong. I've seen people report identifications based on a single matching peak without checking if the isotope pattern fit. The monoisotopic peak matches, but the M+1 and M+2 intensities are completely off. That's not your compound. It's something else with a similar nominal mass.
When Mass Spectrometry Fails You
It doesn't work for everything. Non-ionizable compounds, extremely nonpolar molecules, and thermally labile substances can be nearly impossible to analyze without derivatization. If your compound won't ionize in ESI or APCI, you're looking at chemical modification or a different technique entirely. MALDI helps with some of these cases but introduces its own problems with matrix interference in the low mass region below 700 m/z. Sensitivity limits are another constraint. Even the best instruments struggle to detect compounds below picomolar concentrations in complex matrices without prior enrichment. If you're working with trace environmental contaminants or early-stage biomarkers, you'll need sample preparation that concentrates your analyte. Solid phase extraction, liquid-liquid extraction, or protein precipitation aren't optional extras. They're the difference between detecting your analyte and detecting nothing. Interpretation complexity scales up faster than most people expect. A single LC-MS run can generate tens of thousands of peaks. Distinguishing real signals from noise, isotope clusters, adducts, and in-source fragments requires understanding what each peak represents. Software helps, but automated peak picking still generates significant false positives that a human needs to review. Budget time for this.
Bottom Line
Mass spectrometry is powerful because it gives you structural information that other techniques can't match. It's frustrating because it demands attention to detail at every step from sample prep to data interpretation. The instrument will give you garbage if you feed it garbage. Clean samples, proper calibration, appropriate controls, and a willingness to question unexpected results will get you further than any expensive add-on or proprietary software package.