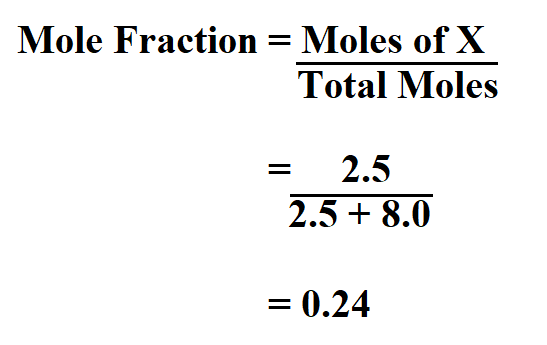Understanding Mole Fraction in Real Practice
Mole fraction is one of those concepts that sounds simple until you're actually using it and the numbers don't behave the way you expect. It's a way of expressing the concentration of a component in a mixture. You take the moles of one substance and divide by the total moles of everything in the mixture. The result is always between zero and one. That's it, fundamentally. But the practical side gets messy fast. I worked on a distillation column design project once where we were calculating mole fractions for a three-component hydrocarbon mix — benzene, toluene, and xylene. The textbook formulas worked fine on paper. Then we ran into a real issue: the light key component kept showing up in the heavy bottoms product at levels that made no physical sense. Turns out we'd been treating the feed as an ideal solution when it wasn't. The activity coefficients were pushing us off. We ended up switching to a NRTL model for the non-ideal behavior and recalculated everything from scratch. That took about three days instead of the four hours I'd originally planned.
What Is Mole Fraction and Why People Get It Wrong
The definition is straightforward. X subscript i equals n subscript i divided by the total n. Nothing fancy there. What trips people up is assuming that mole fraction and mole percent are interchangeable without converting. If your lab report says 25 percent, that's 0.25 in fraction form. Mess that conversion up and your downstream calculations — vapor-liquid equilibrium, partial pressures, everything — cascade into garbage results. Another thing that catches people: mole fraction changes with temperature and pressure for gas mixtures, even though the actual number of moles doesn't change. This is because for gases, when you switch to volume-based measurements, the volume shifts with temperature. If you're working with liquid solutions, the effect is much smaller but still present in precision work. I've seen process engineers ignore this in ambient lab conditions and then get burned when the plant ran at elevated temperature. When dealing with electrolyte solutions, you also need to decide whether you're counting ions separately or as a formula unit. Sodium chloride dissociates into two particles. If your mole fraction calculation treats NaCl as one unit but your colligative property equations expect ions, you'll be off by roughly a factor of two. It depends on what you're actually measuring. Always clarify whether you're working with apparent mole fraction or true mole fraction based on dissociation.
The sum of all mole fractions in any mixture must equal one. This seems obvious but I've checked work where someone calculated individual fractions that added up to 1.04 or 0.91. Usually a rounding error propagated through multiple steps, or one component was accidentally left out of the denominator. Double-check this every time before moving forward. It takes ten seconds and will save you from chasing phantom problems later.
Get the Full Details

Working With Mole Fraction in Vapor-Liquid Equilibrium
Raoult's Law is where mole fraction becomes practically useful for most chemical engineers. For an ideal solution, the partial pressure of each component equals its mole fraction in the liquid phase multiplied by its pure component vapor pressure. P sub i equals X sub i times P sub i star. Simple enough. The total pressure is the sum of all partial pressures. Y sub i, the vapor phase mole fraction, comes out to P sub i divided by the total pressure. So if you know the liquid composition and the temperature, you can calculate what the vapor above it looks like. This is the foundation of distillation design. You build stage by stage from this relationship. Real solutions deviate from Raoult's Law, sometimes significantly. Positive deviations mean the components want to escape the liquid more than they would ideally — think ethanol and water, which form an azeotrope. Negative deviations mean they stick together more — chloroform and acetone is a classic example. When deviations are large, you need activity coefficients. Gamma sub i multiplies into the equation: P sub i equals X sub i times gamma sub i times P sub i star.
I remember running into trouble with a methanol-water system where the azeotrope was right in the middle of our separation target. Standard distillation couldn't break it. We had to switch to pressure-swing distillation, exploiting the fact that the azeotropic composition shifts with pressure. The mole fraction calculations still worked, but the constants changed enough that you have to rebuild your equilibrium data at each operating pressure. Don't just copy and paste K-values from a different pressure condition. For solid solutions, mole fraction appears in freezing point depression equations and phase diagrams. The relationship is linear only in the ideal dilute limit. Once concentrations get high, you need the same activity coefficient treatment. I once saw someone use the simple freezing point depression formula for a 30 mole percent solute solution and wonder why the prediction was wildly off. It only works well below about five percent solute. One thing people consistently overlook is that mole fraction is dimensionless. It carries no units. If you're reporting it with units attached, something is wrong. Percent composition by mole is technically still dimensionless too — it's just scaled by 100. Being clear on this matters when you're setting up spreadsheets or writing equations. Mixing mole fraction with molality or molarity without converting is a common error source, especially when switching between literature values and your own calculations.
If you need to convert from mass fraction to mole fraction, you do it component by component. Divide each mass fraction by the molecular weight of that component, then normalize by the sum of all those terms. It's a five-step process on paper but it's easy to fat-finger in a spreadsheet. I set up a small lookup table for the common components I work with so I don't have to recalculate the inversion every time.

Practical Considerations and Where the Method Breaks Down
Mole fraction works well for homogeneous mixtures where the components are well-mixed at the molecular level. It does not work for heterogeneous systems. If you have a slurry or an emulsion, the concept breaks down because there is no single meaningful composition — different phases have different compositions and the interface region adds complexity that mole fraction can't capture. Use volume fractions or mass fractions for those cases, or better yet, model the phases separately. For gas mixtures at high pressure, the assumption that mole fraction directly gives you partial pressure through Raoult's or Dalton's Law starts failing. You need fugacity coefficients. The higher the pressure, the bigger the deviation. Above about 10 atmospheres for most systems, you should be checking whether your ideal gas assumption is still valid. I've seen people apply ideal mole fraction calculations to supercritical fluid systems and get results that were qualitatively wrong. Another limitation: mole fraction doesn't tell you anything about the spatial distribution of components. In a non-uniform mixture or a gradient system, the local mole fraction varies across the volume. Global mole fraction is just an average. If you're doing diffusion calculations or reaction engineering, the local value matters more than the bulk average. Make sure you're calculating the right thing for your application.
When working with ionic liquids or deep eutectic solvents, the standard mole fraction framework gets awkward because the "components" aren't always clearly defined. Are you counting the cation and anion separately? What about hydrogen bonding networks that create transient species? In these cases, researchers sometimes use solvent-free mole fractions or reference-state conventions that differ from textbook definitions. Read the methodology section of any paper you're pulling data from carefully before plugging numbers into your own equations. For quick estimates during early-stage design, mole fraction is fast and intuitive. For production-level simulation work, you're better off using specialized software like Aspen Plus or ChemCAD that handles the thermodynamic models internally. The manual calculations are still useful for sanity checks and understanding what's happening, but they're not practical for large multicomponent systems with non-ideal behavior. I typically spend about ten minutes per stream doing hand calculations to verify that the simulator isn't converging to some numerically correct but physically nonsense solution.