Understanding the shape of molecules
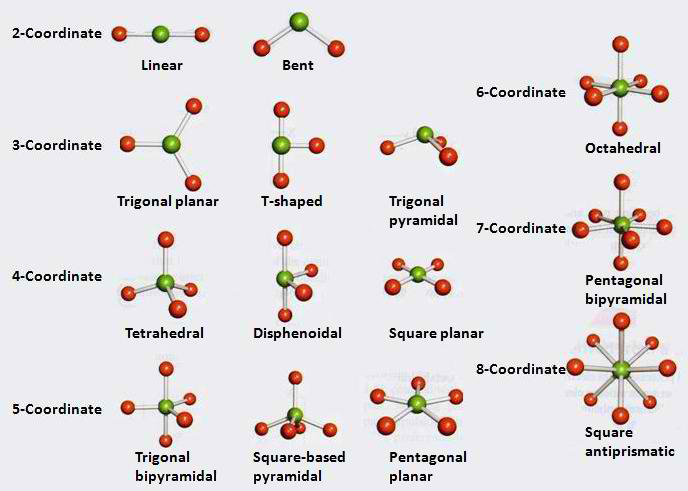
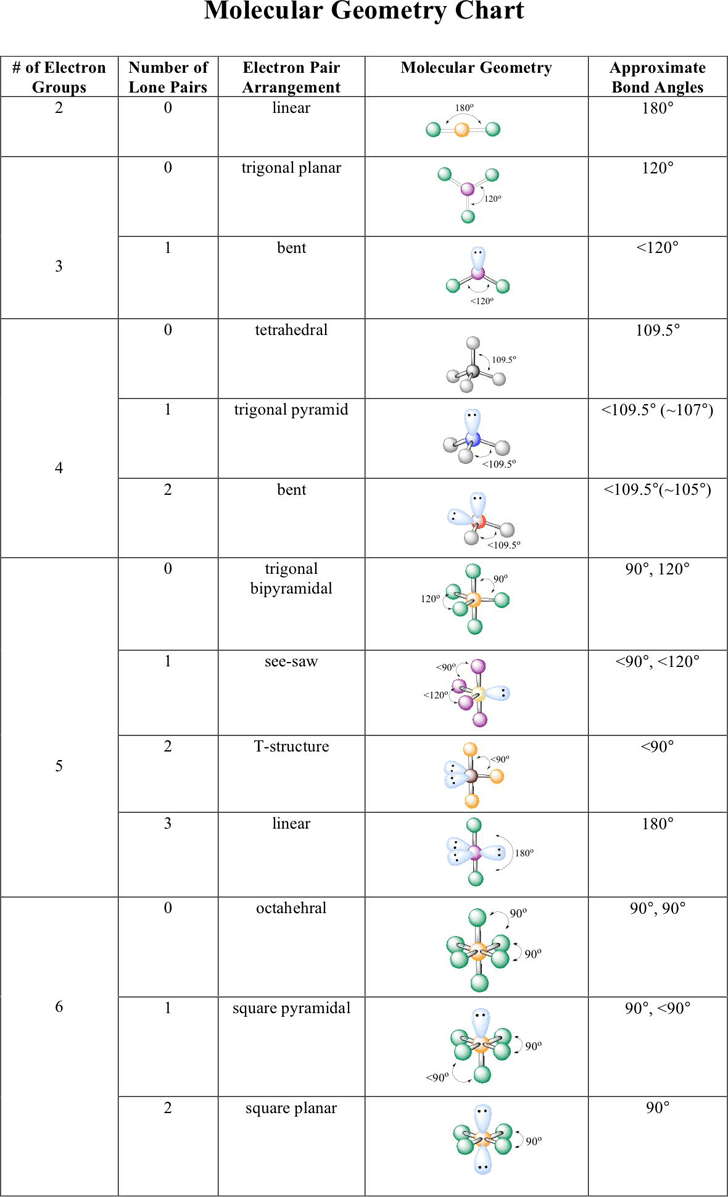
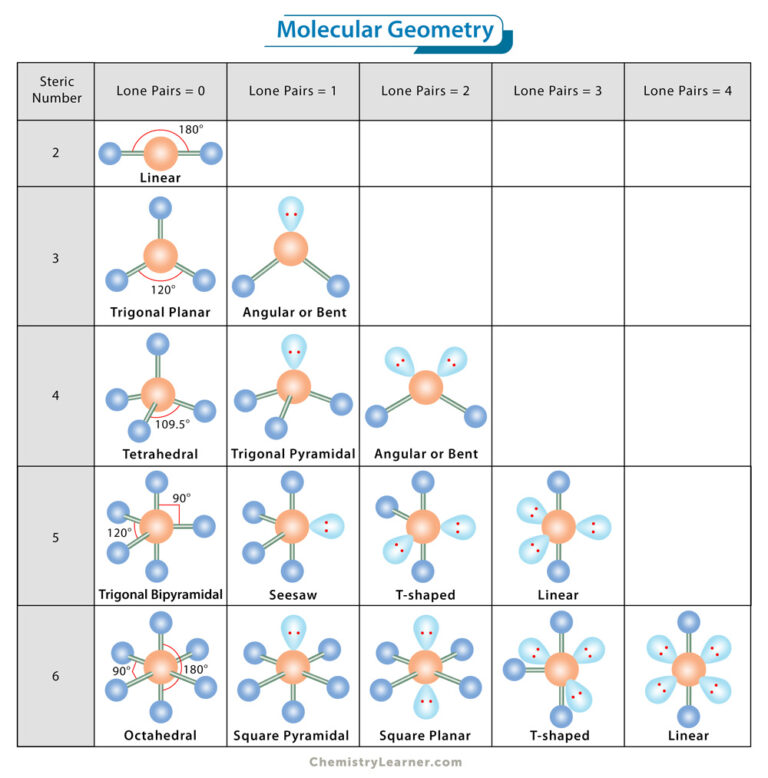
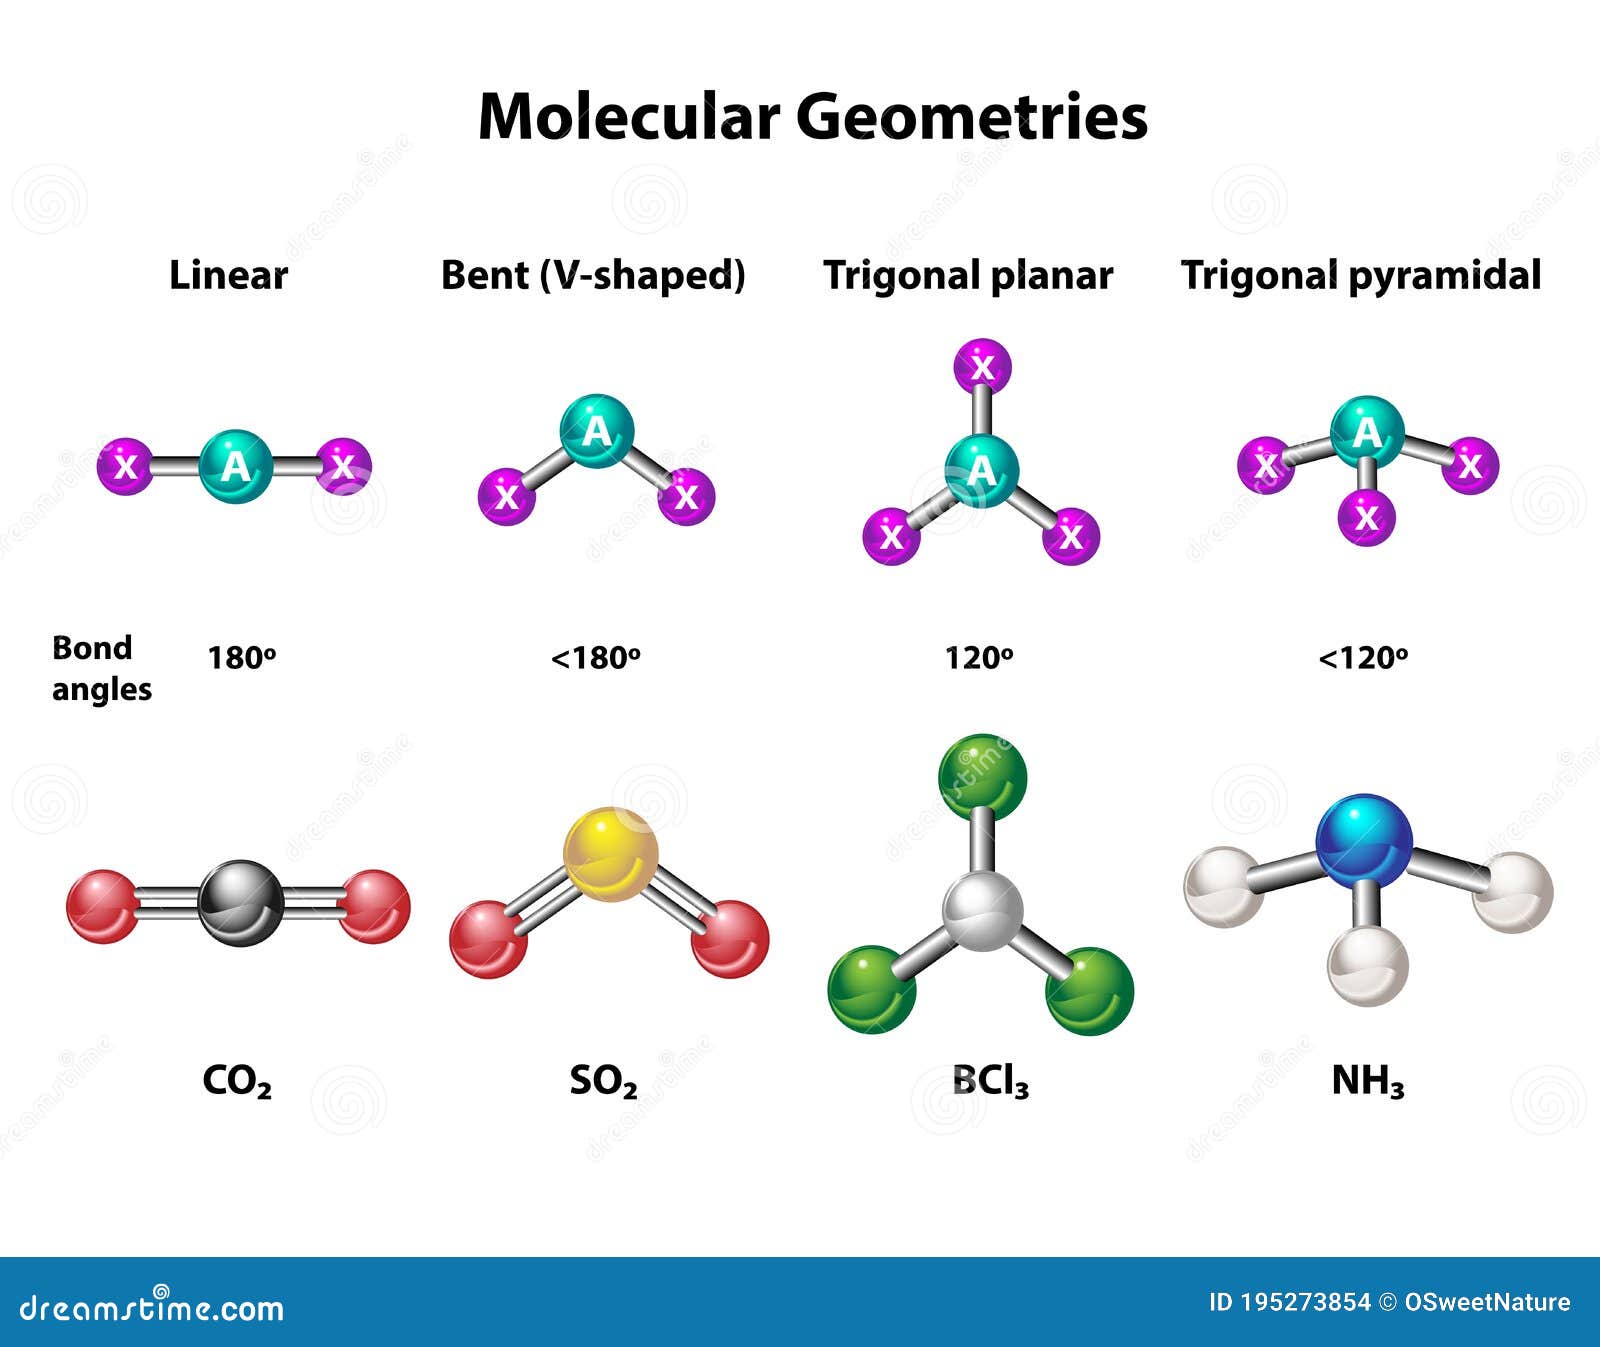
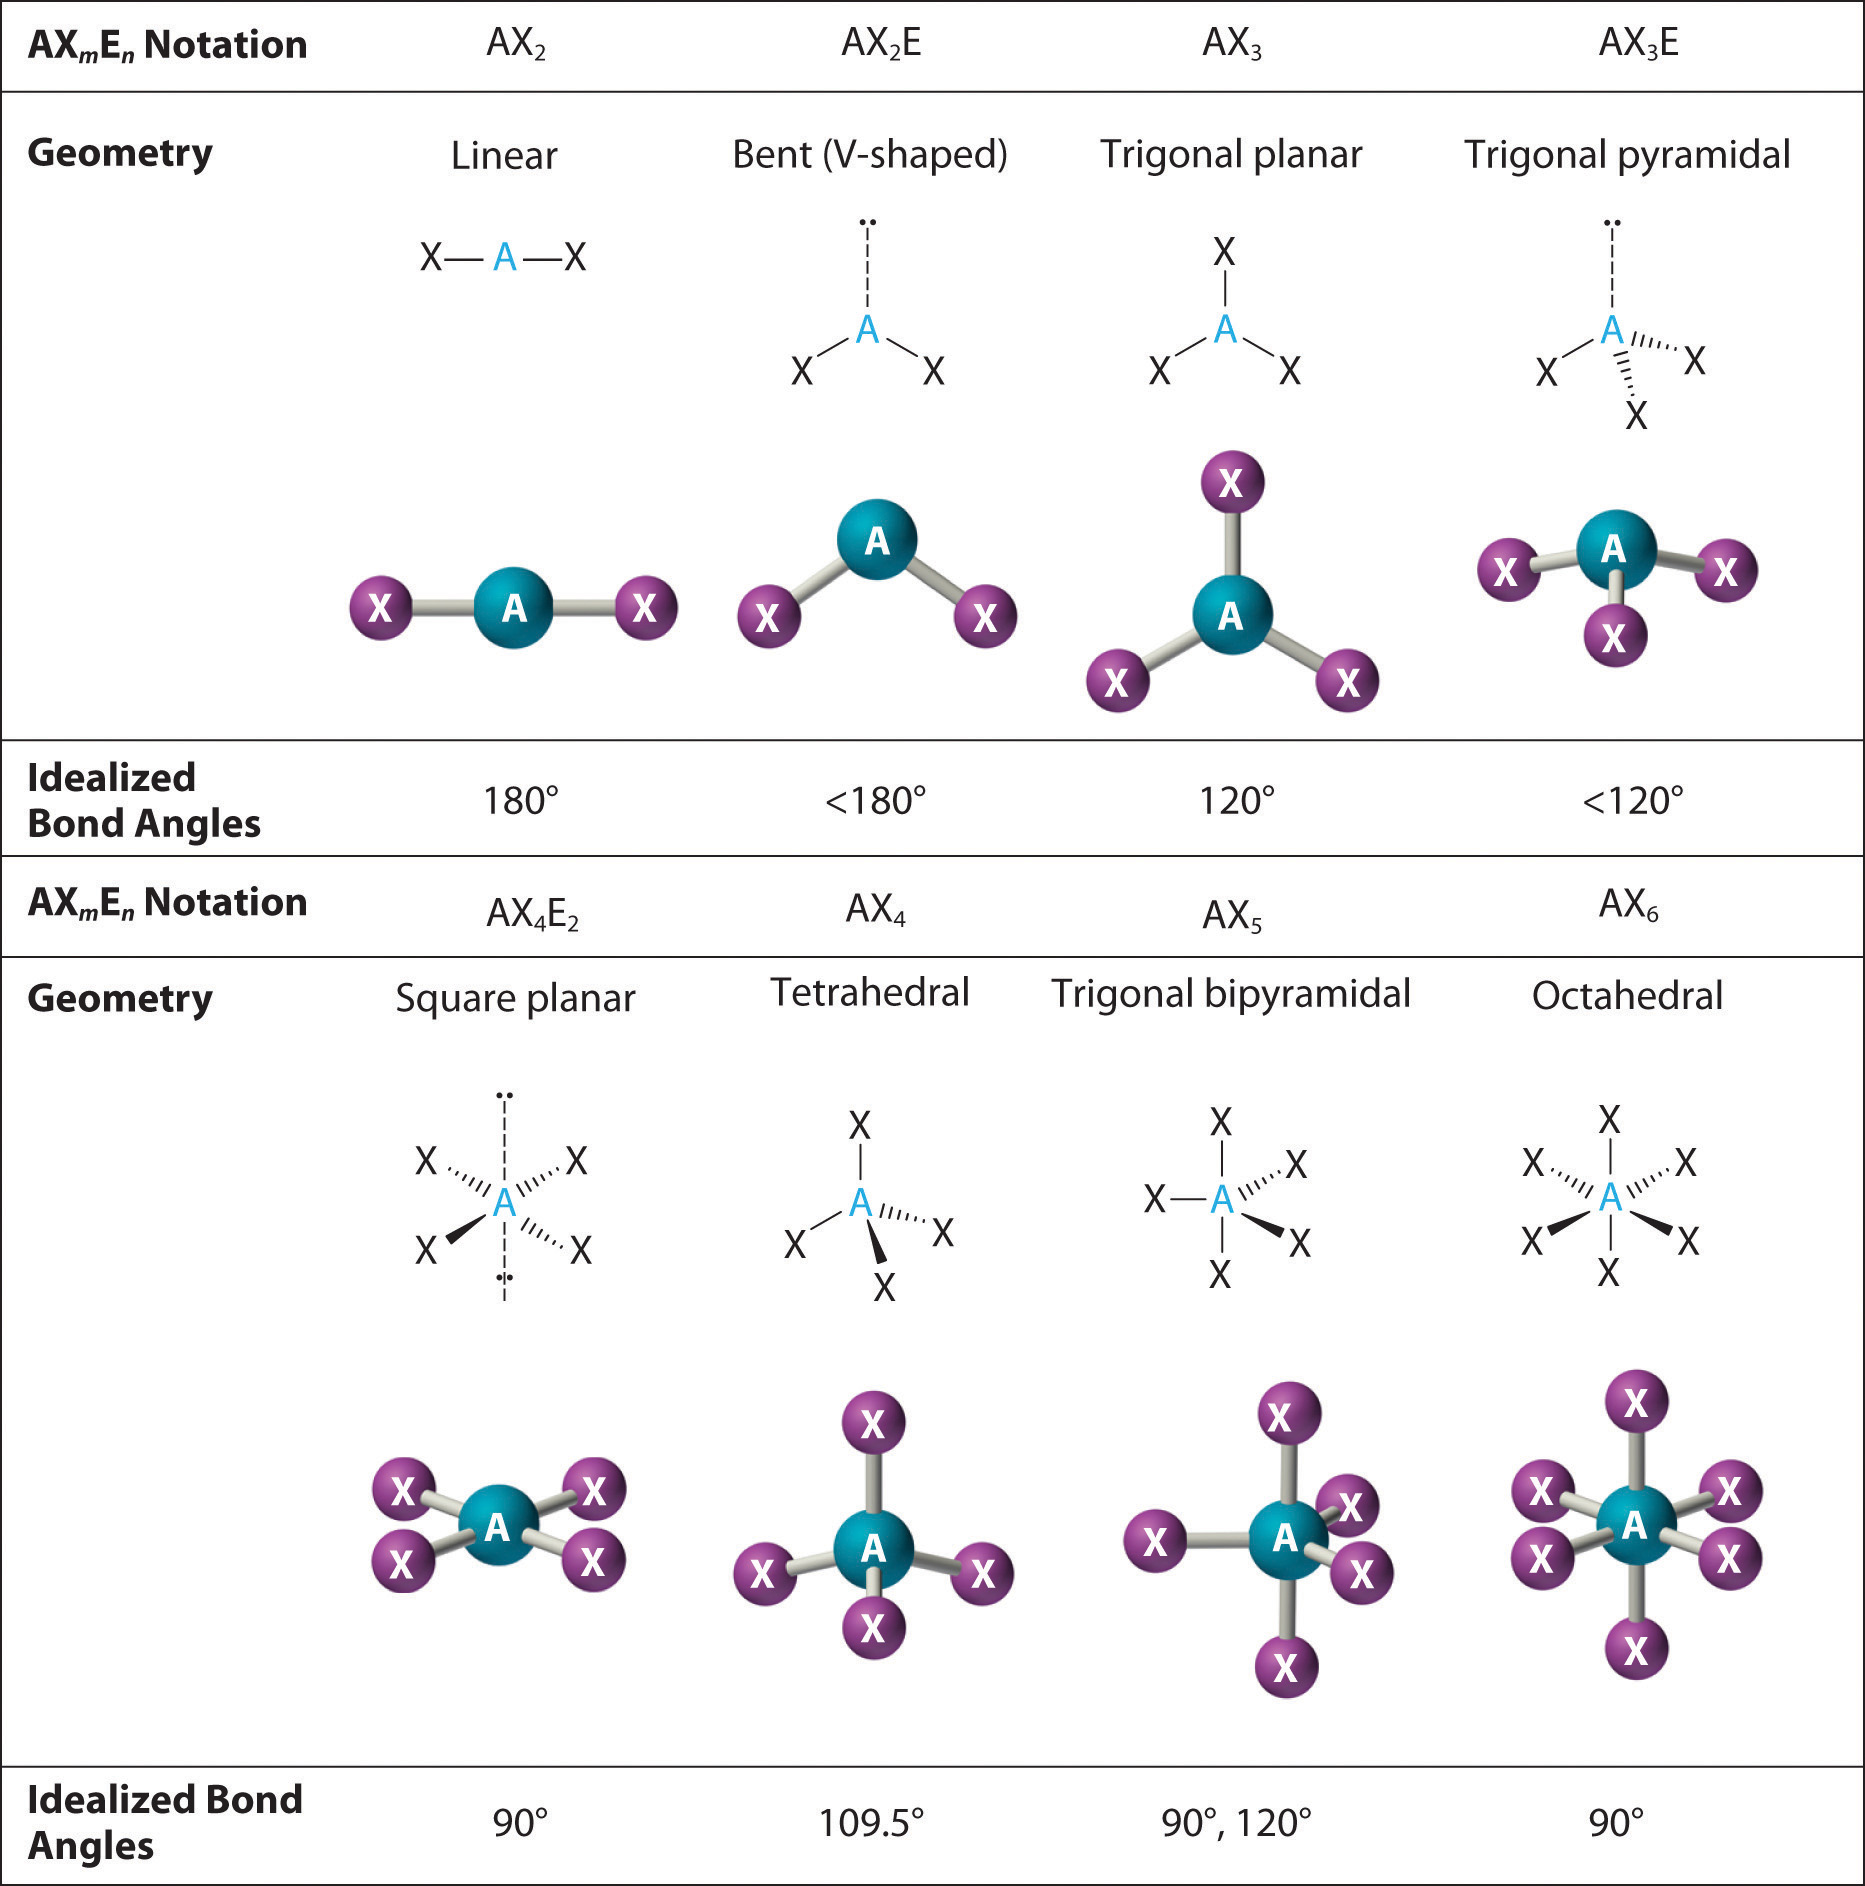
Molecular geometry describes the three-dimensional arrangement of atoms within a molecule. This isn't just academic curiosity. The shape determines reactivity, polarity, boiling points, and how drugs fit into protein binding sites. If you're working in chemistry, materials science, or pharmaceutical development, you need to predict or interpret these shapes accurately. Here's what actually works and where the standard models break down. The foundation is VSEPR theory — Valence Shell Electron Pair Repulsion. The basic premise is straightforward: electron pairs around a central atom repel each other and arrange themselves as far apart as possible. Lone pairs take up more space than bonding pairs. This is why water is bent instead of linear, and why ammonia has a trigonal pyramidal shape rather than trigonal planar. The theory gives you a fast mental framework for predicting geometry from a Lewis structure. You count electron domains, assign the corresponding arrangement, and account for lone pair effects. Most introductory courses stop here. That's where the real problems begin.
What Is Molecular Geometry Beyond the Basics
Standard VSEPR covers the main cases — linear, trigonal planar, tetrahedral, trigonal bipyramidal, octahedral — and their derivatives when lone pairs are involved. But the moment you step outside simple main-group compounds, the model gets messy. Transition metal complexes are the first place where things get complicated. d-orbital participation, crystal field effects, and Jahn-Teller distortions all play roles that VSEPR simply doesn't address. A classic example is square planar geometry in d8 complexes like Pt(II) and Pd(II). VSEPR would predict tetrahedral, but the actual geometry is square planar due to d-orbital energy splitting that the theory ignores entirely. Hybridization theory sits alongside VSEPR and provides a different way to rationalize the same geometries. sp hybridization gives linear geometry, sp2 gives trigonal planar, sp3 gives tetrahedral. This framework maps nicely onto organic chemistry, where carbon compounds dominate. But hybridization is a mathematical construct, not a physical observable. It's useful for drawing and thinking about molecules, but it doesn't predict anything VSEPR can't already tell you. The real predictive power comes from computational methods when you encounter something outside these neat categories. I ran into a specific problem recently that exposed the limits of both VSEPR and simple hybridization. I was working with a bismuth-containing compound — BiF5 — and trying to predict its geometry. The central atom has five fluorine ligands, which from a basic standpoint suggests trigonal bipyramidal. The experimental structure, confirmed by X-ray crystallography, shows a distorted trigonal bipyramid with significant asymmetry in the axial positions. The lone pair on bismuth is stereochemically active, and the sheer size and relativistic effects of the heavy element make standard models unreliable. What I ended up doing was running a DFT calculation with a relativistic effective core potential for the bismuth and comparing the computed geometry against the crystallographic data. The calculation reproduced the distortion accurately. For heavy main-group compounds like this, no amount of hand-waving with electron domains will get you the right answer.
Beyond transition metals and heavy elements, there are other common pitfalls. Hypervalent molecules like SF6 or PCl5 were historically problematic for VSEPR. The theory handles them fine in practice — six domains give octahedral, five give trigonal bipyramidal — but the underlying bonding description is fuzzy. Three-center four-electron bonds explain the bonding without invoking d-orbital participation, which was the original interpretation. Modern computational chemistry confirms that d-orbital contributions are minimal for second-row hypervalent compounds. This means when you're learning this material, the "expanded octet" explanation you might see in older textbooks is technically wrong even though the predicted geometry is correct. Aromaticity and conjugation also create geometries that don't fit standard VSEPR predictions. The benzene ring is planar because of pi delocalization, not because of any simple electron pair repulsion argument. Cyclooctatetraene avoids planarity and adopts a tub shape precisely to avoid the antiaromatic destabilization that a planar geometry would introduce. These are electronic effects that dominate over steric considerations, and VSEPR has nothing to say about them. For practical purposes, here's how I approach geometry prediction depending on the system. Simple organic and main-group molecules — use VSEPR and hybridization. It's fast and accurate enough. Transition metal complexes — use crystal field theory and ligand field theory. Know your d-electron count and the geometry preferences of common coordination numbers. Heavy or exotic compounds — run a computation. Gaussian, ORCA, or even semi-empirical methods in GAMESS can give you reliable geometries in a fraction of the time it takes to troubleshoot an incorrect prediction by hand. For quick checks, xtb (extended tight-binding) is free and surprisingly accurate for organic and organometallic systems.
Get the Full Details
The biggest limitation of all these approaches is that they treat molecules as static structures. Real molecules vibrate, rotate, and sample multiple conformations. A single optimized geometry is a snapshot at the bottom of a potential energy surface. When you're studying reactivity, you need to consider transition states and reaction pathways, not just the minimum energy structure. Software like Gaussian or ORCA can do transition state searches, but they require a decent initial guess and can get stuck in local minima if the potential surface is complex. For large systems, conformational sampling tools like ConfAb or RDKit's conformer generator can help map out the accessible geometry space before you commit to expensive DFT calculations. Crystallographic data is the gold standard for determining molecular geometry experimentally. Single-crystal X-ray diffraction gives you atomic positions with sub-angstrom precision. Powder diffraction works for cases where you can't grow single crystals but gives lower resolution. Electron diffraction is useful for gas-phase molecules. Neutron diffraction is the only method that can directly locate hydrogen atoms, which matters when hydrogen bonding patterns influence the overall structure. If computational and experimental geometries disagree, the experiment wins — but make sure the refinement was done properly and the resolution is sufficient. Poor quality data can produce misleading bond lengths and angles that look reasonable but aren't trustworthy. There's also the matter of solvent and environmental effects. Gas-phase computations don't account for solvation, which can significantly distort geometry in polar molecules. Implicit solvent models like PCM or COSMO add computational cost but capture bulk solvent effects reasonably well. For hydrogen-bonded systems or specific solvent interactions, explicit solvent molecules in the model may be necessary, which increases complexity substantially. In crystal structures, packing forces can distort bond angles and lengths compared to the isolated molecule. These differences are usually small for rigid molecules but can be significant for flexible or weakly bound systems.
The bottom line is that molecular geometry prediction is a spectrum of approaches with different accuracy and cost trade-offs. VSEPR is a teaching tool and a quick mental check. Hybridization is a descriptive framework that maps well onto organic chemistry but doesn't predict new geometries. Crystal field and ligand field theories handle transition metals but require understanding d-orbital splitting patterns. Computational chemistry is the most general approach but requires setup time and validation. Experimental determination remains essential whenever you need absolute certainty about a structure you haven't seen before.