Protein synthesis isn't as clean as the textbooks make it look
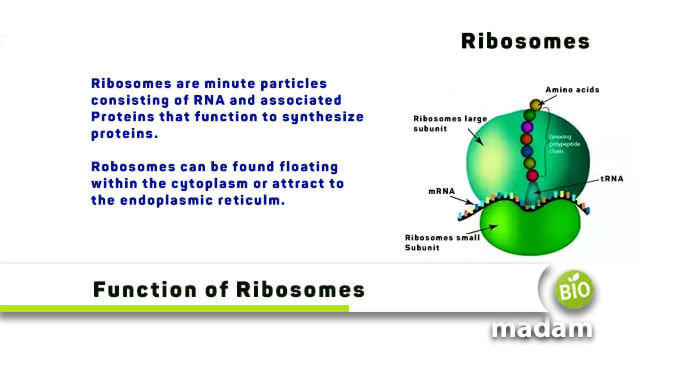
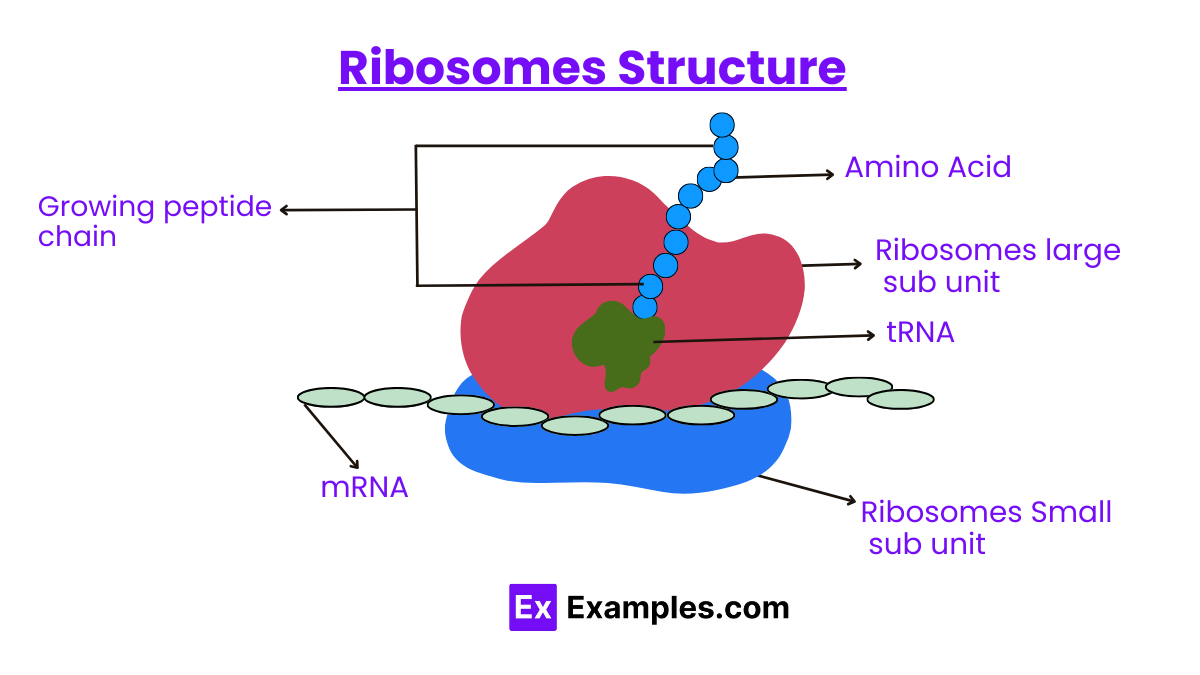
I spent six months debugging weird translation artifacts in a bacterial expression system before I really understood what ribosomes actually do under pressure. The short version: they polymerize amino acids into polypeptide chains using mRNA as a template. That's the textbook answer. The real answer is messier. Ribosomes are ribonucleoprotein complexes. They've got two subunits - the small one handles mRNA binding and codon recognition, the large one catalyzes peptide bond formation through its rRNA. Yeah, the catalysis happens in RNA, not protein. That's why we call them ribozymes and it matters when you're trying to design antibiotics that target bacterial translation without hitting human cells.
What Is The Function For The Ribosomes
Translation. Decoding mRNA into protein. That's the job description. But here's what most guides skip: ribosomes don't just chain amino acids together mechanically. They proofread. Every incoming aminoacyl-tRNA gets checked at the A site before the peptide bond forms. The ribosome spends roughly 0.1 to 0.5 seconds per codon in bacteria, and eukaryotic ribosomes move even slower. That speed variation matters enormously when you're dealing with rare codons or structurally complex proteins that need co-translational folding time. I learned this the hard way working with a recombinant protein that kept precipitating. The sequence was fine, the plasmid was verified, the induction conditions were optimal. The protein just wouldn't stay soluble. Turns out there were clusters of rare arginine codons - AGA and AGG in E. coli K-12 strains - that made the ribosome stall repeatedly. Each stall gave the N-terminal domain too much time to misfold before the rest of the chain was available. Switching to a strain with extra arginyl-tRNA genes or codon-optimizing those regions fixed it completely. Took about three days of troubleshooting that should have taken three hours if someone had warned me about ribosome stalling dynamics upfront. There's also the issue of ribosome queuing on highly expressed mRNAs. When you overexpress a gene from a strong promoter, multiple ribosomes load onto the same transcript simultaneously, forming polysomes. If translation is slow at certain points - rare codons, secondary structure in the mRNA, charged tRNA depletion - ribosomes pile up behind each other. This causes traffic jams that can trigger premature termination, frameshifting, or even ribosome collision pathways that activate stress responses like the stringent response in bacteria. I've seen cultures crash just from overexpression constructs that created enough stalling to trigger robustness pathways.
The peptidyl transferase center is another thing worth understanding practically. It's located entirely in the 23S rRNA of the large subunit. No proteins participate directly in catalysis. This is evolutionary holdover from an RNA world, and it's exactly why certain antibiotics work. Chloramphenicol blocks the PTC in bacteria but not in mitochondria or cytoplasm. Linezolid targets the same region but binds differently. Understanding the geometry helps you predict cross-resistance patterns when you're designing combination therapies. Here's a counter-intuitive point most beginners miss: ribosome density on an mRNA doesn't always correlate with protein output. Sometimes adding more ribosome binding sites or strengthening the RBS actually decreases yield because of the queuing problem I mentioned. The optimal RBS strength is context-dependent. Too weak and you get no translation. Too strong and ribosomes collide, trigger stress pathways, and the cell shuts down expression or degrades the mRNA. I've optimized expression systems where dialing back the RBS strength by half doubled the final protein yield. Another practical nuance: ribosome profiling, or Ribo-seq, reveals that a significant fraction of cellular ribosomes aren't translating coding sequences. They're sitting on non-coding RNAs, stalled at structural elements, or engaged in regulatory pausing that controls co-translational processes. The active pool is smaller than you'd expect from total ribosome counts measured by ultracentrifugation.
Get the Full Details

If you're working with eukaryotic systems, there's the added complication of the Shine-Dalgarno alternative. Eukaryotic ribosomes use the 5' cap and scanning mechanism instead of direct mRNA-rRNA pairing. This makes them slower to initiate but better at dealing with structured 5' UTRs. The tradeoff is that you get more leaky scanning and reinitiation events, which complicates operon-like expression constructs in yeast or mammalian cells. For practical lab work, I'd recommend checking codon usage tables for your host organism before ordering synthetic genes. Most gene synthesis companies do this automatically now, but if you're working with unusual hosts - Pichia pastoris, Bacillus subtilis, insect cells - the recommendations differ. And if you're seeing heterogeneous product or truncation bands on gels, ribosome stalling at rare codons or mRNA secondary structure should be your first suspect, not protein degradation.