The Basics of Getting Polymers to Form
Polymerization is one of those terms you see everywhere in materials science, but most people treat it like it is some kind of black art. It is not. At its core, you are taking small molecules called monomers and linking them together into long chains. That is about it. The complexity shows up in the details — how fast the reaction goes, what byproducts you get, and whether your chains end up straight or tangled. I have spent years troubleshooting polymer batches that went wrong for reasons nobody expected. One thing I learned early is that lab-scale success rarely translates directly to production. The heat profile changes. Mixing isn't the same. Impurities that were negligible at 50 milliliters become dealbreakers at 50 liters. If you are working with this stuff, you need to respect the scale.
What Is The Polymerization Process Actually About
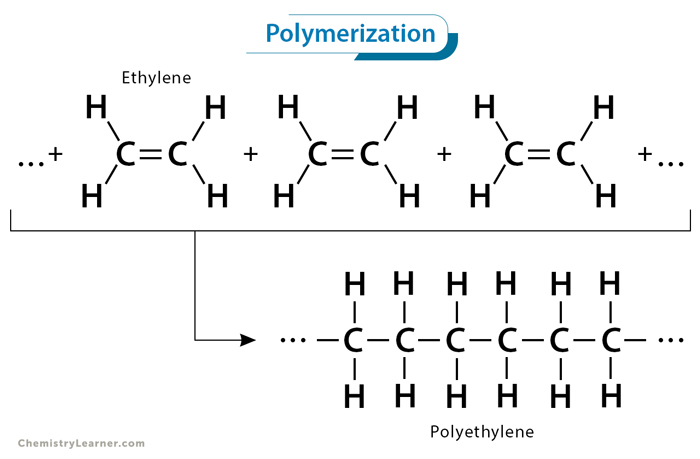
There are really two main families here, and they behave very differently. In addition polymerization, monomers with double bonds open up and connect one after another. Think ethylene turning into polyethylene. No atoms are lost along the way. The other family is condensation polymerization, where monomers join together and release a small molecule as a byproduct — usually water or methanol. Nylon and polyester fall into this second category. Both types can be run via different mechanisms. Free radical polymerization is the most common industrial approach because it works under relatively mild conditions. You throw in an initiator, heat things up, and the radicals start the chain reaction. But free radical processes come with their own headaches — branching can occur, molecular weight distribution gets wide, and controlling the exact chain length is basically impossible without specialized techniques like RAFT or ATRP. Living polymerization techniques give you much better control over molecular weight and structure. They cost more, they are slower, and they require cleaner reaction conditions. For research work they are wonderful. For manufacturing, the economics usually don't justify them unless the final product demands very specific properties.
How to Run a Basic Free Radical Polymerization
Let me walk through a straightforward styrene polymerization. This is the kind of reaction you can set up with standard glassware and reasonable safety precautions. You need styrene monomer, an initiator like benzoyl peroxide or AIBN, and a solvent if you want to control viscosity. Styrene tends to be shipped with a small amount of inhibitor to prevent premature polymerization, so you will likely need to remove that. A simple wash with sodium hydroxide solution followed by distillation works fine. If you skip this step, your reaction will sit there doing nothing for hours while the inhibitor burns itself out. The procedure itself is straightforward. Dissolve your initiator in the monomer under nitrogen atmosphere. Heat to around 70 to 80 degrees Celsius for AIBN, or a bit higher for benzoyl peroxide. The reaction will start on its own once the initiator decomposes. You should see the mixture thicken as the chains grow. Once the conversion hits where you want it, cool the batch and precipitate the polymer into a non-solvent like methanol or water. Dry it under vacuum and you have your polystyrene.
Get the Full Details

Conversion rates in typical lab runs with styrene and AIBN at 70 degrees reach somewhere around 60 to 70 percent before the gel effect kicks in. After that point, the reaction rate spikes because the growing chains trap radicals and termination becomes diffusion-limited. This auto-acceleration is called the Trommsdorff effect, and it is why reactions sometimes run away if you are not paying attention. I lost one batch because I left it overnight without monitoring temperature. The flask cracked from the exotherm.
Common Problems and What Actually Works
Oxygen is the enemy in free radical polymerization. It reacts with radicals to form peroxy species that are far less reactive, which means your initiation drops and molecular weights suffer. Always degas your reaction mixture. Vacuum-nitrogen cycling three times is the standard approach. If you are working on a larger scale, sparging with inert gas for 20 to 30 minutes works adequately. Molecular weight control is another persistent issue. With conventional free radical polymerization, you are essentially at the mercy of the kinetic chain length, which depends on initiator concentration, monomer concentration, and temperature. Double the initiator concentration and you roughly halve the molecular weight. Raise the temperature and you get faster reaction but lower molecular weight. There is no way around these trade-offs in a standard setup. Here is something most beginner guides do not mention: solvent choice matters more than you would think. Bulk polymerization — running the reaction without any solvent — gives the highest molecular weights but creates enormous viscosity problems as conversion increases. The mixture becomes so thick that heat transfer breaks down and you get hot spots. Dilute the reaction with a solvent like toluene or THF and you get better heat control and narrower molecular weight distributions, but you have to deal with solvent removal afterward.
I found that for most practical purposes, a 30 to 40 percent solids content in toluene hits a sweet spot. The reaction stays manageable, heat dissipation is adequate, and the solvent comes off cleanly under vacuum without degrading the polymer.

Characterization and Quality Checks
Once your polymer is made, you need to verify what you actually got. Gel permeation chromatography, also called size exclusion chromatography, is the standard method for determining molecular weight and the distribution width. This typically takes 30 to 45 minutes per sample on a well-maintained instrument. You will get values for Mn, Mw, and the polydispersity index. A PDI below 1.5 indicates reasonably controlled polymerization. Above 2.0 means you have a broad, uncontrolled distribution. Fourier transform infrared spectroscopy will tell you whether your reaction proceeded as expected. Look for the disappearance of vinyl peaks around 1638 inverse centimeters and the appearance of backbone C-H and C-C signals. NMR gives you conversion data and can detect residual monomer if you quantify the ratio of monomer vinyl peaks to polymer aromatic peaks. Differential scanning calorimetry measures your glass transition temperature. For polystyrene, expect around 100 degrees Celsius. If your Tg is significantly lower than literature values, you probably have residual solvent or low molecular weight oligomers in your sample. A higher Tg could indicate crosslinking, which happens when there is significant branching or when di-functional monomers are present as impurities.
When Conventional Polymerization Fails
Sometimes you need polymers with very narrow molecular weight distributions, specific architectures like block copolymers, or functional end groups. Conventional free radical polymerization cannot deliver these. You need controlled radical techniques or ionic polymerization methods. RAFT polymerization has become the most accessible controlled radical method. It works with a wide range of monomers including acrylates, methacrylates, and styrenics. The trade-off is that you need a chain transfer agent, which can be expensive and sometimes difficult to remove from the final product. The dithioester end groups from common RAFT agents also leave a yellow color that may be unacceptable for optical applications. ATRP requires a transition metal catalyst, usually copper based. It gives excellent control but leaves metal residues that are problematic for electronic or biomedical applications. There are copper-free variants, but they tend to be slower and less robust. I switched to photo-mediated RAFT for most of my recent work because it avoids both metal contamination and high temperatures.
For applications where absolute purity matters, anionic polymerization is still the gold standard. But it requires rigorously dry and oxygen-free conditions, low temperatures often below minus 70 degrees Celsius, and monomers that are exceptionally pure. One droplet of water can kill the entire living chain. It is reliable if you have the equipment and patience, but it is not something you set up casually.

A Practical Note on Scaling
If you move from a 100 milliliter flask to a 10 liter reactor, everything changes. The surface area to volume ratio drops dramatically, which means heat removal becomes much harder. The exotherm that was manageable at small scale can cause thermal runaway at larger scale. I once scaled a methyl methacrylate polymerization by a factor of 100 and nearly had a safety incident because I underestimated the heat generation. The reaction temperature spiked to 140 degrees instead of staying at the intended 80 degrees, and the polymer degraded badly. The workaround is stepwise addition of monomer rather than charging everything at once. You start with a small amount of monomer and most of the solvent, let the reaction establish a steady state, then add the remaining monomer slowly over several hours. This keeps the exotherm spread out and the temperature controlled. It also tends to give more consistent molecular weight because the radical concentration stays relatively constant throughout the reaction. Another consideration is mixing efficiency. In a large vessel, dead zones can form where the reaction mixture sits and heats up without circulation. This leads to localized high conversion and potentially crosslinking. Impeller design and placement matter more than people usually account for. A standard pitched blade turbine is adequate for most polymerizations, but highly viscous systems may need anchor or helical ribbon impellers.
Bottom Line
Polymerization is not mysterious. It is chemistry, and like most chemistry, it rewards people who understand the fundamentals and respect the process. Start with small scale. Monitor temperature carefully. Characterize your product. Expect that things will go wrong, and plan for the obvious failure modes before they happen. The difference between a successful batch and a ruined one is often something as simple as forgetting to degas or miscalculating the initiator amount. Most of the time, the practical issues are straightforward to solve once you know what to look for. Molecular weight too low — check your initiator concentration and reaction temperature. Reaction did not start — verify your initiator is active and your atmosphere is truly inert. Product discolored — consider whether oxidation or thermal degradation is the culprit. Each problem has a diagnostic path, and following it systematically is faster than guessing.


