Measuring Vapor Pressure Without Losing Your Mind
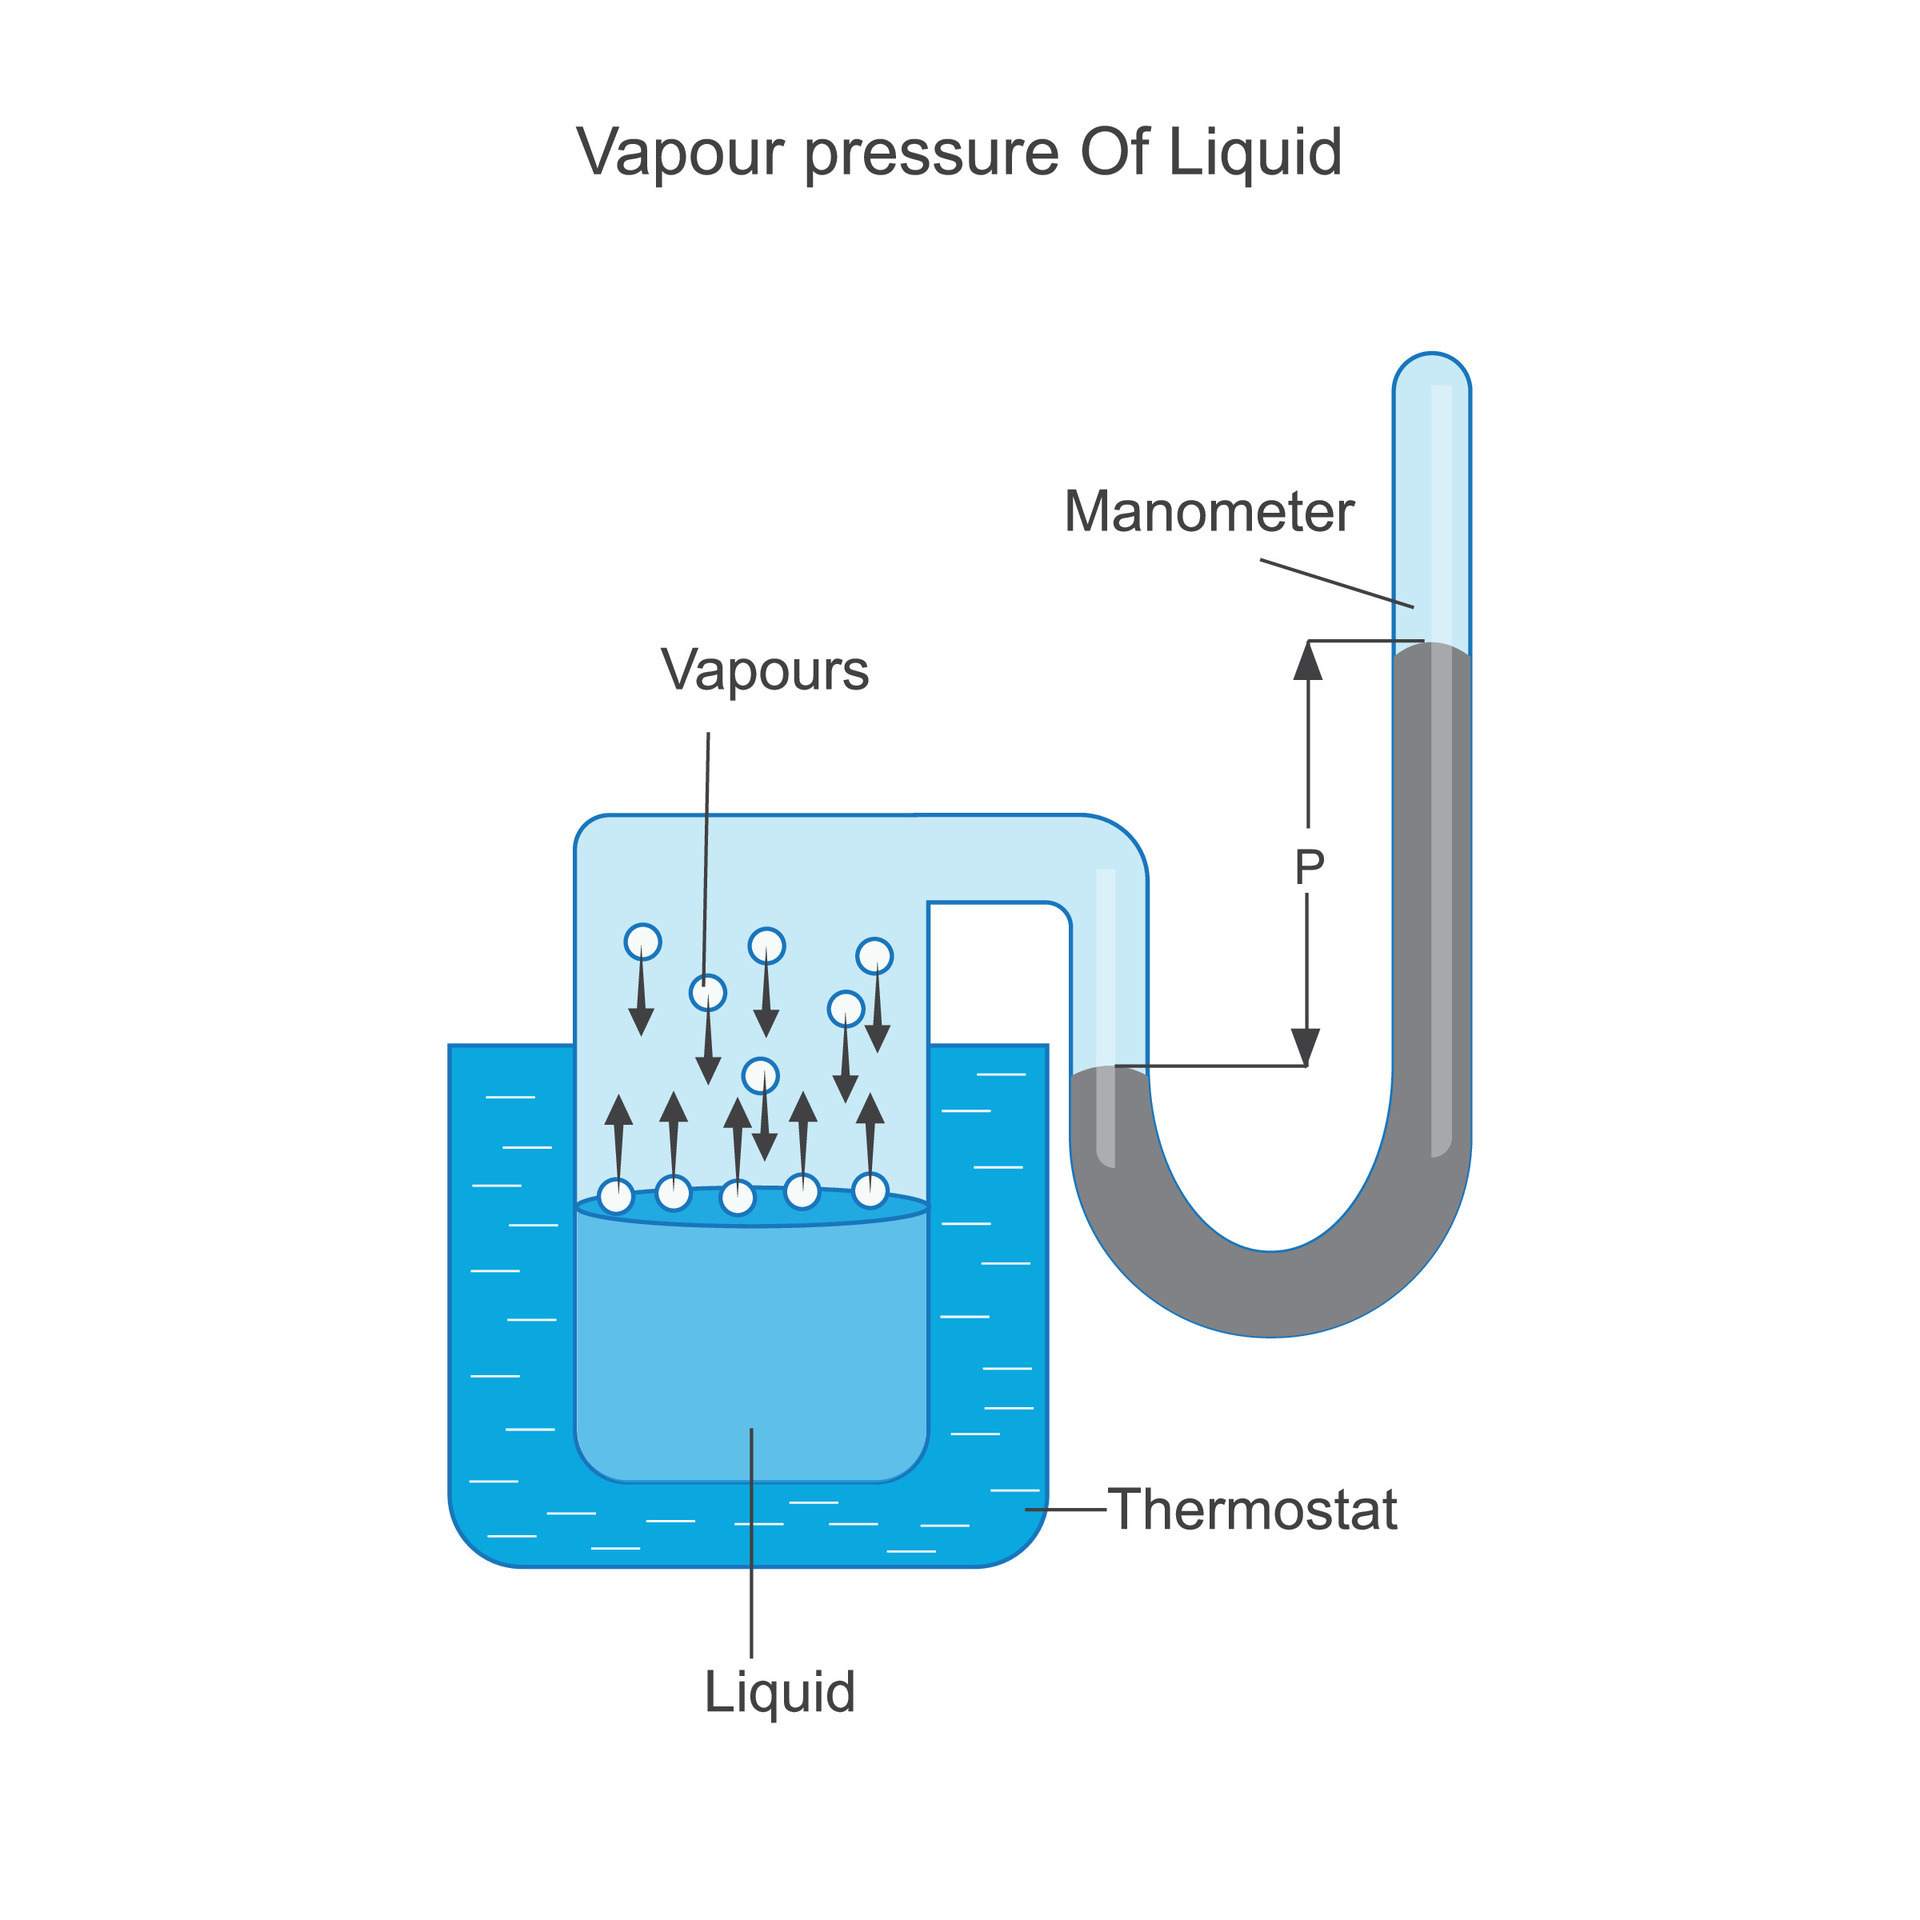
Most people reach for a textbook definition when they need to understand vapor pressure, but that rarely helps when you're standing in front of a pressurized vessel or trying to size a relief device. Start with the practical method: heat the liquid in a closed system and measure the pressure that develops at each temperature point. Run it slowly. Equilibration takes longer than people expect, especially with viscous or mixed compositions. You will see the gauge drift for ten to fifteen minutes after you think things have settled. Let it run. Record the stable reading. That number, plotted against temperature, is your vapor pressure curve. There are several standard methods, and picking the right one matters more than you might think. The ASTM E680 procedure uses a static apparatus where you gradually heat a sealed chamber containing the pure liquid and watch the pressure climb. It is accurate for clean substances but takes about four to six hours for a single data point set. The dynamic method, like ASTM D2879, is faster but less precise for low-volatility materials. For high-pressure applications such as refrigerant blends or supercritical fluids, a visual meniscus disappearance method gives you the critical point directly, which static methods sometimes miss by a degree or two. I recommend the static approach for anything under 50 bar and the dynamic method when you need throughput and the material is relatively clean.
What Is Vapor Pressure and Why Does It Matter in Practice
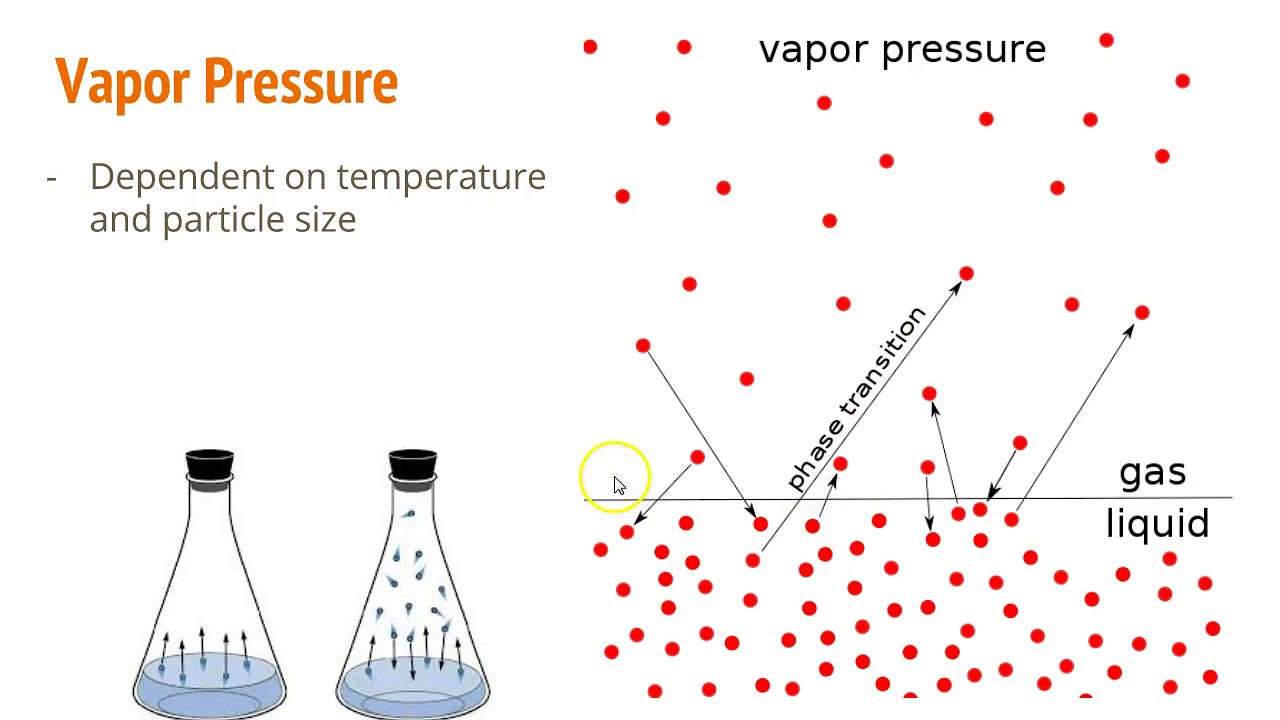
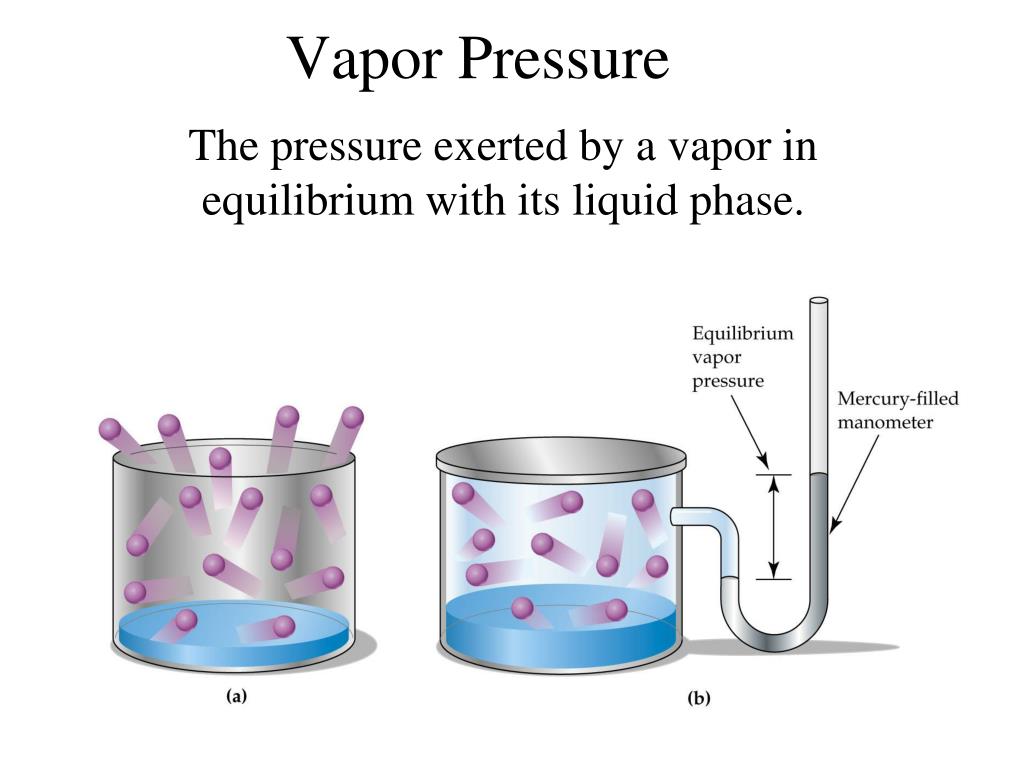
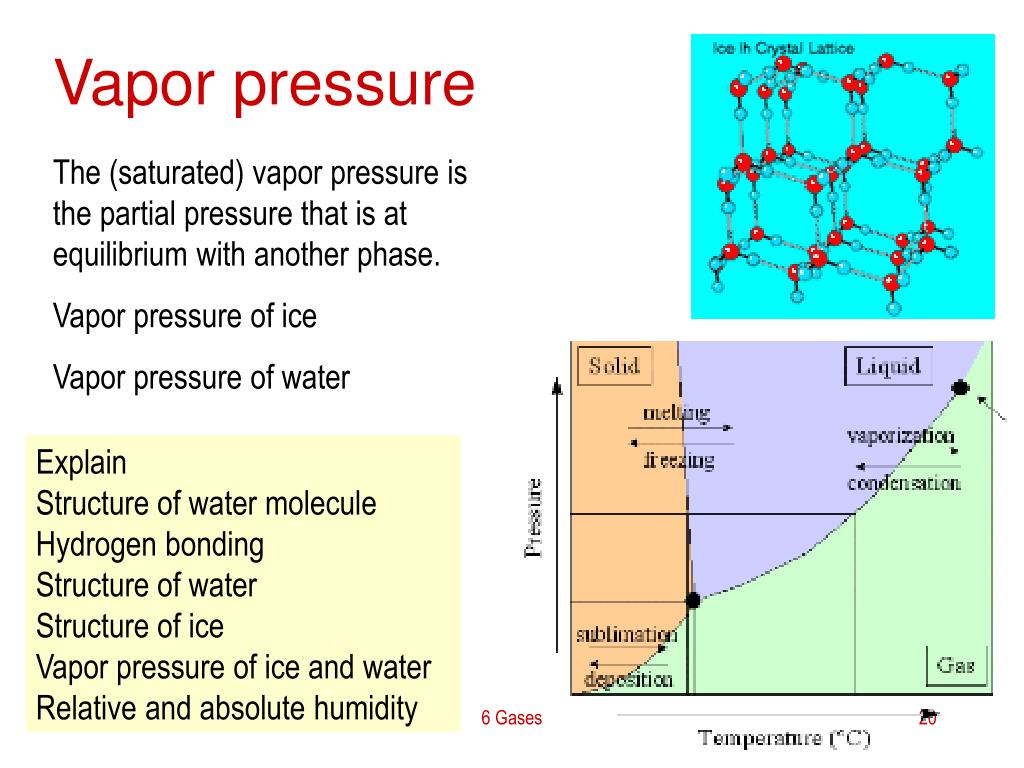
Vapor pressure is simply the pressure exerted by a vapor in equilibrium with its liquid phase at a given temperature. That equilibrium means molecules are leaving the liquid and entering the gas phase at the same rate they condense back. It is not the total pressure in a container unless the container holds only that substance and its vapor. When you have air or another gas present, you are dealing with partial pressure, which follows Dalton's law. The distinction trips up everyone at least once. I once saw a process engineer treat the reading on a vent line as the vapor pressure of a solvent blend and proceed to size relief equipment based on a number that was twenty percent too low because the headspace contained significant amounts of nitrogen from an inerting sweep. The calculation was wrong from the start. The reason vapor pressure matters is that it controls phase behavior across almost every industrial process. Distillation columns rely on relative volatility, which is derived from vapor pressures. Pump suction designs depend on net positive suction head available, which comes directly from the liquid's vapor pressure at operating temperature. Storage tank pressure ratings, pipeline classification, and flare system sizing all trace back to this single property. Get it wrong and you get cavitation, overpressure events, or product loss through vents. Temperature dependence is nonlinear and steep. The Clausius-Clapeyron relation describes it, and for rough estimates you can use the integrated form, but the real world is messier. Mixtures do not follow a simple equation. Each component contributes according to its mole fraction and activity coefficient, which means you need Wilson, NRTL, or UNIQUAC parameters if you want accuracy. Raoult's law works for ideal solutions, and very few industrial mixtures are ideal. Benzene and cyclohexane deviate enough to cause serious errors in column design if you assume ideality. Water and ethanol form an azeotrope that distillation cannot break without an entrainer or pressure swing. These are not edge cases. They are routine.
I ran into a specific problem last year involving a hydrocarbon cut that contained trace amounts of C2 through C5 components dissolved in a C8+ stream. The vendor supplied vapor pressure data from a Reid method test, which measures at 37.8 degrees Celsius with a vapor-to-liquid ratio of four to one. The specification called for a maximum Reid vapor pressure of 10 psi, so the material passed inspection. We stored it in a atmospheric tank at ambient temperature and still had periodic pressure relief valve lifts during afternoon heat exposure. The Reid test does not capture the true equilibrium vapor pressure at storage conditions because it forces a large vapor space and a fixed temperature. The actual bubble point of that mixture at 25 degrees Celsius was closer to 8 psi, and under direct sunlight the tank wall temperature reached 50 degrees Celsius, pushing the equilibrium pressure well past the relief set point. The workaround was straightforward: we switched to pressure testing using a constant-volume cell at the actual storage temperature range instead of relying on the Reid value alone. We also added a floating roof to eliminate the headspace vapor volume. The relief events stopped.
Get the Full Details
Counter-Intuitive Points Beginners Miss
First, vapor pressure has nothing to do with how fast a liquid evaporates under open conditions. Evaporation rate depends on surface area, airflow, humidity, and the difference between the saturation pressure and the actual partial pressure in the gas phase. A liquid with high vapor pressure can evaporate slowly in still, saturated air. A liquid with moderate vapor pressure can evaporate quickly in a breeze. People conflate these because both relate to volatility, but they are different phenomena governed by different equations. Second, adding a non-volatile solute to a liquid raises its boiling point but also changes the vapor pressure in a way that is not always obvious. Freezing point depression and boiling point elevation are colligative properties, meaning they depend on the number of solute particles, not their identity. But the vapor pressure lowering follows the same logic only for ideal dilute solutions. In concentrated electrolyte solutions, ion pairing and activity coefficient effects dominate, and the simple mole-fraction relationship breaks down. I have seen formulations fail because someone applied Raoult's law directly to a brine system without accounting for activity coefficients. The predicted vapor pressure was off by nearly forty percent. Third, vapor pressure is independent of the amount of liquid and the size of the container, provided both phases are present. This sounds trivial, but it causes real mistakes in scale-up. A small batch in a lab flask and a large tank of the same material at the same temperature will show the same equilibrium vapor pressure. However, the time to reach equilibrium scales with volume and surface area, and the heat capacity of the system affects how quickly temperature changes propagate. Operators sometimes assume that because a lab-scale test showed stable pressure, a production vessel will respond the same way. It will not. The thermal lag in a large vessel can create transient pressure differences that exceed the equilibrium value temporarily during heating or cooling cycles.
Limitations and When This Approach Fails
Static methods fail above the critical point because there is no distinct liquid-vapor interface. Dynamic methods struggle with high-viscosity materials because mass transfer within the liquid becomes the rate-limiting step, and the measured pressure lags behind the actual equilibrium value. Mixtures with wide boiling ranges require incremental removal of the more volatile component during measurement, which changes the composition and invalidates a single reading. Azeotropes create local maxima and minima in the vapor pressure curve that standard correlation methods smooth over unless you have experimental data to anchor them. If you need vapor pressure data for a novel compound with no published information, the best approach is to measure it yourself rather than relying on group contribution methods like Joback or Lydersen. Those methods typically have errors in the range of fifteen to thirty percent, which is unacceptable for relief device sizing or column design. UNIFAC can predict mixture vapor pressures with reasonable accuracy for thermally stable systems, but it breaks down for associating fluids like alcohols and water at high concentrations. In those cases, you need experimental vapor-liquid equilibrium data from the literature or your own measurements. For routine engineering work, the DIPPR database and NIST Chemistry WebBook are reliable sources, but always check the original reference. Data has been transcribed incorrectly multiple times, and temperature units are sometimes converted wrong in secondary sources. A value listed in kelvin instead of celsius or vice versa will throw off every calculation that follows. I have caught this error in vendor-provided datasheets on more than one occasion.