The Claisen Condensation: What Actually Works
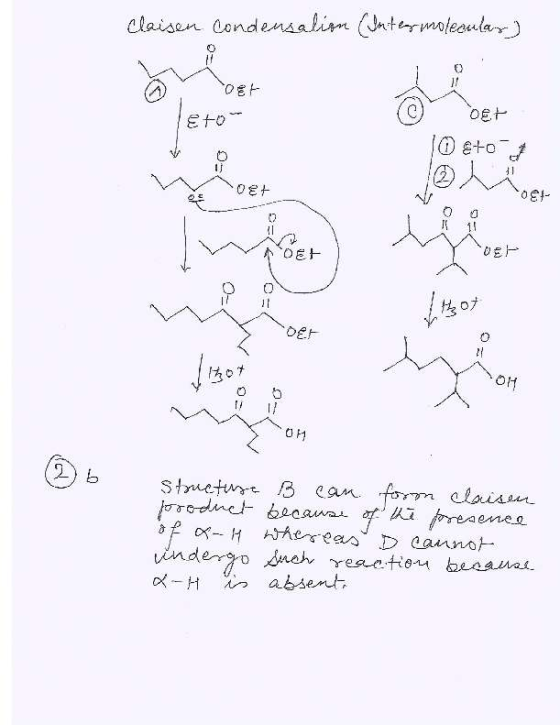
The Claisen condensation requires two specific things from your ester: it needs at least one alpha hydrogen, and the ester oxygen should ideally come from an alkoxide that matches the leaving group to avoid transesterification messes. The reaction joins two ester molecules together, forming a beta-keto ester. That's the basic definition. But the practical details are where things get fussy. Self-Claisen reactions work cleanly with esters that have two or more alpha hydrogens on the same carbon. Diethyl adipate cyclizes nicely to form 2-carbethoxycyclopentanone under standard conditions. Diethyl succinate gives you a six-membered ring product. These intramolecular versions are actually more reliable than the intermolecular ones because the entropy factor works in your favor during ring closure. Simple esters like ethyl acetate and ethyl propionate undergo regular Claisen condensation without trouble. You dissolve them in their corresponding alcohol, add sodium ethoxide, and the reaction typically completes within an hour or two at room temperature. The product precipitates when you acidify the reaction mixture because the beta-keto ester is insoluble in aqueous acid. That's one of the few easy workups you'll encounter in organic synthesis.
Crossed Claisen reactions between two different esters are where things get complicated. If both esters have alpha hydrogens, you get four possible products instead of one. That's not a practical synthesis. The workaround is to use one ester that has no alpha hydrogens as the electrophile partner. Ethyl benzoate and ethyl formate are the standard choices here. You add the enolizable ester slowly to a mixture containing the non-enolizable ester and base, which keeps the concentration of the enolate low relative to the electrophile and suppresses the unwanted self-condensation pathway. Carbonate esters like diethyl carbonate also work well as the non-enolizable partner. I've used them repeatedly when I needed to introduce a carboxylic acid group onto a ketone product through subsequent decarboxylation. Special case: esters with bulky groups near the reaction site tend to give poor yields even when they have alpha hydrogens. The steric hindrance slows down the initial deprotonation and the subsequent nucleophilic attack. I had a substrate with a tert-butyl group three bonds away from the carbonyl and got maybe 30% yield instead of the expected 70%. Switching from sodium ethoxide to potassium bis(trimethylsilyl)amide and running the reaction at -78°C improved things enough to make the reaction viable, though the isolation was still tedious.
The Mechanism and Why It Matters for Substrate Selection
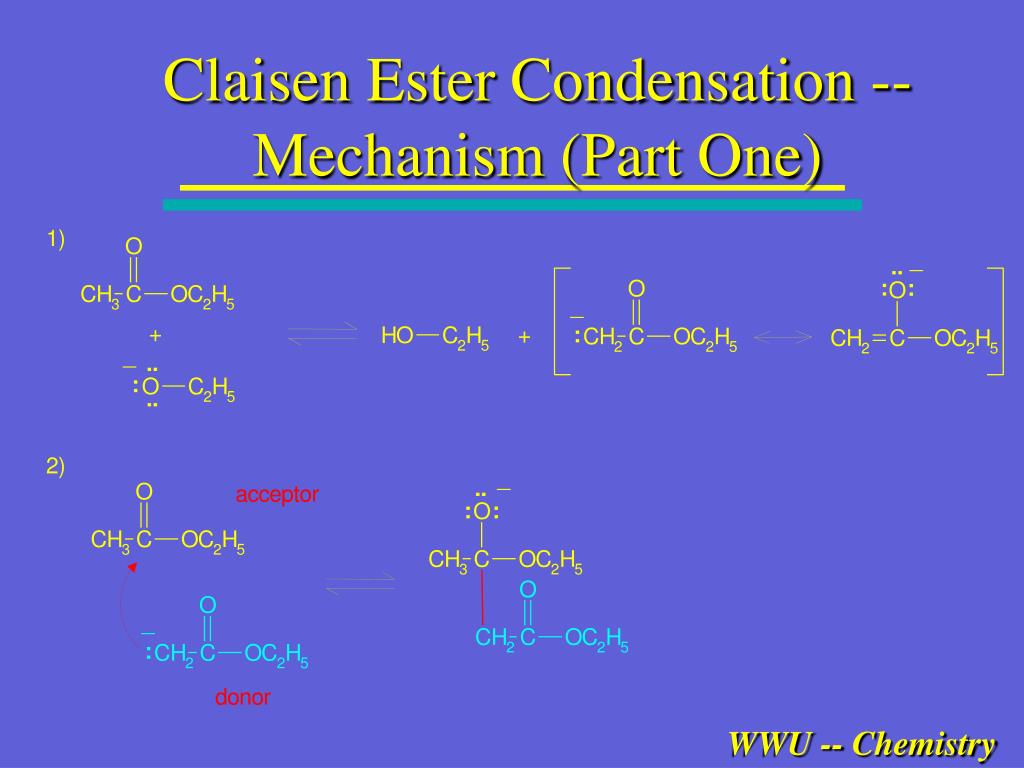
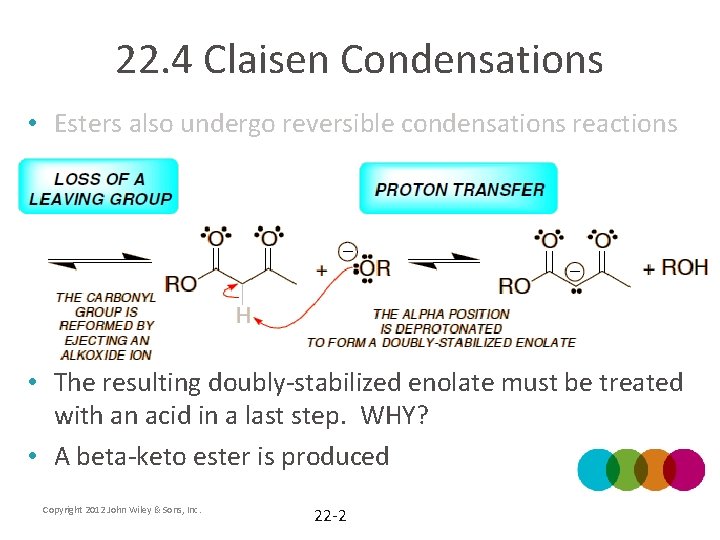
The reaction proceeds through deprotonation of the alpha carbon to form an enolate, nucleophilic attack of that enolate onto the carbonyl carbon of a second ester molecule, and loss of the alkoxide leaving group. The final step is a driven deprotonation of the beta-keto ester product by the alkoxide base. This last step is crucial because it's essentially irreversible and pushes the equilibrium toward product formation. Without that final deprotonation, the reaction reaches an equilibrium that heavily favors the starting materials. The pKa of a beta-keto ester is around 11, so any alkoxide base stronger than ethoxide will drive the reaction to completion. Sodium methoxide, sodium ethoxide, sodium hydride, and potassium tert-butoxide all work. The choice mostly affects whether transesterification becomes a side reaction. Here's something most textbooks don't emphasize enough: the choice of solvent matters more than people realize. Dry ethanol is standard for ethyl esters, but if your ester is expensive or scarce, using excess ester as both reactant and solvent eliminates the need for an added solvent entirely. This is the classic procedure for ethyl acetate self-condensation and it works because you're not diluting the reaction mixture unnecessarily.
Get the Full Details

For less common esters, THF or toluene with a stoichiometric base is better. I switched from ethanol to THF when working with an ester bearing a silyl ether protecting group because the basic ethanol was slowly cleaving the TMS group over several hours. The reaction took longer in THF but the protecting group stayed intact and the yield jumped from 45% to 72%.
Common Pitfalls and How to Avoid Them
The biggest issue is incomplete deprotonation. If you use less than one full equivalent of base relative to your ester, the reaction stalls partway through because there isn't enough enolate formed. I've seen people use 0.5 equivalents and then wonder why the yield is terrible. Use at least one equivalent. Better yet, use a slight excess like 1.1 equivalents to account for any moisture or protic impurities in your reagents. Water is the enemy here. Even small amounts of moisture consume the base and quench the enolate before it can attack the ester carbonyl. Keep your glassware dry, use anhydrous solvents, and work under inert atmosphere if your ester is sensitive. I learned this the hard way when a reaction that usually gave 80% yield dropped to 15% because I skipped the flame-drying step on my flask. Took me three attempts to figure out what was going wrong. Another issue specific to certain substrates: esters with electron-withdrawing groups on the alpha carbon are overactivated and can undergo side reactions like elimination or polymerization under basic conditions. I worked with an ethyl 2-chloroacetate derivative and got significant amounts of vinyl chloride byproduct alongside the desired Claisen product. Lowering the temperature to 0°C and adding the base more slowly reduced the elimination pathway enough to make the reaction workable.
When the ester has only one alpha hydrogen, the final deprotonation step of the product doesn't occur, which means the equilibrium doesn't favor products. The reaction becomes reversible and you get poor yields. This is why diesters and monoesters with two alpha hydrogens on the same carbon are the most reliable substrates. If you must use a monoester with only one alpha hydrogen, consider using a stronger base like NaH in THF and driving the reaction with extended time and elevated temperature, though even then the yields are usually mediocre at best.
Intramolecular Variants and Beyond
The Dieckmann condensation is simply the intramolecular version of the Claisen reaction. Diesters with the right chain length cyclize to form cyclic beta-k keto esters. The 1,6-diester gives a five-membered ring product most readily. The 1,7-diester gives a six-membered ring. Five- and six-membered rings form much faster than three-, four-, or seven-membered rings due to ring strain and entropy factors. If your diester would form a seven-membered ring, the intermolecular pathway usually wins and you get polymer or dimer instead of the desired cycle. Malonic esters and similar 1,3-diesters behave differently. They can undergo Claisen-type condensation but the product often decarboxylates under the basic reaction conditions if the temperature gets high enough. That's not always a problem. Sometimes you want the decarboxylation to happen in situ and you just isolate the final ketone product after acid workup. It saves you a separate step. The Rosenberg modification uses sodium amide in liquid ammonia instead of alkoxide bases. This is useful when your ester contains base-sensitive functional groups that would decompose under traditional conditions. The reaction is faster at -33°C than at room temperature with alkoxide, and the non-nucleophilic amide anion avoids transesterification entirely. I used this approach for a substrate containing an acetalsensitive protecting group that survived the sodium amide conditions but would have been destroyed by sodium ethoxide in ethanol.
When the Claisen Is Not the Right Tool
If your ester lacks alpha hydrogens entirely, like ethyl pivalate or ethyl benzoate, you cannot run a self-Claisen. These can only participate as the electrophilic partner in a crossed Claisen. If you need to couple two identical esters that lack alpha hydrogens, you'll need a different strategy entirely, like a ketone-ester condensation or a directed ortho metalation approach followed by ester trapping. For esters that are highly sterically hindered, the Stork enamine synthesis followed by acylation and hydrolysis sometimes gives better results than direct Claisen condensation. I swapped to the Stork method for a diisopropyl-substituted ester and went from 20% yield to 65% in a single step. The enamine intermediate bypasses the need for strong base deprotonation of a crowded alpha carbon. Similarly, if you're working with a substrate that has multiple reactive sites and you need chemoselective ester condensation at only one position, modern alternatives like organocatalytic esterase-mediated coupling or enzymatic resolution might be worth exploring, though those are in a different ballpark cost-wise and scale-wise from a standard Claisen.