Working With Yeast in the Lab
Yeast As A Model Organism
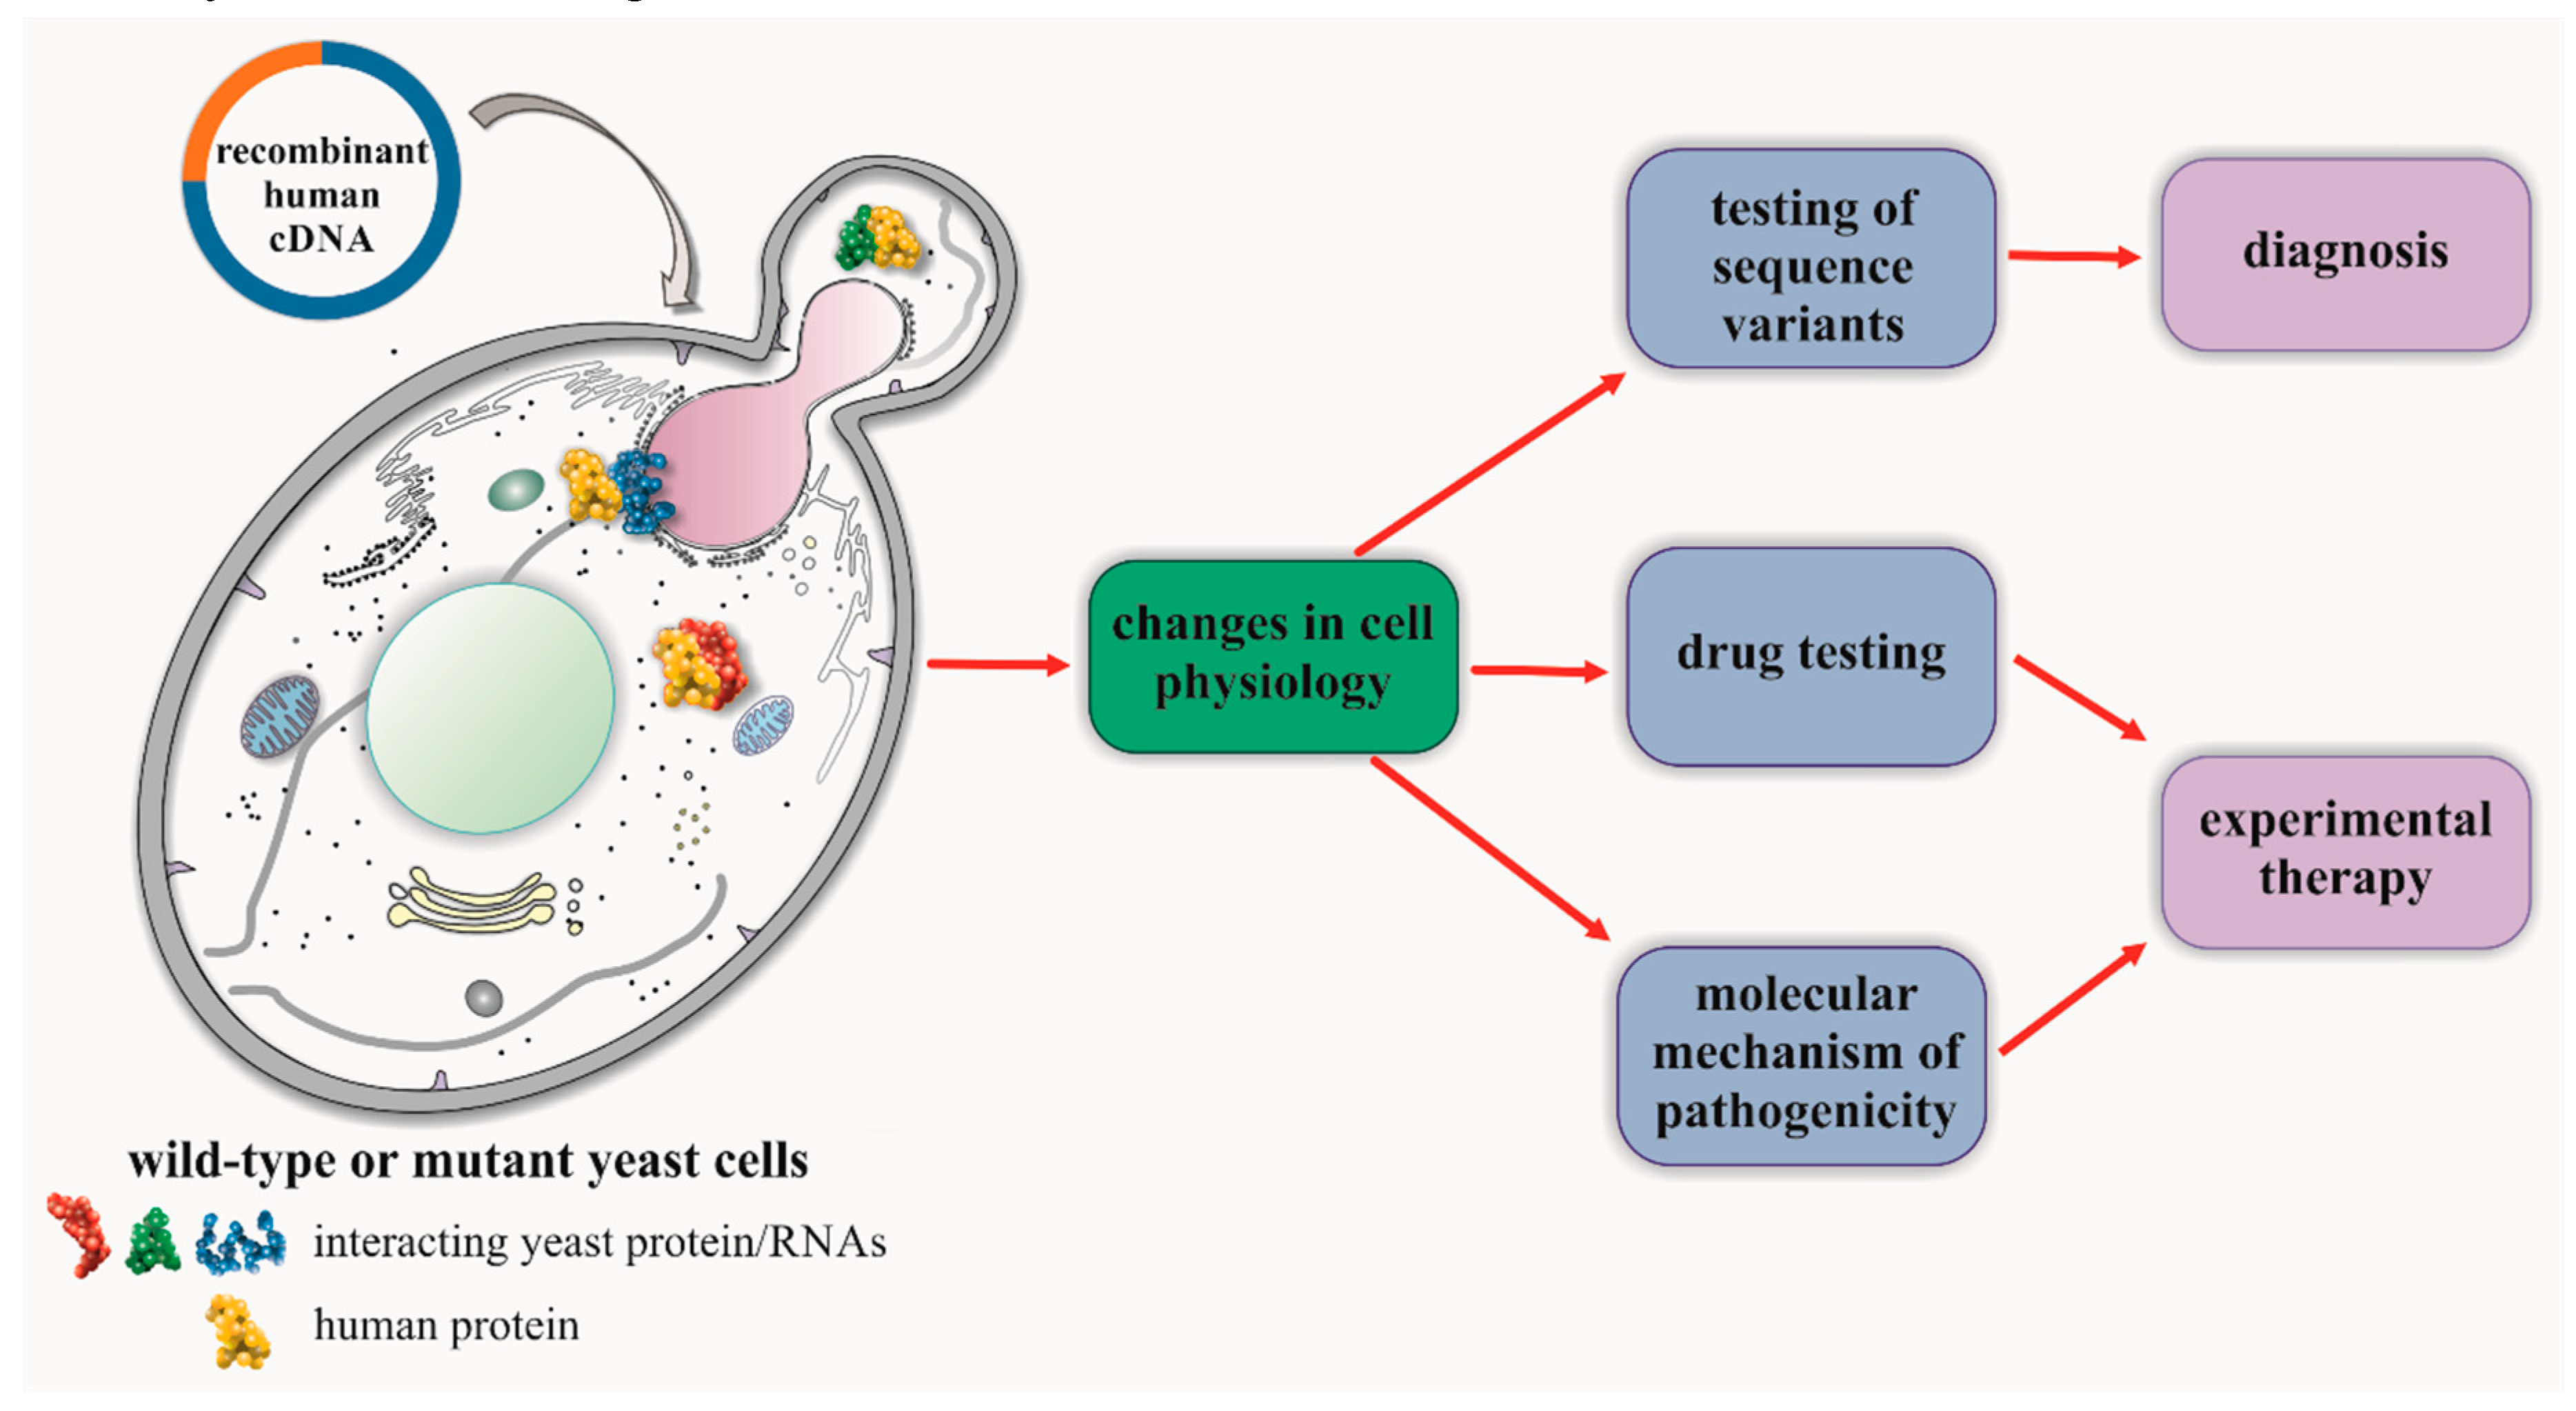
Yeast is small, grows fast, and has been genetically manipulated for decades. When you pick it as a model organism, you are choosing one of the most thoroughly characterized eukaryotic systems available. Saccharomyces cerevisiae has a complete genome sequence, a vast collection of knockout strains, and well-established protocols for everything from standard cloning to CRISPR-based genome editing. The main reasons people use it are speed and relevance. It divides every 90 minutes under optimal conditions, which is slow compared to bacteria but still practical for eukaryotic work, and its cellular machinery is closer to human cells than prokaryotic systems, making findings more translatable. I keep a few stock plates of S. cerevisiae on the bench because they are reliable. The standard laboratory strain is S288C and its derivatives. If you order gene knockout collections, Euroscarf and the Yeast Knockout Collection are the usual sources. Transformation is generally done with the lithium acetate method. You grow cells to mid-log phase, wash them in lithium acetate solution, mix with carrier DNA and PEG, heat shock briefly, and plate on selective media. The whole process takes about two hours from an overnight culture. One practical detail that catches people off guard: yeast do not grow well on rich media if you are trying to maintain plasmids without selection pressure. I once spent three weeks wondering why my transformants kept losing a URA3-containing vector. It turned out the lab's YPD plates had been sitting in the incubator for two weeks past their expiration date, and the dropout mix had degraded enough that the selective pressure was essentially gone. New plates solved the problem immediately. This is the kind of issue that does not appear in protocols but costs real time in practice.
Counter-selection is one of the more useful features of yeast genetics. The URA5 gene allows you to select for loss of function using 5-fluoroorotic acid. This is how golden gate-style assembly and scarless editing often work in yeast. You clone your construct into a URA3 marker, select for integration, then use FOA plates to cure the marker. It is efficient but not foolproof. Background can be an issue. Some strains naturally carry suppressor mutations that allow growth on FOA even without the expected gene disruption. If you are working with an unfamiliar strain, always include a no-template control on your FOA plates to check for background. CRISPR editing in yeast is straightforward if you follow the standard protocol. You design a guide RNA targeting your locus, prepare a linear donor oligonucleotide or PCR fragment with homology arms of about 40 base pairs, co-transform with Cas9 mRNA or a plasmid expressing Cas9, and plate on selective media. Transformation efficiency for CRISPR edits typically lands between 100 and 1000 colonies per microgram of DNA, which is reasonable. Screening is usually done by colony PCR. A single correct edit takes about two days from design to confirmation. There are things yeast will not do well for. Membrane protein expression is one. Yeast have different lipid compositions and glycosylation patterns than mammalian cells, so proteins that require specific post-translational modifications for proper folding or function often misfold or get hyperglycosylated. If your project involves a human GPCR or a complex membrane transporter, yeast may not be the right system. Use a baculovirus or mammalian expression system instead. Another limitation is that yeast do not naturally support large plasmid maintenance. If you need to clone fragments larger than 10 kilobases, BACs or yeast artificial chromosomes are possible but significantly more work than standard plasmid cloning in E. coli. The maintenance becomes unstable without constant selection, and recombination between repetitive sequences is common.
Media and Strain Selection
For routine growth, YPD is the standard. Yeast extract, peptone, and dextrose at 1 percent each. It supports fast growth but is not defined. If you need reproducible experiments, especially for metabolic studies or protein expression where you want to control every variable, switch to YEPD with defined salts or use synthetic complete medium. SC medium requires you to add the right dropout mix. The typical composition is 0.67 grams of yeast nitrogen base without amino acids per liter, supplemented with the amino acids your strain needs minus the one you are selecting for. Strain choice matters more than most people realize. Common lab strains like W303 and S288C derivatives have different backgrounds that affect transformation efficiency, sporulation rates, and tolerance to stress. W303 is known for higher recombination rates, which is useful for certain genetic crosses but problematic if you are doing stable long-term expression. The BY4741 and BY4742 strains from the knockout collections are homozygous disruptants in an epsilon lysing background. They are convenient but more fragile than some wild-type isolates. If you are doing industrial fermentation or stress-response work, consider a non-S288C background strain that has been selected for robustness over many generations. Sporulation and tetrad dissection are where yeast really shines as a genetics tool. You take a diploid heterozygote, grow it on potassium acetate plates for 24 to 48 hours, induce meiosis, and then dissect the resulting asci under a microscope to separate the four spores. This gives you direct access to Mendelian segregation ratios. I have used this approach to map point mutations that were difficult to track with molecular methods alone. The trick is getting the sporulation efficiency high enough. Some strains sporulate poorly on standard acetate plates. Switching to potasse tartarate or adding a small amount of glucose before the shift can improve sporulation rates from 20 percent to over 80 percent in my hands.
Get the Full Details

Protein expression in yeast is another area where people have strong opinions. The pGAP and pYES2 vectors are the most widely used. pGAP uses the glyceraldehyde-3-phosphate dehydrogenase promoter for constitutive expression. pYES2 uses the GAL1 promoter for inducible expression with galactose. The inducible system gives you tighter control over timing, which is important when your protein is toxic or when you want to maximize cell density before inducing. Expression levels can vary widely depending on the integration site. A single-copy integration gives lower but more consistent expression. Multi-copy arrays or integration near strong chromosomal elements can boost yield but increase instability. I usually test three to five independent clones and pick the best performer rather than assuming a single clone represents the construct. If you are doing high-throughput work, automated liquid handling and plate readers have changed the workflow significantly. You can grow yeast in 96-well format, measure optical density at 600 nanometers, induce expression, and harvest cells all in one rack. The data quality is comparable to shake flask cultures for most applications. The main disadvantage is evaporation. Wells on the outer rim of a plate dry out faster, so I avoid using those positions or seal the plate with a breathable membrane. Growth curves in microtiter plates typically take four to six hours from inoculation to mid-log phase, which is faster than parallel liquid cultures in flasks because of the higher surface-area-to-volume ratio. Storage is another area with small but important details. Glycerol stocks at minus 80 degrees Celsius are standard. The typical ratio is one part overnight culture to one part 40 percent glycerol. If you freeze at too high a concentration, viability drops sharply. Thawing on ice and pipetting gently rather than vortexing preserves the culture. Aliquot your stocks so you never thaw and refreeze. A single freeze-thaw cycle can reduce viable counts by half, and repeated cycles make the stock unreliable within weeks.
The bottom line is that yeast is a powerful system when you understand its constraints. It is not a universal solution. For basic molecular biology and genetics, it is hard to beat. For structural work on membrane proteins or for studying eukaryotic pathways that differ significantly from yeast, other systems may be more appropriate. The key is matching the biology of your project to the strengths and weaknesses of the organism.