Understanding the 3 Laws Of Thermodynamics
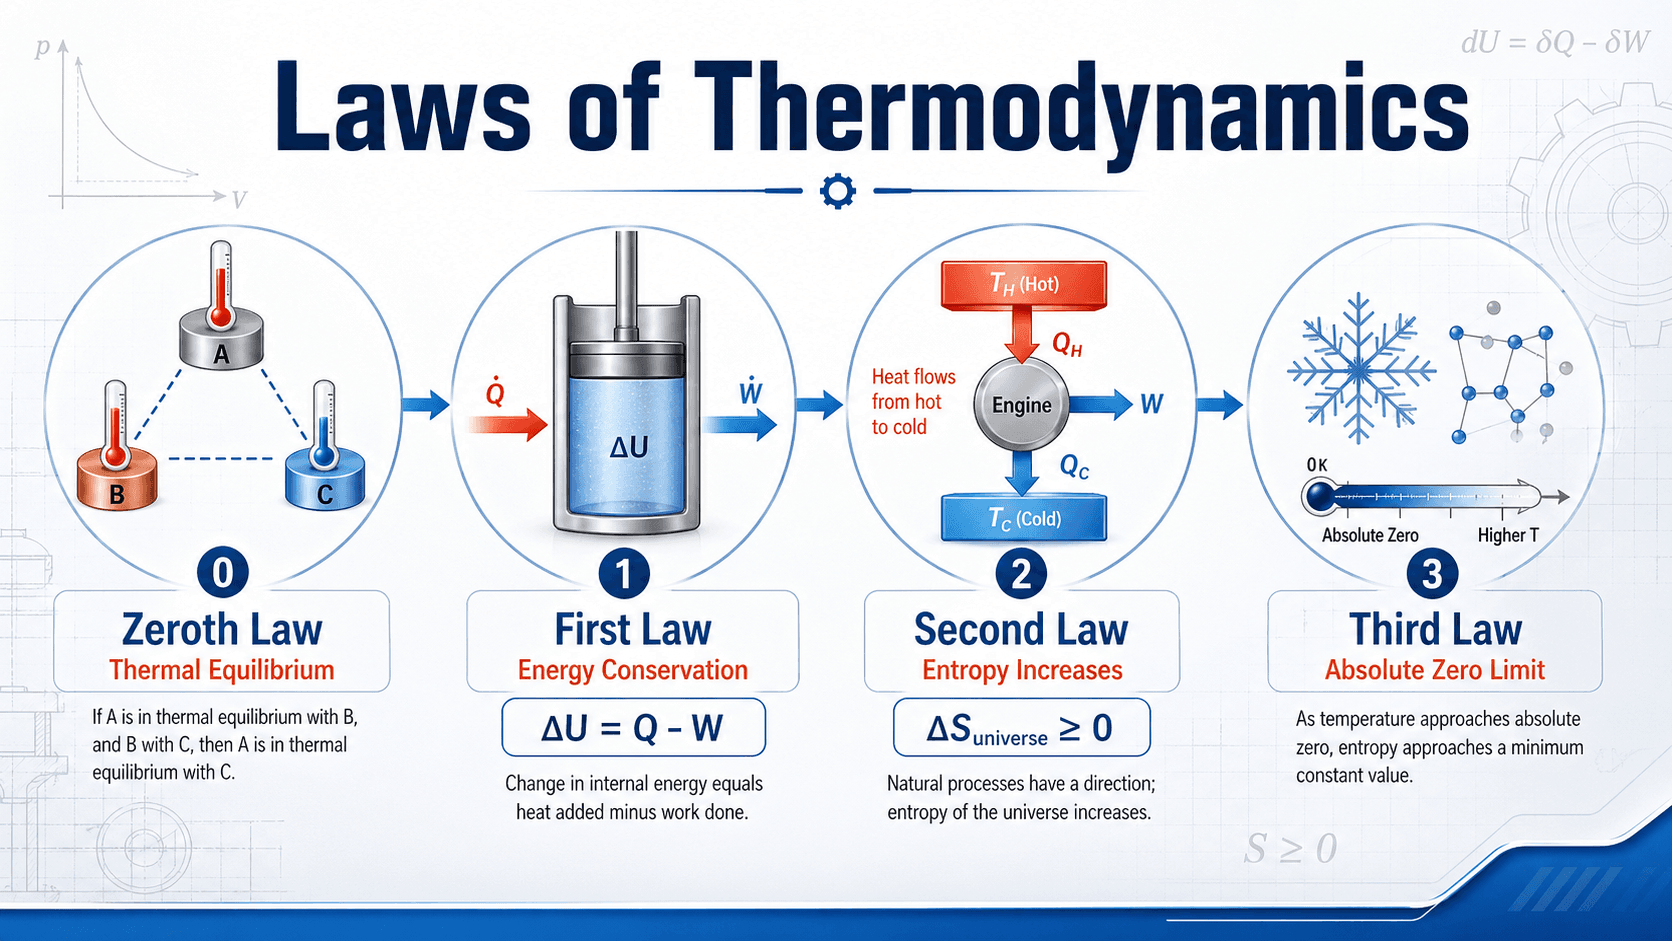
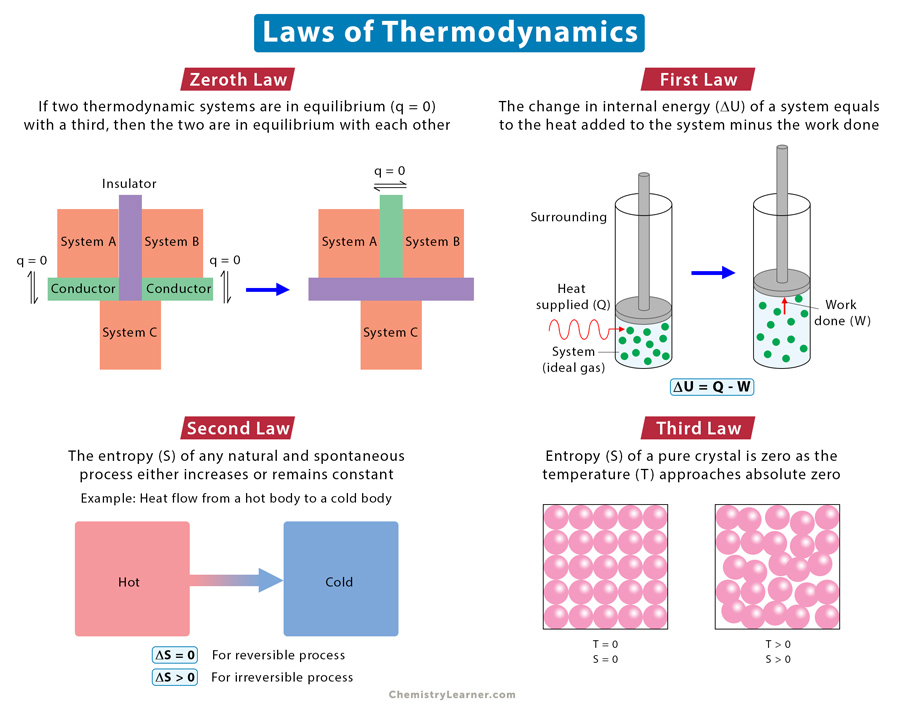
Every time you've seen a steam engine diagram or tried to calculate the efficiency of a refrigeration cycle, you've run into these laws. They aren't abstract philosophy. They're the reason your HVAC system has a COP of maybe 3 or 4 and never 10. Let me walk through what each one actually means and where people regularly mess up the application. First Law: Energy cannot be created or destroyed. In a closed system, any heat added equals the change in internal energy plus the work done by the system. Mathematically: dU = dQ - dW. This sounds simple until you try to apply it to an open system with mass flowing in and out, where you suddenly need enthalpy terms and control volume analysis. People forget the sign convention constantly. If work is done ON the system, it's positive. If the system does work on its surroundings, it's negative. Get that wrong and your energy balance equation gives you nonsense results every time. Second Law: Entropy of an isolated system never decreases. Heat flows from hot to cold, not the reverse, without external work input. The Clausius inequality states that for any cycle, the integral of dQ/T is less than or equal to zero, with equality holding only for reversible processes. This is where most students hit a wall because entropy is not directly measurable — you have to calculate it from property tables or equations of state. I've seen engineers try to skip this step and just assume constant specific heats across wide temperature ranges, which introduces errors of 15 to 30 percent in efficiency predictions for combustion cycles.
Third Law: The entropy of a perfect crystal at absolute zero is exactly zero. This establishes an absolute reference point for entropy calculations, which is why we can look up standard molar entropies in tables and meaningfully compute reaction entropy changes. It also tells us that reaching absolute zero is physically impossible in a finite number of steps, which matters less in engineering practice but becomes critical in cryogenics research where you're already operating at 4 K and trying to get to millikelvin temperatures. Now here's the thing nobody emphasizes enough: these three laws form a hierarchy. The first law tells you what's energetically possible. The second law tells you how much of that energy transfer you can actually harness as useful work. The third law gives you the reference frame. When you're designing something real, you work through all three sequentially, and skipping even one will give you answers that look plausible but are fundamentally wrong.
How I Actually Use This in Practice
I spent years working on heat exchanger design for industrial processes, and the second law was where everything fell apart most often. There's a specific problem that comes up repeatedly: when you're analyzing a real heat exchanger with finite temperature differences, the entropy generation method seems like the cleanest approach on paper. You calculate the entropy change of the hot stream, the entropy change of the cold stream, and the difference is your irreversibility. Simple enough. The edge case I keep running into is when one of the streams undergoes a phase change — say, condensing steam heating a liquid stream. The temperature of the condensing steam stays nearly constant, which makes the logarithmic mean temperature difference calculation straightforward but the entropy calculation trickier. If you approximate the phase change stream as having infinite heat capacity (constant temperature), you'll underpredict the entropy generation by roughly 8 to 12 percent compared to using the actual property tables with temperature-dependent enthalpy and entropy values. That might sound small, but when you're optimizing a network of six or seven heat exchangers, those errors compound and your pinch point analysis shifts by 5 to 8 degrees Celsius. The workaround I use is to split the phase change region into small temperature intervals even though the temperature change is minimal, and integrate the entropy calculation across those intervals using the actual saturation properties from steam tables rather than assuming constant values. It adds maybe ten minutes to a calculation that would otherwise take two, and it saves you from having to redo the whole analysis when the design review team asks why your predicted duty doesn't match the simulation. I also learned the hard way that using average temperature for the entropy integral instead of the log-mean temperature difference in these cases produces errors that get worse the larger the temperature glide is on the other stream.
Get the Full Details

Common Pitfalls and Where the Laws Break Down
The first law is almost never wrong in classical thermodynamics, but people misuse it by treating open systems as if they were closed. If you're analyzing a turbine or a compressor without accounting for the flow work term (the Pv contribution to enthalpy), your energy balance will be off by an amount that depends on the pressure and specific volume — easily hundreds of kilojoules per kilogram for gas systems. Enthalpy exists precisely to fold that flow work into a single property, so use it. The second law is where the real subtlety lives. A common mistake is confusing entropy with energy degradation. Entropy isn't energy — it's a measure of energy dispersal at a given temperature. When you calculate exergy destruction as T times entropy generation, you're converting an entropy problem into an energy problem, and that conversion factor (the dead state temperature) is easy to get wrong if your reference environment isn't clearly defined. I've seen projects use 25°C as the dead state when the actual ambient conditions were 35°C, which shifts every exergy calculation by about 3 percent. Another counter-intuitive point: entropy can decrease locally. In a refrigeration cycle, the entropy of the refrigerant decreases in the condenser as it rejects heat. That's fine — the total entropy of the system plus surroundings still increases. What trips people up is when they see entropy values drop in a component and assume they've made a calculation error. Check your property tables first, then check your assumption about whether the process is adiabatic. If heat is leaving the control volume, entropy can absolutely go down.
The third law is mostly a theoretical convenience in everyday engineering, but it becomes practically important in cryogenic processes and in calculating absolute entropy values for chemical reactions. If you're working with substances near absolute zero — liquid helium systems, for example — the heat capacity drops dramatically and the standard approximation that Cp is constant with temperature fails completely. You need Debye model corrections or tabulated low-temperature data, and if you don't have that data available, your entropy calculations for those systems are essentially guesswork. There are also regimes where classical thermodynamics itself breaks down. Near critical points, the distinction between liquid and vapor phases disappears, and properties become extremely sensitive to small changes in temperature and pressure. The ideal gas law and many of the standard property correlations lose accuracy in this region. I've seen designs fail because someone used ideal gas assumptions for CO in a transcritical cycle operating near its critical point of 31°C and 7.38 MPa. The compressibility factor deviates significantly from 1.0 in that region, and ignoring that deviation introduced errors in the predicted coefficients of performance that were large enough to make the system uneconomical.
A Practical Framework for Applying These Laws
When I'm starting a new thermodynamic analysis, I follow a sequence that keeps the three laws in the right order. First, I define the system boundaries clearly — closed or open, steady or transient, and what crosses those boundaries in terms of mass, heat, and work. Second, I apply the first law to get the energy balance. This usually gives me one equation with multiple unknowns, so I need property relationships. Third, I check whether the second law imposes any constraints — is the proposed process reversible or irreversible, and what's the entropy generation? Fourth, if absolute entropy values matter for my application, I anchor them using the third law reference point. The property data is where most delays happen. If you're working with water or steam, IAPWS-IF97 is the standard formulation and it's reliable across the full industrial range. For refrigerants, the NIST REFPROP database is the go-to, though it requires a license. For air and common gases, the JANAF tables or the older NASA polynomials work fine up to about 2000 K, beyond which you need to be more careful about dissociation effects. None of these sources are free, and none of them are instantaneous to look up when you're in the middle of a calculation. Building a quick-reference sheet with the property values you use most often saves considerable time over a project lifecycle. I also recommend keeping a separate spreadsheet for entropy calculations. The first-law energy balances tend to be straightforward enough to do in your head or on scrap paper, but entropy is where rounding errors accumulate. If you're summing several entropy changes and each one is rounded to three significant figures, your final entropy generation value could be off by 2 to 5 percent depending on the magnitude of the individual terms. Running the numbers through a spreadsheet with consistent precision throughout the calculation chain eliminates that source of error.
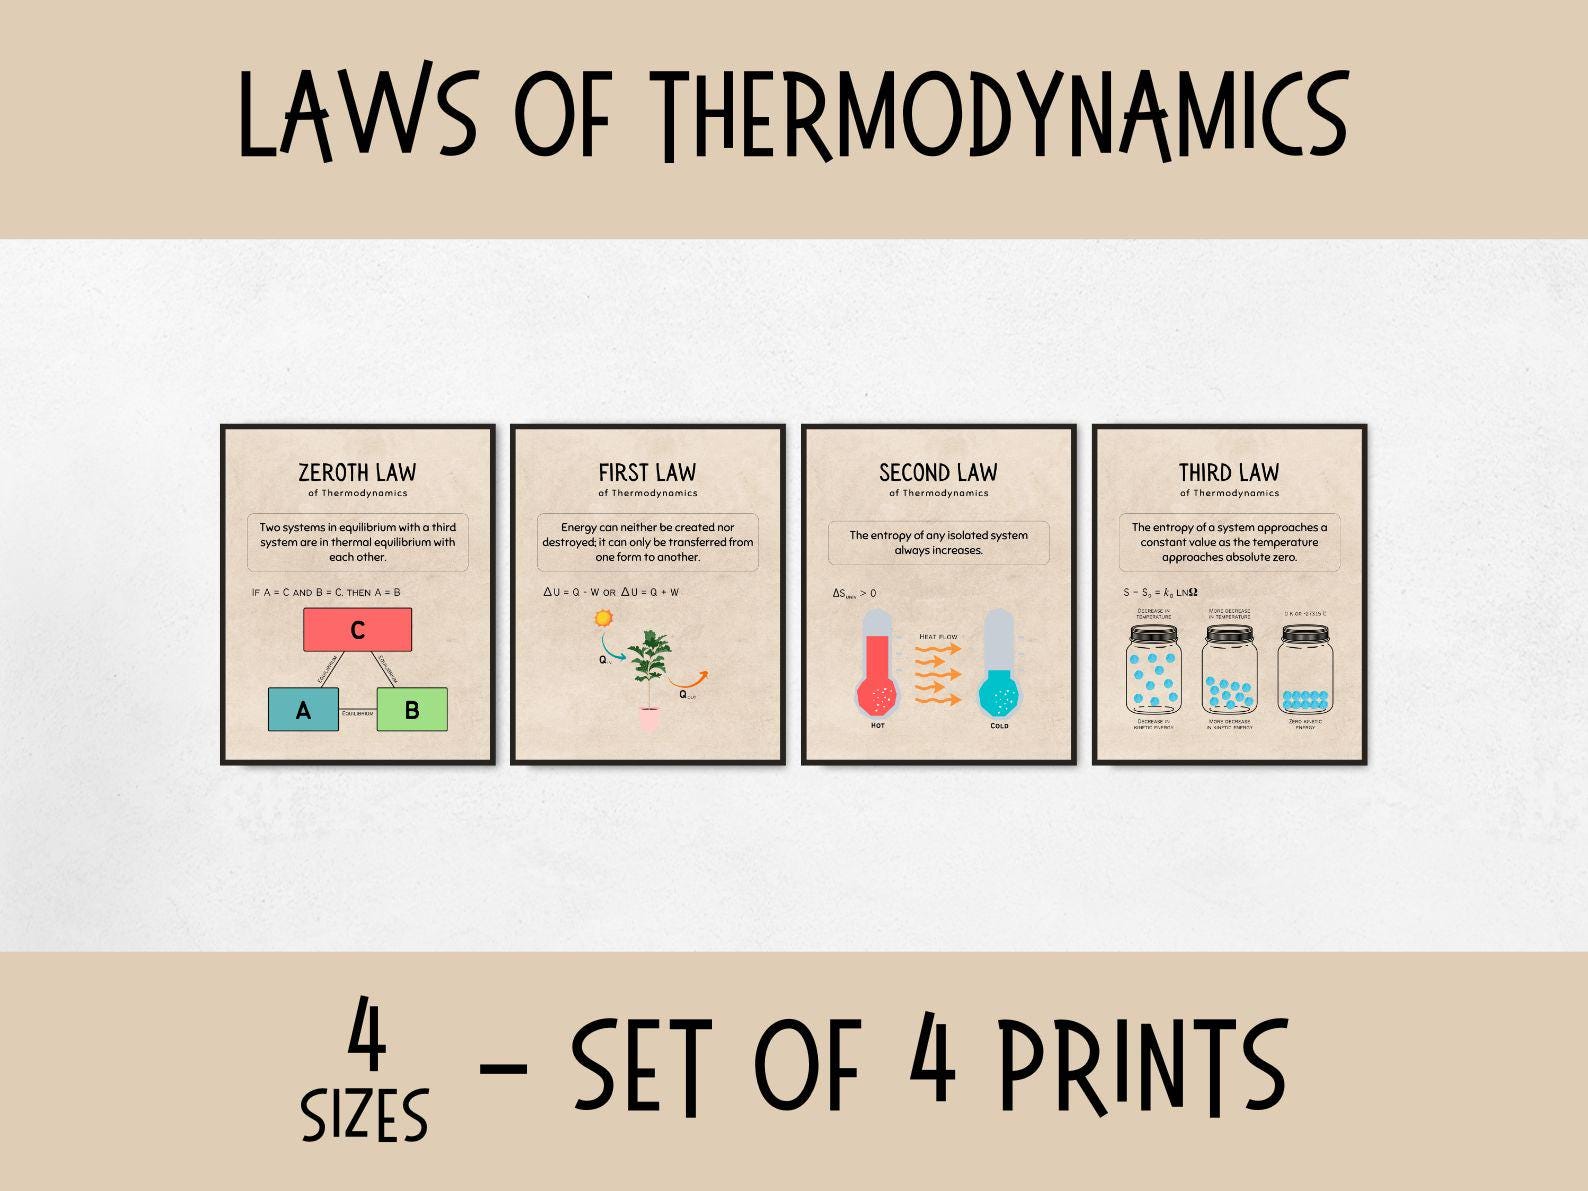
The bottom line is that the 3 Laws Of Thermodynamics aren't just textbook material. They're the operating constraints on everything that involves energy transfer, and getting them wrong in a real design doesn't just give you a bad grade on an exam — it gives you a system that doesn't meet its performance targets or, in worse cases, operates unsafely. The laws themselves don't change, but your ability to apply them correctly depends on paying attention to the details: the system boundaries, the property data quality, the sign conventions, and the assumptions you're making about reversibility.