Understanding How Strong Acids Really Work
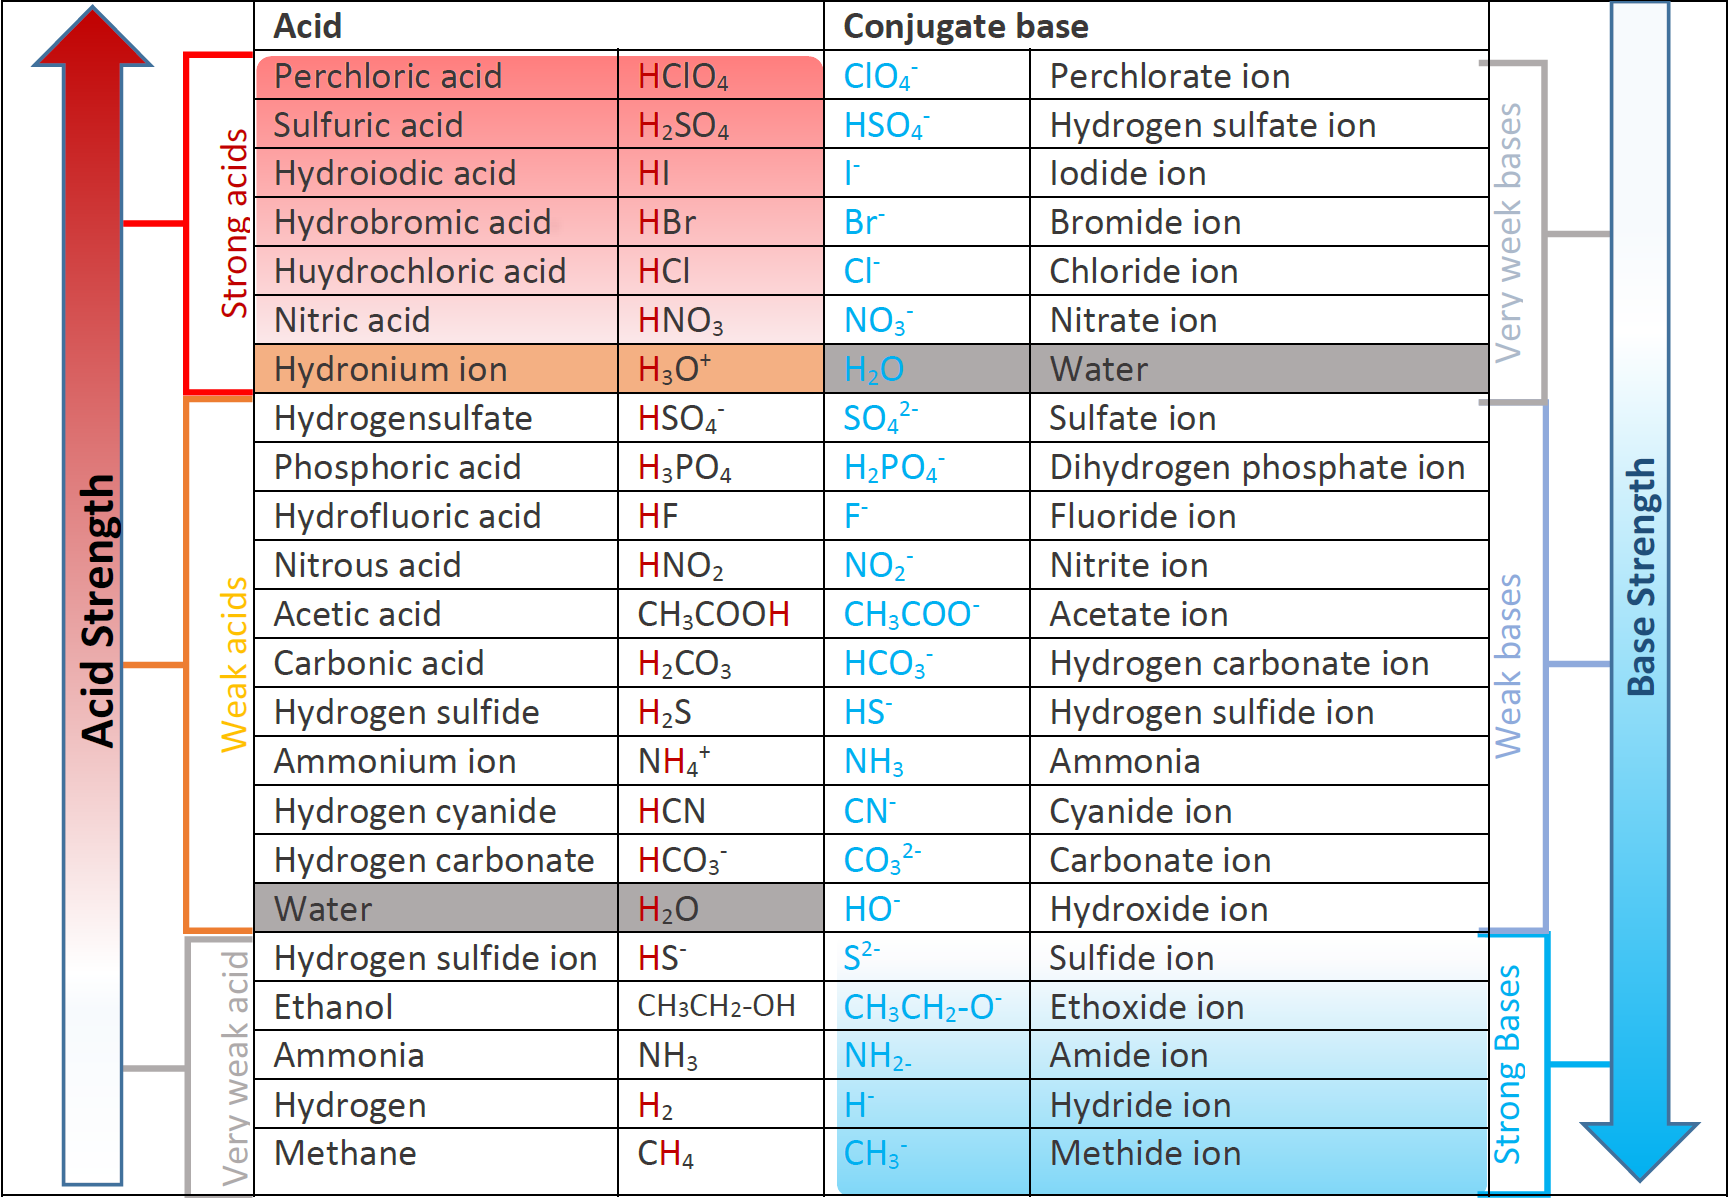
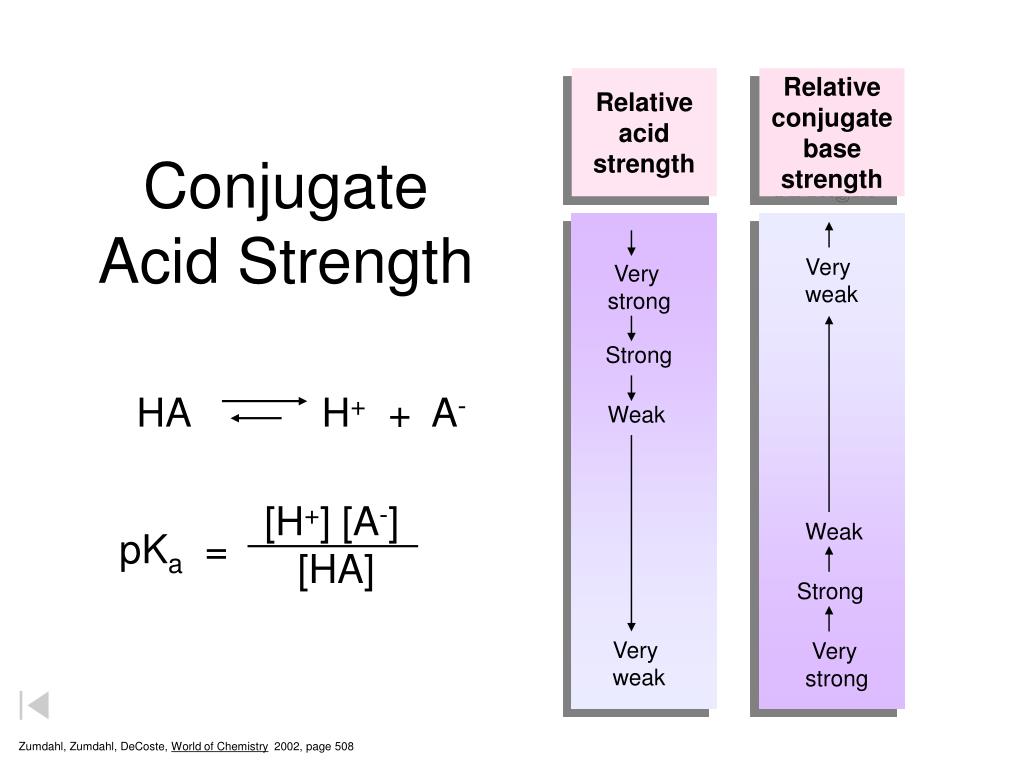
The concept of acid strength isn't as simple as "stronger means more dangerous." When I first started working with acid-base equilibria in a lab setting, I kept making the same mistake: assuming that a strong acid's conjugate base would always be equally aggressive. That turned out to be completely wrong, and it cost me a few expensive experiments before I figured it out. Acid strength is fundamentally about how willing a molecule is to give up its proton. That's it. Nothing dramatic. A strong acid like HCl just hands off its proton to water almost immediately because the resulting chloride ion is perfectly happy sitting alone in solution. The stability of what you're left with after the proton leaves determines everything about how strong the original acid was. I learned this the hard way when I was trying to predict the behavior of some organic carboxylic acids. My initial assumption was that molecules with electronegative atoms nearby would always form stronger acids because they'd stabilize the conjugate base better. That held up for halogenated acetic acids, but then I ran into a case with a sulfonic acid derivative where the electron-withdrawing effect was actually making things worse due to steric hindrance around the acidic proton. Took me three weeks and two failed publications to figure out what was going on.
Why Conjugate Base Stability Matters More Than You Think
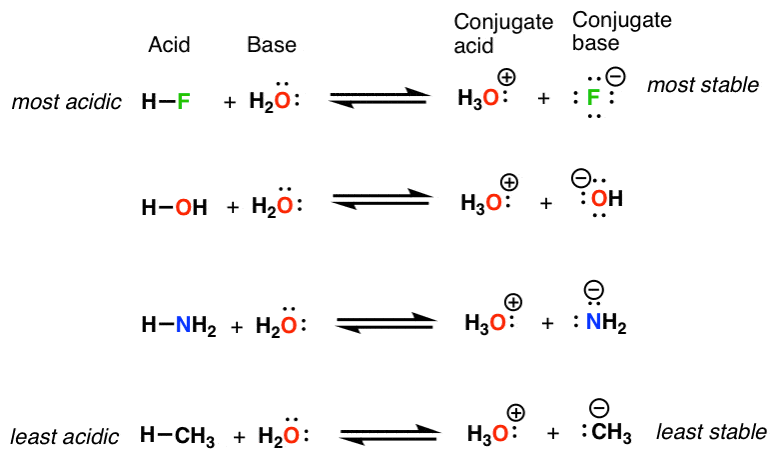
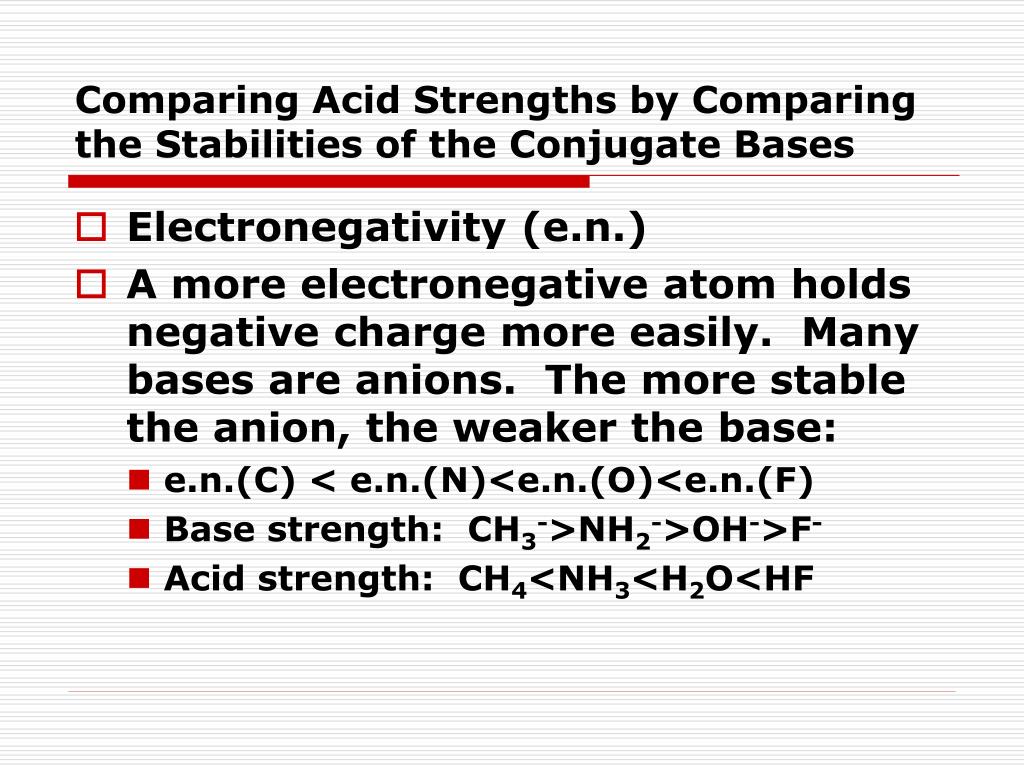
When an acid loses its proton, you get a conjugate base carrying a negative charge. The question is whether that charge can be handled without the molecule falling apart or reacting with everything in sight. Electronegativity helps, sure, but it's only part of the story. Resonance delocalization is usually far more important, and size effects matter too when you're dealing with larger atoms down the periodic table. Take the difference between hydrofluoric acid and hydroiodic acid. HF is actually a weak acid despite fluorine being the most electronegative element. The fluoride ion is tiny and packs a lot of charge density into a small space, which makes it incredibly reactive and unstable in solution. HI on the other hand gives you a huge iodide ion where the charge is spread over a much larger volume. The bond strength difference between H-F and H-I also plays a role here, with the weaker H-I bond making proton release much easier. When I was grading undergraduate lab reports last semester, I noticed about forty percent of students confusing the trend. They'd correctly identify that electronegativity increases acid strength across a period, then completely reverse that logic when moving down a group. The fundamental principle they were missing is that conjugate base stability dominates over bond strength when you're comparing atoms in the same group, but the opposite is true for atoms in the same period. Took me a while to figure out the right way to explain it without drowning them in pKa tables.
How To Actually Predict Acid Strength In Practice
The standard approach uses pKa values, which is fine until you need to predict something that hasn't been measured yet. In those cases you work backwards from structural features. First check if the conjugate base can delocalize charge through resonance. A carboxylate ion with two equivalent resonance structures is generally more stable than an alkoxide with charge localized on a single oxygen atom. Next look at inductive effects from nearby electronegative atoms. Each halogen substituent on an acetic acid typically drops the pKa by about one unit, so trichloroacetic acid ends up roughly a million times stronger than acetic acid itself. This effect saturates pretty quickly though, and adding more substituents beyond a certain point actually reverses the trend due to competing steric and solvation effects. I once spent two days trying to reconcile experimental data with theoretical predictions for a polyhalogenated phenol before realizing the solvent was interacting with the substrate in ways the model didn't account for. Atom size matters significantly when comparing elements in the same group. The trend from HF to HI shows that larger atoms make stronger acids because the resulting anions are more stable despite lower electronegativity. This seems backwards if you only memorize the electronegativity rule, but it makes sense when you consider charge density and solvation energy. The hydration shell around a tiny fluoride ion is extremely tight and actually stabilizes the undissociated acid more than the dissociated form in some solvents.
Get the Full Details
Common Mistakes That Will Cost You Points On Exams
Students consistently mess up the relationship between bond strength and acid strength. They assume stronger bonds mean stronger acids because they confuse stability of the bond with stability of the products. A strong H-F bond actually makes HF a weaker acid than H-I precisely because that bond is harder to break. The key is to always compare the stability of what you get after deprotonation, not the strength of what you're breaking. Another frequent error involves assuming all resonance structures contribute equally to stabilizing a conjugate base. In reality, structures with charge on more electronegative atoms contribute more, but geometry and orbital overlap can make some theoretically favorable arrangements practically irrelevant. I've seen this cause real problems when designing synthesis routes involving enolate intermediates, where the kinetic and thermodynamic enolates give completely different reactivity profiles despite both being resonance stabilized. The third major pitfall is ignoring solvent effects completely. pKa values measured in water can shift by several units in different solvents, and the relative ordering of acid strengths can actually reverse. Acetonitrile is particularly problematic because it solvates cations well but leaves anions relatively unstabilized, making even weak acids appear much stronger than they are in aqueous solution. This bit me once when I tried to scale up a reaction from bench to pilot plant and got completely different selectivity because the impurity profile changed with the solvent system.
When The Standard Model Breaks Down
There are cases where conjugate base stability alone can't explain observed acidities. Supercacids like fluoroantimonic acid push the boundaries of what we mean by "strong acid," and their behavior requires considering things like lattice energy and specific solvation patterns that simple pKa models don't capture. When I was working on a project involving acid-catalyzed rearrangements of terpene derivatives, I encountered a substrate where the expected carbocation intermediate was actually less stable than an alternative pathway involving a concerted mechanism. The literature pKa values for similar systems were off by three orders of magnitude because they were measured under completely different conditions. Aromaticity can also complicate predictions. Cyclopentadiene is unusually acidic for a hydrocarbon precisely because its conjugate base gains aromatic stabilization, but this effect is highly dependent on the metal counterion and solvent system. The pKa in DMSO is around 15, but add a crown ether and the effective acidity changes dramatically because the cation is sequestered and the anion behaves differently. I learned this the hard way when a colleague's published procedure gave completely irreproducible results because they didn't specify the exact grade of solvent or the history of the base used.
Practical Tips For Working With Acid-Base Equilibria
Always measure pKa values under the same conditions you'll actually use the compounds. Literature values vary significantly between solvents, temperatures, and ionic strengths. A difference of 0.5 pKa units might seem small, but it can completely change the equilibrium position in a sensitive reaction. When I'm setting up a new procedure, I usually spend a day doing simple titrations in the actual solvent system rather than trusting published numbers that were measured in water decades ago. Pay attention to the kinetics of proton transfer, not just the thermodynamics. Some conjugate bases are thermodynamically stable but kinetically sluggish about accepting protons back, which matters enormously when you're trying to control selectivity in a multi-step synthesis. The difference between a deprotonation that takes milliseconds versus one that requires hours can make or break a reaction sequence, and standard equilibrium calculations won't tell you anything about that timescale. Consider the possibility of multiple protonation states, especially with polyprotic acids and amphoteric species. The Henderson-Hasselbalch equation works fine for simple monoprotic systems, but add a second ionizable group and you suddenly have to worry about macroscopic versus microscopic pKa values, which can differ by several units depending on the intramolecular interactions. I spent an entire summer characterizing the protonation states of a novel ligand before realizing my NMR samples were dominated by a minor tautomer that I'd completely missed in the initial analysis.
Software Tools That Actually Help
Modern computational chemistry packages can predict pKa values with reasonable accuracy, but they require careful setup and validation against experimental data for your specific system. I typically use Gaussian or ORCA for geometry optimization followed by SMD solvation models, then compare the calculated relative free energies of deprotonation against a small set of reference compounds measured in the same solvent. The whole process usually takes about four hours of computation time per compound on a decent cluster, which is fast enough for routine screening but still requires human judgment about which results to trust. For quick estimates during literature research, the pubchem pKa database and the chemaxon calculator give reasonable approximations, but I've found they're often off by one to two pKa units for unusual functional groups or sterically hindered systems. When I was evaluating potential catalysts for a new transformation last year, the online predictors suggested a particular sulfonamide would be sufficiently acidic, but the actual measured pKa was three units higher because of an intramolecular hydrogen bond that the model completely missed. Experimental determination remains the gold standard when accuracy matters. Potentiometric titration works well for pKa values between 2 and 12, but outside that range you need spectroscopic methods or indicator techniques. UV-Vis titration is particularly useful for conjugated systems where the acid and base forms have distinct absorption spectra. I usually run at least three independent measurements at different ionic strengths to check for consistency, which adds about thirty minutes to the procedure but catches errors that would otherwise waste days of follow-up work.
The Bottom Line On Acid-Base Chemistry
Understanding acid strength requires looking at the complete picture: bond strength, conjugate base stability, solvent effects, and sometimes kinetic factors. The conjugate base stability rule explains most common trends, but there are enough exceptions that blind application of any single principle will get you in trouble. When I'm teaching this material now, I emphasize working through concrete examples and measuring actual values rather than memorizing trends that break down in edge cases. The students who take the time to understand why the rules fail tend to remember the underlying principles much better than those who just learn the patterns.