Getting the Aldol Reaction to Actually Work
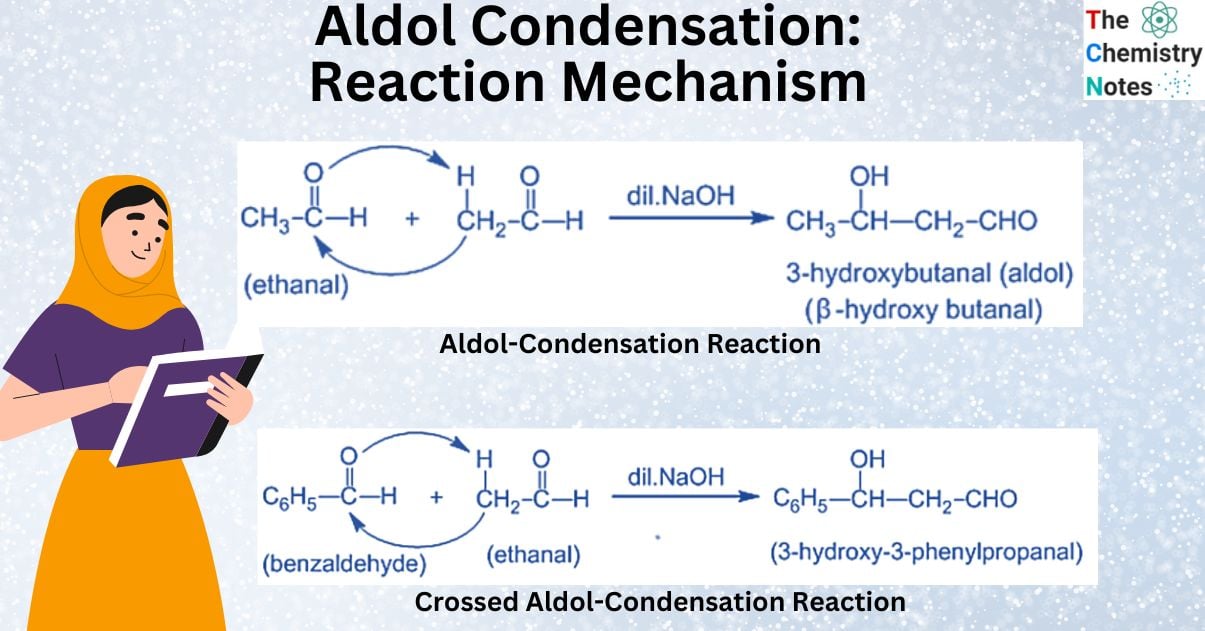
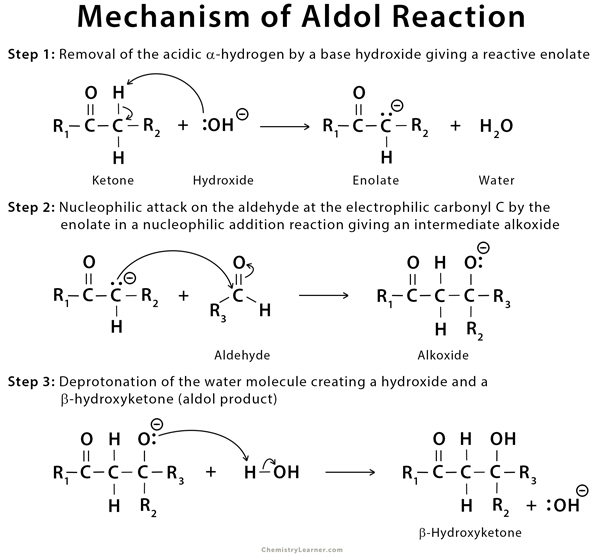
The aldol condensation is one of those reactions that looks simple on paper but gets messy fast in practice. You combine two carbonyl compounds, usually with a base or acid catalyst, and you get a beta-hydroxy carbonyl product that can lose water to form an alpha,beta-unsaturated carbonyl. That is the textbook version. The reality involves enolate equilibration, self-condensation side reactions, and E/Z selectivity that nobody warns you about until your TLC plate looks like a crime scene. Here is how I break it down when teaching this or running it myself. You start with a carbonyl compound that has at least one alpha proton. A base removes that proton to form an enolate. The enolate then attacks the electrophilic carbon of another carbonyl molecule. This gives you an alkoxide intermediate, which grabs a proton from solvent or water to yield the aldol addition product. If you apply heat or continue basic conditions, the hydroxyl group and a neighboring hydrogen are eliminated to give the conjugated enone. That dehydration step is what separates an aldol addition from an aldol condensation. The mechanism changes slightly depending on whether you are running base-catalyzed or acid-catalyzed conditions. Under base, the enolate forms first and then attacks. Under acid, you generate an enol instead, which is less nucleophilic but still reactive enough to attack a protonated carbonyl. Both paths lead to the same class of products, but the selectivity profiles are different. Base conditions tend to give cleaner dehydration because the resulting conjugated system stabilizes the product. Acid conditions can be trickier since the protonated intermediate is more prone to polymerization if you are not careful with temperature.
I once ran a crossed aldol between cyclohexanone and benzaldehyde using sodium hydroxide in ethanol. On paper this should give a single product. Instead I got a mixture of self-condensation products from cyclohexanone, the desired crossed product, and a significant amount of unreacted starting material. The problem was that cyclohexanone can both form the enolate and act as the electrophile, so it outcompetes benzaldehyde for the enolate attack. Benzaldehyde has no alpha protons so it cannot self-condense, but it also does not easily form an enolate to attack itself. The solution was switching to a directed aldol approach using LDA at low temperature to form the cyclohexanone enolate quantitatively before adding the benzaldehyde. That eliminated the self-condensation pathway and gave me the crossed product in about 82% yield after workup. It took me three separate failed runs to figure that out, so I do not take simple aldol conditions for granted anymore. The key practical insight most people miss is that not all aldol condensations proceed through the same rate-determining step. In dilute base with simple aldehydes, the C-C bond formation step is usually rate-limiting. But with ketones, the dehydration step can become slower than the addition step, which means you can sometimes isolate the beta-hydroxy ketone if you keep the temperature low and avoid prolonged heating. I have seen students assume that every aldol reaction automatically goes to the condensation product, but that is only true when the product is sufficiently stabilized by conjugation or when you deliberately drive the elimination with heat. Another thing nobody emphasizes enough is the reversibility of the aldol addition step. The initial C-C bond formation is actually an equilibrium process. The condensation product pulls the equilibrium forward by precipitating out or by the irreversible loss of water, but if you run the reaction in a solvent that promotes retro-aldol conditions, you can undo months of synthetic work in a single afternoon. I learned this the hard way when I accidentally ran an aldol in aqueous dioxane at elevated temperature and watched my yield disappear back into starting materials over several hours. Switching to a non-protic solvent like THF and running the dehydration separately with a mild dehydrating agent fixed the problem entirely.
When discussing the mechanism in detail, the stereochemistry deserves attention. The Zimmerman-Traxler model describes the six-membered cyclic transition state that forms during the enolate addition step. This model predicts whether you get syn or anti diastereomers based on whether the enolate is Z or E and whether you are running a chair-like transition state. Most undergraduate courses skip this part because it complicates the basic mechanism, but if you are actually trying to control stereoselectivity in your product, the Zimmerman-Traxler framework is what you need to understand. It explains why certain base and solvent combinations favor one diastereomer over another, and it is not optional knowledge if you plan to use aldol chemistry for anything beyond simple textbook examples. The limitations of this reaction are real and worth stating plainly. Crossed aldol reactions between two different carbonyls with alpha protons are essentially uncontrollable without a directed strategy. You will get a mixture of four possible products including both self-condensation and crossed variants. The reaction also struggles with steric hindrance around the carbonyl carbon, meaning bulky ketones react very slowly or not at all under standard conditions. Electron-deficient aldehydes work well as electrophiles, but electron-rich ones can be sluggish. And the dehydration step requires that you have a hydrogen on the carbon alpha to the newly formed hydroxyl group. If that position is fully substituted, you will get the aldol addition product and nothing else, regardless of how much heat you apply. For cases where the standard aldol condensation with mechanism approaches fails, alternatives like the Mukaiyama aldol reaction using silyl enol ethers and a Lewis acid catalyst give you far better control over crossed selectivity and stereochemistry. It costs more in reagents and adds a preparation step, but it avoids the equilibrium problems and mixture issues that plague traditional base-catalyzed methods. I use the standard aldol when the substrate is simple and cost matters. I switch to the Mukaiyama version when the molecule has sensitive functional groups or when I need specific stereocontrol that the classical method cannot provide.
Get the Full Details