Working with Alpha Carbon Amino Acid in Peptide Synthesis: What Actually Happens
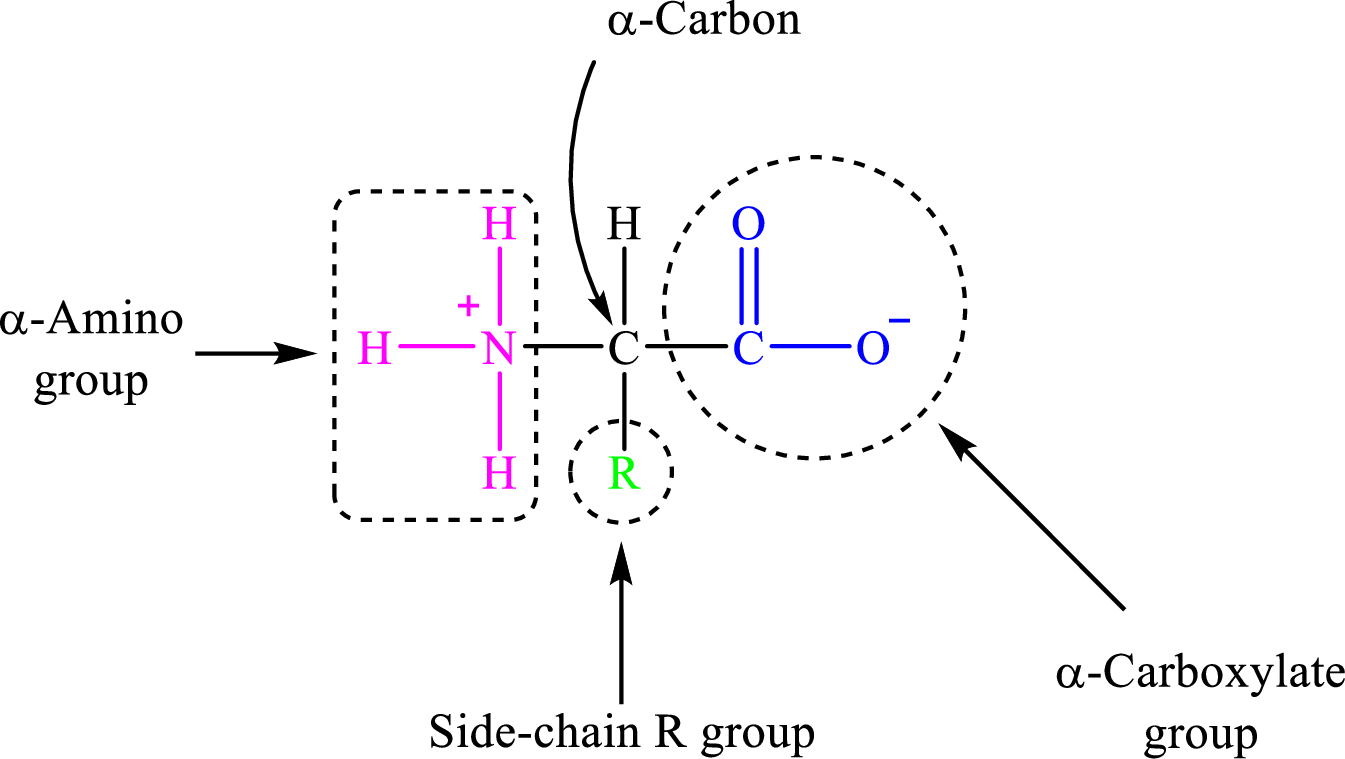
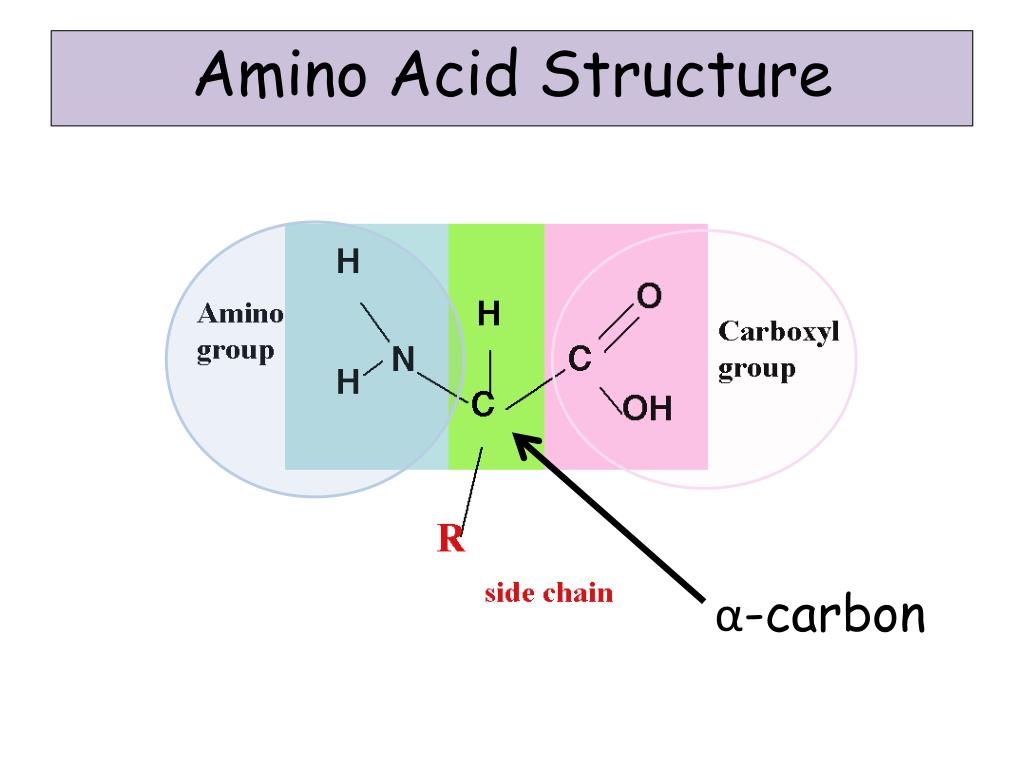
When you're running Fmoc-based solid-phase peptide synthesis, the stereochemistry around the alpha carbon is where everything can quietly go wrong. It's not dramatic until it's too late. You've got your resin loaded, your coupling reagents mixed, and then HPLC shows a second peak you didn't expect. That's usually epimerization at the alpha carbon, and it tends to sneak in during activation steps, especially with sterically hindered residues like phenylalanine or leucine when the coupling time drags on past twenty minutes without proper monitoring. The basics are simple enough. An Alpha Carbon Amino Acid has the amine group and the carboxylic acid attached to the same central carbon. That central carbon is also bonded to a hydrogen and a variable side chain. This is what makes the 20 standard proteinogenic amino acids what they are. But the structural simplicity hides a lot of practical headaches, particularly when you're working with protected intermediates or trying to couple difficult sequences.
Common Pitfalls with the Alpha Carbon in Coupling Reactions
Here's the thing most protocol templates don't emphasize: the rate of racemization depends heavily on the base you choose and the activation method. Using DIC alone with Oxyma gives you relatively clean coupling for most standard residues, but switch to HBTU with DIEA and you're looking at measurable epimerization on second positions in sensitive sequences. I ran into this with a twelve-residue therapeutic peptide where the third position was phenylalanine. After switching from HBTU/DIEA to DIC/Oxyma and cutting the activation time to five minutes before dispensing onto the resin, the impurity dropped from eight percent to under one percent on the final cleavage cocktail. The difference wasn't in the amino acid itself. It was in how long the activated species sat before hitting the nucleophile. Another detail people gloss over is the effect of the N-terminal protecting group. Fmoc is generally gentler on the alpha carbon than Boc, but Fmoc removal with piperidine creates a highly nucleophilic free amine that can sit around and degrade if you're not moving fast. I've seen batches where the resin swelled improperly because the DMF wasn't anhydrous, and the alpha carbon ended up with hydrolyzed side products. Running Karl Fischer titration on your solvents before you start a long synthesis is probably the single most overlooked quality check. If water content is above five hundred ppm, you're rolling the dice.
Analysis and Quantification
Once your peptide is cleaved, you need to verify that the alpha stereochemistry held up. Acid hydrolysis followed by chiral HPLC is the standard approach. You'll typically use a column like the Chiralpak AD-H or a similar cellulose-based stationary phase with a mobile phase of hexane and isopropanol containing a small amount of diethylamine. The elution order matters here. D and L isomers come off at different retention times, and for some amino acids like cysteine the separation is tight enough that you need to be precise with flow rate and temperature. I keep my column at thirty degrees Celsius rather than room temperature because it stabilizes the retention times and makes integration more reproducible. If you're working with modified amino acids or non-natural analogs where the side chain is bulky or charged, standard hydrolysis conditions might not give clean results. In those cases, derivatization with OPA or FMOC-Cl before chiral analysis can sharpen the peaks significantly. The derivatization step adds about ten minutes to each sample but improves detection limits by roughly an order of magnitude. That's useful when your epimer content is below one percent and you need to confirm whether it's actually there or just noise in the detector baseline.
Get the Full Details

When Alpha Carbon Chemistry Fails Completely
Sometimes the substrate itself is the problem. Amino acids with electron-withdrawing groups on the side chain, like DOPA or phosphoserine, are much more prone to racemization under standard coupling conditions. The alpha proton becomes more acidic, and even weak bases can abstract it during activation. If you're working with these residues, consider using pre-activated esters like Pfp esters that bypass the carbodiimide activation step entirely. They're more expensive per gram, but the yield improvement on difficult couplings usually justifies the cost. I once spent three days troubleshooting a coupling that turned out to be a straightforward racemization issue with a phosphorylated serine. Switching to the pre-formed Pfp ester cut the synthesis from seven failed cycles to one clean run. For absolute configuration assignment when chiral HPLC isn't available, capillary electrophoresis with cyclodextrin additives can separate D and L enantiomers. It's slower than HPLC but requires less specialized column inventory. The tradeoff is that method development takes longer, and you'll need to validate it against a known standard before trusting any quantitative results.