Working With Alpha Carbon Chemistry Enols And Enolates In The Lab
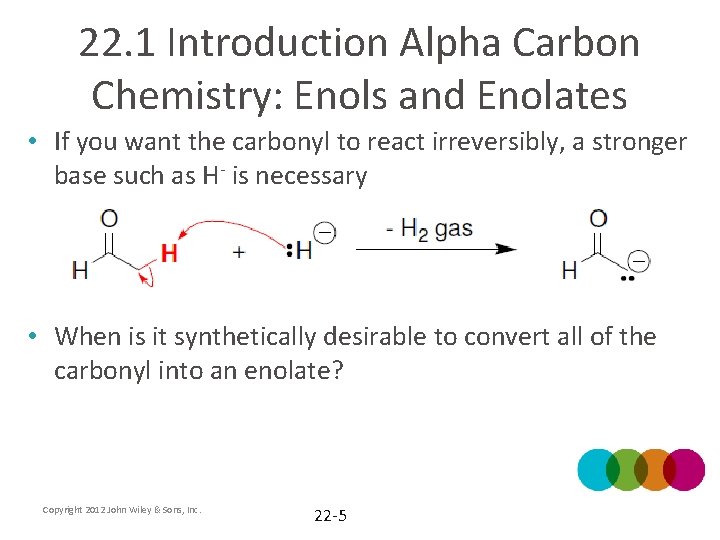
The alpha proton is acidic because the resulting carbanion is stabilized by resonance with the carbonyl. That is the basic mechanism. The practical problem is that controlling which alpha carbon gets deprotonated, and whether you get kinetic or thermodynamic product, takes real attention to detail. Most people mess this up on their first try. An enol forms when the alpha hydrogen moves to the oxygen and a double bond creates between the alpha and carbonyl carbons. An enolate is the deprotonated version, carrying a negative charge that delocalizes onto the oxygen. The oxygen end is more electronegative but the carbon end is usually the nucleophilic site you want for C-C bond formation. This distinction matters more than textbooks make it sound. Here is how I actually set up an enolate alkylation. Take a ketone, dissolve it in dry THF, cool to -78°C under nitrogen, then add LDA dropwise. Wait twenty minutes for complete deprotonation before adding the alkyl halide. If you skip the cold temperature step, you will get a messy mixture of self-condensation products and polymers. I learned this the hard way with a substrate that had two different alpha positions.
The real complexity comes with unsymmetrical ketones. Consider 2-methylcyclohexanone. It has two distinct alpha carbons, and each gives a different enolate depending on conditions. Using LDA at low temperature with slow addition gives the kinetic enolate, deprotonating the less hindered alpha position. Switching to a weaker base like sodium methoxide in methanol at room temperature gives the thermodynamic enolate, favoring the more substituted double bond. This is not optional. Both products will form simultaneously unless you control the conditions carefully. I ran into a specific issue last year with a bicyclic ketone that had a bridgehead alpha position. The expected kinetic enolate was not forming at all because the geometry prevented proper orbital overlap for deprotonation at that site. Instead of getting the clean single product I was after, I got a mixture. The workaround was switching to potassium bis(trimethylsilyl)amide in hexane, which is bulkier and more selective, and running the reaction at -100°C using a dry ice/acetone bath with careful temperature monitoring. The selectivity improved dramatically, though the reaction time roughly tripled.
Choosing Your Base Matters More Than You Think
LDA is the standard kinetic base but it is not the only option. Sodium hydride is cheaper and easier to handle, though it tends to give more thermodynamic mixtures unless you are working at elevated temperatures with long reaction times. Potassium t-butoxide in DMSO can push things toward kinetic control in some cases because DMSO does not solvate the cation well, leaving a more reactive free anion. This is the so-called "superbase" condition that some papers reference, and it works but it is also harsh on sensitive functional groups. If your molecule has an ester alongside a ketone, and you want to selectively deprotonate the ketone alpha position without touching the ester, choose your base carefully. LDA will preferentially deprotonate the ketone because the resulting enolate is more stabilized. But if you use something like NaH, the ester alpha protons may start competing, especially if you let the reaction warm up. I have seen this cause problems in multistep syntheses where the next step involves an electrophile that reacts with both enolates. Another thing people miss is that the solvent choice changes reactivity. THF is standard for LDA reactions. Diethyl ether works too but gives slower deprotonation. DMF and DMSO increase the reactivity of the enolate significantly but also make quenching and workup messier. If you are running a large scale, the extra solvent removal cost in rotovap time adds up quickly.
Get the Full Details

Trapping and Alkylation Practical Notes
Once your enolate is formed, you need to trap it with an electrophile. Alkyl halides are the most common choice. Methyl iodide and ethyl iodide work well for primary alkylations. For secondary halides, expect elimination competition. The enolate acts as a base as much as a nucleophile in those cases, and you will see the alkene byproduct along with your desired alkylation product. This is why primary halides and tosylates are preferred when possible. Silyl enol ethers are an alternative trapping method. You form them using a mild base like triethylamine with TMSCl, then isolate or use them directly. The advantage is you can store them and use them later with a Lewis acid promoter. This is useful when your enolate is unstable or your electrophile is sensitive to strong base. The downside is an extra synthetic step and lower overall yield due to the isolation. Aldol reactions are where enolates really show their value. You can do direct aldol with preformed enolates, or use boron enolates for better stereocontrol. The Mukaiyama aldol, which uses a silyl enol ether and a Lewis acid, is more forgiving than direct aldol because you avoid the strong base entirely. This is the method I reach for when my substrate has sensitive functional groups that would decompose under typical enolate conditions.
There are also cases where enolates undergo conjugate addition instead of direct alkylation. If your electrophile is an alpha-beta unsaturated carbonyl, the enolate can do 1,2-addition or 1,4-addition depending on conditions. Hard nucleophiles like simple enolates tend toward 1,2. Softer conditions or copper catalysis can shift things to 1,4. I once ran a reaction expecting conjugate addition and got mostly direct addition because I used a lithium enolate at low temperature instead of a cuprate. The product analysis took a day to figure out.
When Enolates Fail You
Enolate chemistry is not a universal solution. If your alpha carbon is sterically shielded, deprotonation can be extremely slow or require forcing conditions that degrade your substrate. If your molecule contains electrophilic sites elsewhere besides your intended reaction center, the enolate will attack those too. Epoxides, acyl chlorides, and even certain protecting groups can get hit. You need to map all the reactive sites before you start. Another limitation is that enolates are strongly basic. If your substrate has any acidic protons other than the one you want to remove, they will be deprotonated first. This includes terminal alkynes, phenols, and even some amide N-H bonds. You may end up with a poly-anionic species that behaves unpredictably. In those cases, protecting groups or alternative methodologies like organocatalysis might be cleaner. The E1cB elimination pathway is another failure mode. If your product has a good leaving group beta to the carbonyl, the enolate can eliminate rather than couple. This is particularly common with beta-halo ketones and substrates that have been alkylated multiple times at the alpha position. Running the reaction colder and shorter helps but does not always prevent it. You may need to redesign the synthesis route entirely.

Workup is another area where things go wrong. Quenching an enolate reaction with water or dilute acid can sometimes reverse the enolization or cause product decomposition, especially for sensitive beta-dicarbonyl compounds. Saturated ammonium chloride is usually safer than strong acid. Ice water works in a pinch but introduces moisture that can hydrolyze sensitive intermediates. I tend to quench with a minimal amount of saturated NH4Cl at low temperature, then extract quickly.
Monitoring and Analysis
You should monitor enolate formation whenever possible. TLC can show disappearance of starting material but it cannot tell you whether you have enol, enolate, or starting ketone. NMR of the quenched reaction mixture gives you more information. Look for the vinyl proton signal around 5.5 to 6.5 ppm for enols and enolates. If you see the carbonyl peak but no vinyl proton, you probably have unreacted ketone. If you see multiple vinyl signals, you have a mixture of regioisomers. GC-MS and HPLC are useful for quantifying the ratio of kinetic to thermodynamic products. This is important when you are optimizing conditions and need to know whether you are getting 80:20 or 50:50 selectivity. Running a few small test reactions before committing to a full scale saves a lot of time and material. Yield expectations with enolate alkylations typically range from 40 to 80 percent for clean reactions. Lower yields usually mean side reactions from competing deprotonation sites or elimination. If you are getting below 40 percent, revisit your base choice, temperature, and solvent. The most common fix is lowering the temperature further and using a stronger, more selective base.
Handling and storage of enolates deserves mention. Most enolates are not isolable and must be generated and consumed in situ. Silyl enol ethers can be stored for weeks in a freezer if kept dry. Lithium enolates in THF at -78°C are stable for hours but will decompose if warmed. Do not leave them sitting in the fume hood overnight expecting them to survive. I have had reactions fail because the cooling bath evaporated before the next step was ready. The bottom line is that alpha carbon chemistry with enols and enolates is straightforward in principle but requires careful execution in practice. Pick your base, control your temperature, know your substrate, and expect to optimize. Most of the problems I have encountered trace back to one of those three factors being off by even a small margin.