What actually happens when you mix acid and base in water
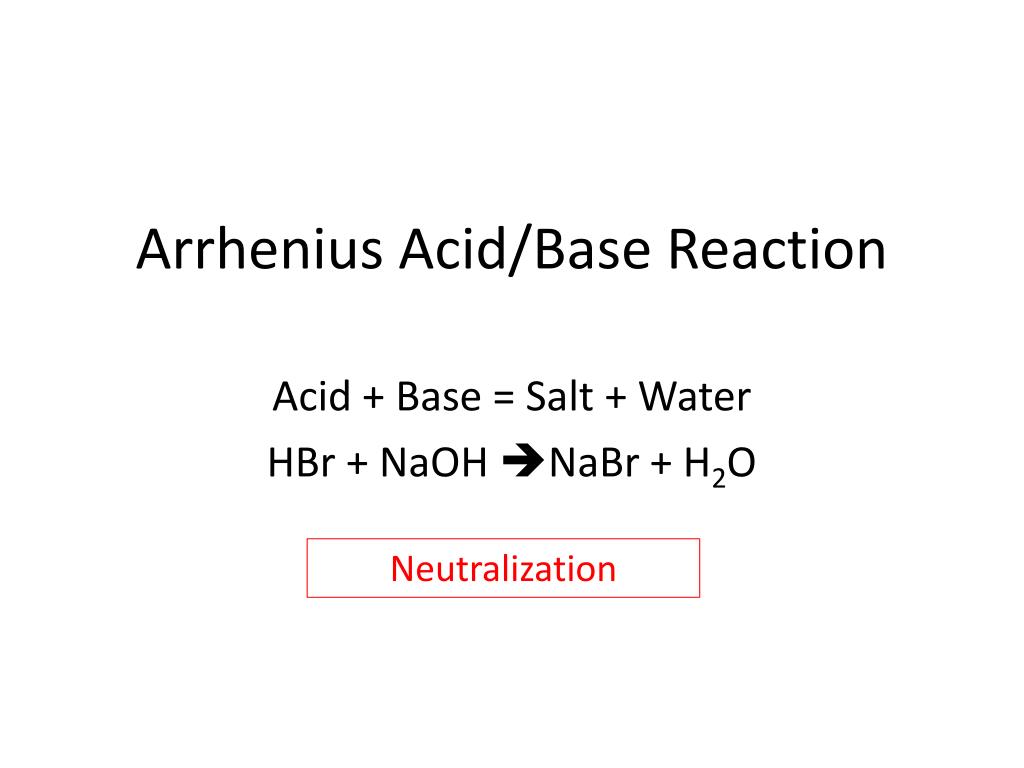
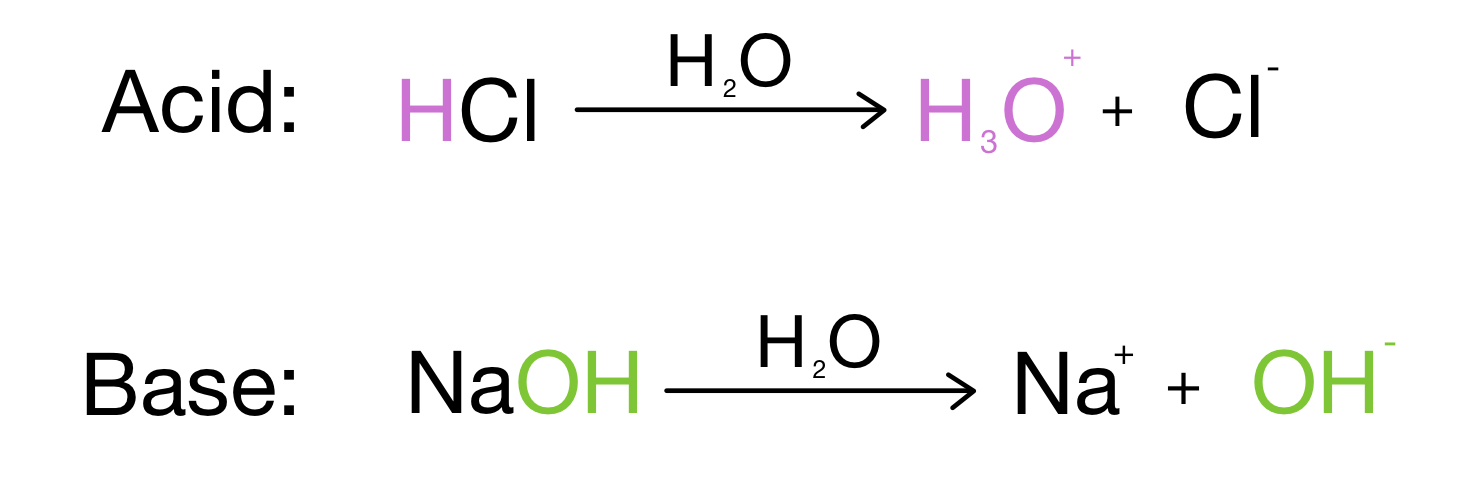
The Arrhenius Acid Base Reaction is straightforward when you strip away the textbook language. An Arrhenius acid is any substance that increases the concentration of H ions when dissolved in water. An Arrhenius base is any substance that increases the concentration of OH ions in water. When you combine them, the H and OH meet and form water. The remaining ions stay in solution as a salt. That is the entire reaction, nothing more. I still see people get tripped up on the simplest part: the water itself. In a typical lab setting, when you titrate 0.1 M HCl against 0.1 M NaOH, the equivalence point lands at pH 7 because the salt (NaCl) is neutral. But that changes immediately if your acid or base is weak. A weak acid like acetic acid with a strong base like NaOH gives an equivalence point around pH 8.7, not 7. The acetate ion that remains hydrolyzes water, pulling H out of solution and leaving excess OH behind. I burned a batch of titration data once because I assumed a neutral equivalence point for a vinegar sample and didn't recalibrate the pH electrode. Took me two hours to re-run the trials. Now I always check whether the acid-base pair is strong-strong, strong-weak, or weak-strong before I assign an expected endpoint pH.
The Arrhenius Acid Base Reaction in practice
Here is the step-by-step process I use when I need to set this up without overcomplicating it. Step 1: Identify your acid and base using the Arrhenius definitions. Does the substance produce H in aqueous solution? That is your acid. Does it produce OH? That is your base. Keep it simple. HCl, HNO, and HSO are strong Arrhenius acids. NaOH, KOH, and Ca(OH) are strong Arrhenius bases. Anything else is weak and will complicate your calculations. This distinction matters more than students are usually told. Step 2: Write the balanced molecular equation. Take the classic example: HCl(aq) + NaOH(aq) NaCl(aq) + HO(l). The net ionic equation strips away the spectator ions and becomes H(aq) + OH(aq) HO(l). This net ionic equation is essentially the same for every strong acid-strong base Arrhenius reaction. That is why the enthalpy of neutralization stays remarkably consistent at about 57.3 kJ/mol across different strong acid-strong base pairs. The spectators don't participate.
Step 3: Account for polyprotic acids if needed. HSO donates two H ions. The first dissociation is strong, the second is weak. If you are doing a full neutralization, you need two moles of NaOH per mole of HSO. Many students miss that and calculate the stoichiometry wrong. I once prepared a standardization solution with the wrong molar ratio because I treated sulfuric acid as monoprotic. My titration curve looked fine but the calculated concentration was off by roughly half. Never skip the proton count. Step 4: Calculate using concentration and volume. The basic equation is MV = MV for strong acid-strong base pairs at equivalence. Adjust for stoichiometry if the mole ratio isn't 1:1. For weak acid or weak base systems, you need the Ka or Kb values and the Henderson-Hasselbalch equation to predict pH at any point along the titration curve, not just the endpoint. Step 5: Verify with a pH measurement. Calculations are theoretical. A calibrated pH meter tells you what is actually happening. If your measured pH at equivalence deviates from your prediction by more than 0.3 units, something is wrong: either your concentrations are off, your acid-base pair isn't what you assumed, or the temperature is affecting the dissociation constants significantly. Temperature matters more than people realize. The Kw of water changes from 1.14 × 10¹ at 0°C to 5.13 × 10¹³ at 100°C, which shifts neutral pH from 7.47 to 6.14. If you are working outside standard room temperature conditions, you need to correct your expectations.
Get the Full Details

The Arrhenius model has real limitations that you should understand before relying on it. It only applies to aqueous solutions. Ammonia gas reacting with hydrogen chloride gas in a dry flask is an acid-base reaction, but neither qualifies as an Arrhenius acid or base because there is no water involved. You have to switch to Brønsted-Lowry definitions for those cases. Similarly, the Arrhenius model cannot explain why substances like NaHCO or NHCl show acidic or basic behavior even though they don't contain OH groups in their formulas. These are salts that undergo hydrolysis, and the Arrhenius framework doesn't account for that mechanism directly. Another practical issue: the Arrhenius definition assumes complete dissociation for strong acids and bases. In concentrated solutions above about 1 M, even strong electrolytes like HCl don't fully dissociate due to ionic strength effects and ion pairing. The activity coefficients deviate significantly from 1, so your calculated pH will be wrong if you use concentration instead of activity. I learned this the hard way when preparing high-concentration stock solutions for industrial pH adjustment. The labeled molarity didn't match the measured pH at all. Switching to activity-based calculations and using a properly calibrated electrode with liquid junction compensation brought the readings into alignment within 0.05 pH units. If you need a reference for the standard enthalpies of neutralization, dissociation constants, or solubility data for common salts produced in these reactions, the CRC Handbook of Chemistry and Physics remains the most reliable source. It has been updated annually and covers the thermodynamic data you need for accurate calculations.
The Arrhenius Acid Base Reaction is a foundational concept, but it is narrow in scope. Use it when you are working with dilute aqueous solutions of classic acids and bases. Move to Brønsted-Lowry or Lewis definitions when your system leaves the aqueous phase or involves species that don't fit the original framework. Both expansions exist for a reason, and trying to force Arrhenius logic onto non-aqueous or gas-phase systems will only give you wrong answers.