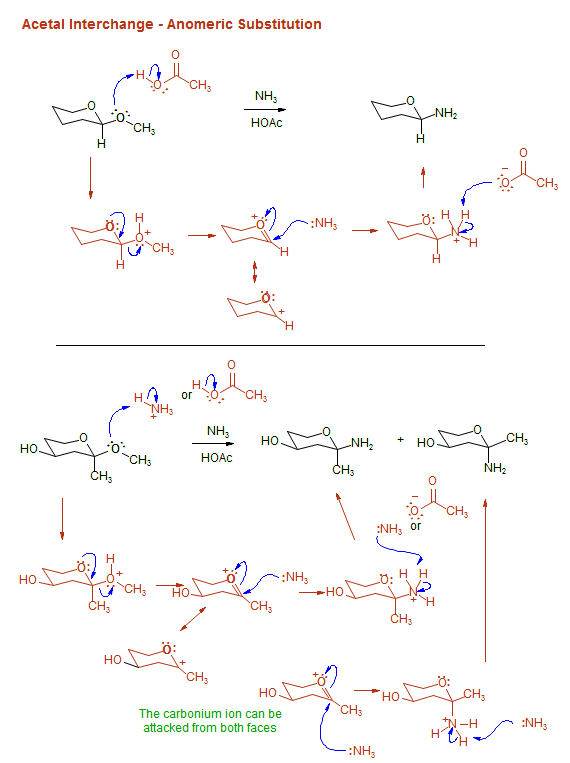
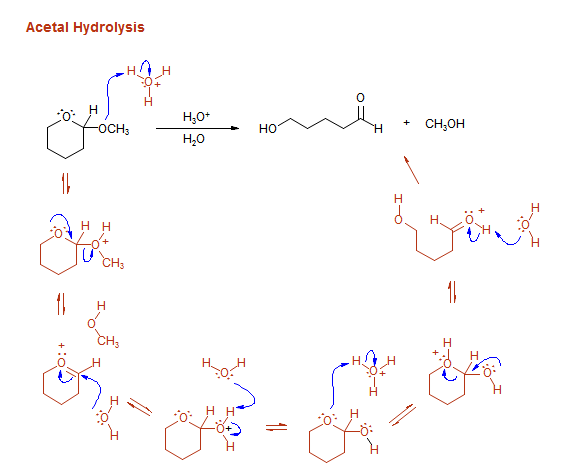
Arrow Pushing In Organic Chemistry
I spent my graduate school years drawing curved arrows until my wrist cramped, then realized I had been doing it wrong for two semesters straight. The core idea is simple: curved arrows track electron movement from source to sink. Electrons flow from regions of high electron density—lone pairs, pi bonds, or negative charges—toward regions of low electron density. That's it. Everything else is just applying that logic to increasingly complex molecules. The way you write arrows shapes how you think about a mechanism. A double-barbed arrow (the standard fishhook) shows two electrons moving. A single-barbed arrow shows one electron in radical chemistry. Most problems students encounter involve double-barbed arrows in polar mechanisms.
Getting started with Arrow Pushing In Organic Chemistry
Before touching a single mechanism, make sure you can correctly count valence electrons and assign formal charges. This sounds obvious, but I've seen advanced undergraduates miss entire reaction pathways because they drew a carbocation where there should have been a neutral amine. The electrons you're tracking need somewhere to go, and if your formal charge bookkeeping is off, your arrows will lead nowhere useful. Start with acid-base reactions. They are the simplest arrow-pushing problems and they teach you the vocabulary. When hydroxide attacks an alpha-hydrogen on a ketone, draw the arrow from the lone pair on oxygen to the hydrogen atom. Then draw a second arrow from the C-H bond to the carbon. Both arrows start at electron sources. Neither arrow points from a positive charge outward—that is a common error that creates physically impossible electron sources. nucleophilic substitution is the next step. SN2 reactions are clean: one arrow from the nucleophile's lone pair to the electrophilic carbon, one arrow from the C-LG bond to the leaving group. Watch out for the stereochemistry consequence. The backside attack inverts configuration, so you cannot represent the mechanism with frontside attack arrows without introducing an incorrect product.
SN1 requires two steps and three arrows total. First, the leaving group departs: one arrow from the C-LG bond to the leaving group. Second, nucleophilic attack: one arrow from the nucleophile to the carbocation carbon. The carbocation intermediate has an empty p-orbital, which is your electron sink in the second step. Here is something most textbooks gloss over: carbocations rearrange. If you generate a secondary carbocation adjacent to a tertiary carbon, a 1,2-hydride shift will almost certainly occur. Draw the arrow from the C-H bond on the neighboring carbon to the cationic center. The resulting tertiary carbocation is more stable. Students who skip this step lose points on exams and make mistakes in the lab. I once set up a reaction expecting a straightforward substitution and got a completely different product because the carbocation had shifted before the nucleophile could attack. Took me an hour to figure out why my NMR didn't match the predicted structure.
Get the Full Details

Common mechanisms and where arrow pushing gets tricky
E1 and E2 eliminations follow similar logic to substitution but produce alkenes instead of substituted products. The key difference is what you aim your arrows at. In E2, the base abstracts a proton while the leaving group departs simultaneously. Draw the arrow from the base to the beta-hydrogen, and another arrow from the C-H bond forming the new pi bond, and a third arrow from the C-LG bond to the leaving group. Three arrows in one concerted step. Make sure the beta-hydrogen and the leaving group are anti-periplanar—that geometry requirement matters for the orbital overlap, and anti-periplanar is the only conformation that works efficiently in most cases. E1 goes through a carbocation intermediate just like SN1. The base removes a beta-hydrogen after the leaving group has already left. Zaitsev's rule applies here: the more substituted alkene forms preferentially. You can predict this by drawing arrows to every possible beta-hydrogen and seeing which product has the most substituted double bond. Electrophilic aromatic substitution is where arrow pushing gets genuinely interesting. The benzene ring acts as a nucleophile, attacking an electrophile. Draw arrows from the pi system to the electrophilic species, then restore aromaticity by removing a proton. The intermediate is a sigma complex with a positive charge delocalized across the ring. Resonance structures matter here—you need to show where that positive charge lands to understand regioselectivity in substituted rings.
Conjugate addition, or Michael addition, involves a nucleophile attacking the beta-carbon of an alpha,beta-unsaturated carbonyl. The electrons flow from the nucleophile to the beta-carbon, then through the pi system to the carbonyl oxygen. Draw the arrow from the C=C bond to the C=O bond. This is different from direct 1,2-addition, and getting the arrow direction right determines which product you predict.
Advanced pitfalls I have encountered
The biggest conceptual error I see is drawing arrows from atoms instead of from bonds or lone pairs. An arrow must originate at an electron source. A carbon with four bonds and no lone pair is not an electron source. If you try to draw an arrow starting from a neutral carbon in a typical organic molecule, you are describing something that does not happen. Electrons do not spontaneously leave a saturated carbon. Another issue: students sometimes draw arrows that create pentavalent carbons. If you push electrons toward a carbon that already has four bonds without simultaneously breaking a bond, you violate basic valence rules. Every arrow pushing sequence must conserve electron count and respect the octet rule. Check your work after drawing each step. Count bonds around every atom. If any carbon has five bonds, you made an error. Radical mechanisms use single-barbed arrows and behave differently from polar mechanisms. The electron flow is half at a time. Homolytic bond cleavage produces two radicals, and each radical participates in chain propagation steps. Do not mix single-barbed and double-barbed arrows in the same mechanism unless you are explicitly describing a radical-polar crossover process, which is rare and requires careful justification.
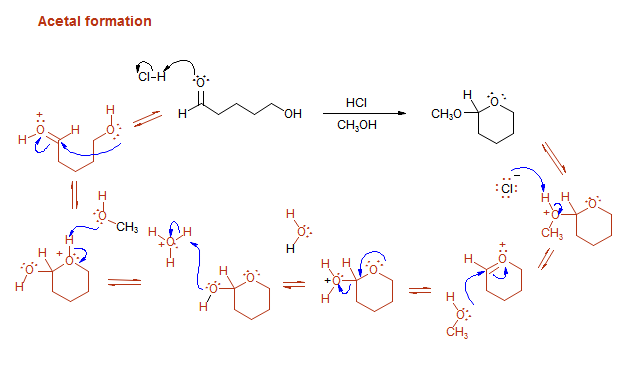
Pericyclic reactions are a special case. The arrows represent the concerted reorganization of electrons in a cyclic transition state. Diels-Alder reactions, electrocyclic reactions, and Cope rearrangements all follow specific arrow patterns that reflect orbital symmetry. The Woodward-Hoffmann rules determine whether a pericyclic reaction is thermally allowed or requires photochemical activation. Drawing the correct arrow pattern for a Diels-Alder reaction means showing three curved arrows moving in a cycle: from the diene's terminal pi bond to form a new sigma bond, from the dienophile's pi bond to form the second sigma bond, and from the remaining diene pi bond to form the new pi bond in the product.
Practical tips that actually help
Practice with real molecules, not generic R groups. When you work with actual compounds like enantiomerically pure substrates or heteroatoms with multiple lone pairs, you encounter edge cases that generic problems skip. I learned more from struggling through mechanisms involving sulfur and phosphorus than from hundreds of carbon-only exercises. Heteroatoms change everything—oxygen and nitrogen have lone pairs that participate differently than carbon pi systems, and sulfur can expand its valence shell, which creates reaction pathways that do not exist for second-row elements. Check your arrows against experimental evidence whenever possible. If your mechanism predicts a product that no one has ever isolated, reconsider your arrow pushing. Mechanisms are models, and models are only useful if they match reality. I have corrected my own drawn mechanisms after running control experiments that showed unexpected products, which forced me to rethink whether I had identified the correct nucleophile or missed a competing pathway. Use molecular models or drawing software to verify stereochemistry. Hand-drawn mechanisms on paper can obscure three-dimensional geometry that matters for the reaction outcome. A mechanism that looks correct on a flat page may be stereochemically impossible when you consider the actual spatial arrangement of substituents around a chiral center or ring system.
When in doubt about arrow direction, ask yourself: where are the electrons coming from, and where do they need to go? High density to low density. If you cannot identify a clear electron source or sink, you probably do not have the right mechanism. Go back to the structure. Check formal charges. Look for lone pairs you missed. There is no shortcut around practice. I spent roughly six months working through mechanism problems daily before drawing arrows became automatic. The skill is procedural, not theoretical. You learn it the same way you learn any physical skill: repetition, feedback, correction. The more mechanisms you draw correctly, the faster you recognize patterns in new reactions.