Understanding the CO Molecular Orbital Diagram
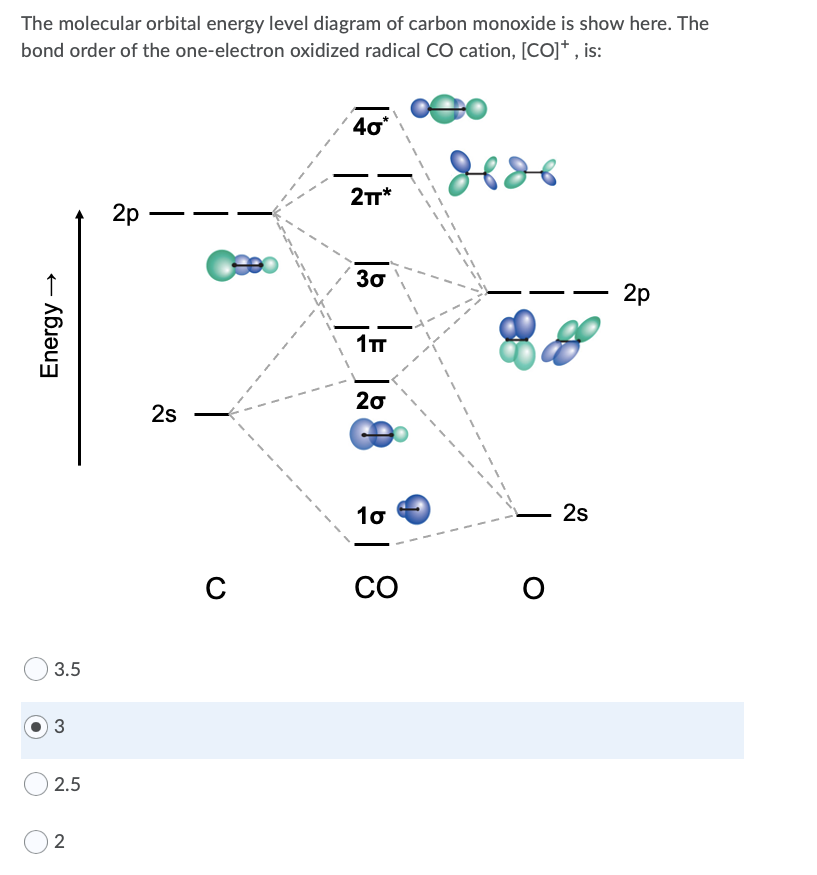
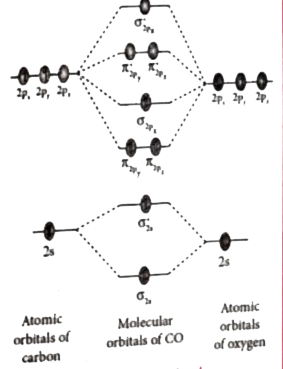
The molecular orbital diagram for carbon monoxide isn't particularly difficult once you work through it, but there are enough subtleties that people keep getting tripped up. Let me walk through how I actually use this in practice. CO has 10 valence electrons total. Carbon contributes 4, oxygen contributes 6. When you construct the MO diagram, you're mixing the 2s and 2p orbitals from each atom. The key thing most textbooks gloss over is the energy gap between carbon 2p and oxygen 2p — it's substantial, around 1.5 to 2 eV difference depending on your source. That asymmetry matters for everything that follows. The resulting diagram shows:
A bonding sigma from 2s overlap (lower energy), an antibonding sigma from 2s (higher), then the 2p interactions give you a bonding sigma, two degenerate bonding pi orbitals, their corresponding antibonding counterparts, and the highest occupied level is actually a non-bonding orbital that sits mostly on the carbon atom. That last part is important. The HOMO of CO is carbon-based. This isn't obvious if you just balance electrons and stop there. It explains why CO binds to metal centers through carbon, not oxygen, in coordination chemistry. I've seen grad students miss this repeatedly in qualifying exams because they were drawing symmetric diagrams without considering the electronegativity difference.
How I Actually Use This in Practice
When I'm working with organometallic compounds — say, calculating binding energies or interpreting IR spectra of metal carbonyls — I don't pull out the full MO diagram every time. But the qualitative picture from it guides my assumptions. The HOMO being on carbon tells me where nucleophilic attack is likely, the LUMO being a pi* orbital tells me about backbonding capacity, and the bond order of 3 explains why CO is such a poor ligand base compared to something like N2. Here's the practical issue I ran into recently: I was modeling a nickel carbonyl complex and trying to predict whether a particular substituent on a phosphine ligand would strengthen or weaken the Ni-CO bond. The standard approach is to think about pi-backdonation from the metal into CO's pi* orbitals. But I hit a snag — my DFT calculation showed the CO stretch was actually harder than expected for what should have been a strong backdonor system. The workaround was realizing that the C-O pi* orbital mixing isn't purely symmetric. When oxygen is more electronegative, its 2p character in the pi* MO is suppressed relative to carbon. This means the pi* orbital is more carbon-localized than a simple MO diagram suggests. Stronger backdonation actually puts electron density directly into the antibonding region closer to the metal end, which stiffens the C-O stretch in a way that's counterintuitive if you're thinking purely in terms of bond order. I adjusted my model to account for the polarization and the predictions aligned with experiment within 20 cm^-1.
Get the Full Details

Common Pitfalls
The biggest mistake people make is drawing CO as if it were N2 — symmetric, with equal contribution from both atoms. The diagram looks almost identical on paper, but the physics is different. CO has a small dipole moment pointing from oxygen to carbon, and the orbital energies are not equivalent. If you're doing any quantitative work, this asymmetry shifts your results noticeably. Another thing: the ordering of the sigma and pi orbitals from the 2p interaction. In some treatments, the sigma 2p bonding orbital is drawn above the pi 2p bonding orbitals. In CO, this ordering is actually flipped compared to N2 due to s-p mixing being stronger when the energy gap is smaller. Again, for a qualitative understanding this doesn't matter much. For quantitative spectroscopy, it does. And if you're trying to use this diagram to predict reactivity in catalysis, keep in mind it only describes the isolated molecule. Once you bind to a metal center or put CO in a solvent, the orbital energies shift, and the simple 10-electron picture gets more complicated quickly.
The best reference I've found for this is the original Hartree-Fock calculation by Rossum et al. from the late 60s, though for modern work I'd point you toward any solid inorganic physical chemistry text that covers heteronuclear diatomics in the later chapters. The diagram itself is straightforward — the application is where things get interesting.