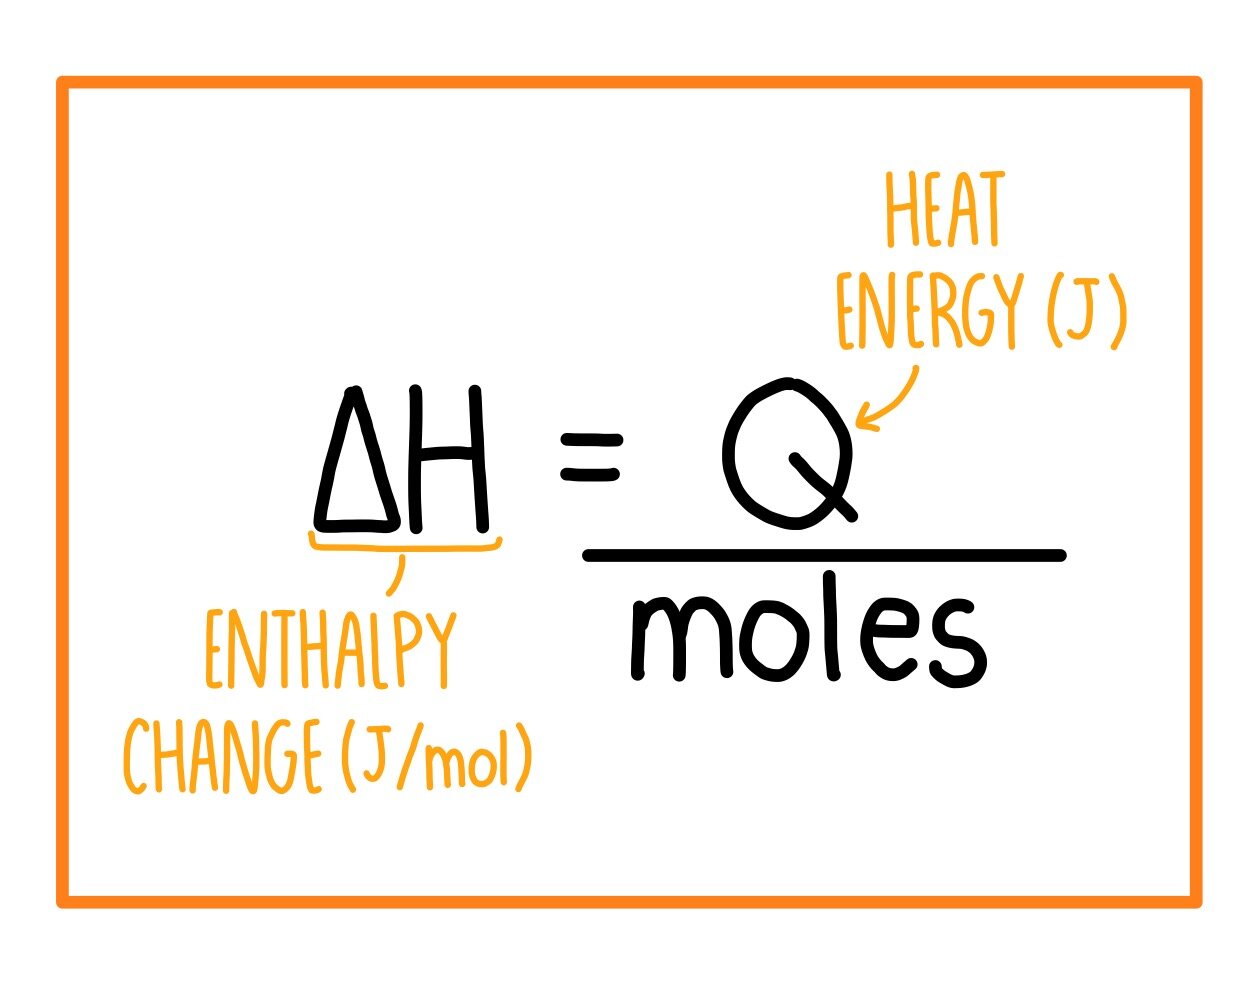Enthalpy Change Isn't as Simple as H = Products Reactants
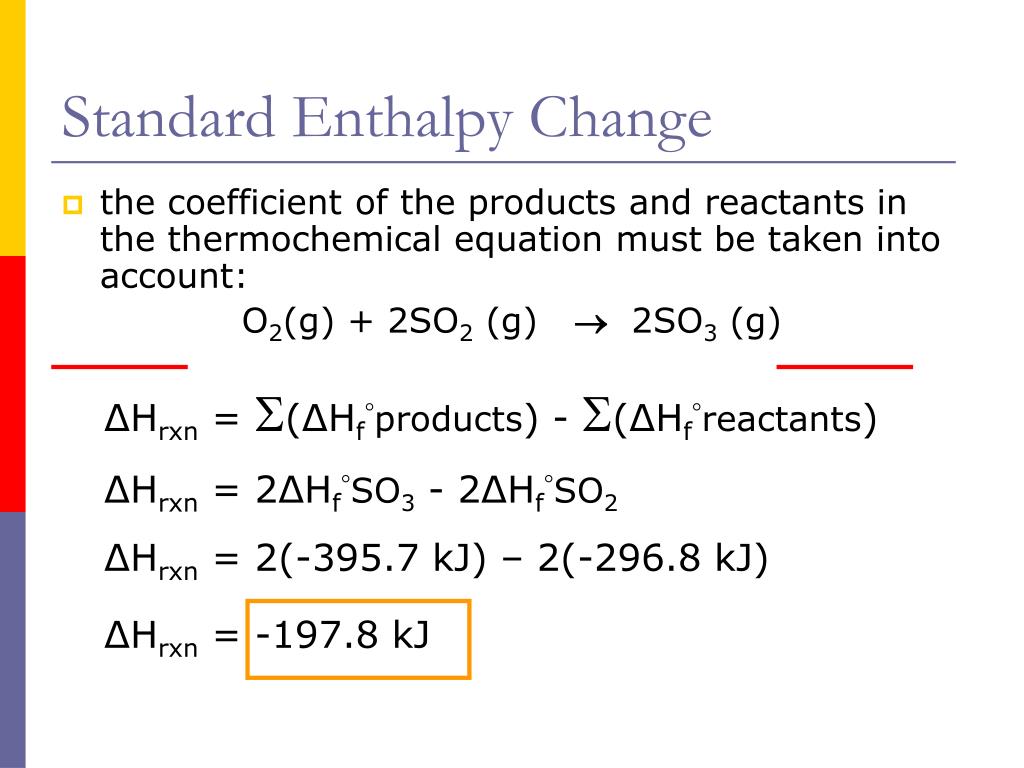
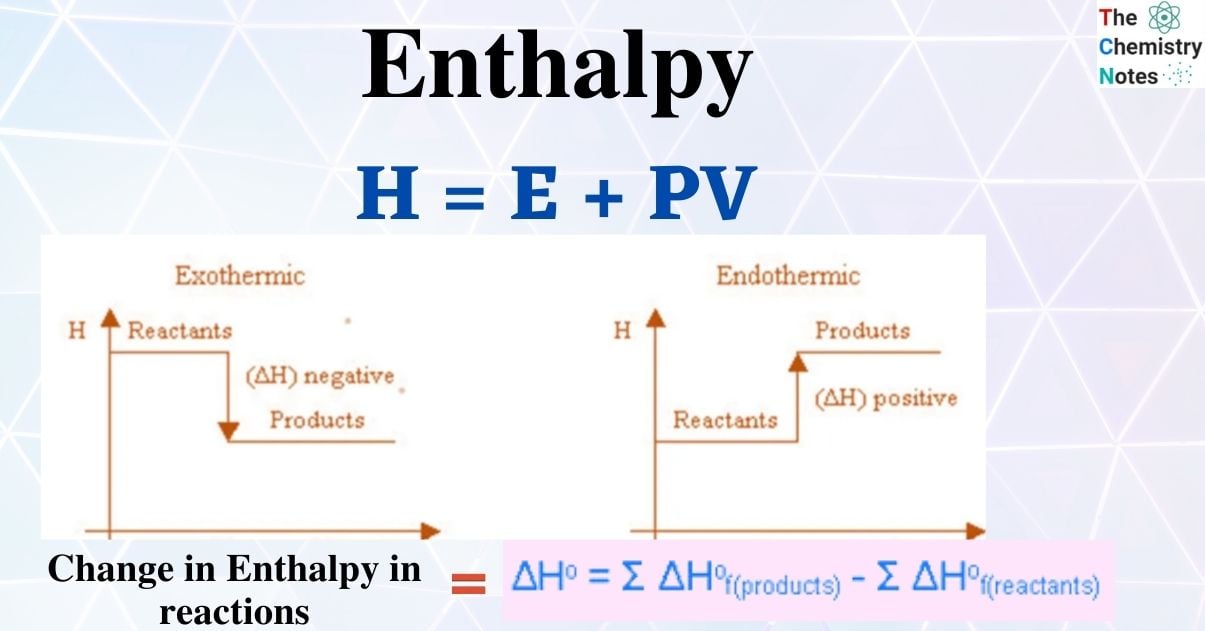
The equation you learned in your first chemistry course works for textbook problems, but it falls apart the moment you deal with real systems. I have spent more time debugging enthalpy calculations for process design than I care to admit. Here is what actually matters when you are working with this stuff. Start with the basic form: H = H_products H_reactants. That is the foundation, but it assumes constant pressure and no phase transitions during the calculation. If your reaction happens at 500°C and involves water switching from liquid to gas, that simple equation gives you garbage numbers. The practical version looks like this:
H_reaction(T) = H°_reaction(298K) + [Cp(products) Cp(reactants)] dT from 298 to T You need heat capacity data for every species involved. The integral accounts for sensible heat changes as temperature shifts away from standard conditions. Skip it and your results will be off by hundreds of kilojoules per mole at elevated temperatures. I ran into this head-on while designing a waste heat recovery system for a chemical plant. The spec sheet said a particular exothermic reaction released 450 kJ/mol at standard conditions. When I actually integrated the Cp curves for the real operating temperature of 723 K, the true enthalpy change came out to roughly 380 kJ/mol. Fourteen percent difference. The heat exchangers we originally specified would have been undersized and we would have had a serious thermal management problem down the line. Integrating Cp properly fixed that before any hardware was ordered.
For reactions involving phase changes, you add the latent heat terms directly into the balance. Melting, vaporization, sublimation — each one contributes a discrete enthalpy step that the temperature integral alone does not capture. Another detail people routinely miss: heat capacity is not linear with temperature. Using a single average Cp value across a wide temperature range introduces error. The proper approach uses polynomial correlations like the Shomate equation or the NASA seven-coefficient form. These give you Cp as a function of temperature and the integral becomes much more accurate.
Get the Full Details
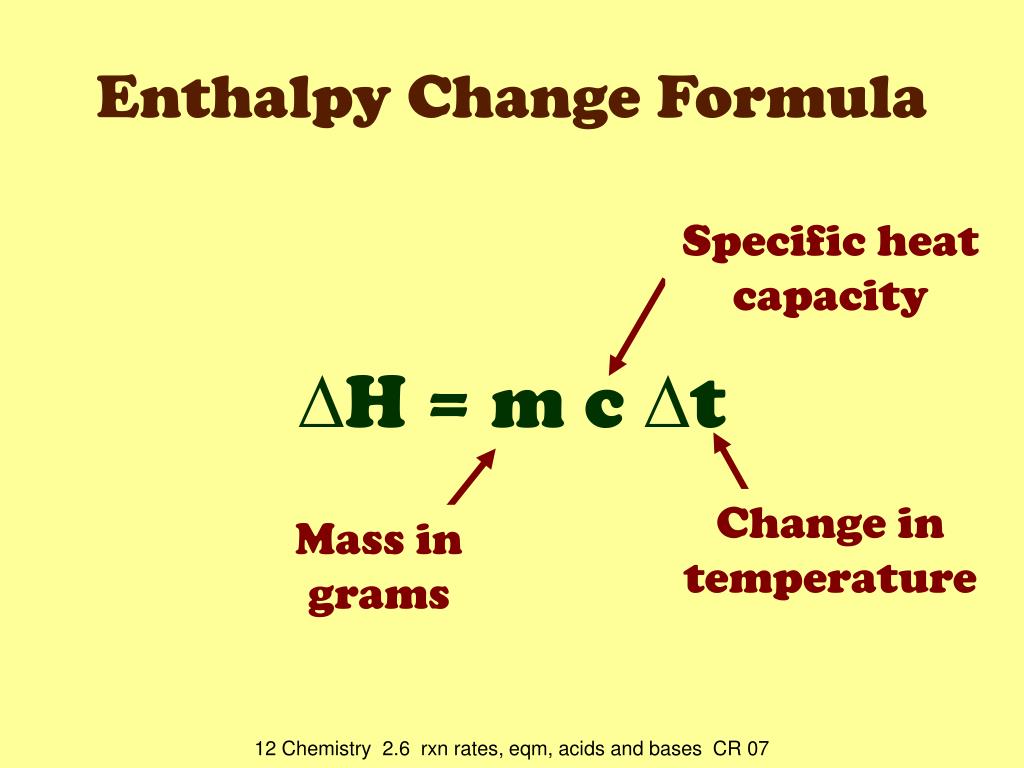
When the Standard Approach Completely Fails
There are scenarios where even the integrated form with proper Cp data breaks down. High-pressure systems above roughly 100 bar require fugacity corrections because the ideal gas assumption underlying the standard enthalpy formulation no longer holds. Real gas behavior changes the enthalpy significantly at those pressures. You need an equation of state like Peng-Robinson or Soave-Redlich-Kwong to correct for it. Mixtures are another headache. The simple subtraction method assumes ideal mixing, which is rarely true. Excess enthalpy of mixing can add or subtract meaningful energy from your balance. If you are working with non-ideal solutions — acids, bases, organic mixtures with hydrogen bonding — you need experimental mixing data or an activity coefficient model like NRTL or UNIQUAC to account for it. For combustion calculations specifically, I usually bypass the manual integration entirely and use a software tool. The NIST Chemistry WebBook provides enthalpy of formation data and heat capacity polynomials for thousands of species. Plugging those into a spreadsheet with the NASA polynomial coefficients cuts the setup time from a couple hours to maybe twenty minutes for a typical reaction.
If you need raw thermodynamic tables, the most reliable source I have found is the DECHEMA Chemistry Data Series. It is not free, but the data quality is substantially better than what you get from random online repositories. The JANAF Thermochemical Tables remain the gold standard for high-temperature data, though they are expensive and dense. The main takeaway is this: the basic equation is a starting point, not an endpoint. Once you move past 298 K and 1 atm, you need temperature-dependent heat capacities, phase change terms, and corrections for non-ideality if your system demands it. Skipping any of those steps is where the errors creep in. For quick reference, the standard enthalpies of formation you need to look up are typically listed in kJ/mol at 298.15 K and 1 bar. Make sure you are consistent with units and pressure standards between your formation data and your heat capacity correlations. Mixing data from sources that use 1 atm versus 1 bar introduces small but unnecessary discrepancies, especially when you are summing many species together.