Change In Entropy Formula
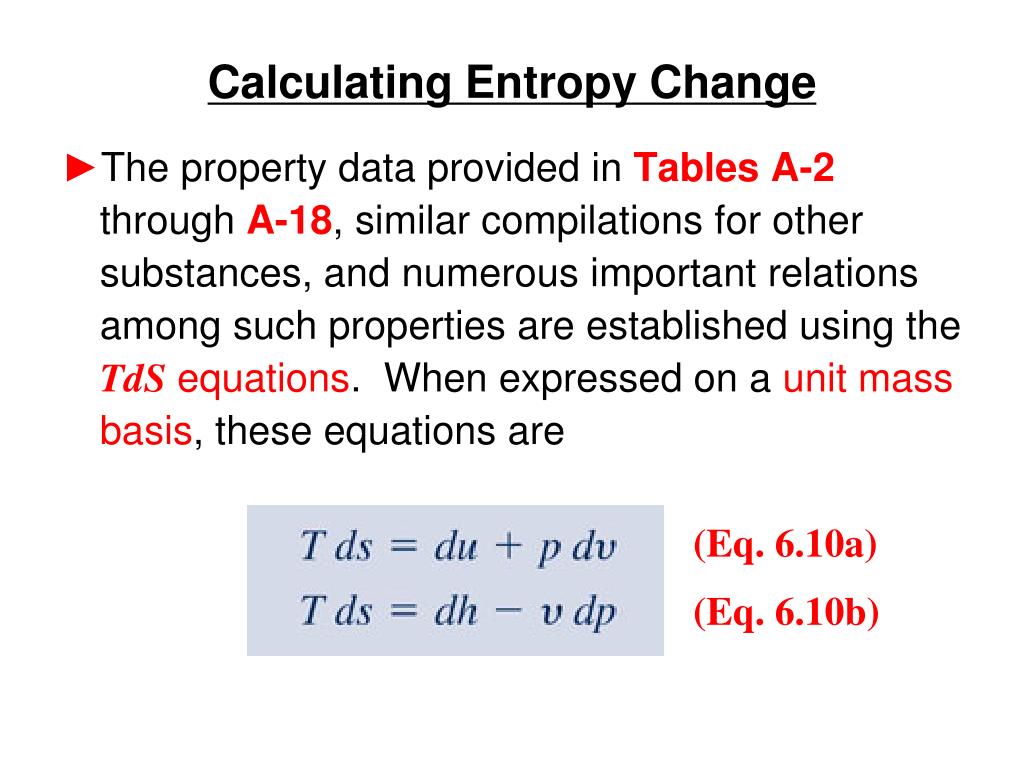
The change in entropy for a closed system undergoing a reversible process is calculated using the basic integral expression, and the version most people actually need in practice is the discrete form where delta S equals the integral of dQ divided by T over the path from the initial to final state. The formula is fundamentally straightforward, but it gets buried under layers of textbook jargon. Delta S equals Q over T only works for isothermal reversible processes where temperature stays perfectly constant. That is a very narrow condition. For anything else, you integrate. The key insight that separates people who understand this from people who memorize it is recognizing that entropy is a state function. The path you use to compute it does not have to match the actual process. You can always design a reversible path between the same two equilibrium states and integrate along that path instead. This matters a lot when dealing with real irreversible processes, because you cannot simply plug dQ over T from the actual process into the formula. The integral would give you the wrong answer.
Practical Cases and Where People Go Wrong
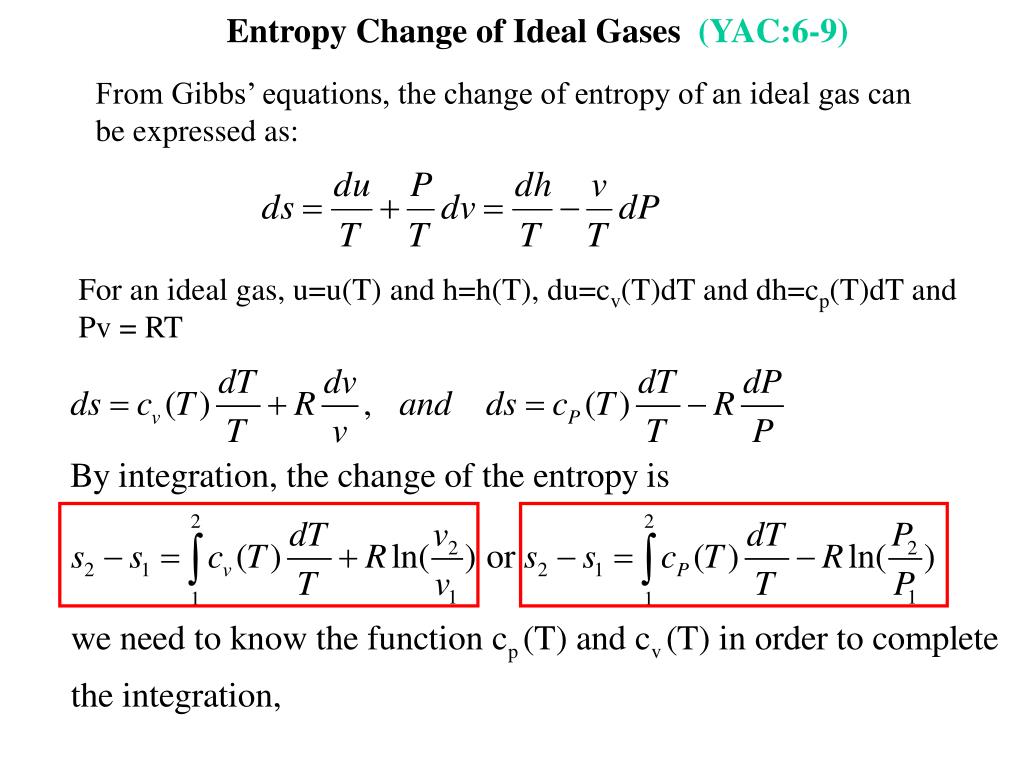
For an ideal gas going from state one to state two, the most useful form of the Change In Entropy Formula is delta S equals n times C sub v times the natural log of T sub f over T sub i plus n times R times the natural log of V sub f over V sub i. This combines the temperature and volume contributions into one expression, which saves you from having to find intermediate states. At constant volume, the volume term drops out. At constant pressure, you substitute C sub p and use the temperature and pressure ratio form. The relationship between C sub p and C sub v for an ideal gas is C sub p minus C sub v equals R. This matters because using the wrong heat capacity will shift your answer by a consistent amount, and on an exam or in a report it will look like a conceptual error rather than a simple mix-up. I ran into a problem last year where a student was calculating the entropy change for free expansion of an ideal gas into a vacuum. The gas expanded irreversibly from volume V to volume 2V at constant temperature. The correct approach is to imagine a reversible isothermal expansion between the same two states and integrate dQ over T along that path. The result is nRln2, even though the actual process had zero heat transfer. Many students try to set Q equal to zero and conclude delta S equals zero, which is wrong. The entropy change of the gas is still positive because the state changed. The total entropy change of the universe is also positive because the process is irreversible, and that is where the real physical content lies.
Phase Transitions and Mixed-System Problems
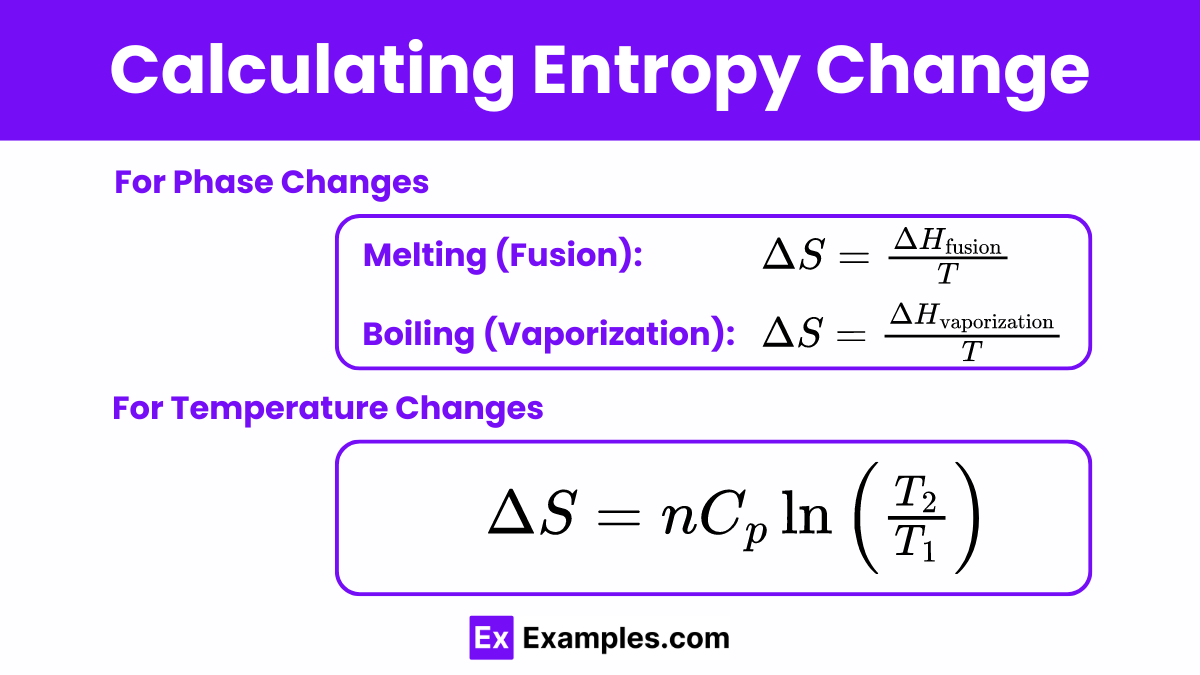
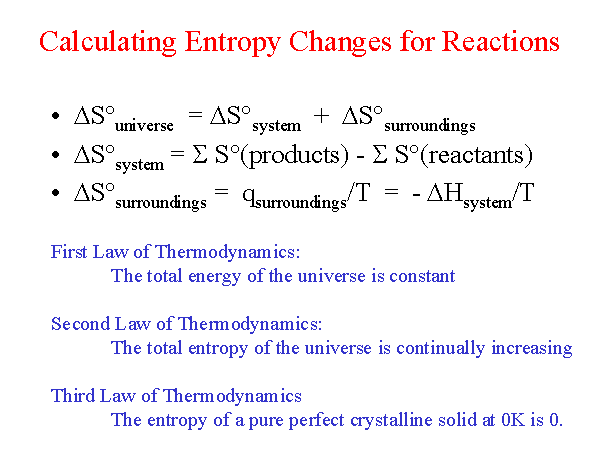
Phase transitions at constant temperature and pressure are where the simple form delta S equals delta H over T becomes genuinely useful. Melting ice at 273 kelvins with a heat of fusion of 333.55 joules per gram gives an entropy change of about 1.22 kilojoules per kelvin per kilogram. The calculation is direct because the temperature does not change during the transition, so the integral collapses to a simple division. When you combine multiple steps, like heating water from 280 kelvins to 350 kelvins at constant pressure, you need to account for the sensible heat in the liquid phase, the latent heat at the boiling point, and then the sensible heat in the vapor phase. Each step uses its own formula, and the total entropy change is the sum. This is routine, but the arithmetic is tedious, and it is easy to miss a step or use the wrong heat capacity for one of the regions. One thing that rarely gets emphasized in textbooks is the role of the surroundings. The formula delta S equals integral of dQ over T applies to the system, but when you want the total entropy change, you must also account for the entropy change of the surroundings, which is negative delta Q system over T surroundings. For an irreversible process, the total is always positive, and this is the actual statement of the second law. Calculating the system entropy using a reversible path and then computing the surroundings contribution separately is the standard workflow, and skipping either part will give you an incomplete picture.
Get the Full Details

Limitations and When the Approach Breaks Down
The Change In Entropy Formula as commonly presented assumes you know the heat capacity as a function of temperature. For many real gases and condensed phases, C sub v and C sub p vary significantly over wide temperature ranges. If you assume they are constant when they are not, your entropy calculation can be off by a meaningful margin, especially over large temperature spans. In those cases, you need tabulated thermodynamic data or an empirical correlation for heat capacity, and you integrate numerically rather than using a simple logarithmic formula. Another boundary condition is that the formula requires well-defined equilibrium states at both ends of the process. If the system passes through non-equilibrium states where temperature or pressure is not uniform, you cannot directly apply the classical entropy formula. Modern statistical mechanics can handle some of these cases through partition functions and molecular distributions, but that is a different framework entirely and not what most engineering or chemistry courses expect. If you are working with open systems where mass crosses the boundary, the entropy balance equation includes entropy transfer terms associated with mass flow, and the simple closed-system formula is no longer sufficient. You need to add the entropy carried in and out by the flowing streams, which introduces mass flow rates and specific entropies at each inlet and outlet.
Worked Example Under Constant Pressure
Consider 2.5 moles of nitrogen heated at constant pressure from 300 kelvins to 500 kelvins. Nitrogen has a molar heat capacity at constant pressure of approximately 29.1 joules per mole kelvin. The entropy change is n times C sub p times the natural log of 500 over 300. That gives 2.5 times 29.1 times the natural log of 500 divided by 300, which is roughly 38.8 joules per kelvin. If you instead used C sub v by mistake, which is about 20.8 joules per mole kelvin for nitrogen, you would get about 27.5 joules per kelvin, and that would be incorrect for a constant-pressure process. The difference is large enough to matter in any quantitative work.
Reversible Adiabatic Processes
A reversible adiabatic process produces zero entropy change in the system by definition, because dQ is zero everywhere along the path. This is a useful check: if you calculate a non-zero entropy change for a process you claimed was both adiabatic and reversible, something is wrong with your assumptions or your arithmetic. Irreversible adiabatic processes, on the other hand, produce positive entropy in the system even though no heat crosses the boundary, because internal irreversibilities like friction and turbulence generate entropy directly. Entropy calculations themselves are not difficult, but the traps are specific and recurring. Using the wrong heat capacity, confusing system and surroundings, treating irreversible paths as if they were reversible, and forgetting to convert temperatures to kelvins in logarithmic ratios are the mistakes I see most often. The formula is stable and well-behaved as long as you respect its assumptions, but it does not protect you from conceptual errors.