Working Through Chemical Kinetics Problems
Most people approach kinetics practice problems backwards. They look for a solution first, copy the steps, and move on without actually understanding why the method works. That strategy falls apart fast once you hit anything that isn't a straightforward first-order decay. The integrated rate laws are where things get messy in practice. You'll see textbooks present the zero, first, and second-order equations as separate topics, but real problems rarely announce which order they are. Your first move should always be figuring out the order, not plugging numbers into whichever equation feels familiar. Here's what I actually do when I'm working through a problem set. I start by checking whether the data gives me concentration versus time directly, or whether I'm dealing with something like absorbance, pressure, or conductivity readings that need conversion first. Half the mistakes I see come from people skipping that step entirely.
Why The Order Determination Step Matters
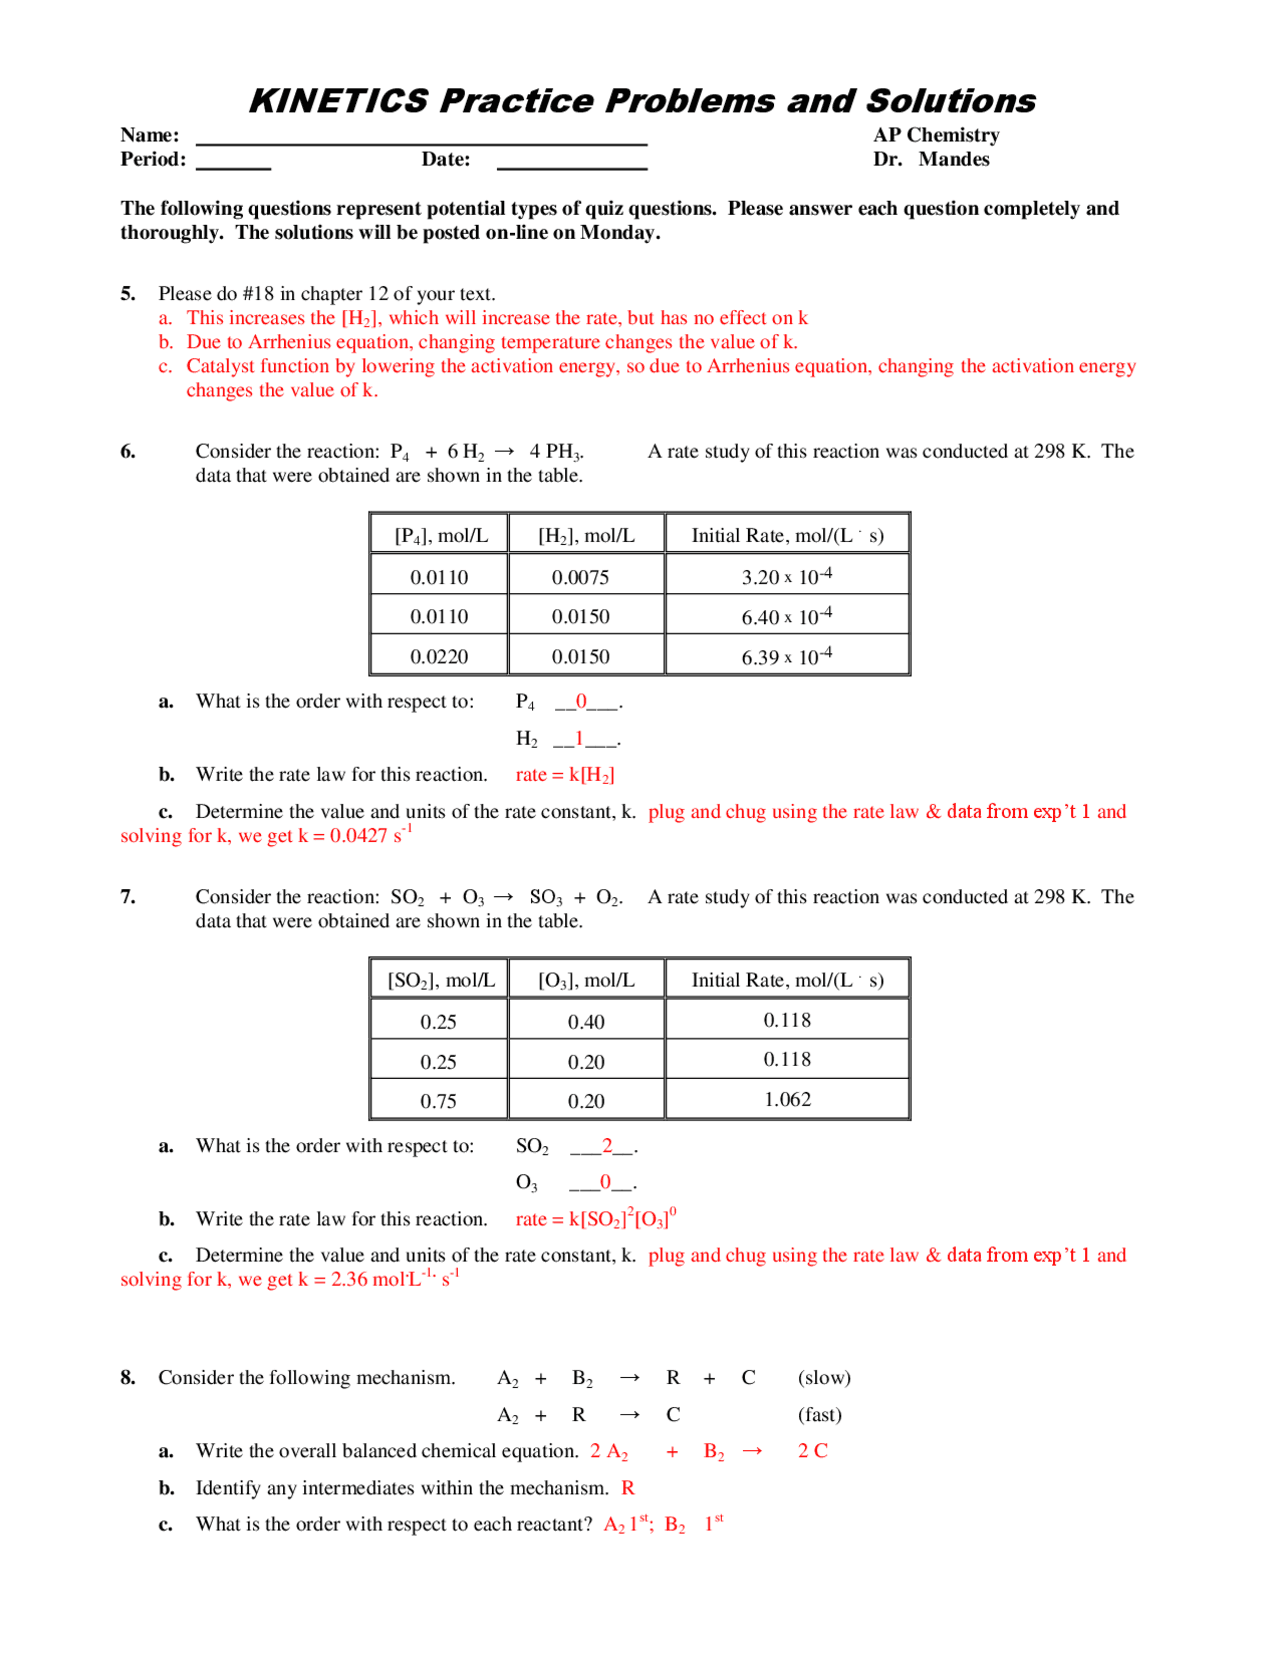
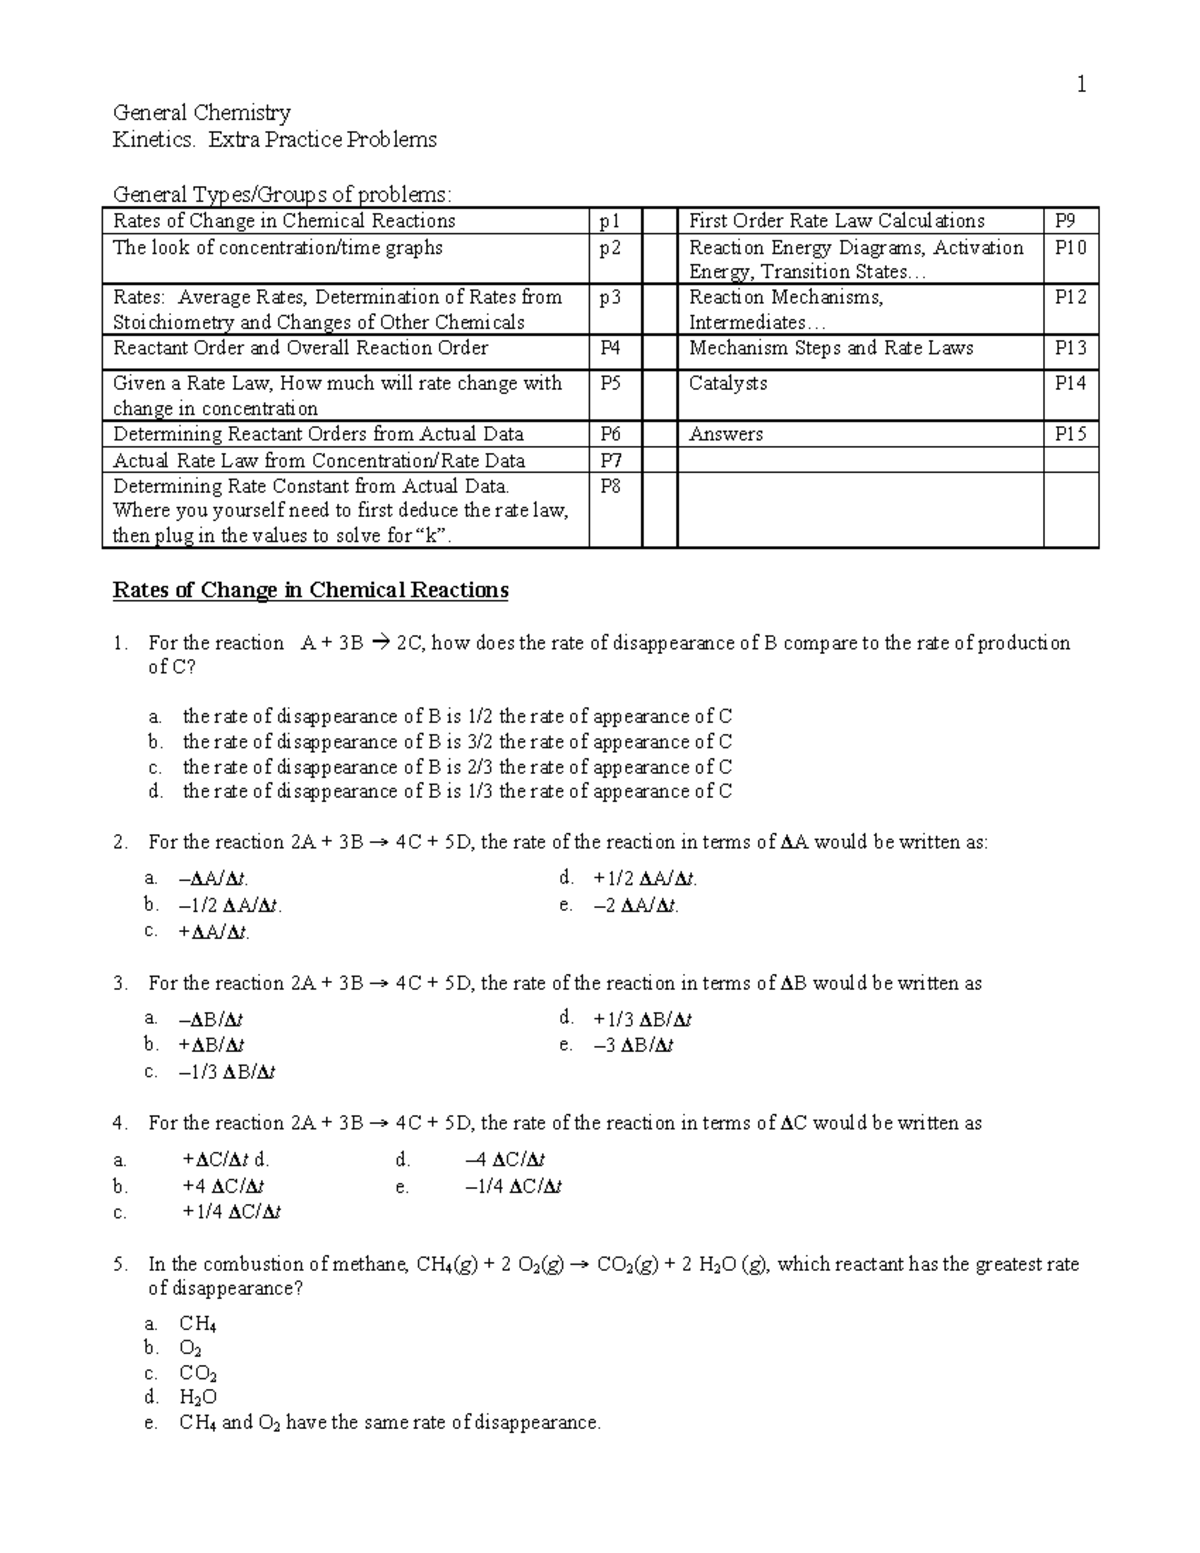
If you're given a table of concentration and time values, plot ln[concentration] versus time. If that line is roughly linear, it's first order. Plot 1/[concentration] versus time instead if you suspect second order. The R-squared value or how closely the points track a straight line tells you more than any guesswork. This is standard practice, not some advanced trick. There's a specific edge case that catches people regularly, and I ran into it just last semester grading exams. You'll get a problem where the stoichiometry doesn't match the rate expression. For example, a reaction like 2A B where the rate law is rate = k[A]², but the problem gives you data for B's formation instead of A's consumption. The relationship between d[B]/dt and -d[A]/dt isn't 1:1 here, it's 1:2. If you ignore that coefficient, your calculated k value will be off by exactly half. I mark that error consistently because it shows students are memorizing procedures instead of reading the equation carefully.
Chemical Kinetics Practice Problems And Solutions
The most common pitfall I see is treating half-life as independent of concentration for everything. That only holds true for first-order reactions. Zero-order half-lives decrease as concentration drops, and second-order half-lives increase. Students will write t/ = 0.693/k on an exam regardless of the reaction order and expect full credit. It doesn't work that way. When you're working with initial rates method problems, the key is changing one reactant concentration at a time while holding others constant. If the problem gives you five different experimental runs but two concentrations shift between any pair, you can't directly compare those two runs to find an individual order. You need to use the ratio method algebraically, which means taking the log of both sides of the rate equation and solving the system. It's tedious but straightforward. Another thing that trips people up is the Arrhenius equation application. The most frequent mistake is using Celsius instead of Kelvin for temperature. A temperature difference between 25°C and 35°C is only 10 degrees, but 298K and 308K are completely different ratios. When you calculate Ea from two rate constants at different temperatures, swapping in Celsius gives you wildly wrong activation energies. I've seen results like -50 kJ/mol for endothermic processes because someone forgot to convert.
Get the Full Details
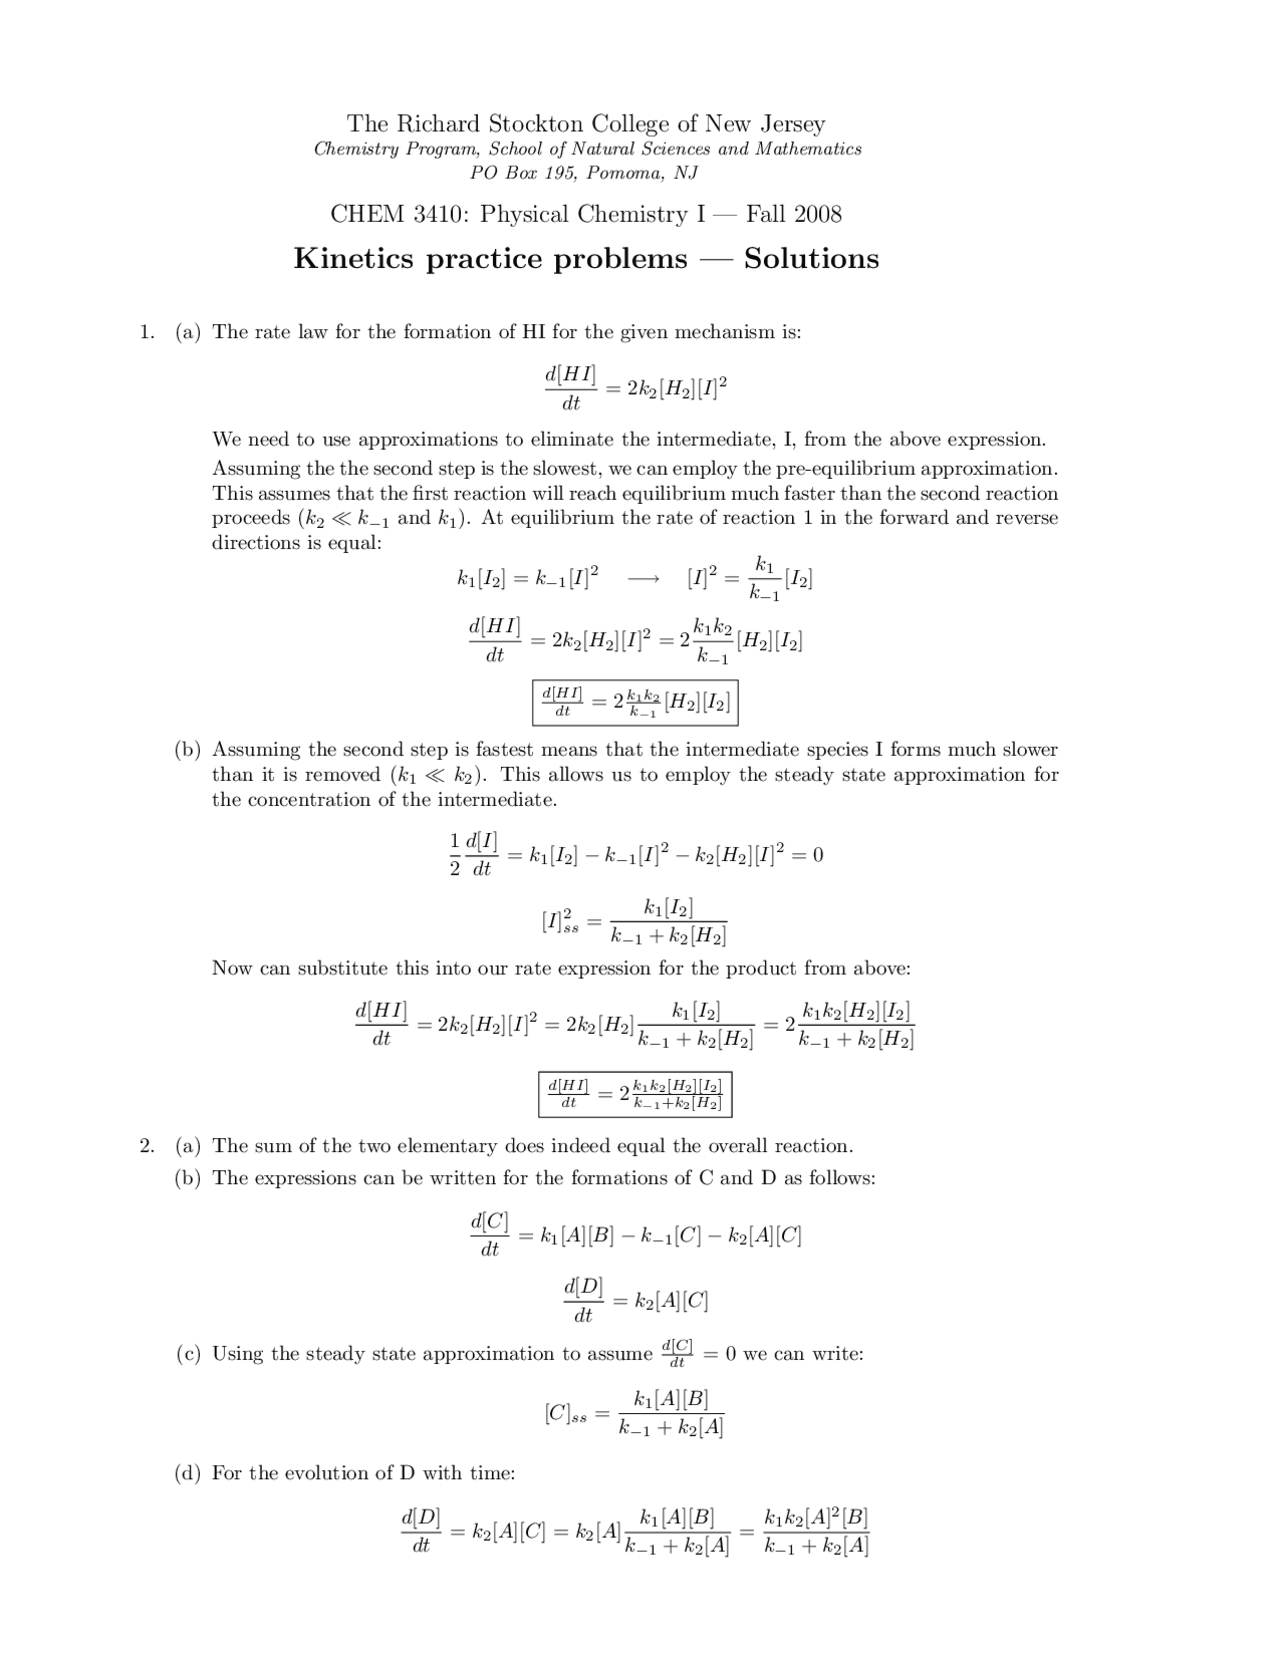
For mechanism problems, the steady-state approximation is the tool you reach for when you have reactive intermediates. The rule of thumb is: if the intermediate appears in the rate law, you need to eliminate it. Set d[intermediate]/dt equal to zero, solve for the intermediate's concentration in terms of the reactants, and substitute back into the rate expression. This can get algebra-heavy fast, especially with multi-step mechanisms, but it's the standard approach and it works reliably. One counter-intuitive insight that takes people by surprise: a catalyst doesn't change the equilibrium constant. It speeds up both the forward and reverse reactions equally. Students sometimes think adding a catalyst shifts the position of equilibrium, which it doesn't. It just gets you there faster. This distinction matters on exams and in real lab work where people confuse kinetic and thermodynamic control. If you're looking for practice material, the OpenStax Chemistry textbooks have a dedicated kinetics chapter with worked examples and problem sets. The University of Texas Chemistry department maintains a free problem bank online that's organized by reaction order and mechanism type. MIT OpenCourseWare also posts midterm problems with solutions from their General Chemistry courses. These are reliable sources because the problems are vetted through actual classroom use, not generated by algorithms.
The real limitation of most practice problem collections is that they oversimplify. Real kinetic data has noise, experimental error, and the occasional outlier point that throws off your linear regression. Textbook problems give you perfect data on a perfect line. When you're actually analyzing experimental results, you'll need to think about confidence intervals and whether your linear fit is statistically meaningful or just visually appealing. That skill doesn't come from working through clean textbook examples. Another practical note: when dealing with pseudo-order kinetics, the trick is making one reactant's concentration so large that it effectively stays constant during the reaction. This lets you isolate the order with respect to the other reactant. The catch is that your observed rate constant k_obs will depend on the concentration of the excess reagent. To recover the true rate constant, you need to run the experiment at several different excess concentrations and plot k_obs against that concentration. Skipping that step means you're reporting an observed constant as if it were intrinsic. For reaction mechanism determination, the slow step approximation and the equilibrium approximation serve different purposes. Use the steady-state approximation when the intermediate is highly reactive and doesn't build up. Use the equilibrium approximation when the intermediate forms and breaks down rapidly compared to the product-forming step. Mixing these up gives you the wrong rate law every time. I've had to redo mechanism problems in office hours because students applied steady-state when equilibrium was the appropriate assumption.
The bottom line is that kinetics problems are solvable with a systematic approach, but the system has to be flexible enough to handle cases that don't match the textbook template. Start every problem by writing down what you know, what you need, and what the units should be. Then figure out the order, choose the right equation, and check whether your answer makes physical sense. If your rate constant has units of M/s for a reaction you just determined is second order, something went wrong.