Setting Up Lab Work for Biology and Biochem Undergrads
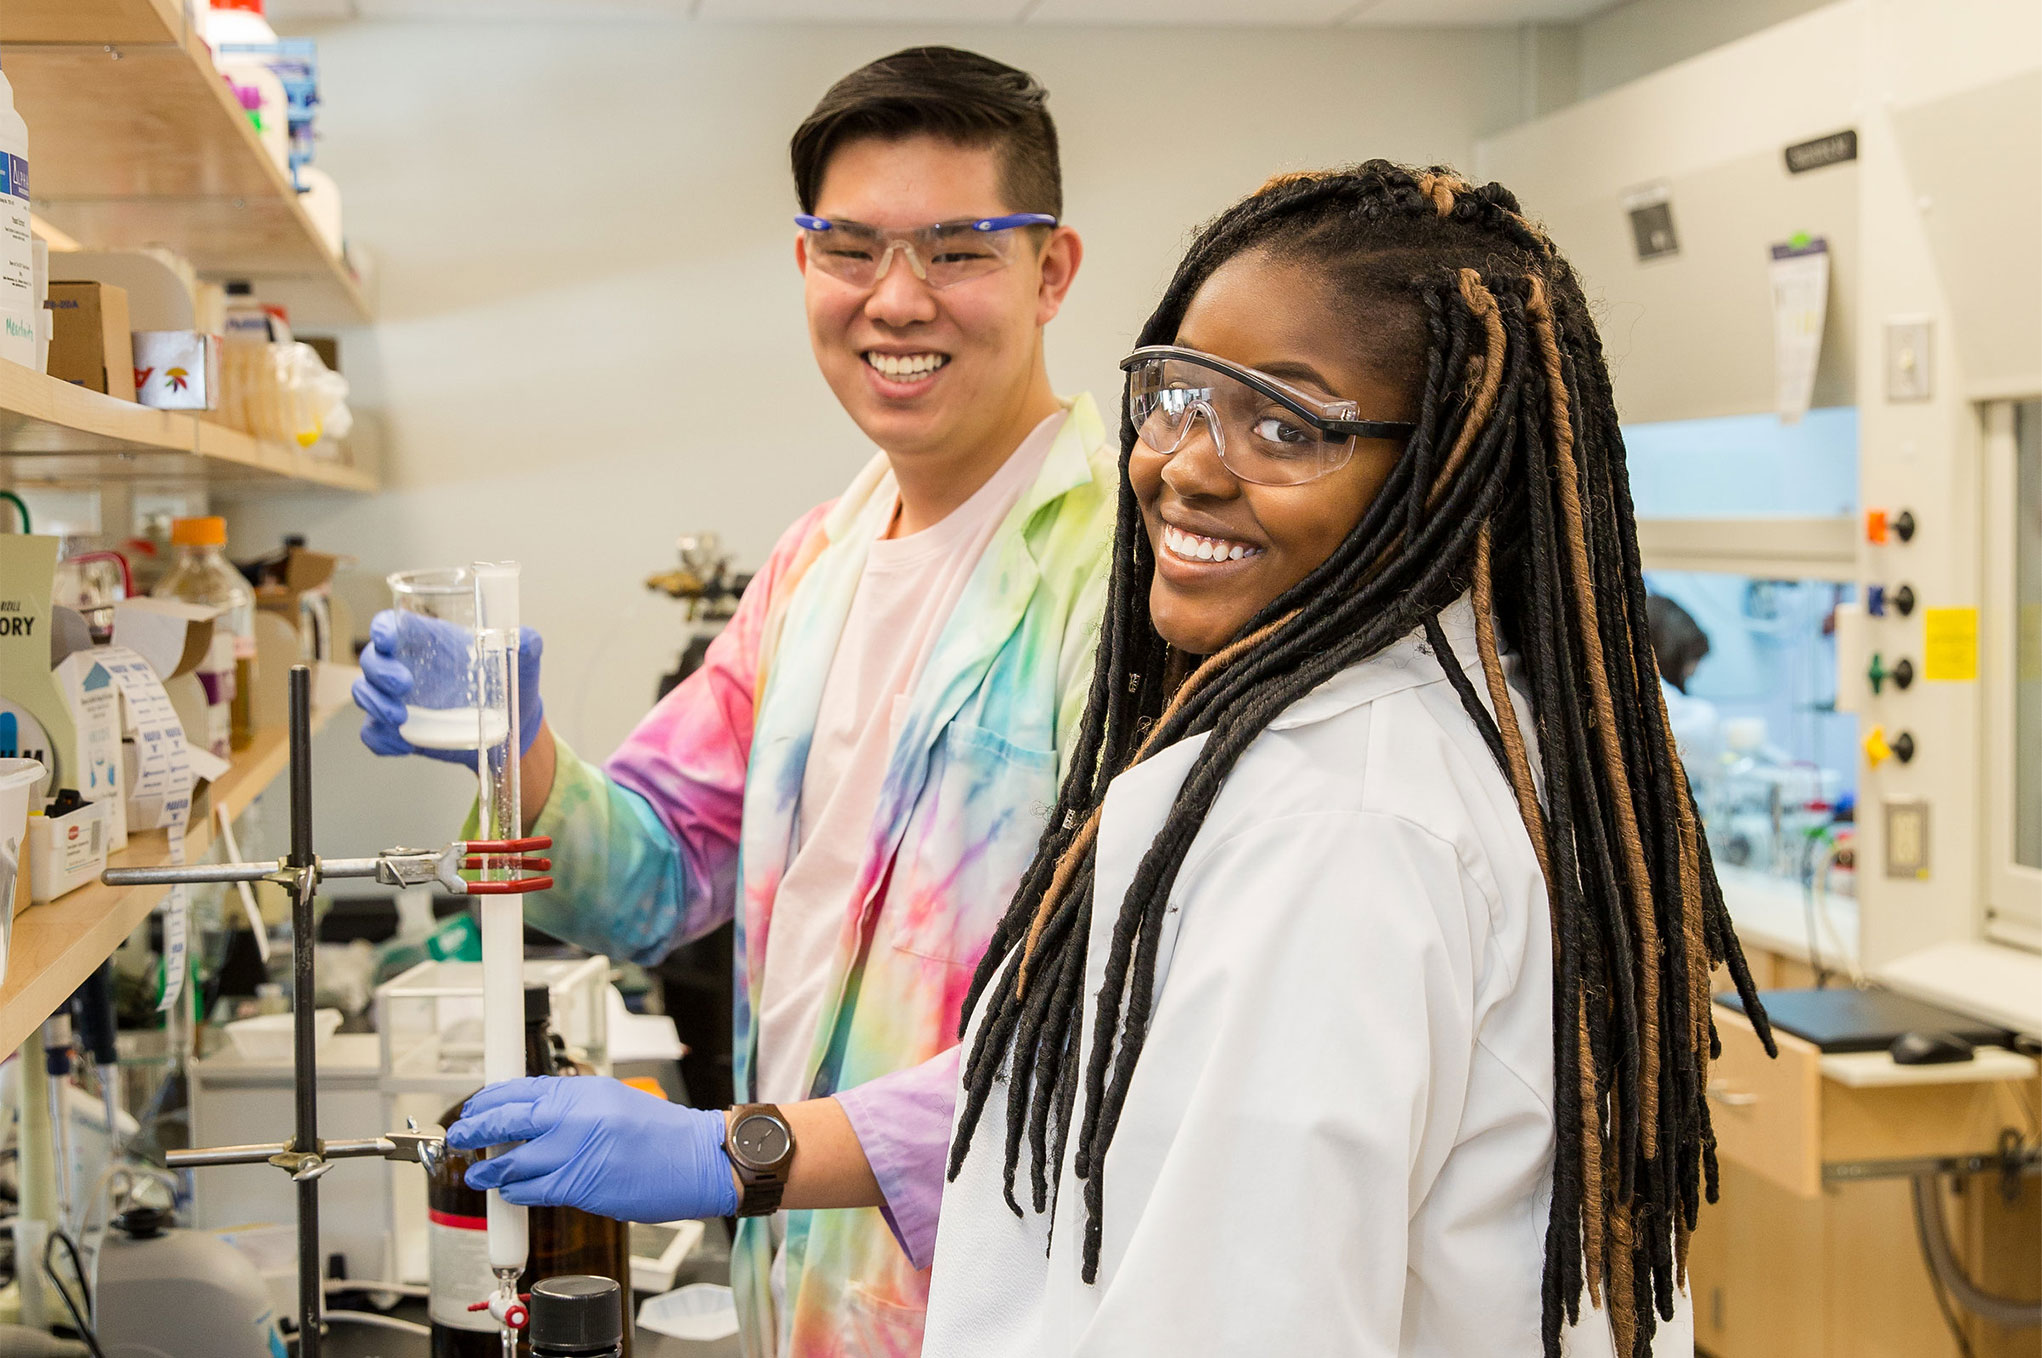
The first time I ran a spectrophotometer assay with a group of life science undergrads, three of them mixed up the cuvette orientation and wasted forty minutes of data collection. It happens constantly. Chemistry experiments for life science majors aren't chemistry majors' chemistry experiments, and the gap between what your lab manual says should happen and what actually happens in a 200-level teaching lab is wider than most students realize. Here is how to actually get usable results without losing your sanity. The standard curriculum tends to revolve around five or six core techniques repeated across different course levels. Acid-base titration with phenolphthalein. A copper cycle where you dissolve copper metal, precipitate it as hydroxide, convert it through several oxidation states, and end up recovering copper. Spectrophotometric determination of an unknown using Beer's Law. Enzyme kinetics with catalase or beta-galactosidase. Paper or thin-layer chromatography of plant pigments. These are standard because they teach transferable skills, not because they are inherently fascinating. I ran a catalase kinetics lab last semester where the hydrogen peroxide solution had partially decomposed on the shelf. The instructor's manual assumed a fresh 3% stock, but ours had aged approximately six weeks. My initial velocity readings were about 40% lower than the expected values. Instead of reporting nonsense or fudging the numbers, I remeasured the actual concentration by titrating the peroxide against potassium permanganate before starting the enzyme assay. The corrected Michaelis-Menten parameters came out within 12% of the literature value, which is acceptable for an undergraduate lab. If your lab partner just reports the raw numbers without checking reagent integrity, their Km value will be garbage and they will get a lower grade than they deserve. Don't be that person, but also don't blame yourself when the textbook answer doesn't match your data.
One thing nobody explains well is buffer preparation. Students treat it like a recipe: add this many grams, adjust pH, fill to volume. They don't understand that the pKa of a phosphate buffer shifts by roughly 0.014 pH units per degree Celsius. If you calibrate your pH meter at 22°C and then run the assay at 37°C, your buffer pH has drifted without you noticing. I learned this the hard way during a protein precipitation lab where my samples kept coming out cloudy instead of forming tight pellets. The issue was that the sodium acetate buffer had been adjusted at room temperature but the experiment required an ice-cold environment. The pH shifted enough to keep some proteins soluble. Recalibrating and readjusting the buffer at the actual working temperature fixed it immediately. This is a small detail that separates students who finish on time from students who spend three lab periods redoing the same procedure.
Practical Workflow for Common Assays
Spectrophotometry is the workhorse technique, and it is also the one where students make the most avoidable errors. The basic principle is simple: measure absorbance at a specific wavelength, plot a standard curve, interpolate your unknown. The actual practice involves more fiddling than the manual suggests. Start by blanking properly. Most students wipe the cuvette with a paper towel and move on. Use a lint-free lens tissue and make sure you are using the same side of the cuvette facing the light path each time. Fingerprint residue on the optical surface can add 0.02 to 0.05 absorbance units, which is negligible for a rough lab but destroys precision on a quantitative assignment. I once had a student spend two weeks trying to figure out why her standard curve had a consistent offset. It turned out she was holding the cuvettes by the frosted sides but occasionally grabbing the optical face. The offset was about 0.03 AU across the board. She felt terrible about it. Don't let that be you. When preparing standard curves, don't just do three points and call it a day. Five concentrations spanning the expected range, plus a zero blank, is the minimum for a reliable curve. The correlation coefficient matters less than you might think. A curve can have an R-squared of 0.99 and still be systematically wrong if your standards were prepared from an incorrect stock concentration. Verify your serial dilutions by checking the math out loud before you pipette. I keep a small notebook at the bench where I write down each dilution factor and the expected final concentration before touching any equipment. It takes ten seconds and has saved me from preparing an entire plate of wrong concentrations at least twice.
Get the Full Details

Common Pitfalls in Quantitative Work
Pipetting technique is where most point deductions happen. The most frequent mistake is pre-wetting the tip. For aqueous solutions this should be standard practice, but students often skip it because the manual doesn't emphasize it enough. A dry tip absorbs a thin film of liquid, and your first dispense will be consistently low by about 1 to 3%. Over a series of serial dilutions, that error compounds. Pre-wet by aspirating and dispensing the liquid two or three times before delivering your actual volume. It adds about five seconds per transfer. Titration is another area where textbook descriptions bear little resemblance to practice. The endpoint in a real titration is not a sharp instantaneous change. It is a range, usually about 0.1 to 0.2 mL for a standard burette. Students who stop the moment they see the first permanent color change are typically overshot. The trick is to slow down to drop-by-drop near the endpoint and swirl continuously. Also, rinse the sides of the flask with deionized water from a wash bottle during the titration. Any droplets of titrant sitting on the walls above the solution level haven't reacted yet, and washing them down ensures accuracy. I had a professor who deducted points specifically for not doing this, and honestly it is good pedagogy. Real analytical work requires this discipline.
When Things Go Wrong and What to Do
Your data will not always look like the textbook example. This is normal. A typical chromatography experiment might show three clean bands in the manual but four smeary ones in your hands. Possible causes include: the solvent front moved too fast because the chamber wasn't properly saturated, the sample was applied too heavily and overloaded the stationary phase, or the developing solvent composition was slightly off because someone used tap water instead of deionized water when making the mobile phase. None of these are catastrophic. Document what happened, note the deviation, and discuss it in your report. Instructors would rather see honest analysis of a messy result than fabricated clean data. One specific scenario that causes recurring headaches is protein assays when your sample contains reducing agents. The Bradford assay, which is the most common undergraduate protein quantification method, gets interfered with by DTT and beta-mercaptoethanol. If your lab protocol requires these reagents and you are using a Bradford kit, your concentrations will be severely underestimated. The workaround is to run a parallel BCA assay, which is more tolerant of reducing agents, or to dilute your sample into a reducing-agent-free buffer before assaying. I mention this because I see it come up every year and students always seem surprised. It is not your fault the protocol didn't warn you, but it is your responsibility to catch it.
Time Management and Lab Efficiency
A standard undergraduate lab period is usually three hours. That sounds generous until you account for setup, cleanup, and the inevitable waiting periods. Plan your workflow so that time-consuming steps happen simultaneously. While your titration is equilibrating, prepare your next set of standards. While your centrifuge is spinning, clean your glassware. I used to watch students sit idle for twenty minutes waiting for a water bath to reach temperature, then rush through the actual measurement because they were behind schedule. Learning to sequence tasks makes the difference between a rushed subpar experiment and a careful reproducible one. Data recording habits matter more than students think. Write down observations in real time, not from memory afterward. I once reconstructed a lab report from memory and got the incubation temperature completely wrong because I hadn't noted that the water bath had fluctuated between 36 and 39°C during the run. That fluctuation explained why my yield was lower than expected. The data was all there, I just failed to record it at the moment. Your lab notebook should include the date, your partner's name, ambient conditions if relevant, and any deviations from the standard procedure. This isn't bureaucracy. It is what lets you actually understand your results when you write the report. The techniques covered in these courses form the foundation for virtually everything else in a life science laboratory. Spectrophotometry underlies enzyme assays, nucleic acid quantification, and cell growth measurements. Titration skills transfer directly to understanding physiological acid-base balance. Chromatography concepts appear again in biochemistry when you purify proteins or analyze metabolites. The reason these particular experiments persist in the curriculum is that they are foundational, not because they are exciting. Treat them as skill-building exercises rather than mysteries to solve, and you will be better prepared for upper-level work.