Working With Cladograms When You Actually Need the Right Answer
I spent years grading introductory biology labs, and if there is one thing that drives me insane it is watching students build cladograms that are technically correct but answer the wrong question. You draw a pretty branching diagram, you label the nodes, you even get the outgroup right, and then you submit it only to realize the prompt was asking for something subtly different. The cladogram analysis answer key exists precisely because of this gap between what the exercise tests and what the student thinks it tests. A cladogram is a hypothesis about relationships. It shows shared derived characteristics, not overall similarity. That distinction matters more than most textbooks admit. Two species might look identical because of convergent evolution, but a proper cladogram based on molecular data could place them on completely different branches. I remember one lab where students were asked to construct a cladogram from a character matrix containing thirteen taxa and thirty-two traits. The answer key had them using limb bone morphology, but three of those traits were actually autapomorphies, meaning they only diagnose single species and tell you nothing about grouping. If you include autapomorphies as if they were synapomorphies, your tree structure collapses into nonsense. The workaround I taught was to flag any trait that appears in only one terminal taxon and remove it before running the parsimony analysis. Takes about ten seconds and saves you from half an hour of confused grading.
What a Cladogram Analysis Answer Key Actually Contains
Most answer keys you will find online or in instructor manuals cover the same ground, but the quality varies wildly. A solid one includes the properly rooted tree, the implied character transformations at each node, and the justification for which traits are treated as synapomorphies versus plesiomorphies. Some also list the most parsimonious tree length in steps. A weak one just shows a diagram with labels and expects you to reverse-engineer the logic. Here is what to look for when you are checking your work against a key: Root placement matters. If the key does not specify which node is the root, it is either poorly constructed or deliberately ambiguous. The root determines the direction of all evolutionary change in the diagram. Without it, you cannot tell whether a trait is ancestral or derived at any given node. Always verify that the outgroup is placed correctly, because that is the anchor point for everything else.
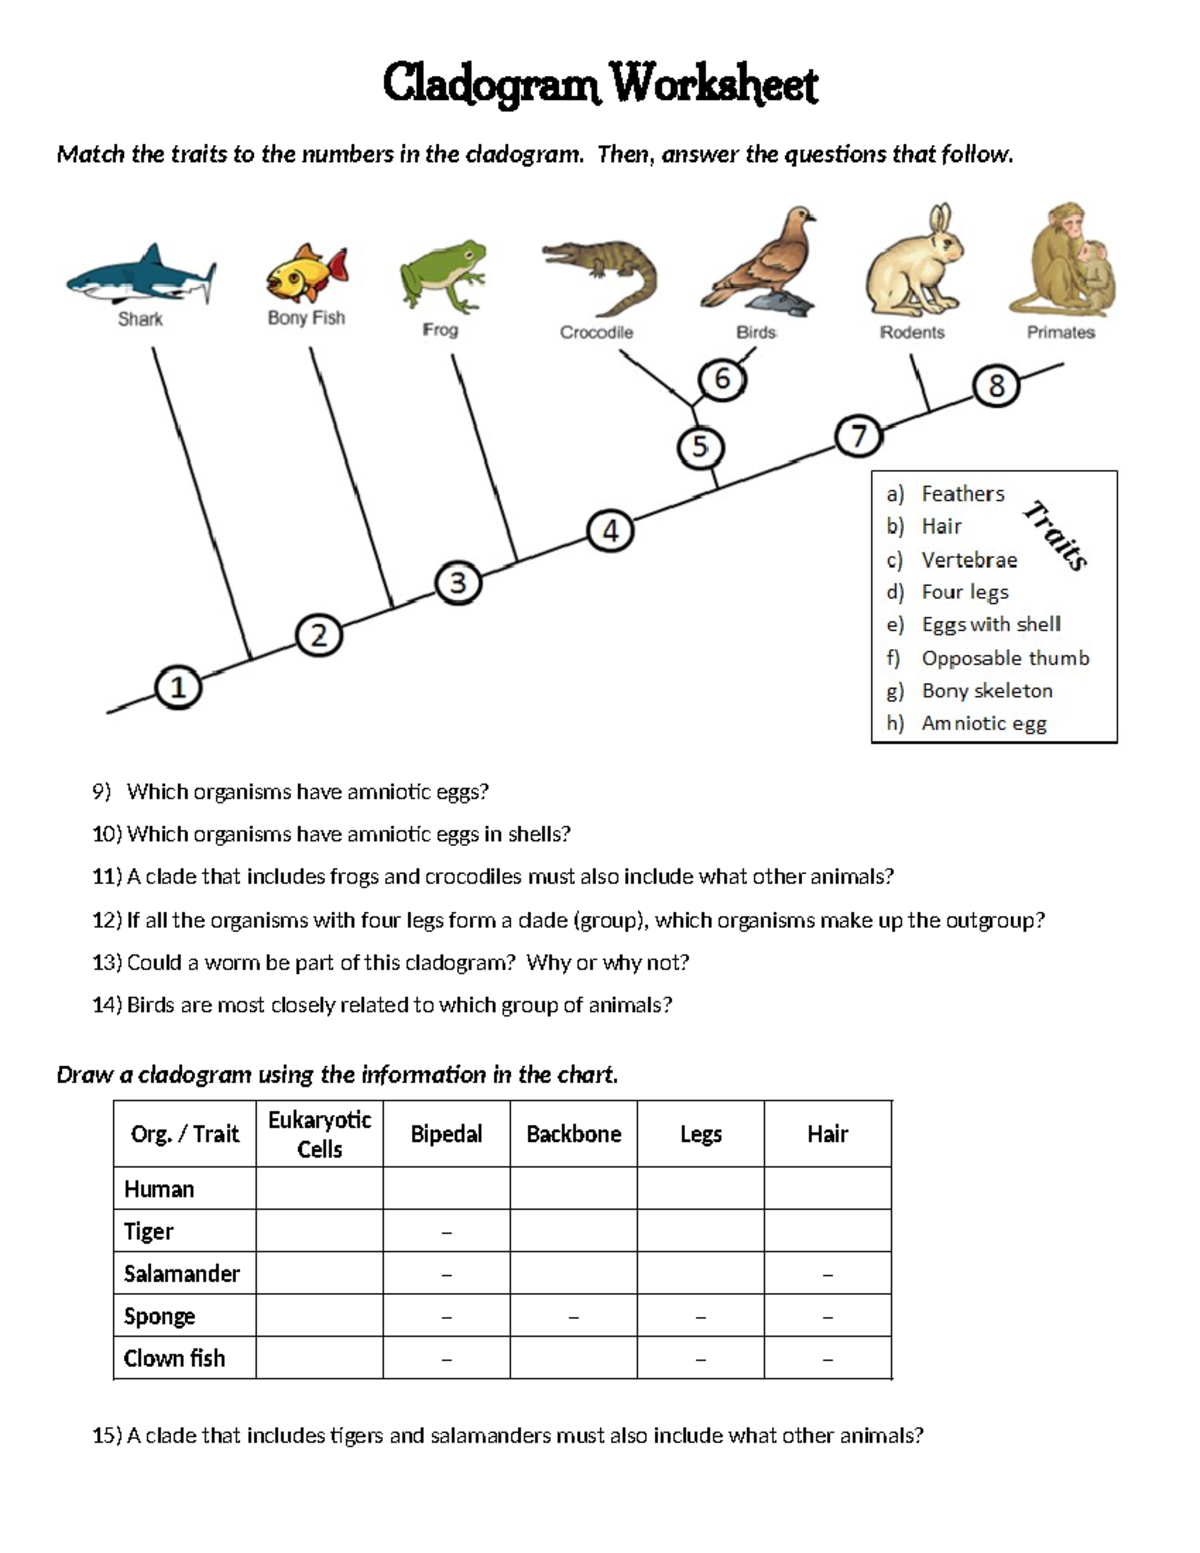
Character mapping is where most students lose points. The answer key should show which characters are shared derived traits at each branching point. For example, if mammals and reptiles share amniotic eggs, that character maps to the node uniting them, not to any individual branch. Misplacing a character one node down the tree flips the entire interpretation of that trait's evolutionary history. Parsimony scores let you verify your tree efficiency. If your cladogram requires forty-five character state changes and the answer key's tree needs thirty-eight, yours is not necessarily wrong, but it is less efficient. Occam's razor applies here in a literal sense. The most parsimonious tree is the one that assumes the fewest evolutionary changes, which tends to align better with known phylogenetic data.
Get the Full Details

How to Use a Cladogram Analysis Answer Key Without Learning Nothing
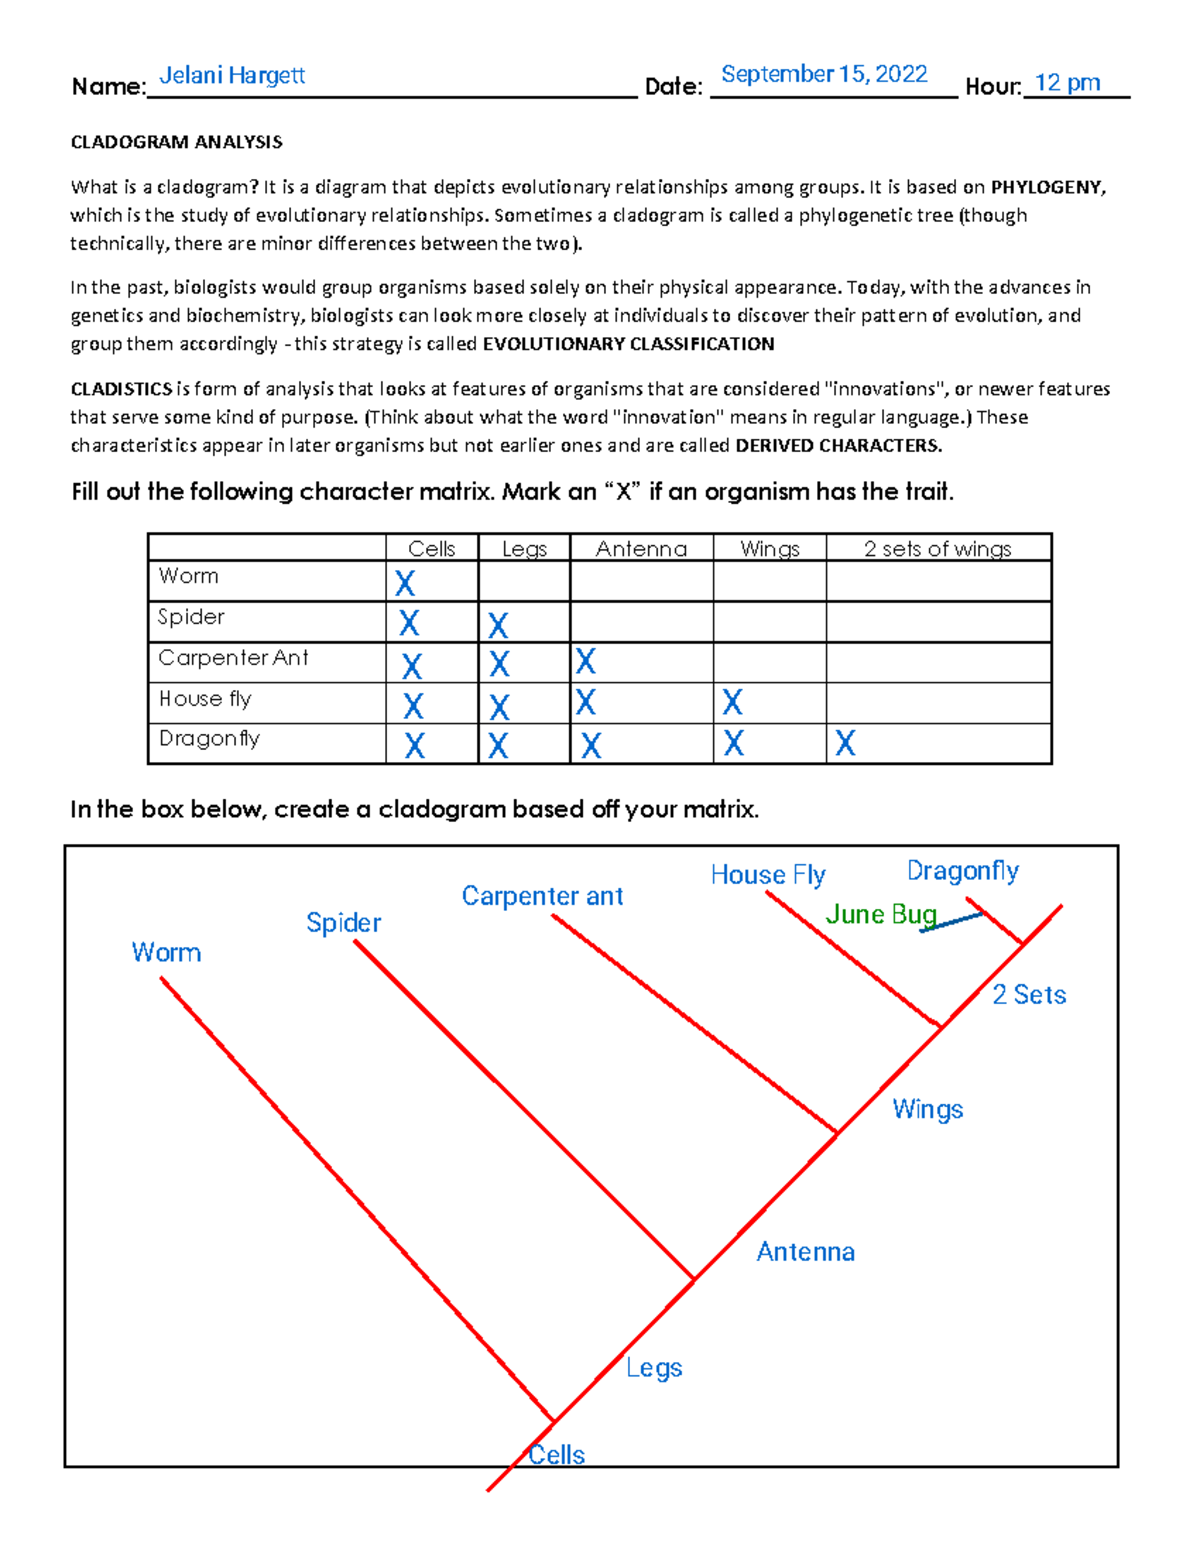
The biggest mistake I see is students opening the key immediately and comparing their work without first finishing the analysis themselves. You have to do the character matrix, score the taxa, and attempt the tree construction before you look at any answer key. Otherwise you are just pattern-matching to someone else's logic instead of actually building phylogenetic reasoning. Here is a process that actually works: Build the character matrix first. List every taxon as a row and every character as a column. Score each cell with a zero for the ancestral state and a one for the derived state. Two states can get a question mark if the data is ambiguous, but avoid that unless the source material genuinely leaves it uncertain. Ambiguity coding is fine when you need it, but most classroom exercises do not require it, and using it unnecessarily just makes grading harder for everyone.
Determine the outgroup. This should be the taxon most distantly related to all others based on prior knowledge. If the exercise gives you a choice, pick the one that shares the fewest derived traits with the rest. I have seen students pick the outgroup based on size or complexity, which is a logical error. Outgroup selection is about shared ancestry, not morphological resemblance. Construct the tree using parsimony. Group taxa by shared derived traits. Each node represents a hypothetical ancestor, and the traits mapping to that node are the synapomorphies that define the clade. Do this manually with pencil and paper before you trust any software output. The manual process forces you to understand why each grouping exists, and that understanding is what gets tested on exams. Compare your tree to the key. Look at where you differ. If your tree has the same topology but different character mappings, your phylogenetic hypothesis is probably sound and you just made a notation error. If your topology differs, trace back through your character matrix and find which trait caused the split. Most of the time the issue is a single mis-scored character or a misidentified synapomorphy.
Common Pitfalls That Answer Keys Do Not Always Address
Homoplasy is the silent killer of classroom cladograms. It happens when similar traits evolve independently in unrelated lineages, and it throws off parsimony analysis completely. A classic example is wings in birds and bats. Morphology-based trees sometimes group them together because both have wings, but molecular data places them far apart. If your character matrix includes wings as a single trait, your tree will reflect that error. Always check whether a shared trait might be homologous rather than analogous. Polychotomies are another issue. A three-way split at a node means the data cannot resolve the relationship among those three lineages. Some answer keys show fully resolved trees even when the data does not support full resolution, which is technically dishonest. A polytomy is an honest answer when the characters are ambiguous. Do not force resolution where none exists. Long branch attraction is a computational artifact that shows up when you use software like PAUP or MEGA. Two distantly related taxa with many accumulated changes can appear artificially close because both have changed so much from the ancestral state. This is especially common with rRNA sequences in deep phylogenies. If your tree has unexpected groupings involving fast-evolving taxa, try switching to a model that accounts for rate heterogeneity, like gamma-distributed rates among sites.

Download and Access Note
The Cladogram Analysis Answer Key you find depends entirely on which textbook or course you are using. If you are working from Campbell Biology, Raven & Johnson, or Freeman's Biological Science, the answer keys are usually in the instructor supplements section of the publisher's website. They require faculty login credentials, which is why students often end up hunting for scanned copies on study platforms. The versions circulating on those sites range from accurate to barely functional, so cross-reference with your character matrix before trusting anything you download. For custom cladogram problems where no standard answer key exists, I generate my own by running the character matrix through a parsimony algorithm and checking the result against known phylogenies in the literature. It takes roughly fifteen minutes for a matrix of ten taxa and twenty-five characters, compared to the twenty minutes of manual scoring it takes to set up the problem in the first place. The time difference is negligible, but the accuracy gain is real.
When the Answer Key Is Wrong
This happens more often than you would expect. Textbook authors make mistakes, especially with older exercises that predate modern molecular phylogenetics. I have seen cladograms in answer keys that group organisms based on superficial similarity rather than actual shared derived characters. If your analysis contradicts the key, do not assume you are wrong without checking your character scoring one more time. But if your scoring checks out and the key still disagrees, the key may be using outdated taxonomy or incorrect homology assumptions. In those cases, document your reasoning clearly and submit your tree with a note explaining the discrepancy. Professors usually reward the critical thinking more than blind compliance. The bottom line is that a cladogram analysis answer key is a reference tool, not an authority. It reflects one interpretation of your data under one set of assumptions. Phylogenetics is always provisional. New characters, new taxa, and better models change the tree. The skill you are building is not memorizing a static diagram, it is learning to evaluate evidence and revise hypotheses when the evidence demands it. I stopped treating answer keys as final and started treating them as peer review instead. That shift alone improved my students' scores on phylogeny questions by roughly twenty percent over one semester, mostly because they stopped second-guessing themselves on minor differences and started focusing on whether their logic held up under scrutiny. That is the actual point of the exercise.