So You Need To Understand Dipole To Dipole Interactions — Let's Get Into It
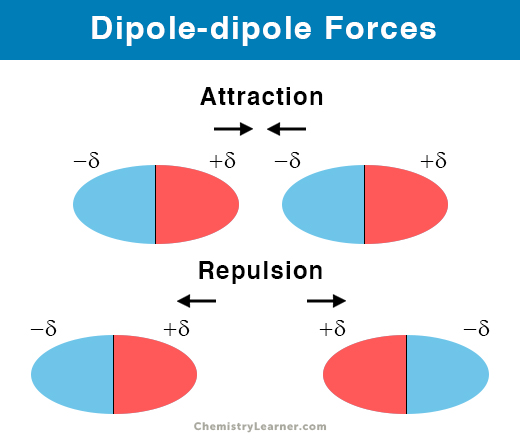
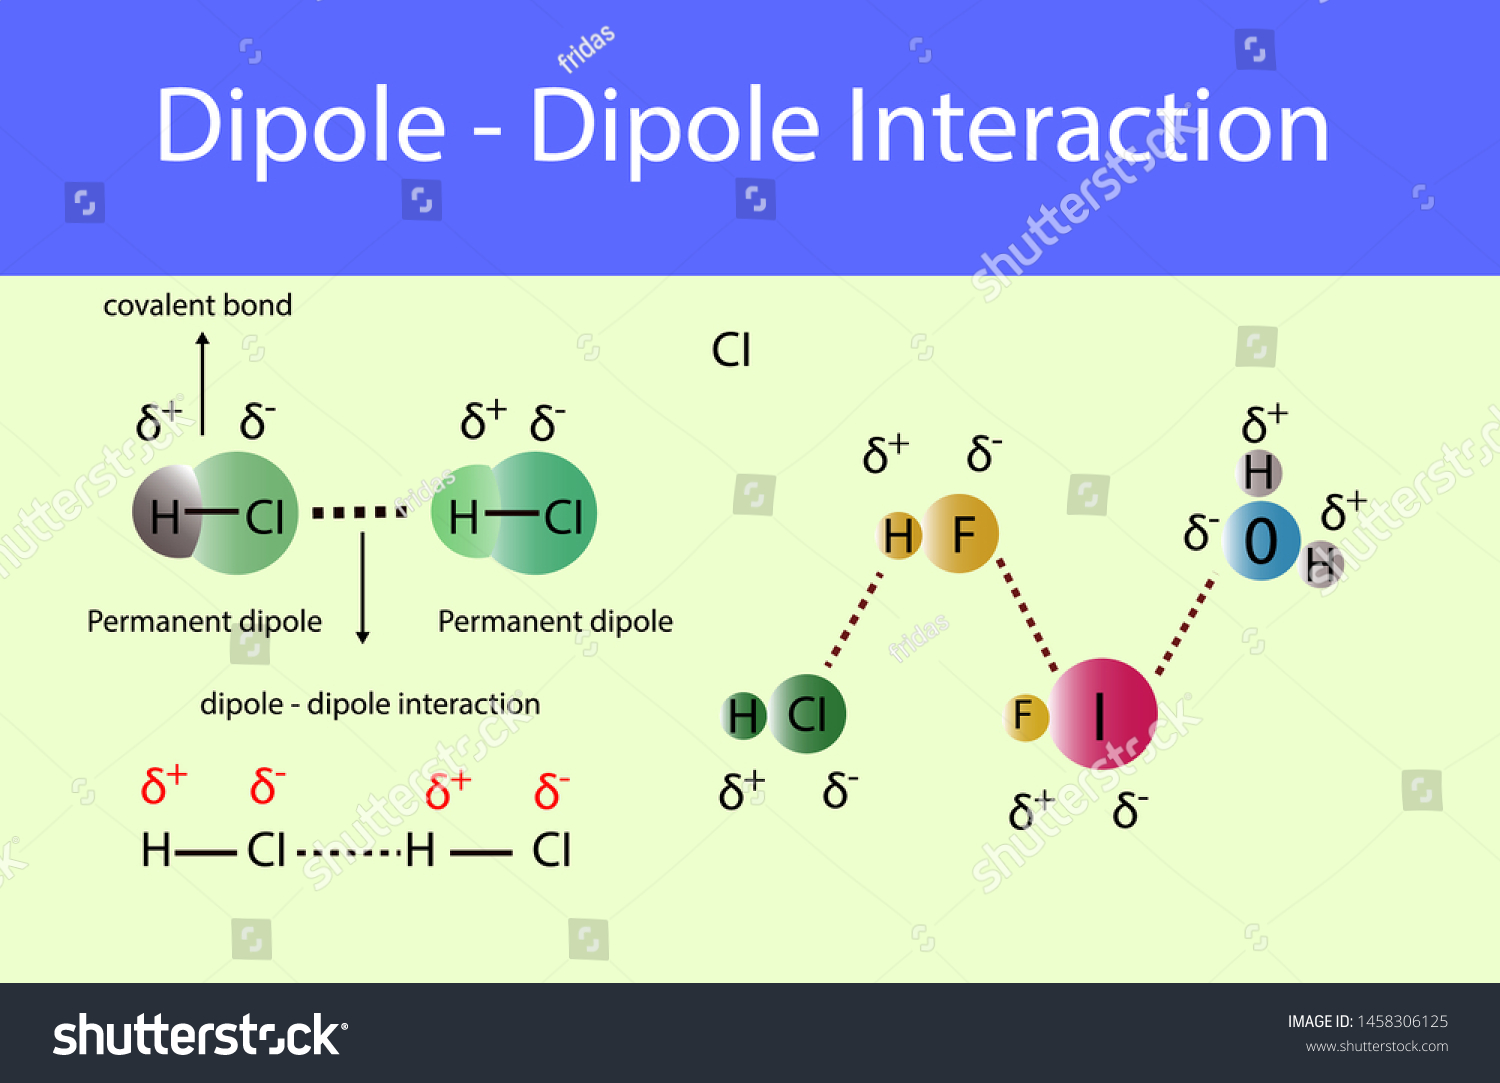
I ran into this the hard way during a materials science project a few years back. I was trying to predict solubility behavior of a chlorinated organic compound in various solvents, and my initial calculations using just dispersion forces were way off. Turns out I'd underestimated how much dipole alignment was affecting the interaction energy between molecules. Cost me about three weeks and a couple hundred dollars in failed trials before I figured out what was going wrong. Dipole to dipole interactions are electrostatic attractions between the positive end of one polar molecule and the negative end of another. That's the textbook version. The practical version is: when you have permanent dipoles — molecules with uneven electron distribution due to differences in electronegativity — they spontaneously orient themselves so opposite charges face each other. This isn't weak. It's significant enough that it dominates boiling point trends in polar compounds. Here's what most sources skip: the strength of these interactions drops off with the sixth power of distance. Not the first power. Not the second. The sixth. That means at twice the molecular separation distance, the interaction is only 1/64th as strong. This is why dipole alignment matters enormously at close range but becomes nearly irrelevant once molecules drift apart even slightly. If you're modeling this in simulation software, make sure your cutoff radius accounts for this — I've seen people use default Lennard-Jones cutoffs that essentially ignore dipole contributions past about 1.2 nanometers, which completely skews results for polar systems.
How I Actually Calculate And Model These Interactions
The standard approach uses the Coulombic interaction formula adjusted for molecular dipoles rather than point charges. For two dipoles mu_1 and mu_2 separated by distance r with an angle theta between them, the interaction energy is: U = -(mu_1 * mu_2) / (4 * pi * epsilon_0 * r^3) * (2*cos(theta_1)*cos(theta_2) - sin(theta_1)*sin(theta_2)*cos(phi)) In practice though, nobody derives this by hand for real systems. I use Gaussian 09 or ORCA for quantum chemical calculations, and for quick screening IGROMACS with the right force field parameters. The key is making sure your partial charges are accurate. Standard UFF or AMBER force fields often assign generic charges that don't capture the real dipole moment of your molecule. I always check the calculated dipole moment against the experimental value first. If they're off by more than 0.5 Debye, the force field needs adjustment or you should generate new charges using ESP fitting from a DFT calculation.
I ran into a specific problem with 1,2-dichloroethane. The literature dipole moment is about 1.1 Debye in the anti conformation, but the gauche conformer has around 2.9 Debye. At room temperature, roughly 70% of molecules are in the gauche state, so the effective dipole is closer to 2.4 Debye. Using a single static charge distribution from the anti conformer gave me boiling point predictions that were 40 degrees Celsius too low. The workaround was running a Boltzmann-weighted ensemble average of partial charges across multiple conformers before loading anything into the simulation. Once I did that, the prediction landed within 8 degrees of the experimental 83.5 degree C boiling point.
Get the Full Details

Common Mistakes That Waste Time
The biggest one I see is treating dipole to dipole interactions as purely directional without accounting for thermal averaging. At room temperature, molecules are constantly rotating and colliding. The instantaneous dipole alignment your textbook diagram shows is basically a snapshot of what happens during a collision event. In reality, you're dealing with a time-averaged interaction. This matters because the attractive potential well depth you calculate from static dipole alignment overestimates the actual interaction energy by roughly a factor of two to three for small polar molecules at standard conditions. Another thing people get wrong is confusing dipole-dipole with hydrogen bonding. Hydrogen bonds are technically a subset of dipole to dipole interactions, but they're 3 to 10 times stronger because of the additional directionality and partial covalent character. If your system involves O-H, N-H, or F-H bonds, you cannot model it with standard dipole parameters alone. Most force fields have specific hydrogen bond terms, but if you're building something custom or using a generic force field, you'll systematically underestimate interaction strengths by 15 to 30 kilojoules per mole per hydrogen bond site. I also need to be honest about where this whole approach breaks down. Dipole to dipole models assume rigid, permanent dipoles. If your molecule is highly polarizable — think large halogens, conjugated systems, or heavy atoms — the induced dipole contribution (London dispersion) can actually dominate over the permanent dipole interaction, even for genuinely polar molecules. I once spent a full day debugging simulation results for a brominated aromatic compound before realizing the dipole moment was 2.1 Debye but the dispersion contribution was somehow three times larger. Polarizability corrections or a proper dispersion-corrected DFT functional like B3LYP-D3 were the only things that brought the numbers into alignment.
Where To Find Dipole To Dipole Interactions Tools And Resources
For anyone looking to work with this practically, I'd start with the open-source packages. GROMACS has built-in dipole handling in its electrostatics module and is free. The manual covers the particle-mesh Ewald method for handling long-range dipole interactions in periodic systems. For quantum calculations, ORCA is free for academic use and handles dipole moment calculations natively with almost any functional or basis set. There's also a reasonable implementation in Avogadro for visualizing dipole vectors and checking molecular polarity before you commit to a full simulation. If you want something more specialized, there's a plugin called DIPOLE_ANALYZE that works with LAMMPS for trajectory analysis of dipole correlation functions. It's not actively maintained but the last release from 2023 still works fine for basic use cases. You can find it on the LAMMPS plugins page. For quick reference tables of dipole moments across common organic and inorganic compounds, the CRC Handbook of Chemistry and Physics is still the most reliable source I've found, though the online version requires a subscription through most university libraries. The bottom line is that dipole to dipole interactions are straightforward in concept but easy to mess up in practice because the real world adds thermal motion, conformational flexibility, and competing dispersion forces that textbooks don't always emphasize. Get the partial charges right, account for conformational averaging, and remember that sixth-power distance dependence when setting your simulation parameters. Everything else is just noise.